Related Experiment Video
Updated: Oct 28, 2026
A Kinetic Fluorescence-based Ca2+ Mobilization Assay to Identify G Protein-coupled Receptor Agonists, Antagonists, and Allosteric Modulators
Published on: February 20, 2018
Diphenylpyraline Derivatives Are Allosteric Agonists of the CB1 Cannabinoid Receptor
Khalil Eldeeb1,2,3, Jill K Harp2,4,5,6, Ryan W Fitzgerald4
1Jerry M. Wallace School of Osteopathic Medicine, Campbell University, Lillington, NC 27506, USA.

Abstract:
Diphenylpyraline analogs JKH1 and JKH10 are monoamine transporter inhibitors and promote relaxation in mouse mesenteric arteries. Here, we evaluated JKH1 and JKH for their interaction with CB1 cannabinoid receptor signaling. In N18TG2 mouse neuroblastoma cells that endogenously express CB1 receptors, JKH1 and JKH10 stimulated [35S]GTPγS binding in a concentration-dependent manner; this stimulation was non-competitively antagonized by the CB1 antagonist-inverse agonist SR141716 (1 µM). JKH10 (but not JKH1) inhibited forskolin-stimulated cAMP accumulation with efficacy as high as the full orthosteric agonist WIN55212-2, and this Gi-mediated inhibition was completely precluded by pertussis toxin treatment. However, JKH1 was a poor inhibitor. The SR141716 failed to completely block the inhibition by JKH10. Both JKH analogs stimulated ERK phosphorylation, but inhibition by SR141716 (1 µM) did not appear to be overcome by increasing concentrations of JKH1 or JKH10. In mouse isolated mesenteric arteries, both analogs (1 µM) attenuated the contractile response, but only the response to JKH10 was blocked by the CB1 antagonist-inverse agonist O-2050 (1 µM). Docking studies to protein data bank CB1 structures 7FEE and 6KQI showed JKH1 was predicted to dock predominantly at a pregnenolone-binding site, whereas JKH10 was predicted to dock at both the pregnenolone and a cholesterol consensus motif at helices 2-3-4 inner membrane leaflet. Thus, a series of diphenylpyraline derivatives can behave as CB1 receptor stimulators, and in silico evidence supports that this could result from interaction with cholesterol and/or pregnenolone protein surface binding sites rather than the orthosteric binding site.
Related Concept Videos
 01:22
Opioid Receptors: Overview
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:25
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:17
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
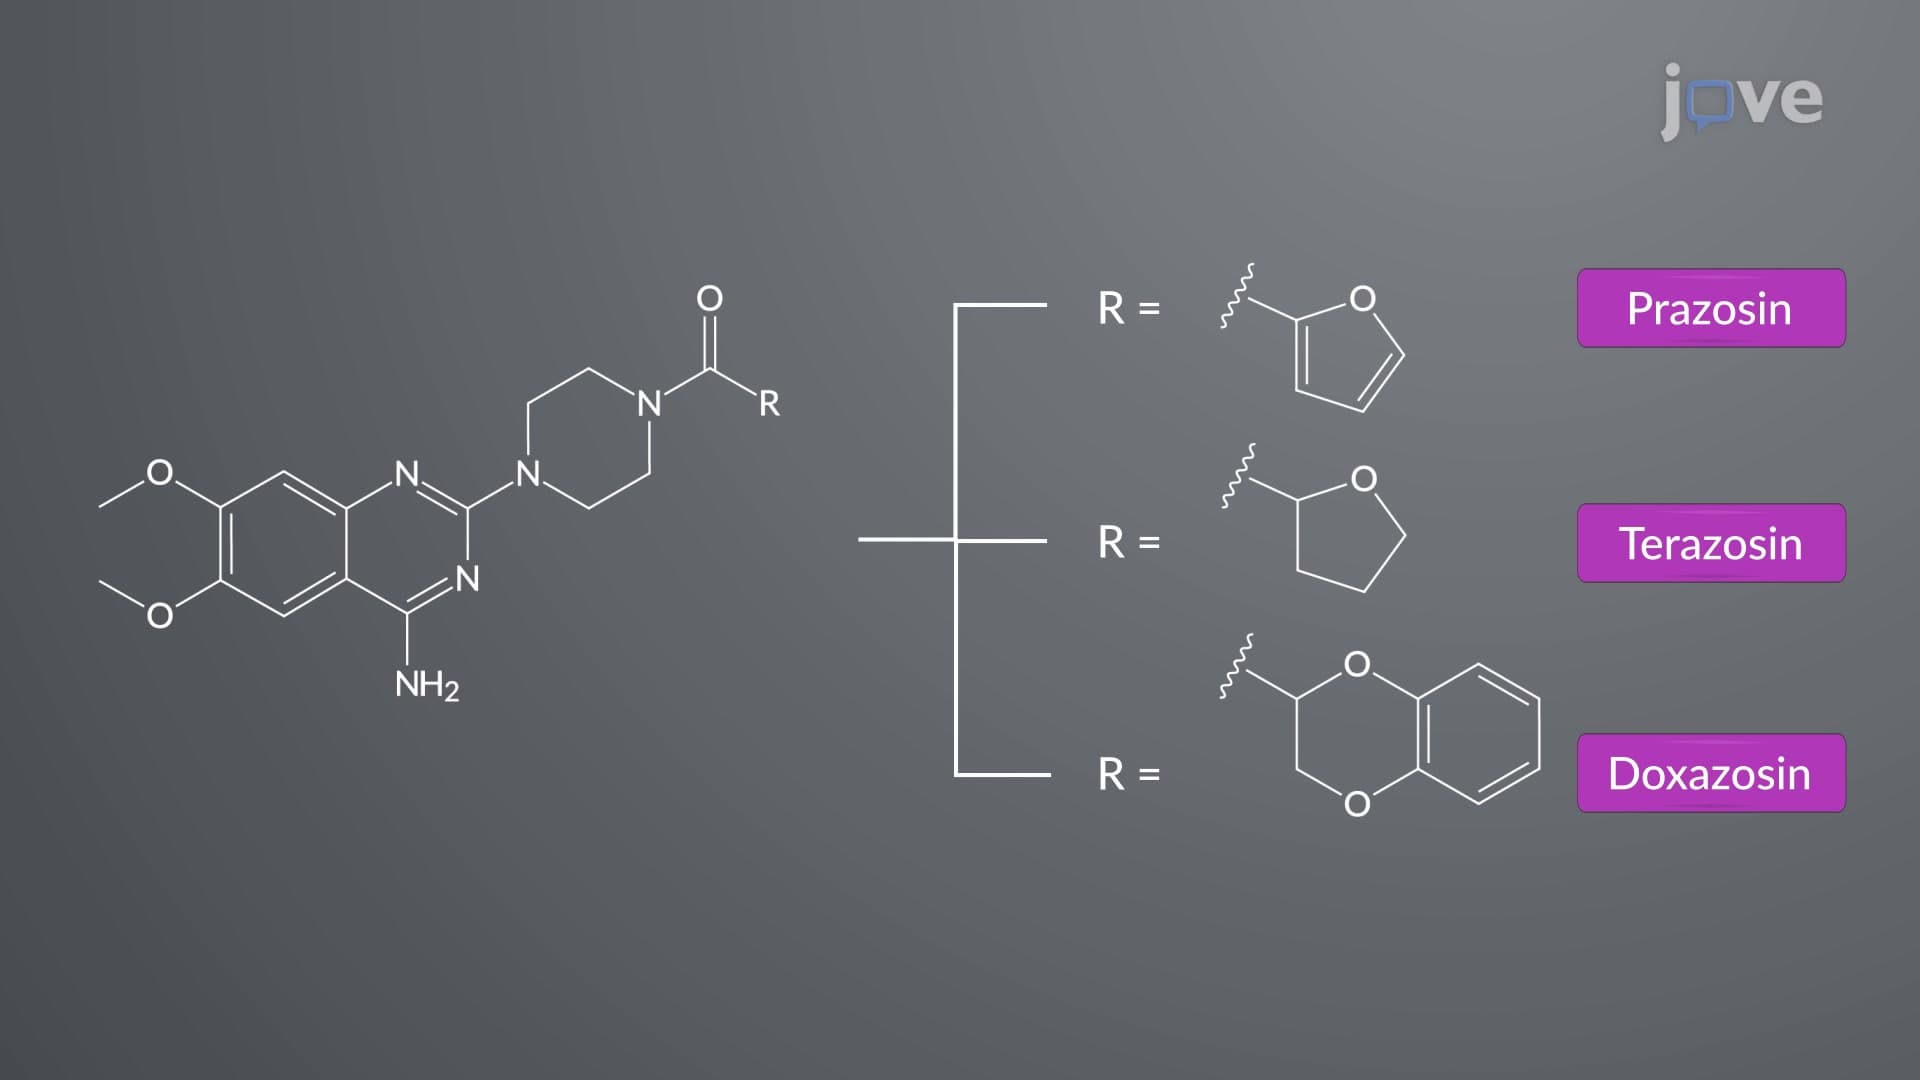
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:25
Adrenergic Agonists: Indirect-Acting Agents
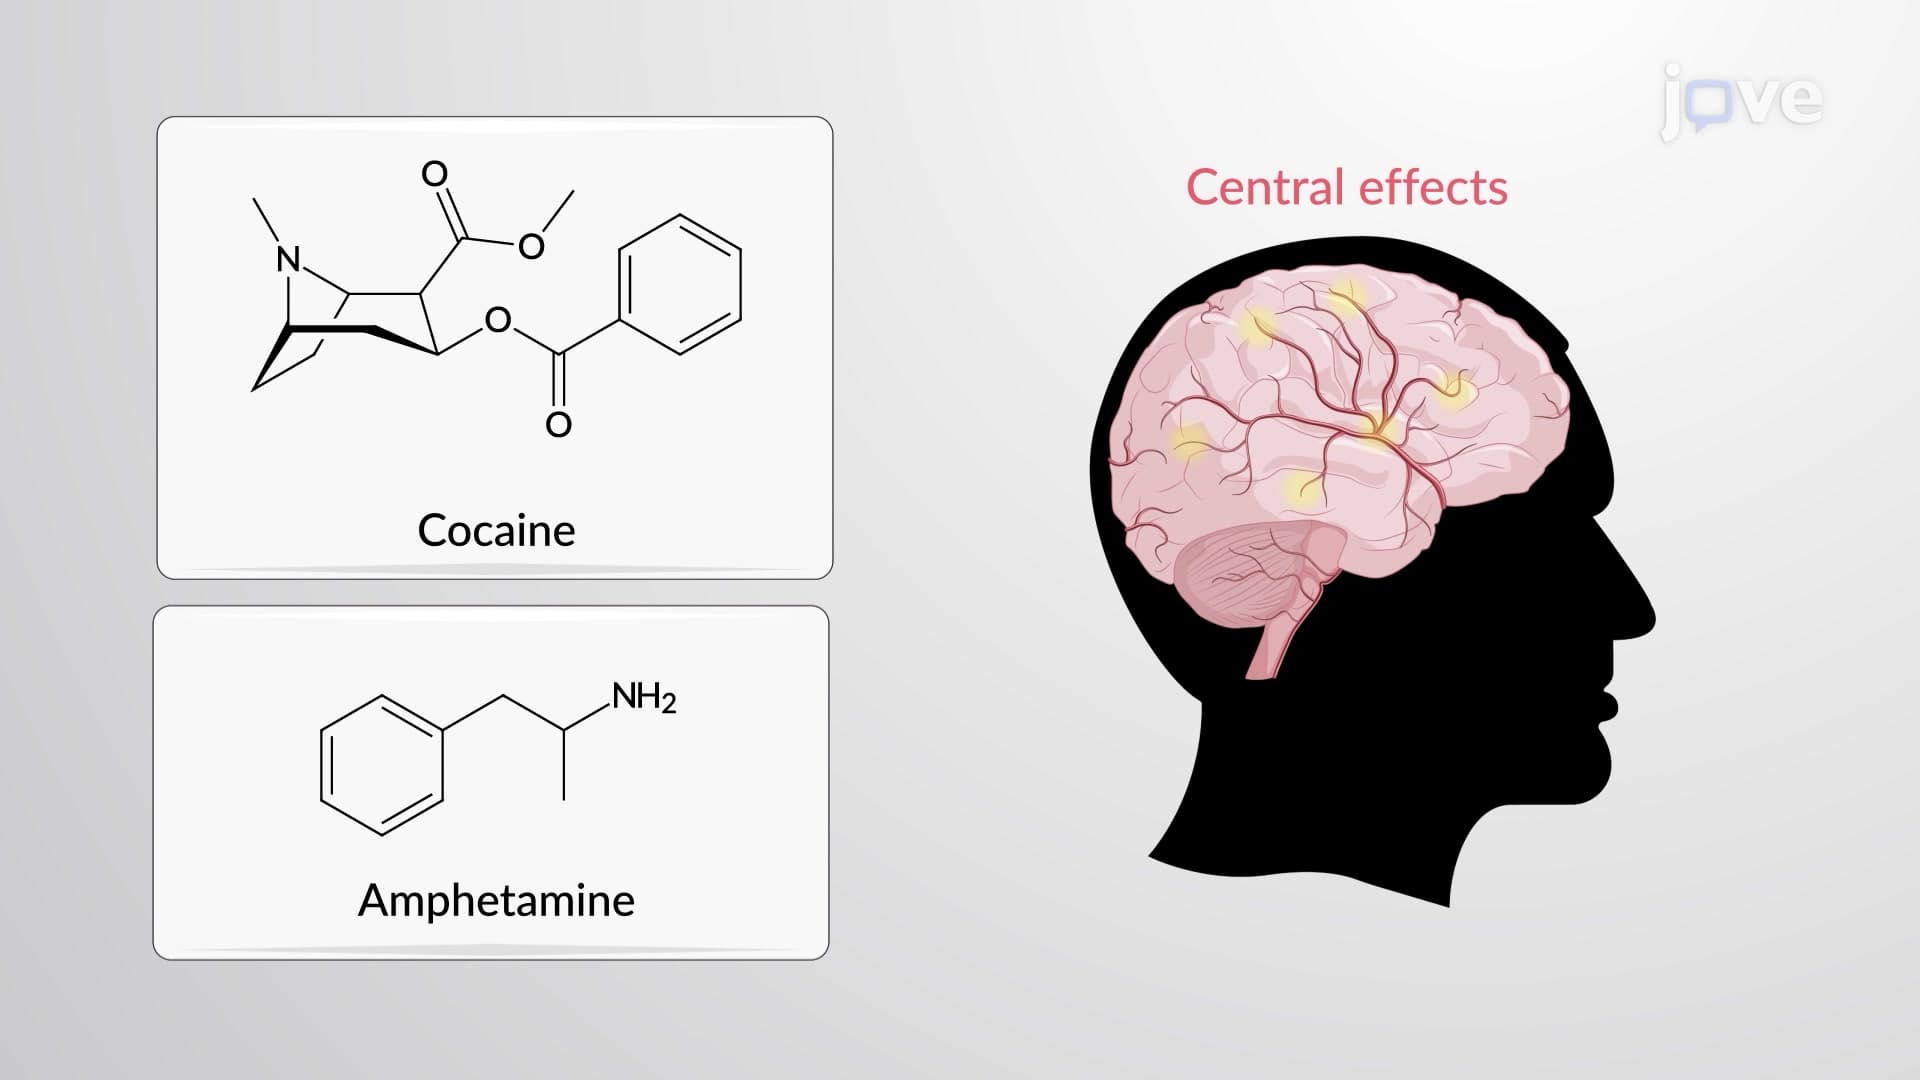
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...