Related Experiment Video
Updated: Oct 29, 2026
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Generative design and computational characterization of pyrazoline-based EGFR kinase inhibitor candidates
Kumud Bhendarkar1, Pramod Khedekar2, Md Ataul Islam3
1Department of Pharmaceutical Sciences, Rashtrasant Tukadoji Maharaj Nagpur University, Nagpur, Maharashtra, India.

Abstract:
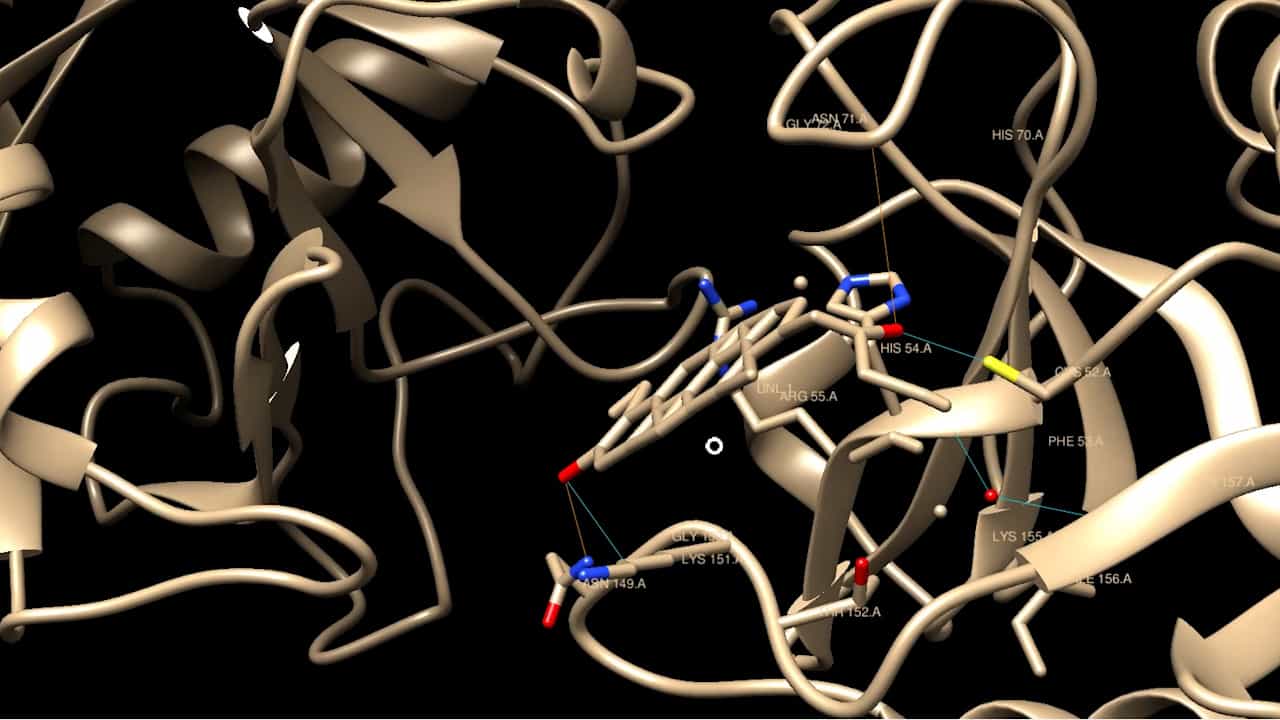
The epidermal growth factor receptor (EGFR) is an established therapeutic target in breast, colorectal, and non-small cell lung cancers, and pyrazoline-containing scaffolds have been repeatedly reported as EGFR inhibitors. This study combined generative machine learning with physics-based modelling to design and computationally characterize novel pyrazoline analogues targeting the EGFR kinase ATP-binding site. A curated set of 50 experimentally validated pyrazoline-based EGFR inhibitors, compiled from ChEMBL and the primary literature, was used to seed the Mol2Mol model in REINVENT4, generating a large library of structurally related analogues. The library was sequentially filtered according to Tanimoto similarity to the seed actives (≥0.6), predicted physicochemical, pharmacokinetic, and toxicity properties, synthetic accessibility (SA score <6), and molecular docking against the EGFR kinase domain (PDB 5U8L), using a self-docking validated protocol (redocking RMSD 1.342 Å). Five candidates, designated PyEG1-PyEG5, achieved docking scores between -9.5 and -10.6 kcal/mol, better than the redocked native ligand (-9.2 kcal/mol). All five complexes remained stable during 100 ns molecular dynamics simulations, with backbone RMSD ≤ 0.27 nm, average ligand RMSD ≤ 0.18 nm, and radii of gyration close to those of the reference complex. MM-GBSA end-point calculations yielded binding free energies of -9.9 to -37.5 kcal/mol, with PyEG5 (-37.5 ± 2.5 kcal/mol) approaching the reference ligand value (-37.6 ± 6.0 kcal/mol). Density functional theory calculations (B3LYP/6-31G(d)) indicated HOMO-LUMO gaps of 5.89-6.65 eV, with PyEG2 and PyEG5 predicted to be the most reactive and PyEG3 the most kinetically stable. Structure-activity analysis based on the predicted binding modes suggests that hinge-directed hydrogen bonding to Met793 and π-π stacking with Phe856 are the principal determinants of predicted affinity, modulated by substituent electronics and steric bulk at the R, R1, and R2 positions. These findings are entirely computational; PyEG1-PyEG5 should be considered prioritized synthetic targets, and their activity requires confirmation through biochemical kinase assays and cell-based testing.