Protein structure-function relationships as the molecular basis of phenotypic diversity in GDAP1-associated Charcot-Marie-Tooth disease
Related Experiment Video
Updated: Oct 31, 2026
Structural Biology and Analytical Chemistry Approaches for Characterizing C-Glycoside Metabolic Enzymes in Human Gut Microbiota
Published on: May 23, 2025
Victoria I Bunik1,2, Alina A Emelyanova2
1Belozersky Institute of Physicochemical Biology, Lomonosov Moscow State University, Moscow, Russia.

Abstract:
Mutations in the GDAP1 gene, which encodes the ganglioside-induced differentiation-associated protein 1 (GDAP1)-a member of the detoxifying glutathione S-transferase (GST) superfamily-cause Charcot---Marie-Tooth disease (CMT) a genetically heterogeneous group of inherited neuropathies. This mini review correlates the clinical phenotypes of CMT with the molecular dysfunctions of GDAP1. Implicated in mitochondrial dynamics, calcium homeostasis, cellular organelle contacts, and the metabolism of mitochondria, lysosomes, and peroxisomes, the known molecular functions of GDAP1 include localization to mitochondrial outer membrane, dimerization, GST activity, and binding of hydrophobic compounds and proteins. Our analysis reveals that mutation-induced perturbations in specific molecular functions correlate with clinical phenotypes better than mutation class (missense vs. nonsense or frameshift), mode of inheritance (dominant vs. recessive), or mitochondrial morphology. The G327D and truncated GDAP1 variants have similar clinical phenotypes because all lack membrane binding. Early disease onset and rapid progression are observed not only with recessive variants (P111, N227, L239, and R282) but also with dominant variants (C240), if the substituted residues are proximal to the hydrophobic ligand. While dominant GDAP1 variants cause axonal CMT, all the recessive variants inferred to affect the hydrophobic ligand binding present with demyelination in addition to axonal damage. These different disease trajectories may rely on the potential formation of fully functional wild-type GDAP1 homodimers in dominant heterozygotes but not in biallelic recessive cases. Thus, elucidating how disease-associated protein variants affect GDAP1 molecular functions advances our understanding of the molecular basis of clinical phenotypes, which is required for tailored therapies.
Related Concept Videos
 01:25
Structure of Cadherins
 02:39
Protein and Protein Structure
 02:18
Intrinsically Disordered Proteins
 01:24
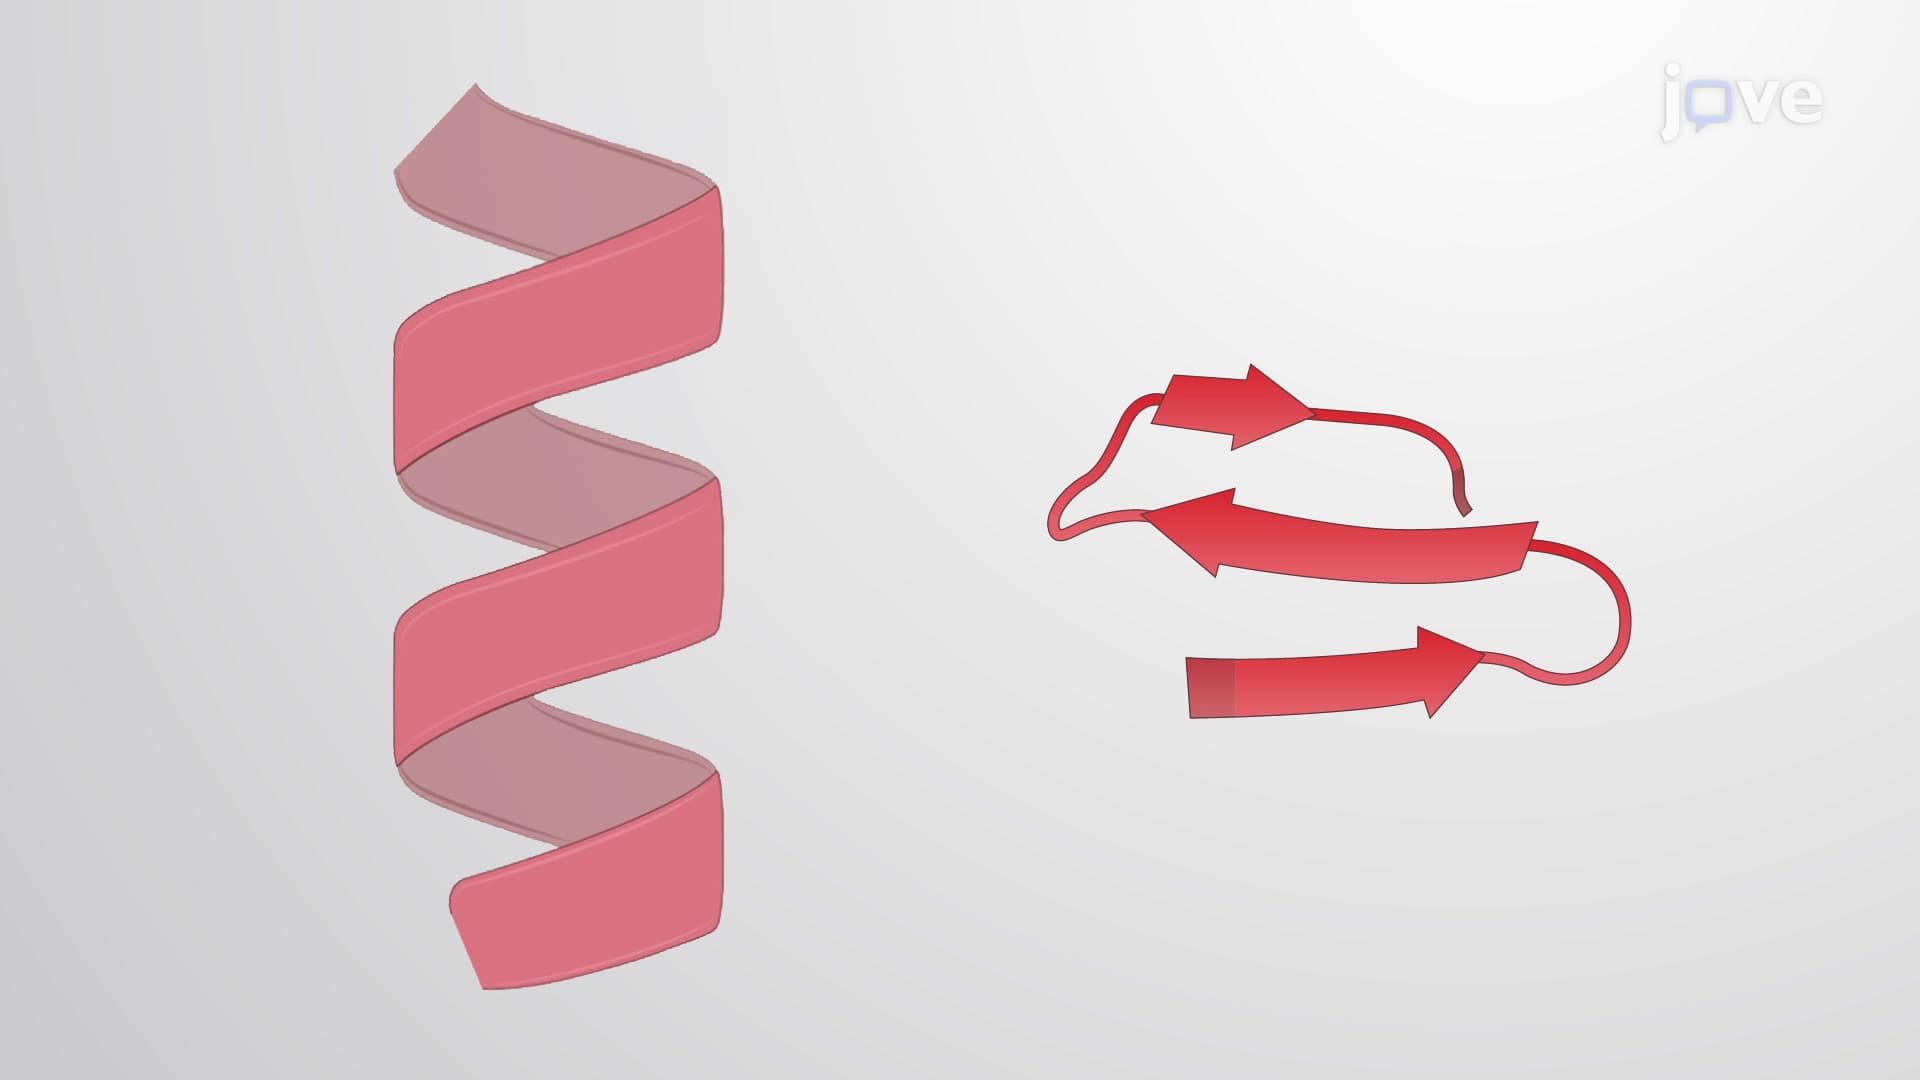
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
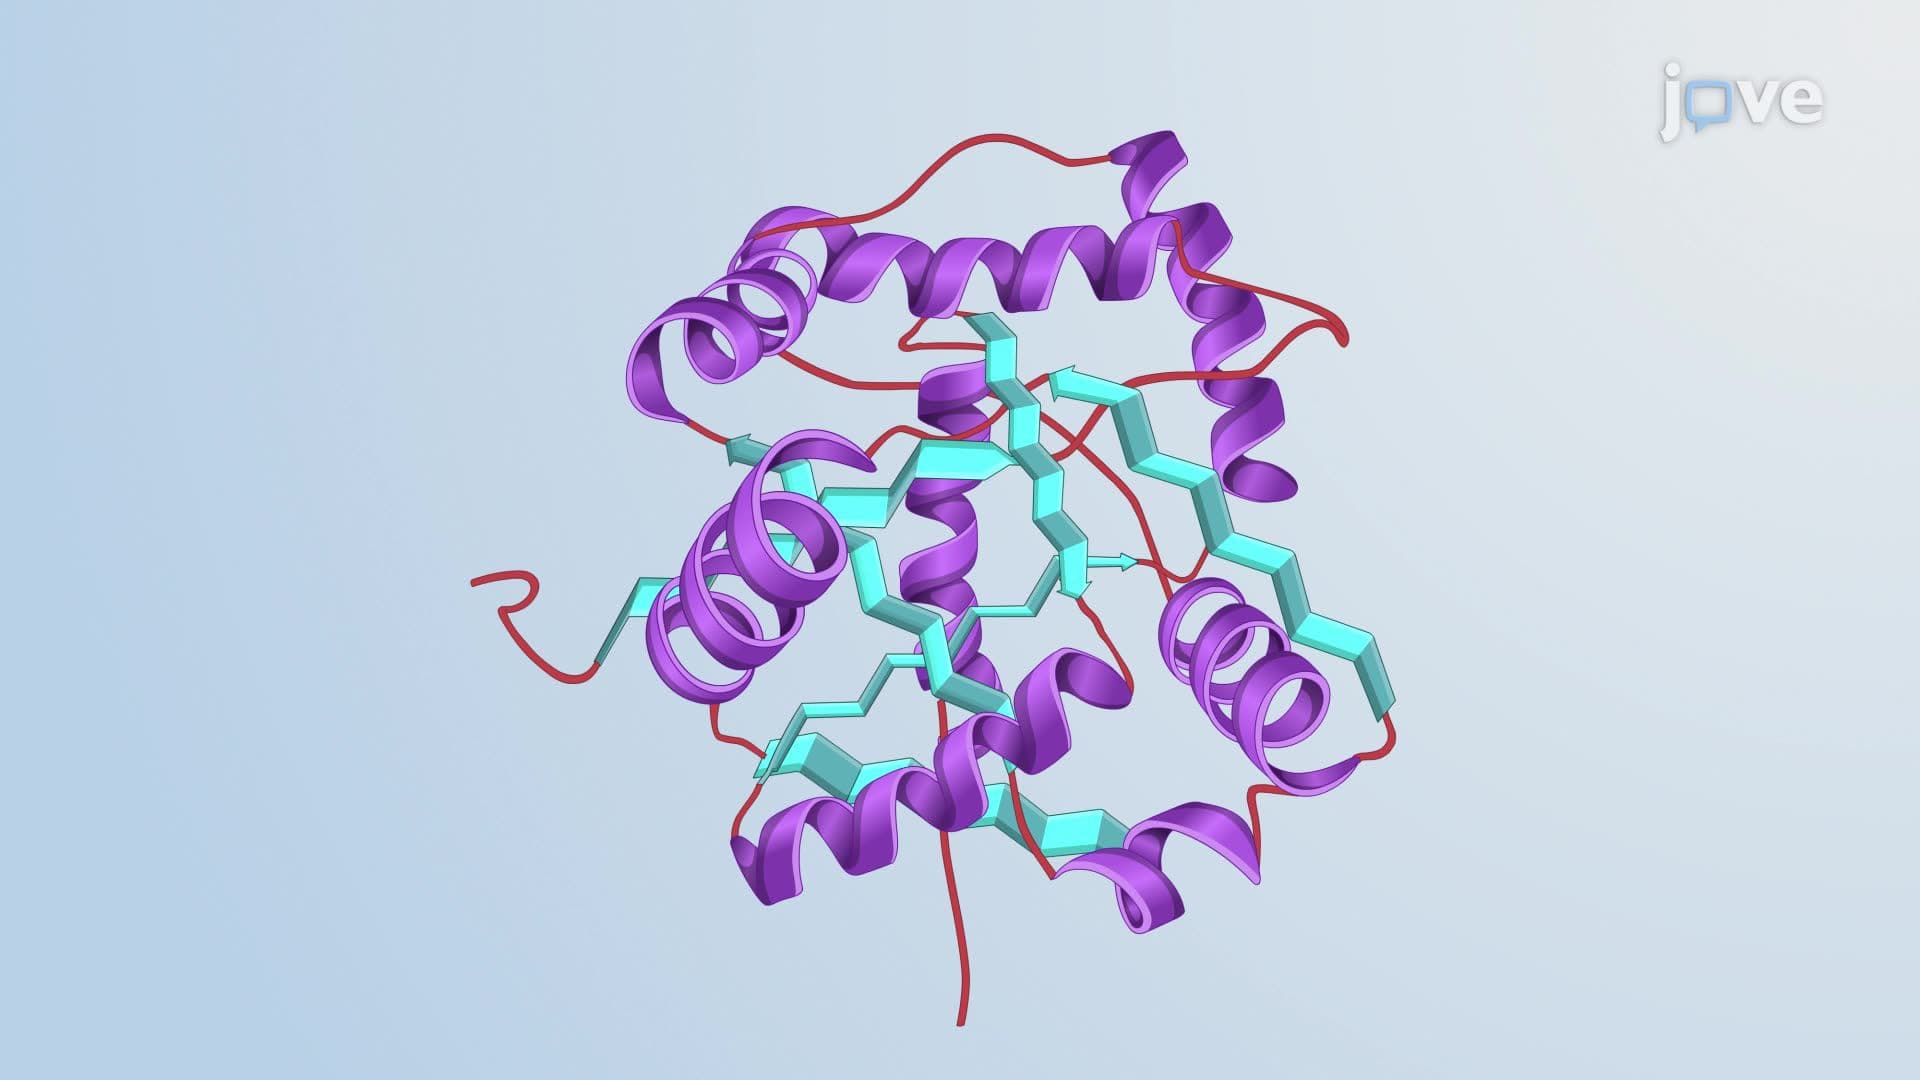 01:49
Protein Folding