An internal -Pro-ΔAA- motif enables catalyst-controlled reduction of dehydropeptides with H2
Related Experiment Video
Updated: Oct 31, 2026
Defining Hsp33's Redox-regulated Chaperone Activity and Mapping Conformational Changes on Hsp33 Using Hydrogen-deuterium Exchange Mass Spectrometry
Published on: June 7, 2018
Pei-Hsuan Chen1, Guangyu Zhang1, Cac Vinh Nguyen1
1Department of Medicinal Chemistry, University of Kansas, Lawrence, KS 66045, USA.

Abstract:
Catalyst-controlled asymmetric hydrogenation of dehydroamino acids (ΔAA) in peptides is a straightforward way to access both (L)- and (D)-diastereomers from a single progenitor peptide. While this approach works well for peptides terminated with ΔAAs, catalyst-controlled reduction of internal ΔAAs is more challenging. They are flanked by neighboring amino acids, greatly reducing their accessibility to metal-based hydrogenation catalysts. Their reactions are also heavily influenced by the innate chirality and shape of the peptide, pitting the stereo-inducing effect of the chiral ligand against the unyielding preference of the peptide. For these reasons, finding a single pair of catalysts that can reliably produce either (L)- or (D)-products by simply changing the ligands stereochemistry is a daunting endeavour. Herein, we show that positioning the ΔAA next to a proline residue enables catalyst-controlled hydrogenation with Rh(*DuanPhos). The origin of this control stems from the unique ability of the -Pro-ΔAA- motif to induce a β turn-like structure that orients the ΔAA away from the peptide, such that catalyst approach can occur from either side of the enamide, unimpeded. This effect can be extended to dehydropeptides with a variety of ΔAA chemotypes, e.g., aliphatic, aromatic, and heteroaromatic. Removing the proline residue diminishes the effect, eroding diastereoselectivity.
Related Concept Videos
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 01:20
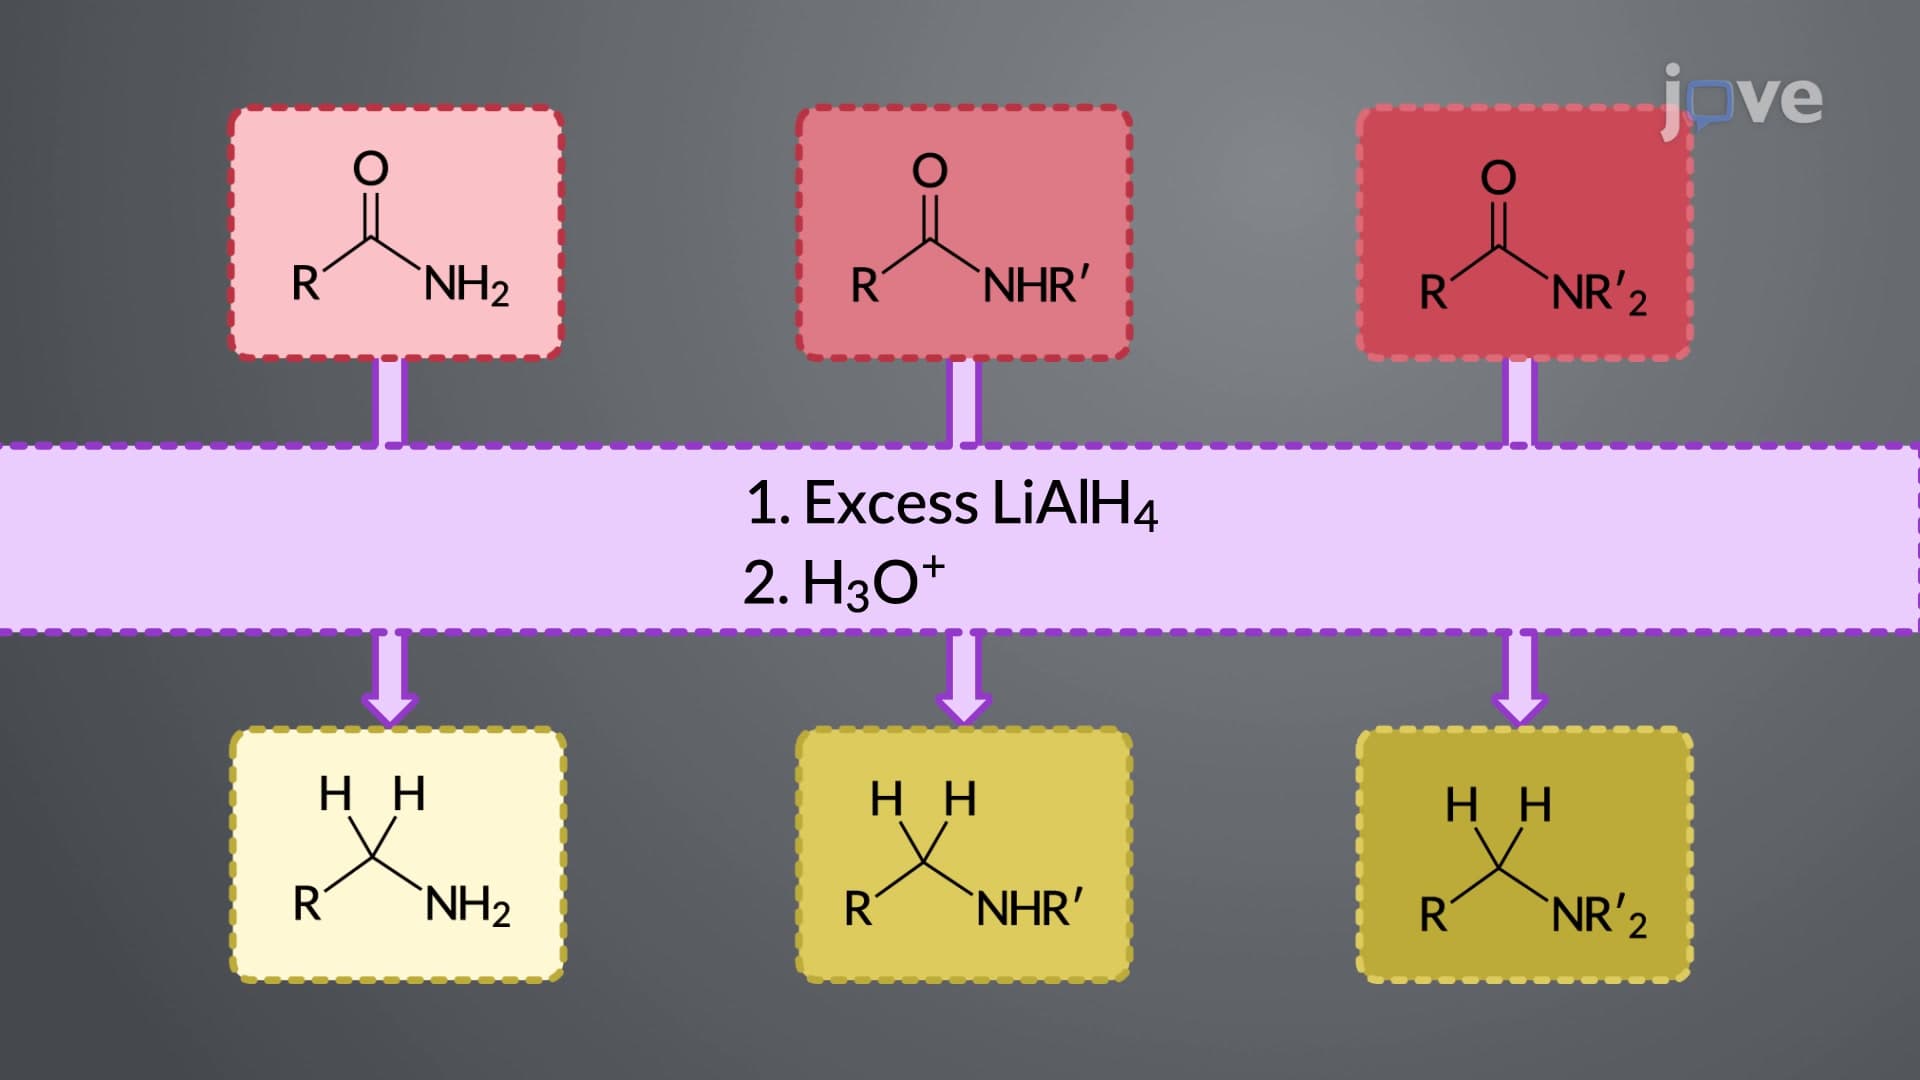
Amides to Amines: LiAlH4 Reduction
Amide reduction requires two equivalents of the reducing agent, acting as a source of hydride ions. As shown in the figure, the reaction is initiated with a nucleophilic attack by the hydride ion at the carbonyl carbon to form a tetrahedral intermediate.
 02:23
Alcohols from Carbonyl Compounds: Reduction
Catalytic hydrogenation is similar to the reduction of an alkene or alkyne by adding H2 across the pi bond in the presence of transition metal catalysts like Raney Ni, Pd–C, Pt, or Ru. Aldehydes and ketones can be reduced by this method, often under mild to moderate heat (25–100°C) and...
 00:55
Nitriles to Amines: LiAlH4 Reduction
As shown below, the mechanism involves three steps. Firstly, the hydride ion acting as a nucleophile attacks the nitrile carbon to form an anion. In the second step, a second equivalent of the hydride ion attacks the anion to...
 01:38
Preparation of Amines: Reductive Amination of Aldehydes and Ketones