Structural evolution of an ancient nucleotidyltransferase underlies divergence of AbiE antitoxins
Related Experiment Video
Updated: Nov 1, 2026
Creating and Applying a Reference to Facilitate the Discussion and Classification of Proteins in a Diverse Group
Published on: August 16, 2017
Tianlang Liu1, Ran Chen2, Kaihao Tang2
1State Key Laboratory of Tropical Oceanography, South China Sea Institute of Oceanology, Chinese Academy of Sciences, No. 1119, Haibin Road, Nansha District, Guangzhou 511458, China.

Abstract:
Type VII toxin-antitoxin (TA) systems neutralize toxins through antitoxin-mediated post-translational modification (PTM), with the oligoAMPylation-based HepT/MntA and phosphorylation-based AbiE systems representing the most prevalent families. Although the AbiE TA system from Mycobacterium tuberculosis is known to employ antitoxin-dependent phosphorylation to antagonize its toxin, the overall distribution and regulatory diversity of AbiE-like systems remain obscure. A comprehensive search of the IMG/M database revealed that AbiE TA systems comprise a conserved toxin family paired with six distinct groups of antitoxins, including four newly identified antitoxin groups. Despite extensive sequence divergence, structural predictions indicate that most of these antitoxins share a conserved winged helix-turn-helix (wHTH) domain and a C-terminal structurally homologous domain (CTD). We further demonstrate a unifying antagonistic mechanism across these diverse systems: cognate antitoxins induce the phosphorylation of conserved serine or threonine residues in their partner toxins, resulting in toxin inactivation in a CTD-dependent manner. Structural analyses reveal that the antitoxin CTD exhibits homology to nucleotidyltransferases (NTases), including the AbiE toxin itself and the MntA antitoxins, suggesting that type VII antitoxins may have evolved from an ancestral NTase superfamily. Moreover, their DNA-binding capabilities are also diverse, with some functioning as transcriptional regulators and others lacking detectable DNA-binding activity. Collectively, this study expands the recognized diversity of type VII TA systems that rely on PTM for antagonism and provides new insights into their evolutionary trajectories.
More Related Videos
 16:02
Demonstration of the Sequence Alignment to Predict Across Species Susceptibility Tool for Rapid Assessment of Protein Conservation
Published on: February 10, 2023
 13:31
Novel Atomic Force Microscopy Based Biopanning for Isolation of Morphology Specific Reagents against TDP-43 Variants in Amyotrophic Lateral Sclerosis
Published on: February 12, 2015
Related Concept Videos
 02:32
Exon Recombination
Exon shuffling follows “splice frame rules.” Each exon has three reading...
 01:28
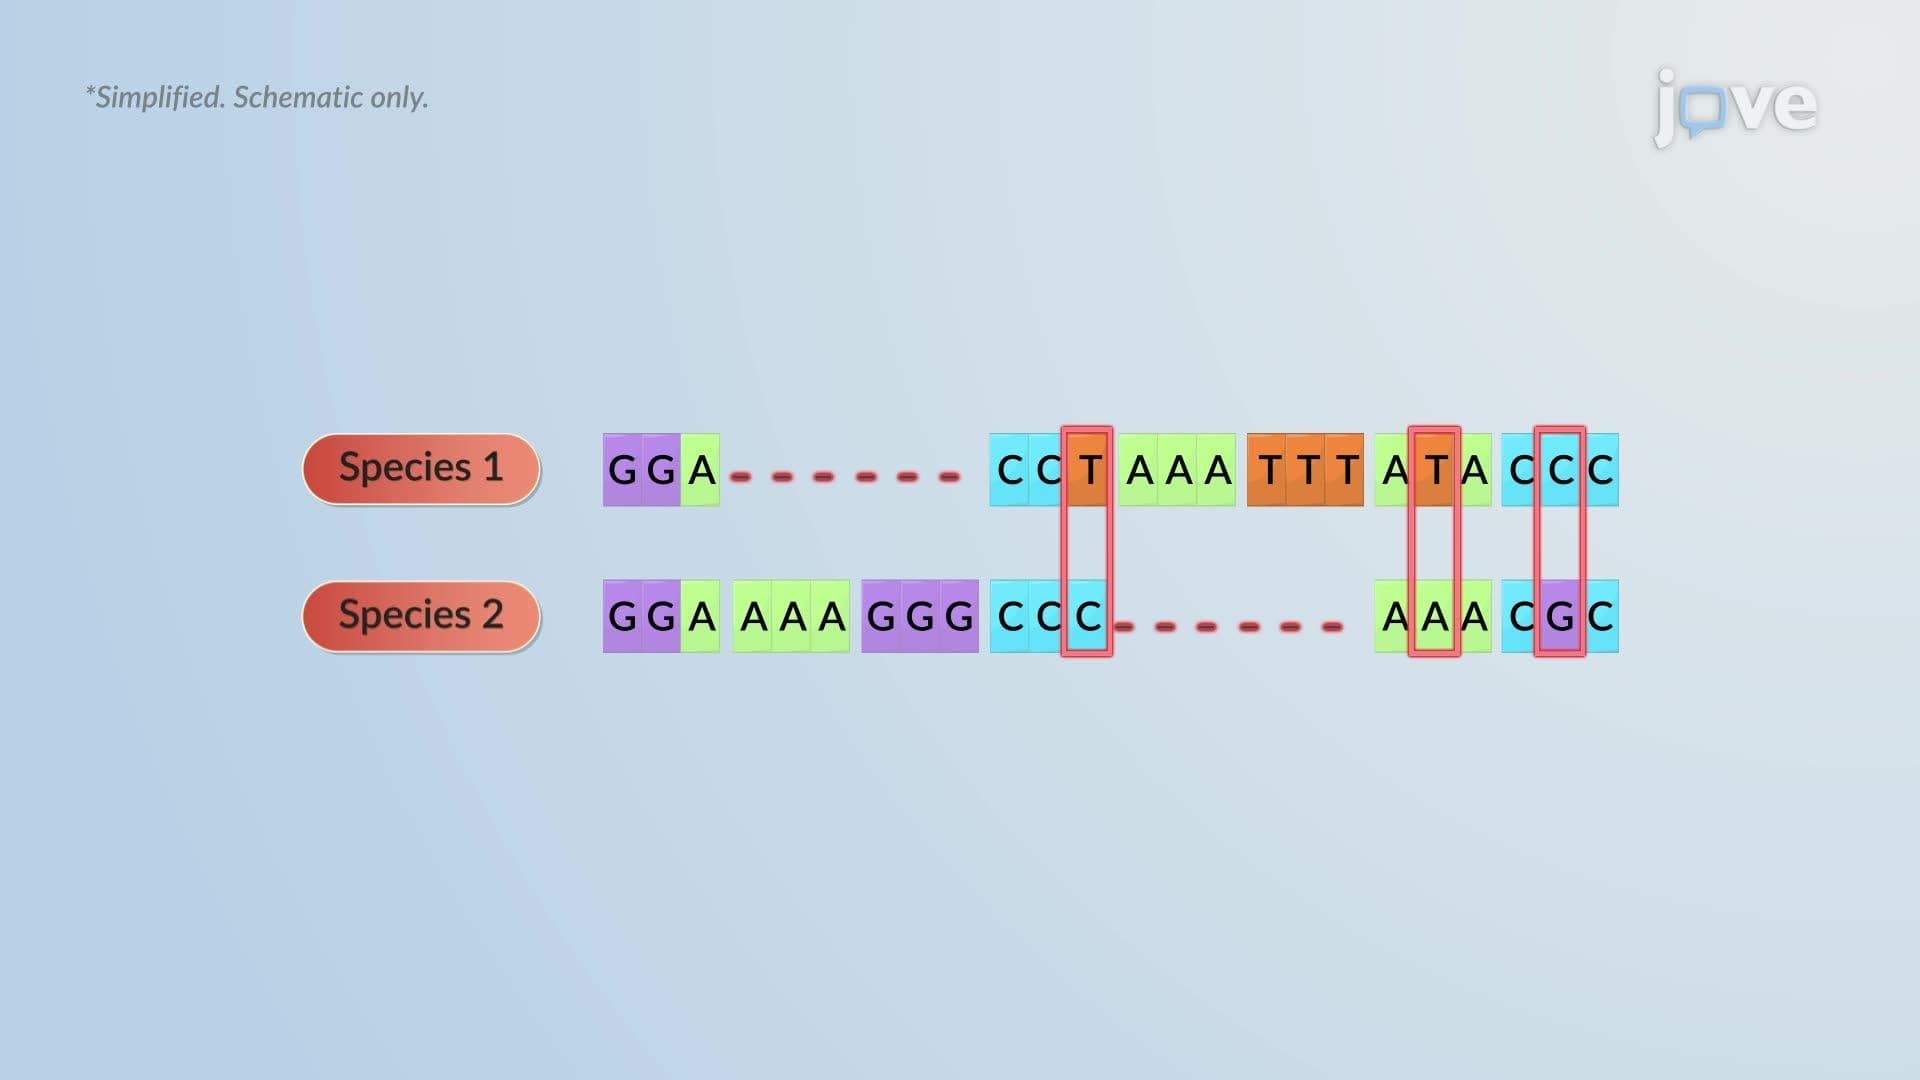
Microbial Phylogeny
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 03:21
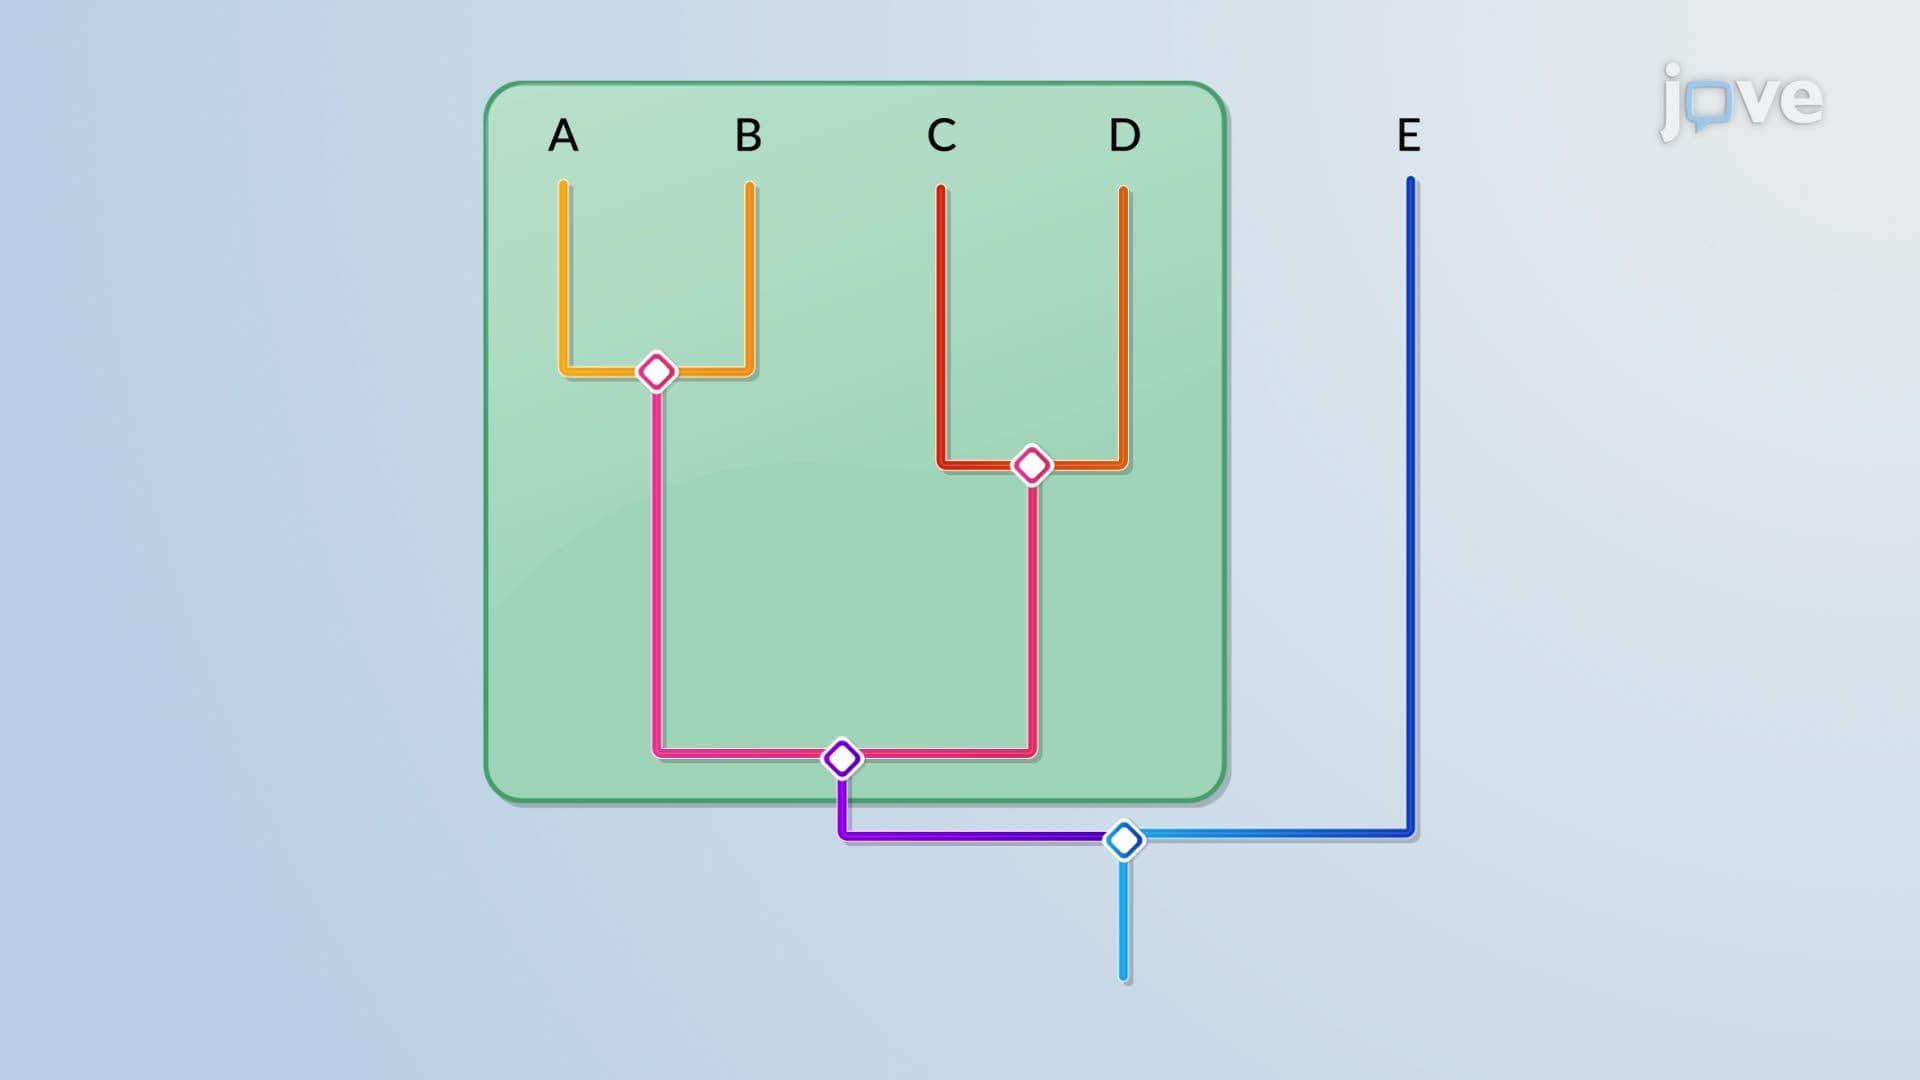
Phylogenetic Trees
 02:54
Evolutionary Relationships through Genome Comparisons
 02:31
Synteny and Evolution
Around 80 million years ago, the human and mice lineages diverged from the common ancestor. During the course of evolution, the ancestral chromosome underwent...