SpatioCAD: Context-aware graph diffusion model for pinpointing spatially variable genes in heterogeneous tissues
Related Experiment Video
Updated: Nov 4, 2026
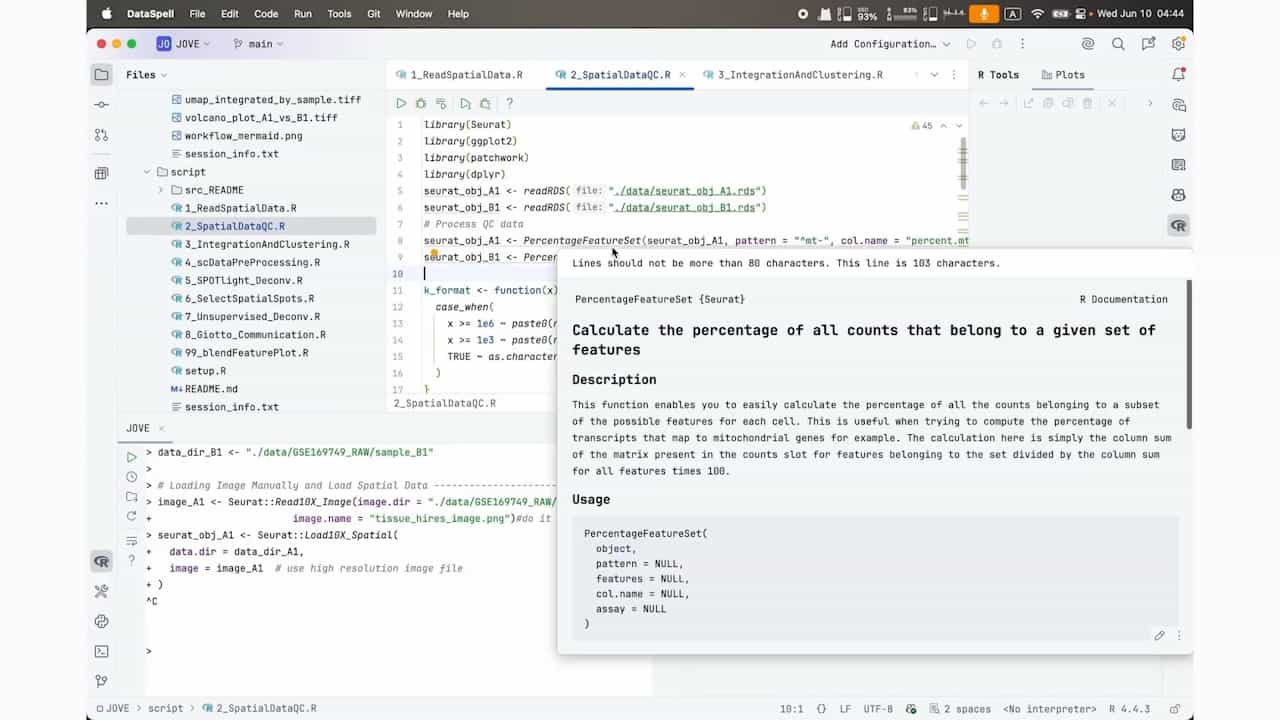
A Practical Workflow for Spatial Transcriptomics Data Analysis: From Data Acquisition to Advanced Analyses
Published on: August 21, 2026
Hao Wen1, Qunlun Shen2, Shuqin Zhang1,3
1School of Mathematical Sciences, Fudan University, Shanghai, China.

Abstract:
Spatial transcriptomics enables comprehensive characterization of tissue architecture, and the identification of spatially variable genes (SVGs) is a critical step for defining region-specific molecular markers and uncovering spatially regulated mechanisms across diverse biological contexts. However, most existing methods for SVG detection overlook cell density variations, a major confounding factor in complex tissues such as tumors, where heterogeneous cellularity frequently introduces false-positive calls. Here we present SpatioCAD, a computational framework that explicitly decouples genuine spatial expression patterns from confounding effects driven by cellularity. SpatioCAD introduces a graph diffusion model that inherently incorporates cell-density-aware conditions to simulate expression propagation, supporting the identification of SVGs across a broad range of expression levels. Systematic evaluations on simulated datasets demonstrate its superior statistical power and specificity. Applied to breast cancer, lung cancer, and glioma datasets, SpatioCAD identifies functionally diverse SVGs, including low‑abundance transcripts with established roles in tumor progression, while also recapitulates biologically meaningful tissue architecture features. Importantly, SpatioCAD is fast and scalable to large tissues.