Periodic Electron Density Overlap Descriptors for Crystal Band Gap Prediction
Related Experiment Video
Updated: Nov 11, 2026
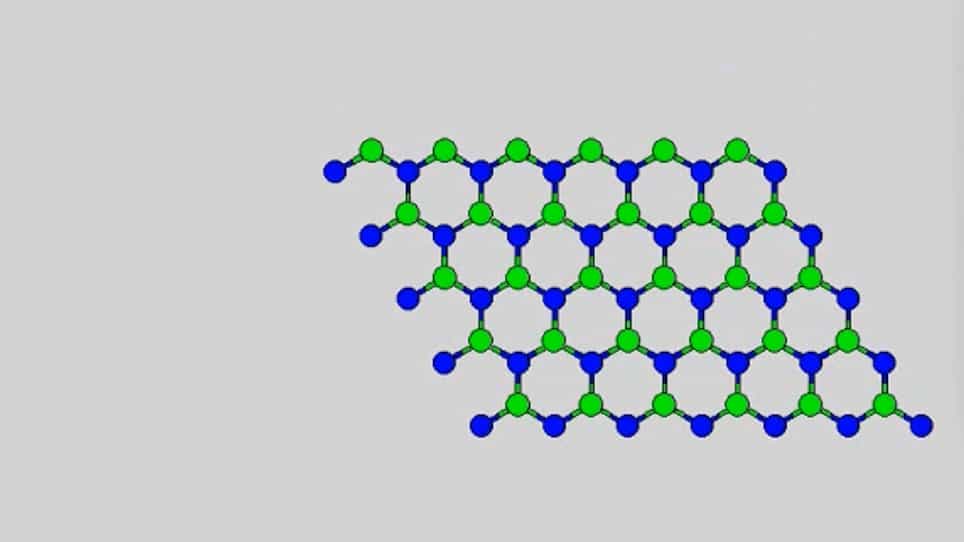
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Zhaosheng Zhang1, Yiheng Xi1, Jiadong Liu1
1College of Chemistry and Materials Science, Institute of Life Science and Green Development, Hebei University, Baoding071002, P. R. China.

Abstract:
A valence electron density encodes spatial electronic structure that complements an atomic position representation. We examine whether a periodic adaptation of the smooth overlap of electron densities (SOED) improves the modeling of density functional theory band gaps in inorganic crystals. Atom-centered neighborhoods are sampled directly from periodic valence density grids, expanded in radial functions and spherical harmonics, and summarized by rotationally invariant power spectra. A scaffold density constructed from atomic positions supplies a SOAP-inspired control with the same overlap machinery. We evaluate 40490 structures from MP-20-Charge after filtering undefined Pauling electronegativities and repartitioning by reduced formula. A validation-selected electronic SOED ensemble attains a held-out RMSE of 0.6179 eV, compared with 0.6595 eV for a local chemistry-matched SOAP control. The reduction of 0.0415 eV has a 95% formula-cluster bootstrap interval of 0.0255 to 0.0578 eV. Five formula-group splits, including the primary split, give mean errors of 0.6315 and 0.6581 eV, respectively. These results support periodic density overlap descriptors for modeling computed band gaps when valence charge densities are available.
More Related Videos
Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states that no two...
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 01:19
Crystal Density