Approaching Coupled Cluster Accuracy with Positive Semidefinite Vertex Corrected Self-Energies
Related Experiment Video
Updated: Nov 12, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Yaroslav Pavlyukh1, Fabien Bruneval2, Arno Förster3
1Independent Researcher , Karlsruhe76351, Germany.

Abstract:
Hedin's formalism of functional derivatives is a successful method for systematically constructing correlated electronic theories, largely due to the success of its lowest-order self-energy expansion, the GW approximation. Beyond GW, diagrammatic resummation schemes attempt to mix correlations simultaneously across all particle-particle and particle-hole channels. Because such a comprehensive treatment is computationally prohibitive for realistic molecular systems, a highly effective alternative is to fully account for electronic correlations in one specific channel, typically the particle-hole channel. This idea was recently implemented for molecular systems, yielding a self-energy expressed in terms of excited-state energies and transition amplitudes from the solution of the Bethe-Salpeter equation, rather than the random-phase approximation used in GW. While this approach is predictive and numerically efficient, it violates the fundamental positive-definiteness constraint on the electron spectral function in certain energy ranges. In this study, we resolve this physical flaw by deriving a positive-semidefinite (PSD) extension of the theory using a rigorous framework based on the nonequilibrium Green's function formalism. The PSD constraint introduces new scattering channels and triplet intermediate states and restores the correct physical behavior. We demonstrate that it consistently improves quasiparticle energies across standard molecular benchmarks, with an accuracy comparable to coupled-cluster reference calculations.
Related Concept Videos
 01:10
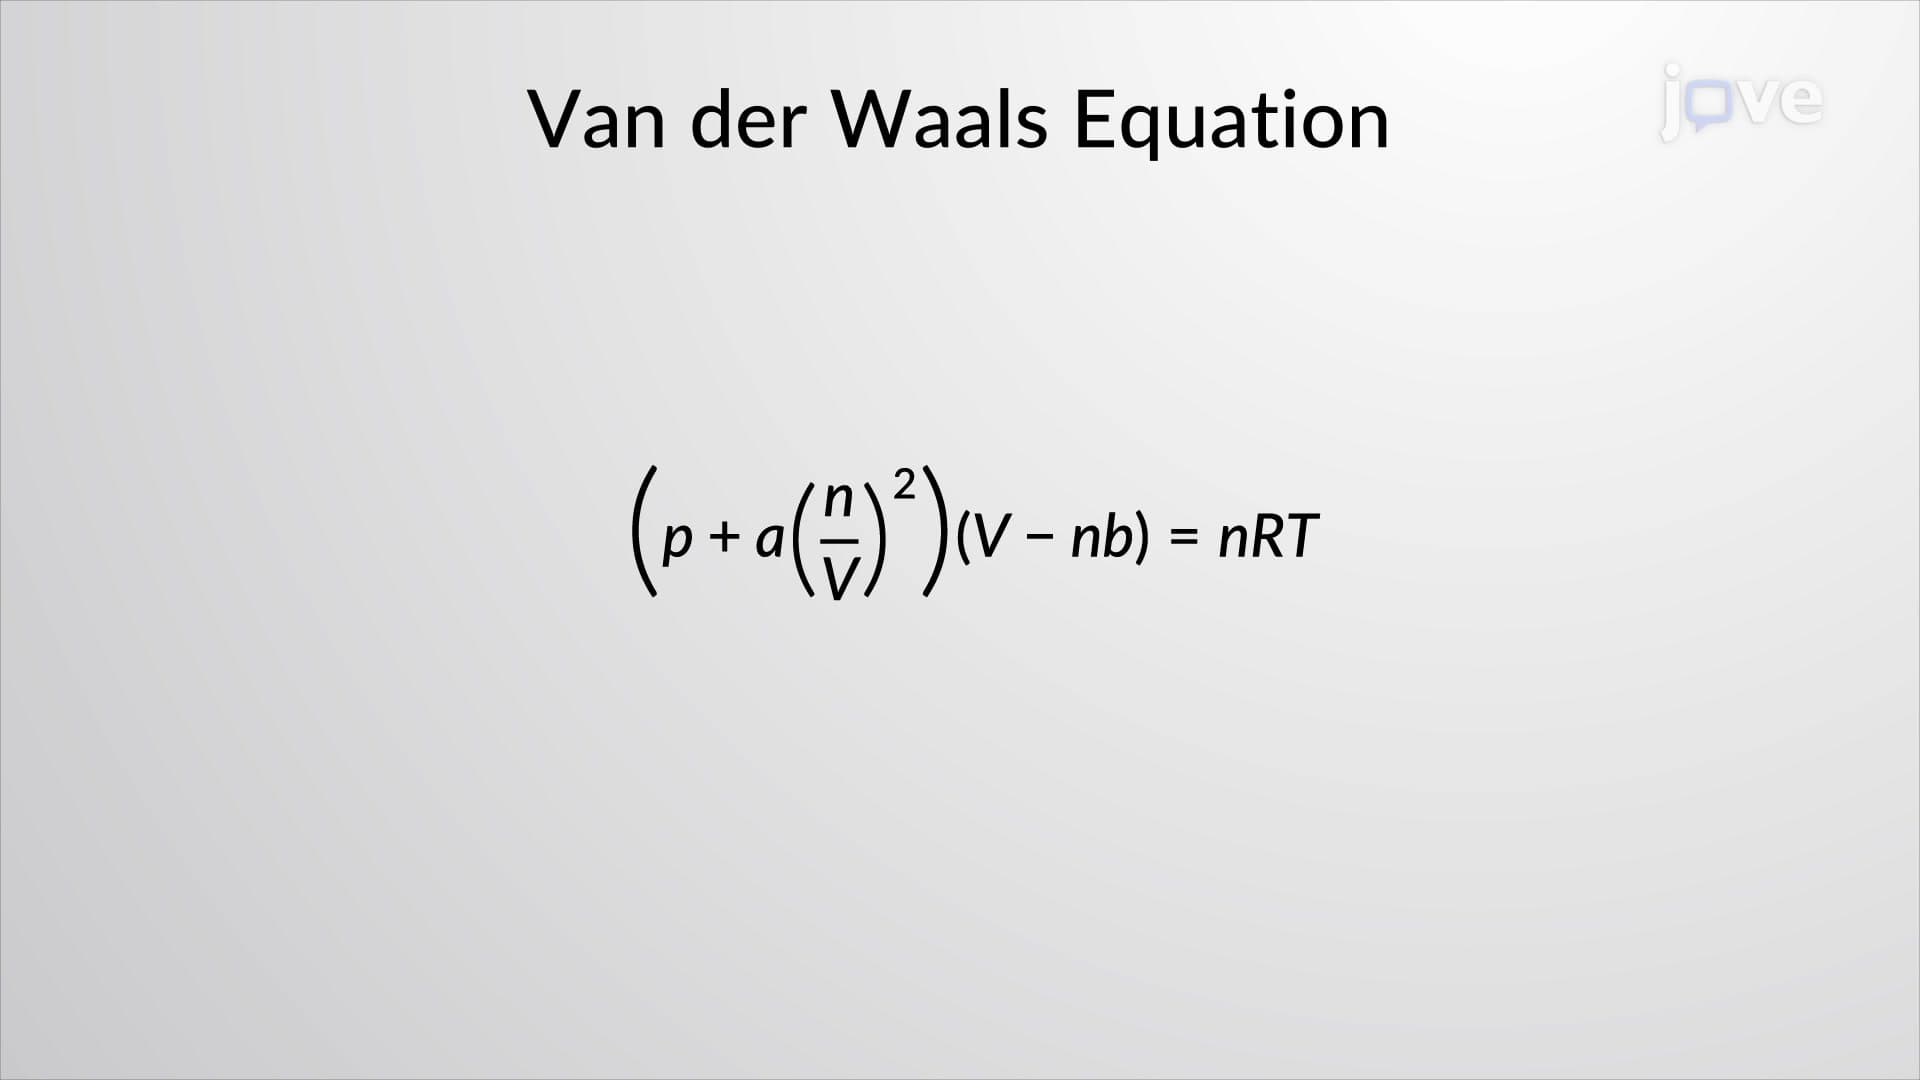
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:35
Hybridization of Atomic Orbitals II
 01:12
Reduced Mass Coordinates: Isolated Two-body Problem
 01:22
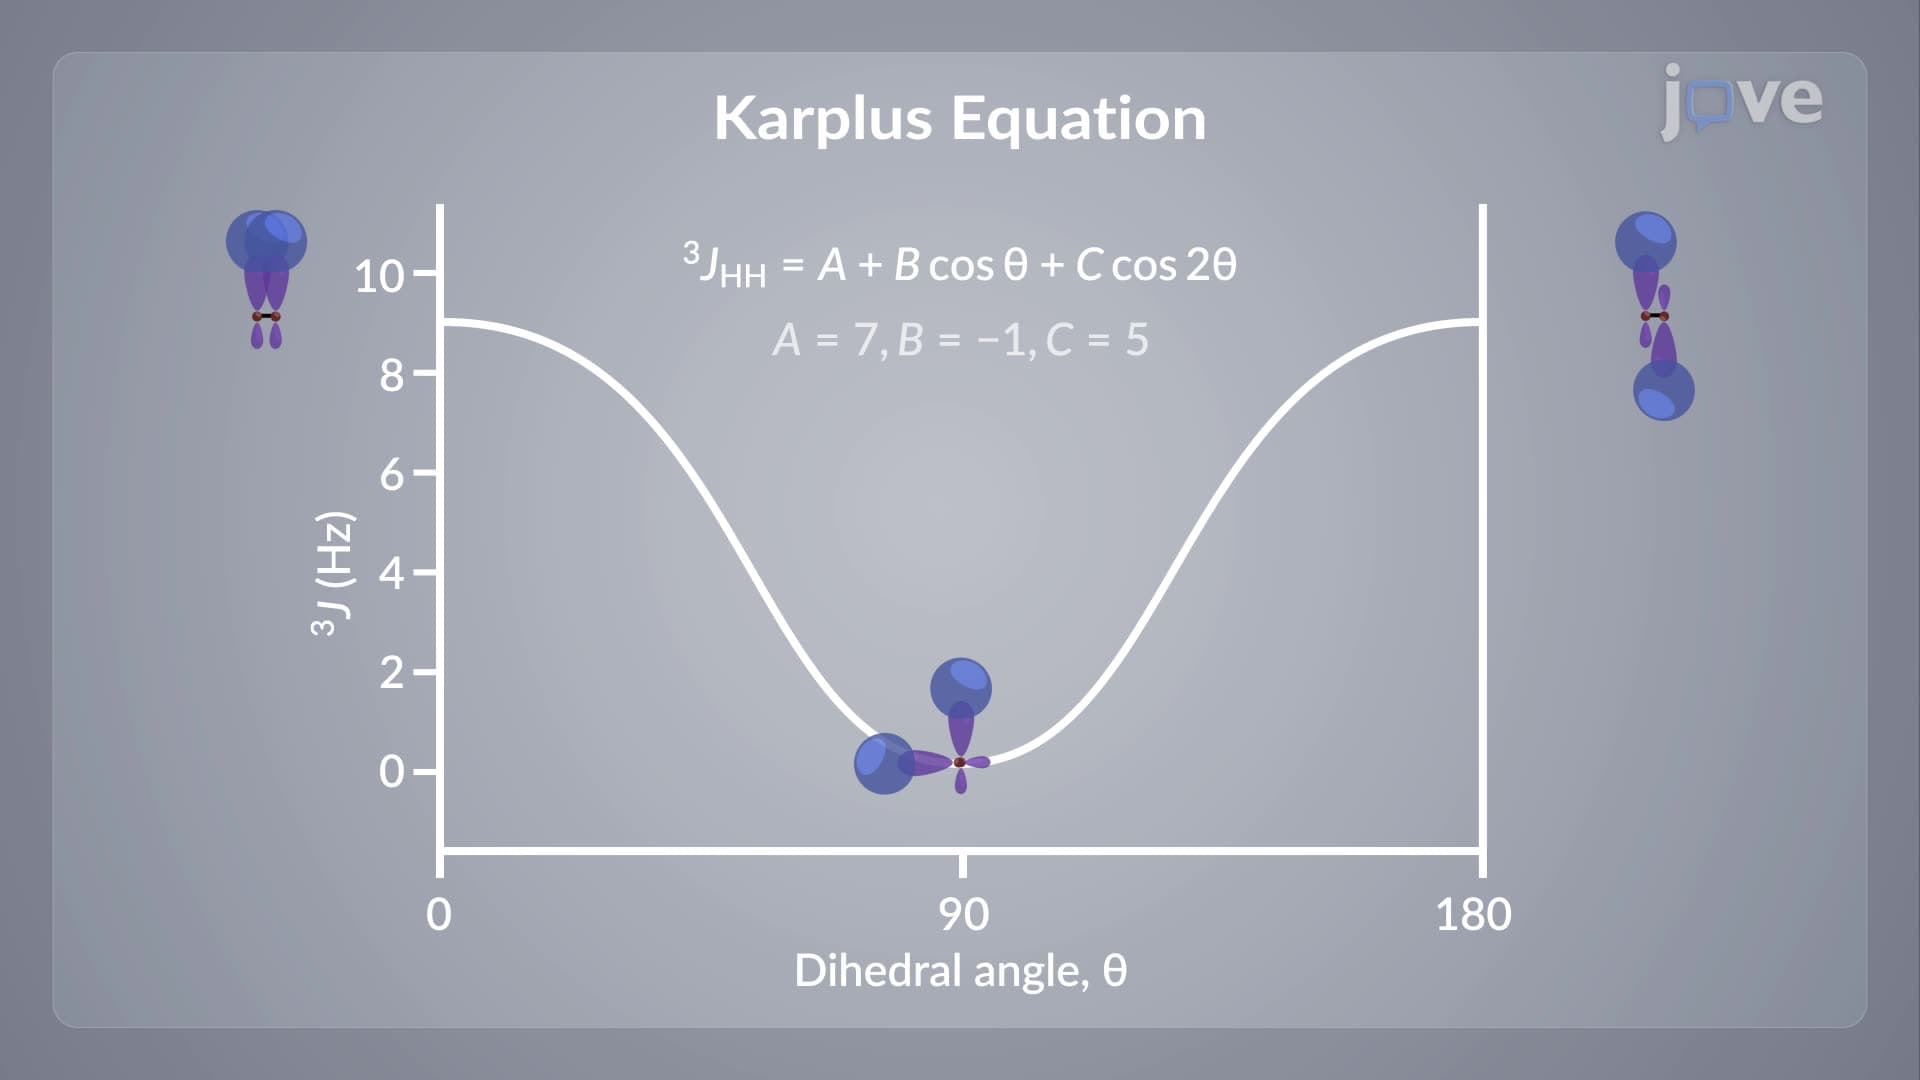
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:24
Van der Waals Interactions
 01:16
Electrostatic Boundary Conditions
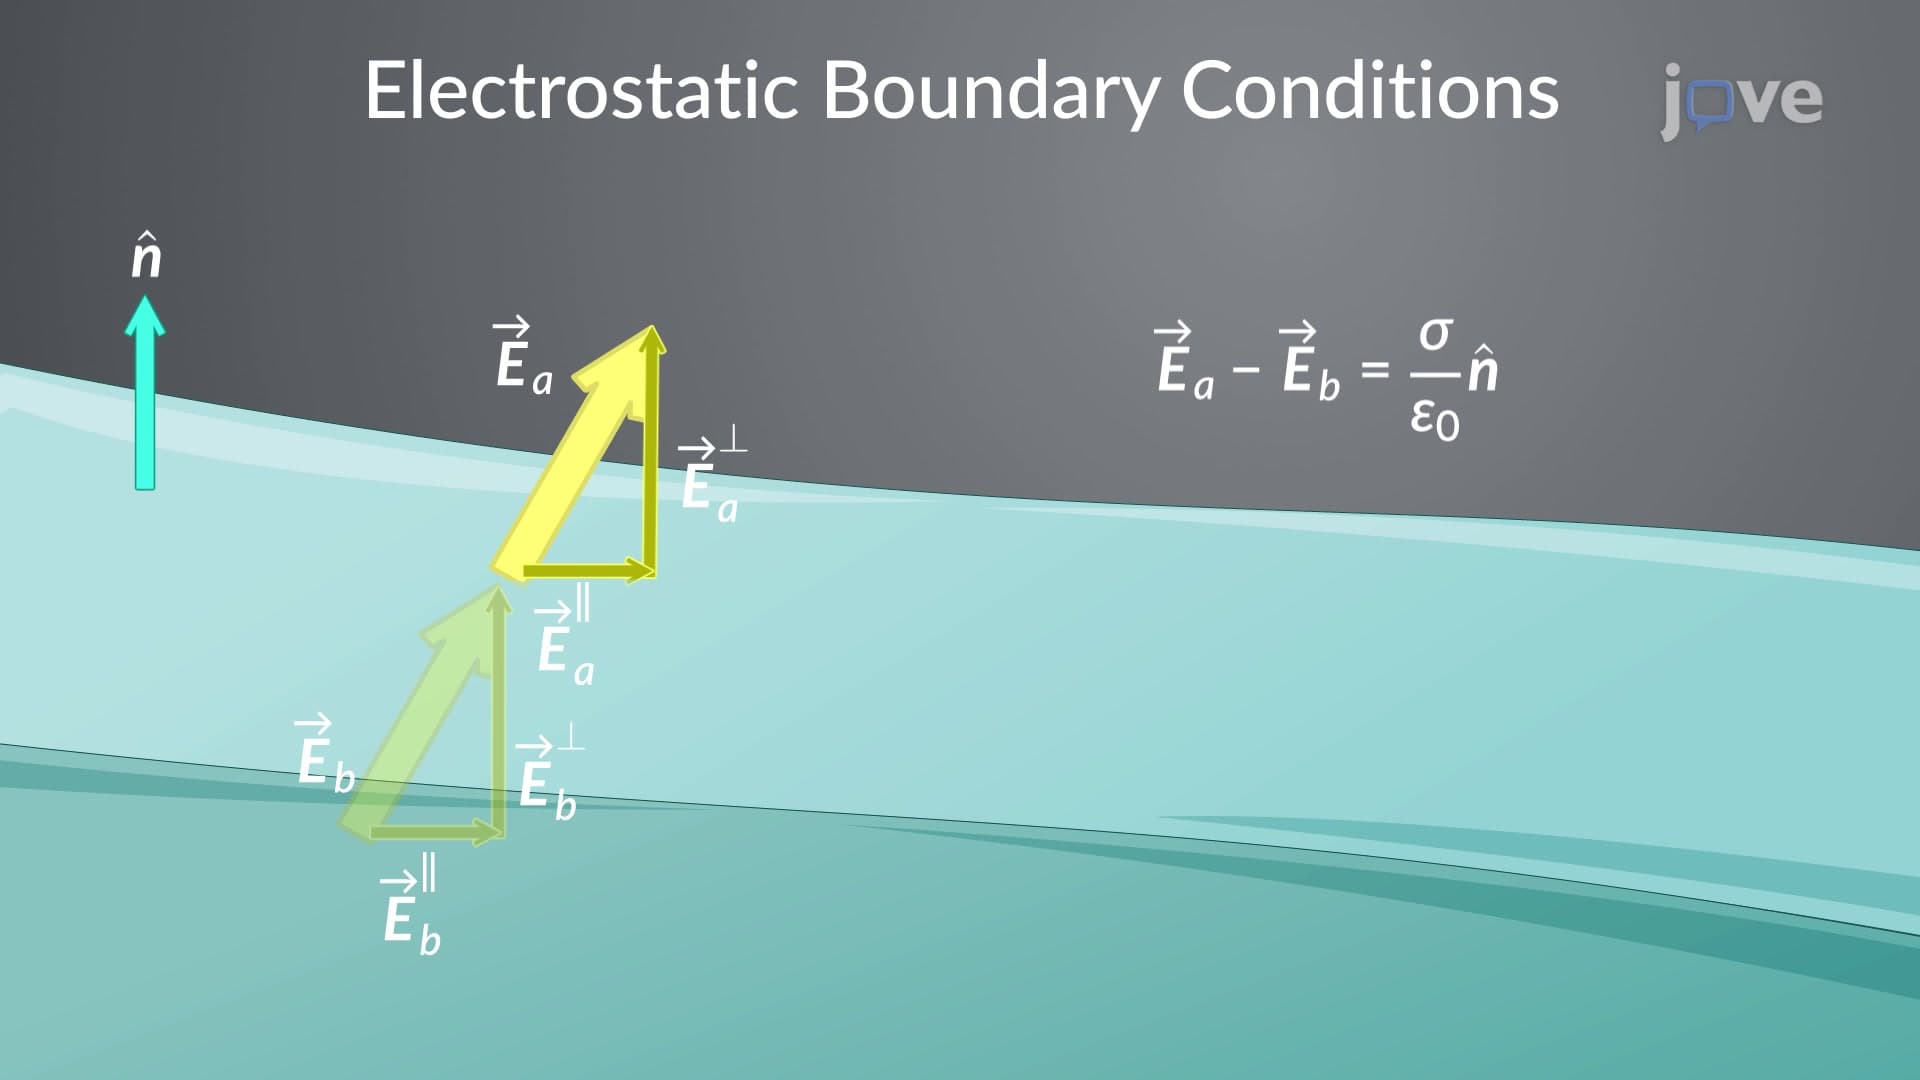
The surface integral of an electric field is given by Gauss's law in integral form and is related to...