Related Experiment Video
Updated: Aug 18, 2026

Preclinical Cardiac Electrophysiology Assessment by Dual Voltage and Calcium Optical Mapping of Human Organotypic Cardiac Slices
Published on: June 16, 2020
Digoxin toxicity compared with myocardial digoxin and potassium concentration

Abstract:
1 Twenty-nine dogs were given digoxin (0.25 mg) by mouth twice daily for eight days. Some of them (group 1) also received diuretics and others (group 2) a mineralocorticoid. The dogs were then given an intravenous bolus injection of digoxin and plasma and cardiac muscle were analysed for digoxin and potassium. 2 In the digitalized dogs, myocardial potassium concentration decreased following the intravenous injection of either 0.05 or 0.15 mg/kg digoxin; in contrast, in those dogs given diuretics or mineralocorticoid the potassium concentration increased. 3 Ventricular arrhythmias occurred after digoxin injection (0.05 mg/kg) in the hypokalemic dogs, in those given a mineralocortocoid and in those dogs which received a toxic digoxin dose (0.15 mg/kg). No arrhythmias where seen in the control (digitalized) group. 4 Myocardial digoxin concentrations were similar in the control digitalized group and in the mineralocorticoid-treated dogs after the intravenous administration of the lower digoxin dose (0.05 mg/kg). The myocardial digoxin concentration was significantly higher in the hypokalemic group and in the group receiving the higher digoxin dose (0.15 mg/kg). 5 There was no obvious relationship between the occurrence of arrhythmias and the myocardial concentration of digoxin or potassium.
More Related Videos
 10:01
High-Throughput Optical Controlling and Recording Calcium Signal in iPSC-Derived Cardiomyocytes for Toxicity Testing and Phenotypic Drug Screening
Published on: March 31, 2022
 08:11
Voltage-Dependent Potassium Current Recording on H9c2 Cardiomyocytes via the Whole-Cell Patch-Clamp Technique
Published on: November 11, 2022
Related Concept Videos
 01:14
Time Course of Drug Effect
 01:12
Antiarrhythmic Drugs: Class III Agents as Potassium Channel Blockers
 01:26
Heart Failure Drugs: Inotropic Agents
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:15
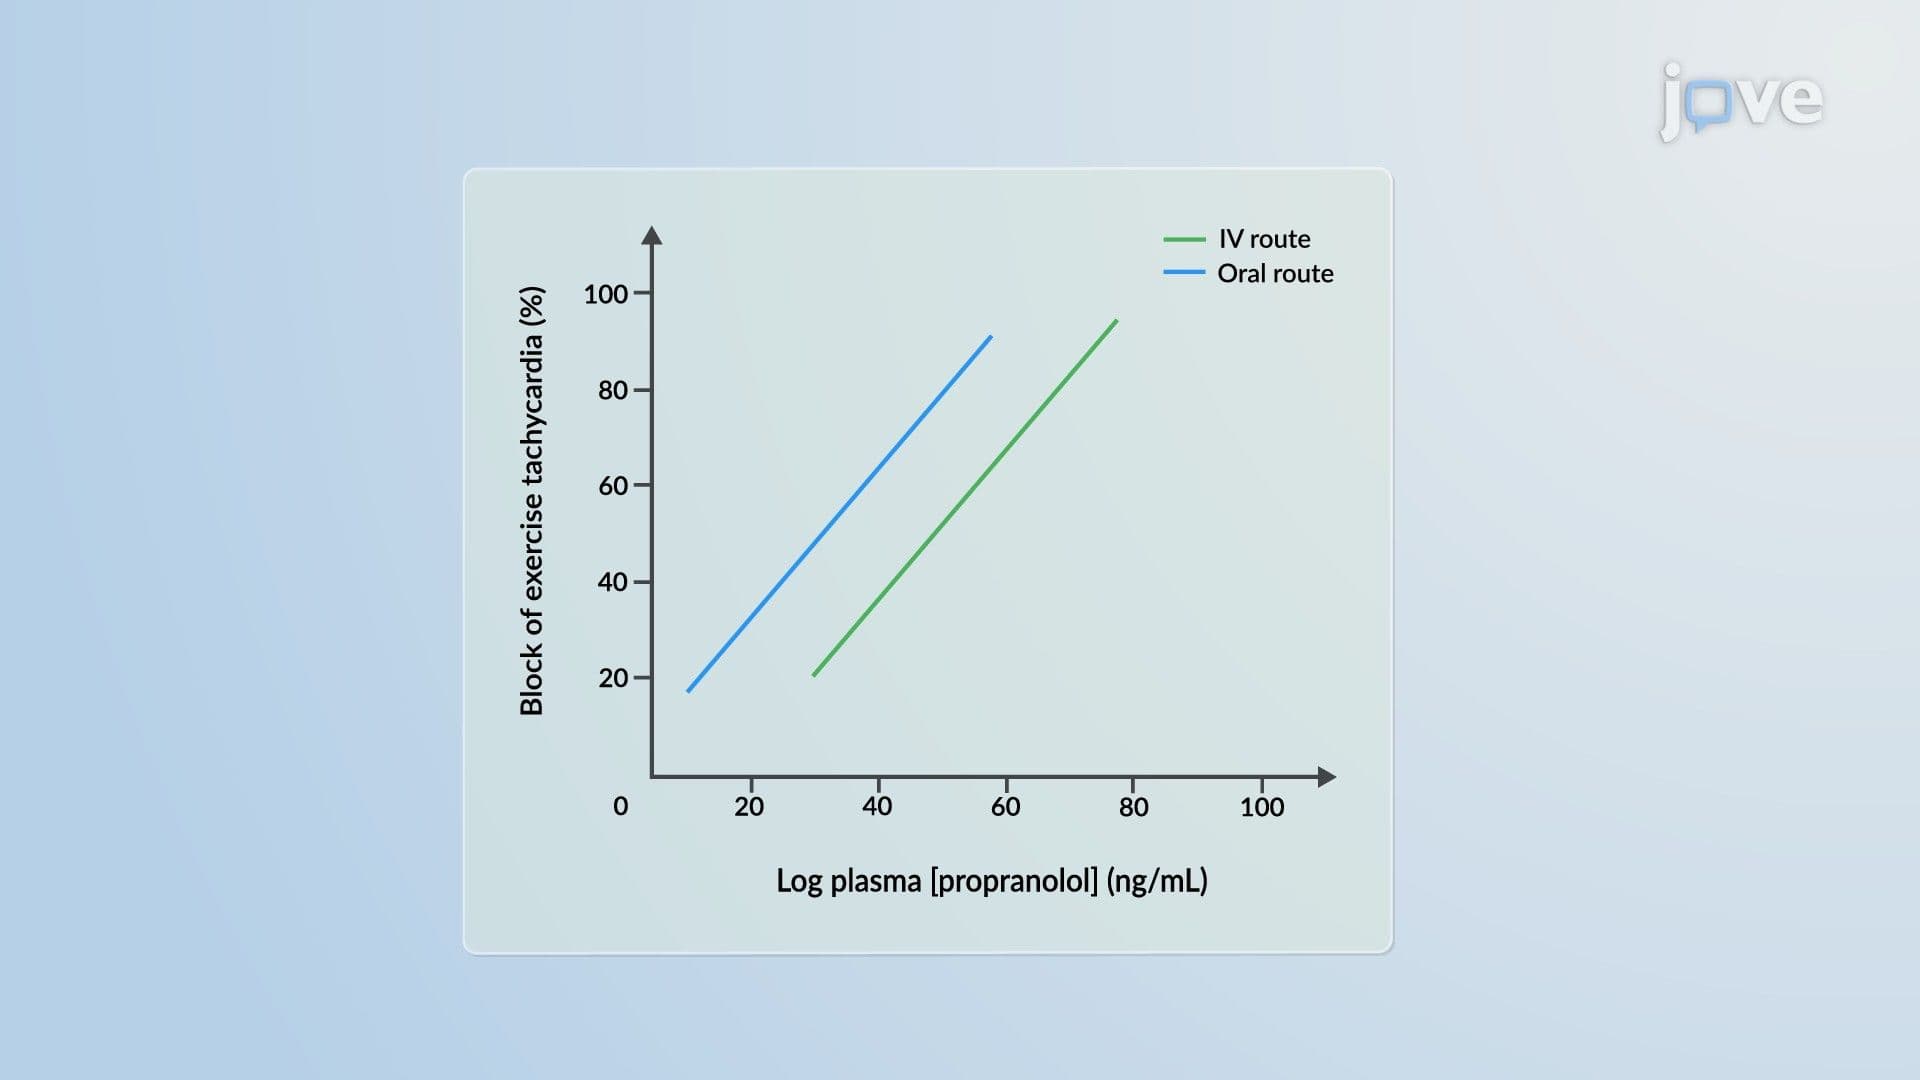
Pharmacodynamic Models: Logarithmic Concentration–Effect Model
 01:24
Drug Toxicity: Dose-Dependent Reactions