Related Experiment Video
Updated: Oct 21, 2026
Evaluation of Drug Sorption to PVC- and Non-PVC-based Tubes in Administration Sets Using a Pump
Published on: March 11, 2017
Effect of concentration-dependent binding to plasma proteins on the pharmacokinetics and pharmacodynamics of

Abstract:
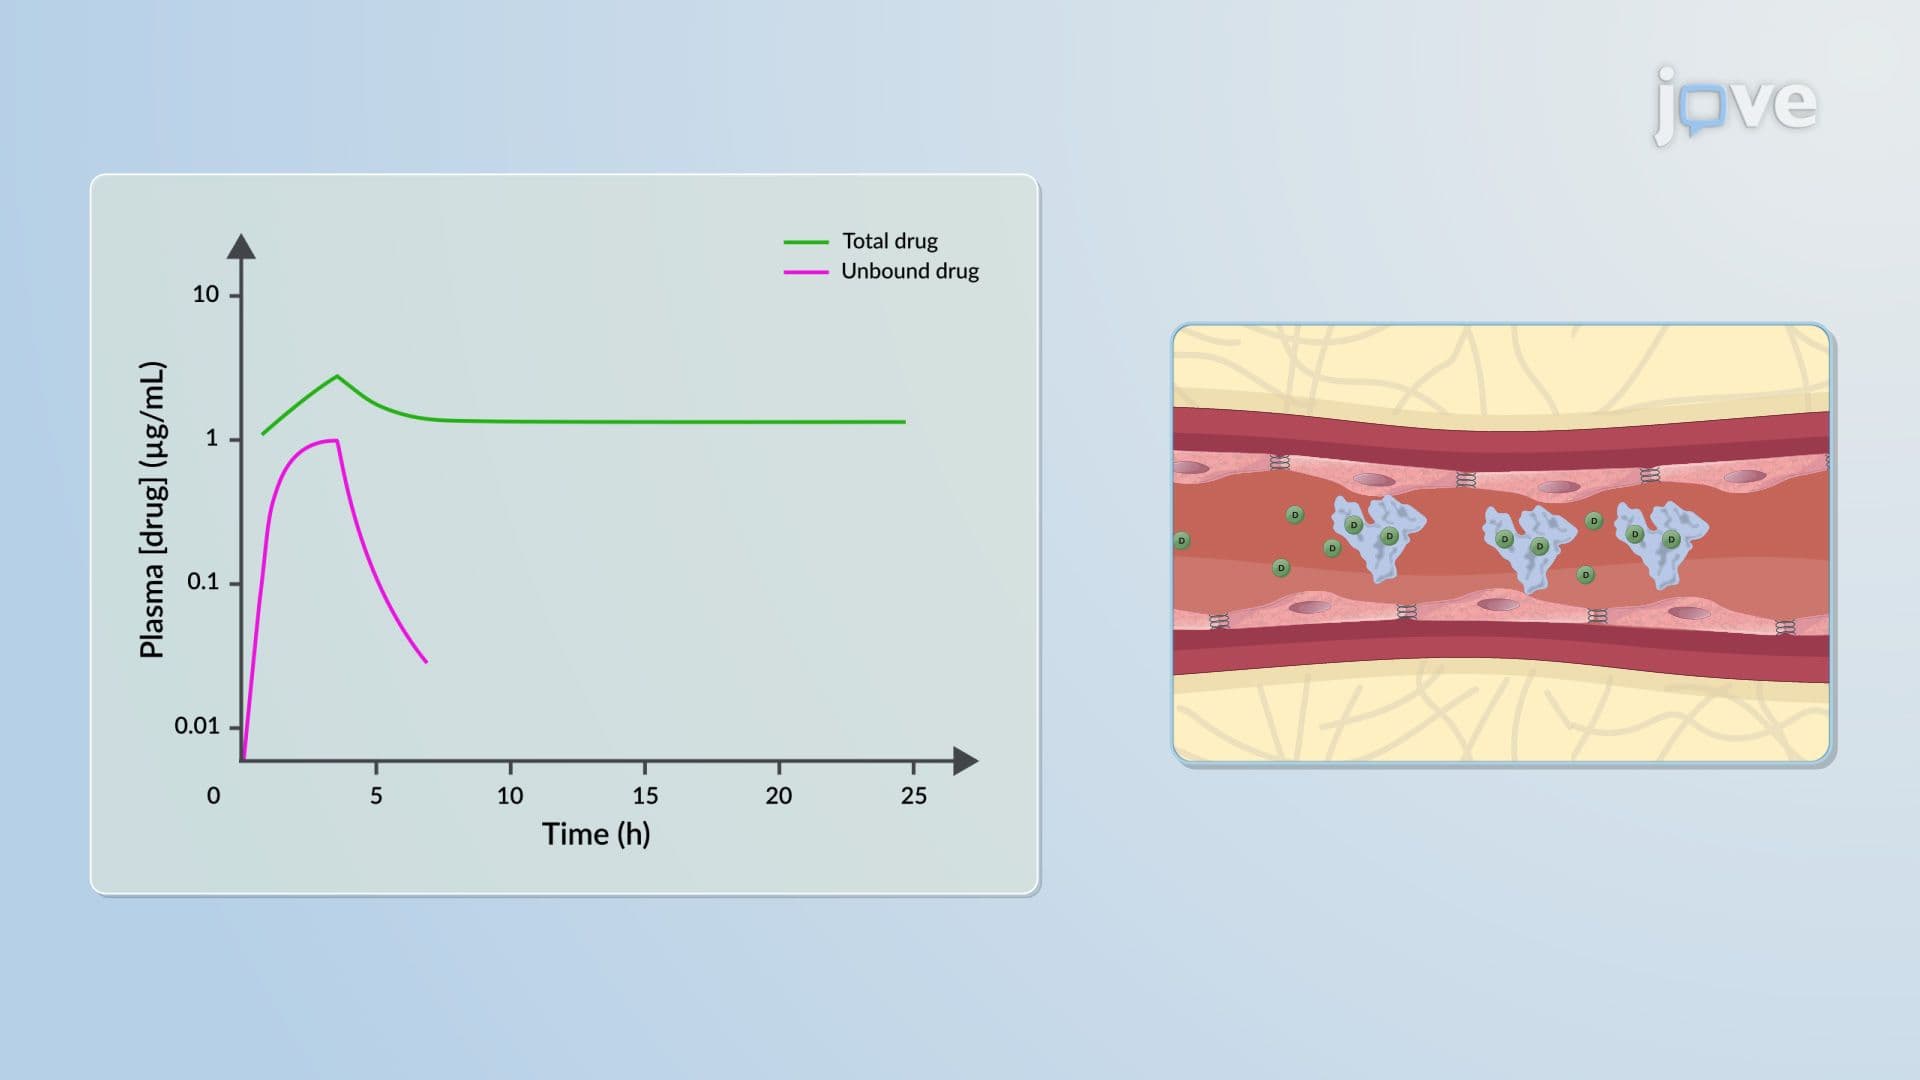
Disopyramide exhibits concentration-dependent binding to plasma proteins at therapeutic plasma concentrations. This paper reviews the effect of this type of binding on both the pharmacokinetic and pharmacodynamic properties of the drug. For a drug with capacity-limited, binding-sensitive elimination like disopyramide, concentration-dependent binding to plasma proteins produces a non-linear relationship between dosing rate and total plasma concentrations of the drug. However, when dosing rate is related to unbound concentrations of drug, the relationship is linear. Renal clearance of total disopyramide has been found to depend on the unbound fraction whereas renal clearance of unbound drug may be dependent upon time after drug administration as well as route of drug administration. However, due to some potential methodological problems, these data need verification. The concentration-dependent binding of disopyramide to plasma proteins, with its resultant effects on clearance and distribution, produces a concave curvature in the log unbound concentration versus time curves and a log-linear decline in total plasma concentration versus time after intravenous administration of the drug. Several pharmacological studies suggest that the unbound drug is active in terms of producing both desirable and undesirable effects. Hence, monitoring unbound concentrations of disopyramide seems more rational for clinical purposes. The few studies that have been carried out examining the optical isomers of disopyramide have suggested that both the pharmacokinetic and pharmacodynamic properties of the drug are stereoselective. Recent data reveal stereoselective binding to plasma proteins in humans.(ABSTRACT TRUNCATED AT 250 WORDS)
More Related Videos
 07:42
Contractions of Human-iPSC-derived Cardiomyocyte Syncytia Measured with a Ca-sensitive Fluorescent Dye in Temperature-controlled 384-well Plates
Published on: October 18, 2018
 10:41
Laser-Induced Action Potential-Like Measurements of Cardiomyocytes on Microelectrode Arrays for Increased Predictivity of Safety Pharmacology
Published on: September 13, 2022
Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:23
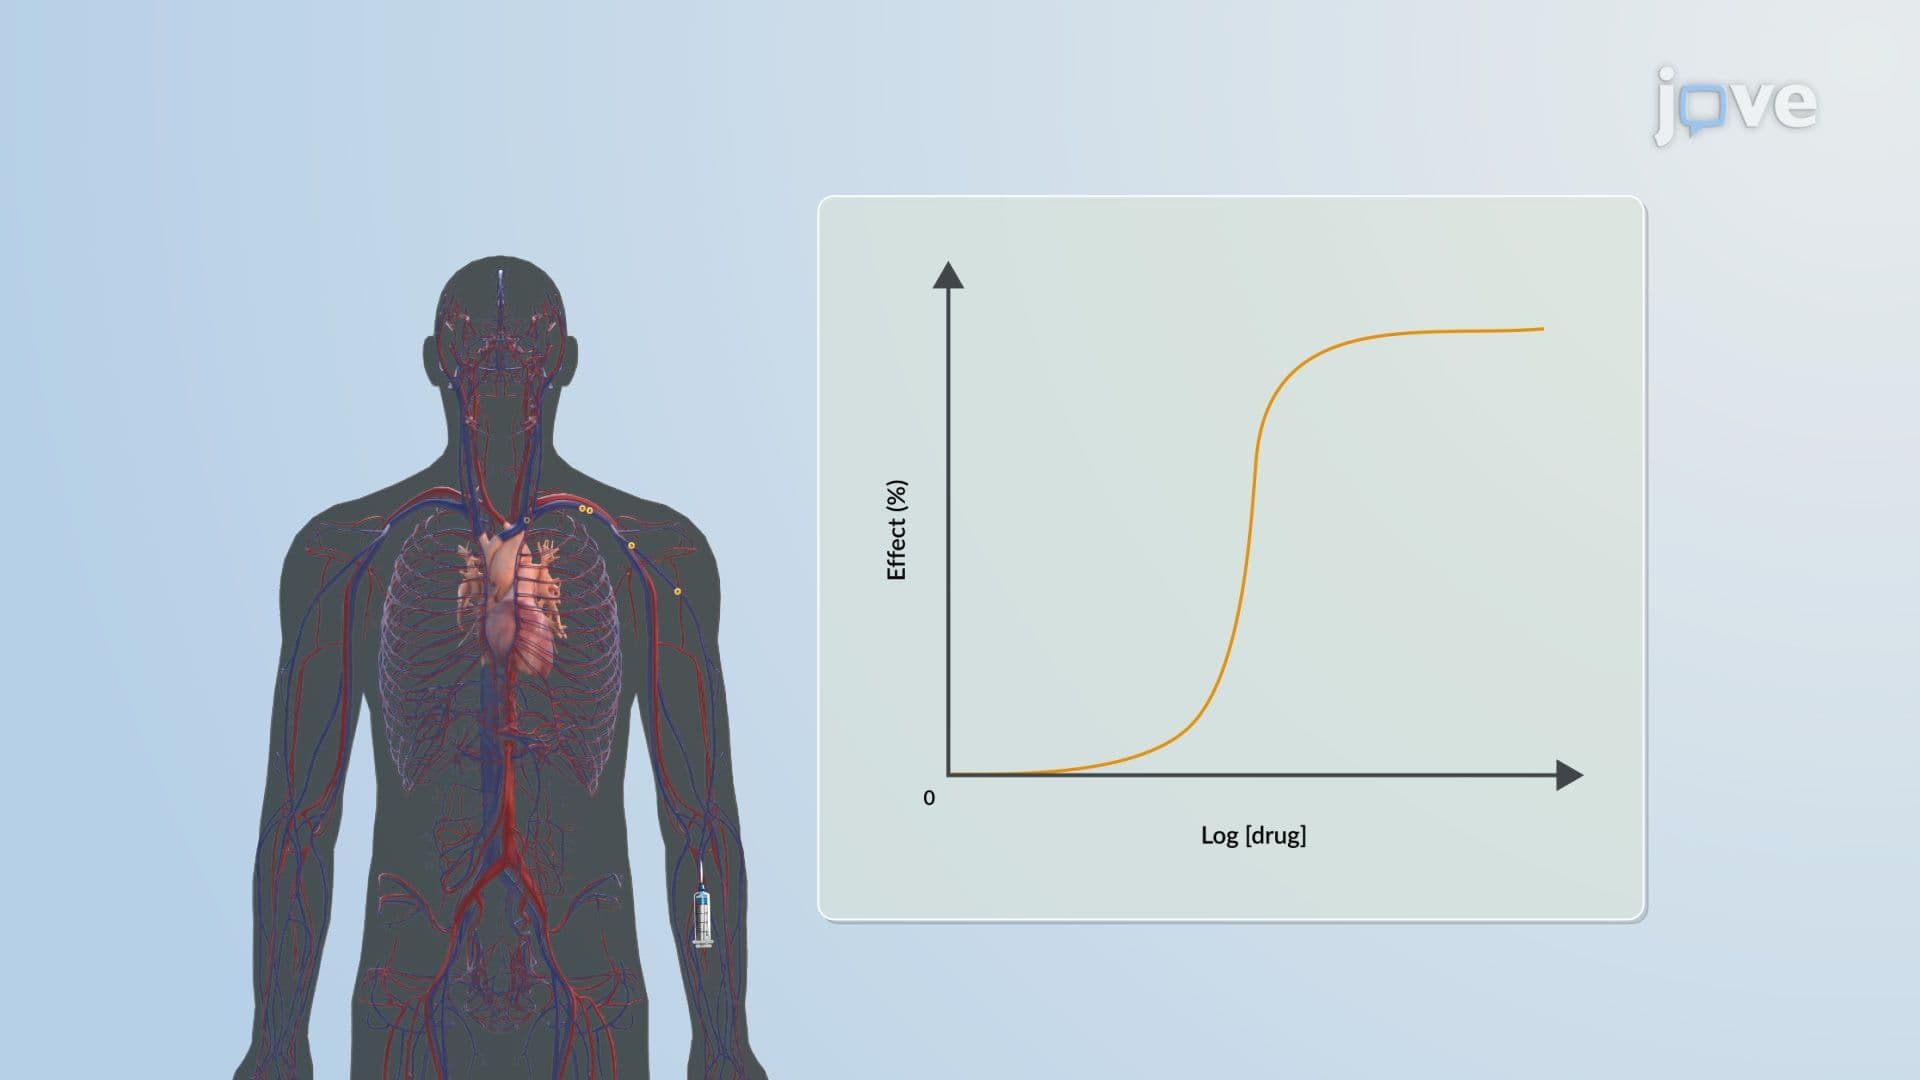
Pharmacokinetic–Pharmacodynamic Relationship: Intensity of Dose-Effect Relationship
 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model