Related Experiment Video
Updated: May 5, 2026

A Precision Medicine Tool for Measurement and Monitoring of Hemoglobin S in Sickle Cell Disease Patients Receiving Transfusion Therapy
Hematologic and biosynthetic studies in homozygous hemoglobin Constant Spring

The elongated alpha-globin chains of hemoglobin Constant Spring (alpha cs chain of HbCS ) are produced in low amounts such that the alpha cs-gene acts as a form of alpha-thalassemia; yet in the homozygous state the pathophysiological effects of this mutant are more severe than in the corresponding conditions that result from alpha-globin gene deletions. In studies designed to examine this discrepancy, we have demonstrated that a significant proportion of red cells produced in an HbCS homozygote has a much reduced red cell life span. Contrary to previous reports, we have been able to demonstrate the expected deficit in alpha-chain production in this condition and have shown that both the cessation of globin chain synthesis in vitro and the destruction of the excess beta-chains occur unusually rapidly. Comparison with various deletion forms of alpha-thalassemia suggests that, in terms of intracellular globin chain precipitates and free beta-chain pool, homozygous HbCS red cells more closely resemble those of HbH disease, with three of the four alpha-genes inactivated, than they do the more comparable alpha-thalassemia carriers with only two genes deleted.
The elongated alpha-globin chains of hemoglobin Constant Spring (alpha cs chain of HbCS ) are produced in low amounts such that the alpha cs-gene acts as a form of alpha-thalassemia; yet in the homozygous state the pathophysiological effects of this mutant are more severe than in the corresponding conditions that result from alpha-globin gene deletions. In studies designed to examine this discrepancy, we have demonstrated that a significant proportion of red cells produced in an HbCS homozygote has a much reduced red cell life span. Contrary to previous reports, we have been able to demonstrate the expected deficit in alpha-chain production in this condition and have shown that both the cessation of globin chain synthesis in vitro and the destruction of the excess beta-chains occur unusually rapidly. Comparison with various deletion forms of alpha-thalassemia suggests that, in terms of intracellular globin chain precipitates and free beta-chain pool, homozygous HbCS red cells more closely resemble those of HbH disease, with three of the four alpha-genes inactivated, than they do the more comparable alpha-thalassemia carriers with only two genes deleted.
Related Concept Videos
 01:49
Multiple Allele Traits
 01:57
Gene Families
Occasionally these regions can be adapted to take on new roles within the organism, becoming novel genes...
 02:21
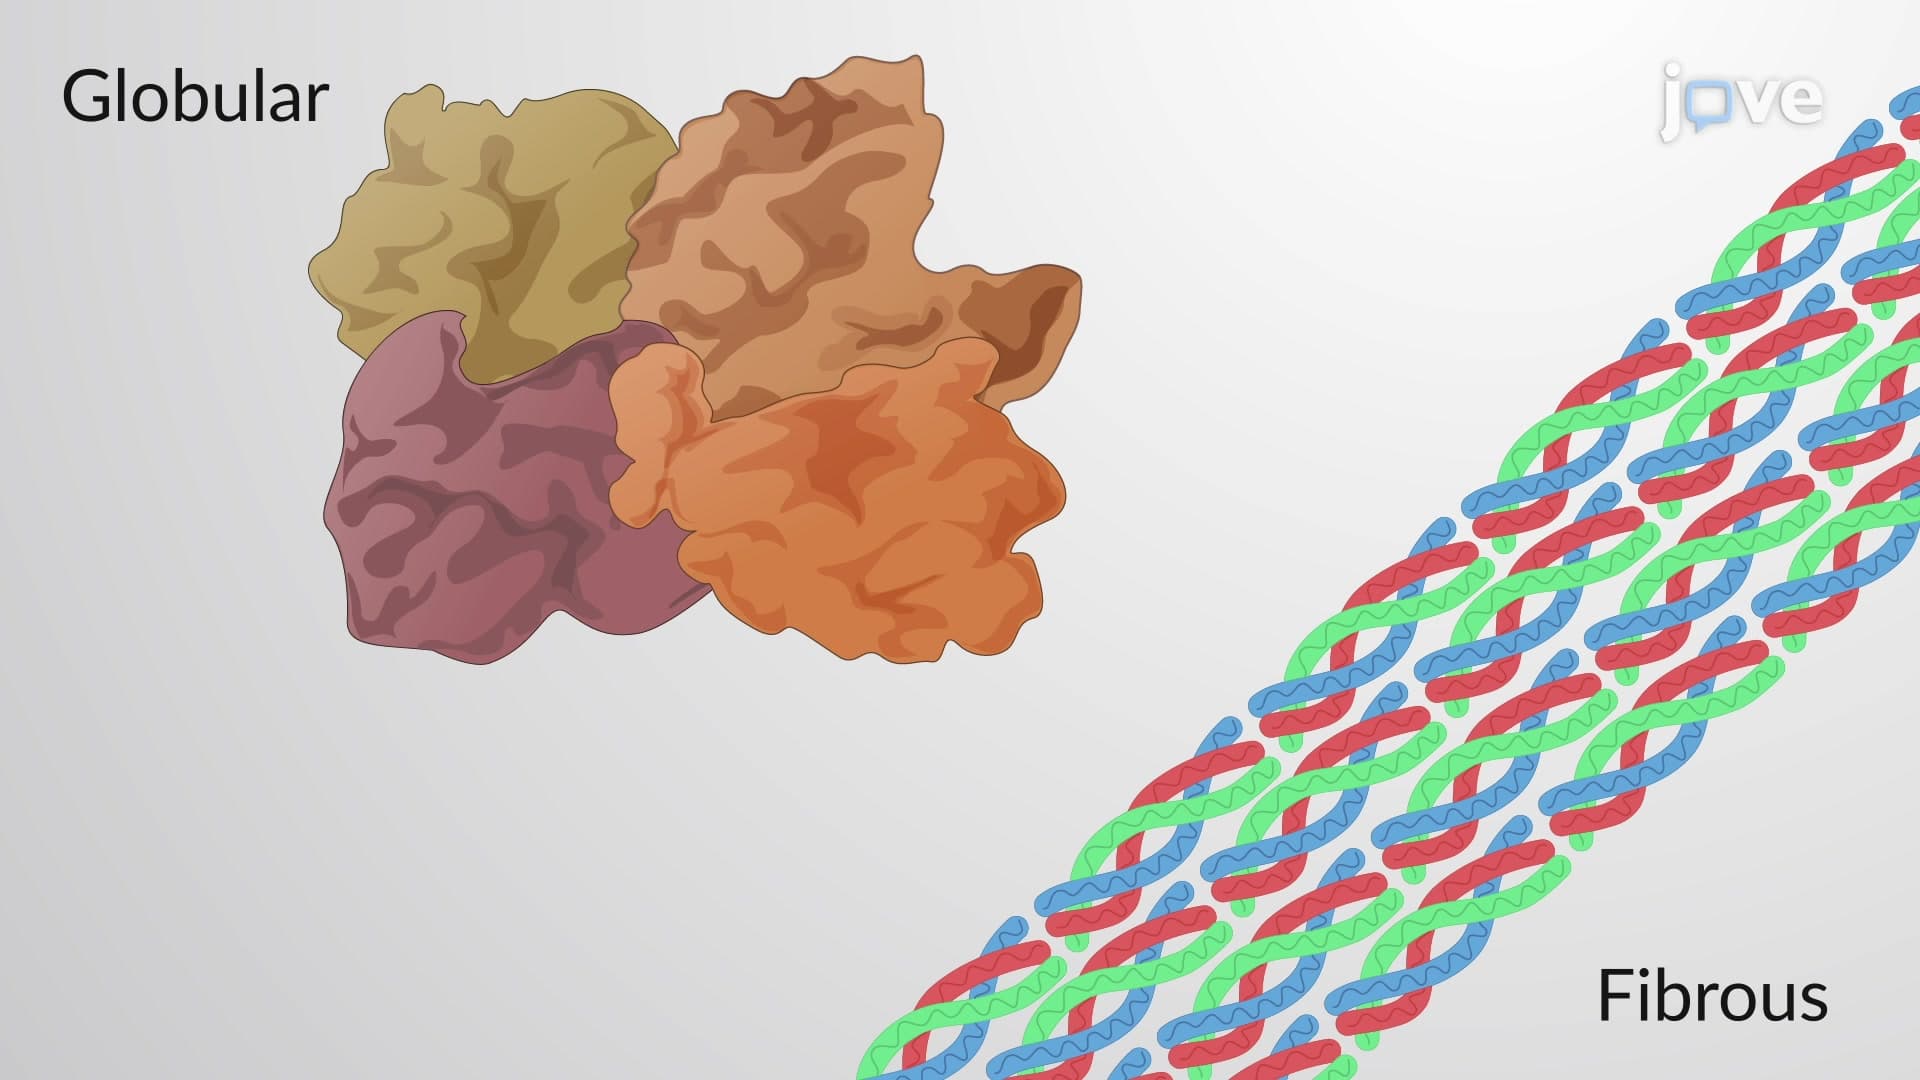
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 01:24
Hemoglobin
When all four heme groups are bound to oxygen, the resulting molecule is called oxyhemoglobin. As a result, arterial blood...
 01:27
Oxygen Transport in the Blood
 01:26
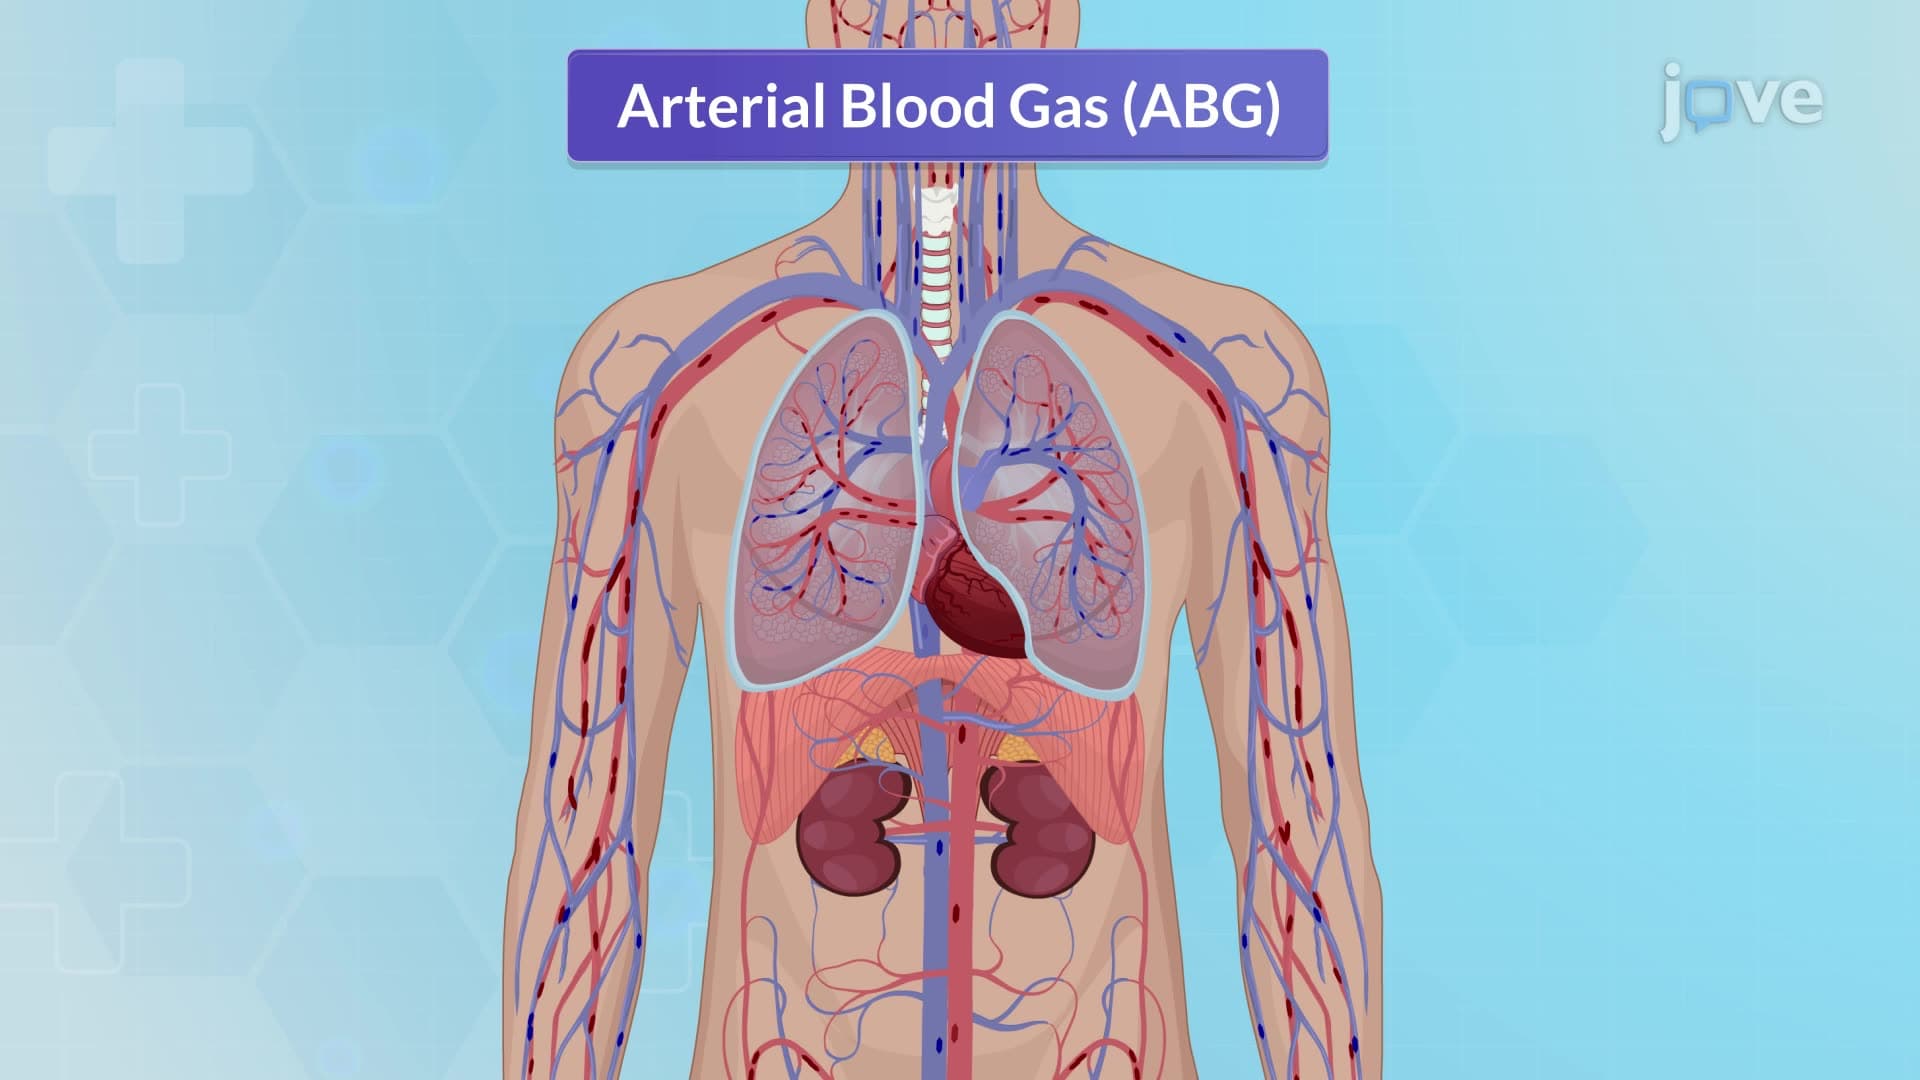
Blood Studies I: ABG and VBG
Arterial Blood Gas (ABG)
Arterial Blood Gas (ABG) studies are crucial for assessing the lungs' ability to supply oxygen and remove carbon dioxide, reflecting the patient's ventilation status. They also help understand the kidneys' capacity to...