4-Hydroxybutyric aciduria
Related Experiment Video
Updated: Sep 27, 2026
Analysis of Interactions between Endobiotics and Human Gut Microbiota Using In Vitro Bath Fermentation Systems
Published on: August 23, 2019
Z Rahbeeni1, P T Ozand, M Rashed
1Department of Pediatrics, King Faisal Specialist Hospital and Research Centre, Riyadh, Saudi Arabia.

Insights
4-hydroxybutyric aciduria is a rare genetic disorder causing severe developmental delay and seizures in infants. Vigabatrine treatment reduced acid levels but showed limited clinical improvement in most patients.
Area of Science:
- Biochemistry
- Genetics
- Neurology
Background:
- 4-hydroxybutyric aciduria is a rare metabolic disorder.
- Characterized by severe neurological impairment in infancy.
Observation:
- Six patients from three families presented with global delay, hypotonia, and seizures.
- Ocular manifestations included optic atrophy and retinitis pigmentosa.
- Movement disorders such as choreoathetosis, myoclonus, and dystonia were observed.
Findings:
- Markedly elevated urinary 4-hydroxybutyric acid (300-1000x normal).
- Associated increases in related organic acids affecting metabolism and beta-oxidation.
- Vigabatrine therapy rapidly decreased 4-hydroxybutyric acid excretion (80-200x normal).
Implications:
- Vigabatrine effectively reduces 4-hydroxybutyric acid levels.
- Clinical outcomes varied, with only partial improvement in some patients.
- Further research is needed to understand the disease's complex pathophysiology and treatment responses.
Abstract:
The clinical findings in six patients from three families with 4-hydroxybutyric aciduria are described. The onset of disease was in early infancy in all cases. All infants presented with severe global delay and severe hypotonia, and all patients had seizure disorder. Eye findings included optic atrophy in two patients, and retinitis pigmentosa in one. Three patients had choreoathetosis, two had myoclonus and one had severe dystonia. The urine 4-hydroxybutyric acid was 300-1000 times that of normal, and other organic acids related to its further metabolism or to its inhibitory effect on beta-oxidation were also increased. The administration of vigabatrine rapidly reduced the excretion of 4-hydroxybutyric acid promptly, and in the long-term its excretion could be kept at 80-200 times that of normal. However, the clinical course of the disease improved in only two, remained the same in two, and worsened in the remaining two patients.
More Related Videos
 10:21
Expression, Purification, Crystallization, and Enzyme Assays of Fumarylacetoacetate Hydrolase Domain-Containing Proteins
Published on: June 20, 2019
 08:56
Synthesis of a Borylated Ibuprofen Derivative Through Suzuki Cross-Coupling and Alkene Boracarboxylation Reactions
Published on: November 30, 2022
Related Concept Videos
 01:23
Urea Cycle
 03:08
Hydroboration-Oxidation of Alkenes
 01:22
Reactions of Aldehydes and Ketones: Baeyer–Villiger Oxidation
The carbonyl center is activated by...
 01:02
Loss of Carboxy Group as CO2: Decarboxylation of β-Ketoacids
 01:16
Nomenclature of Carboxylic Acid Derivatives: Acid Halides, Esters, and Acid Anhydrides
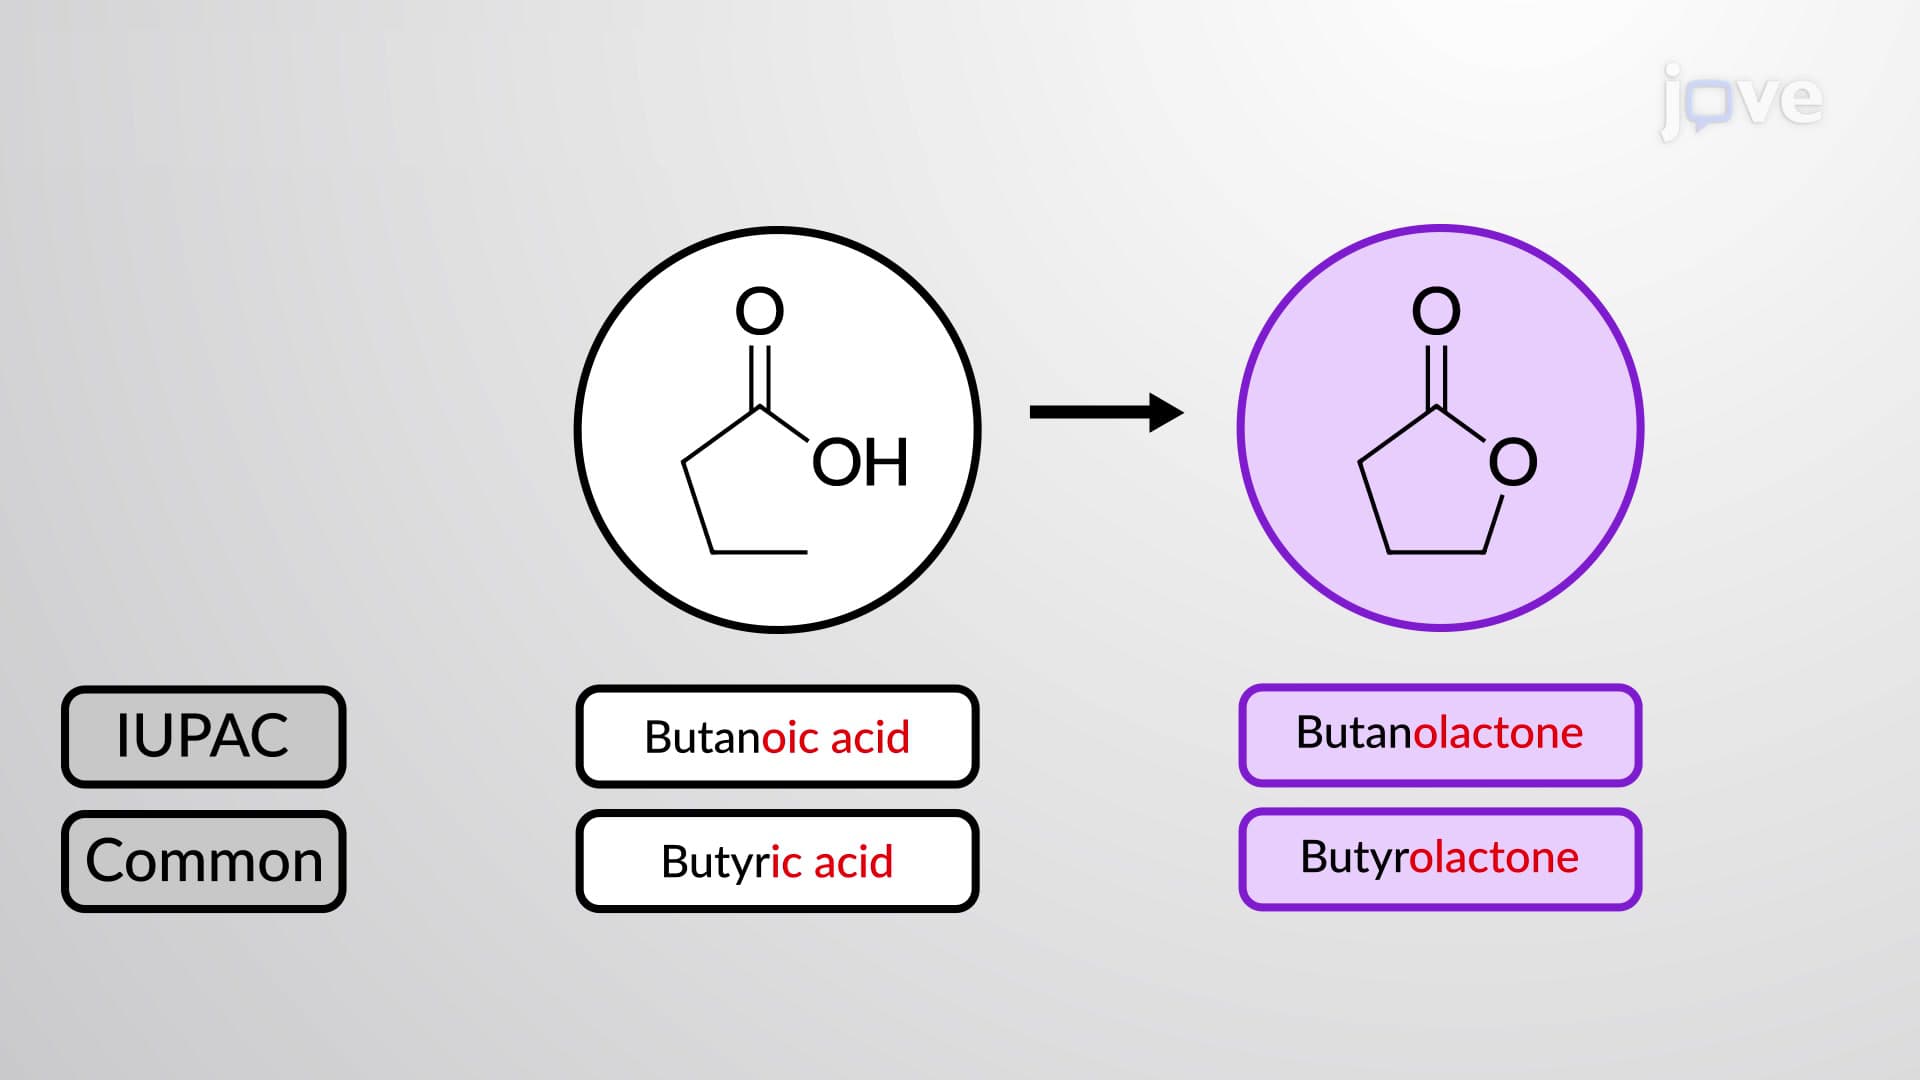
The IUPAC and common names of acid halides are derived from the corresponding carboxylic acids, by changing “ic acid” to “yl halide.” For example, as shown below, the IUPAC name ethanoyl chloride is derived from ethanoic acid, and the common name, acetyl chloride, is obtained from acetic acid.
 01:20
Inborn Errors of Metabolism