Probing structure-function relationships in human immunodeficiency virus type 1 protease via molecular dynamics simulation
Related Experiment Video
Updated: Nov 12, 2026
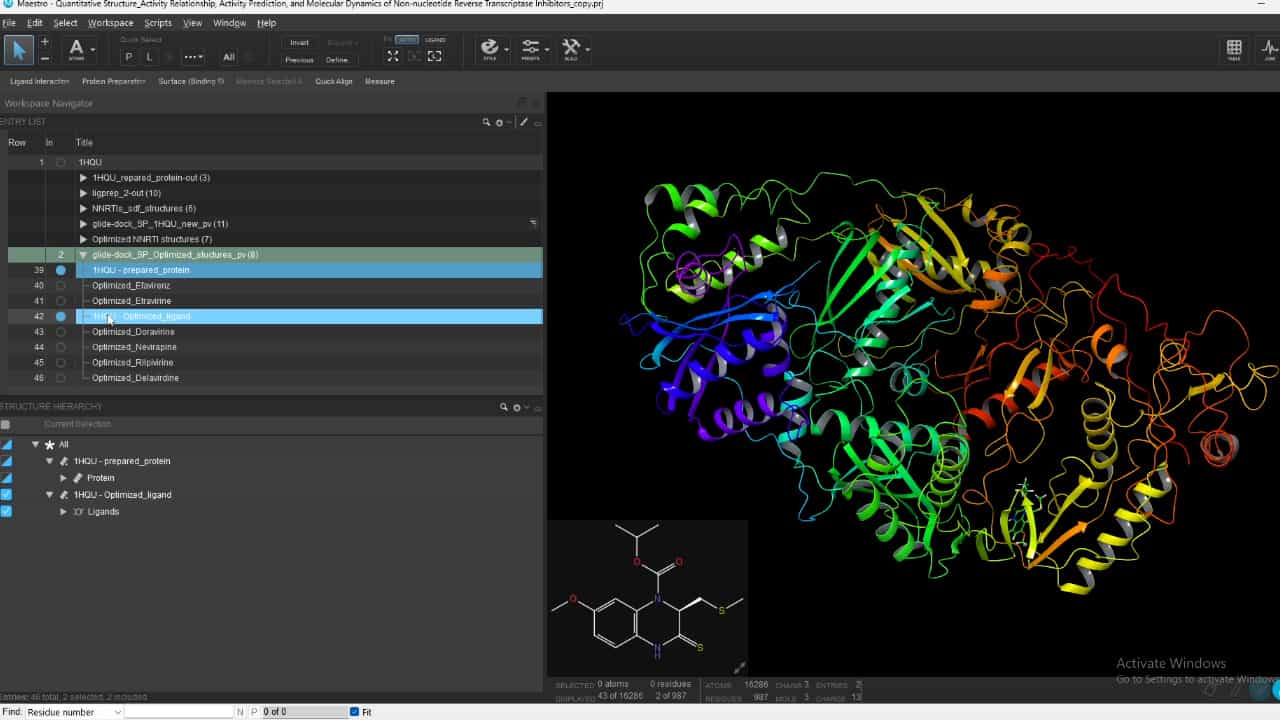
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
1Bristol-Myers Squibb, Pharmaceutical Research Institute, Wallingford, Connecticut 06492.

Abstract:
This chapter has focused on the application of molecular dynamics computer simulations and related molecular modeling techniques to the study of HIV protease structure and structure-function relationships. The abundance of crystallographic data provides ample experimental quantities (average structures, temperature factors, and hydrogen bond topography) to validate the computational techniques employed. Furthermore, these studies provide insight into the structure and functional energetics of HIV-1 protease that would be difficult or impossible to study experimentally. This chapter covers studies that investigate correlated motion between and within subunits of the protease, mutants of the protease that disrupt the tertiary structure and dimer formation, and studies of HIV-1 protease-inhibitor complexes that rationalize both the protonation state of the active site and the observed binding strength of these complexes. These studies demonstrate that MD is capable of contributing to our understanding of structure-function relationships and may aid in the design of potential therapeutics.