Related Experiment Video
Updated: Aug 19, 2026
Evaluation of Drug Sorption to PVC- and Non-PVC-based Tubes in Administration Sets Using a Pump
Published on: March 11, 2017
Pharmacokinetics and plasma binding of diazepam in man, dog, rabbit, guinea pig and rat

Abstract:
The pharmacokinetics and plasma binding of diazepam were compared in man, dog, rabbit, guinea pig and rat. Diazepam (D) and its major metabolite, desmethyl-diazepam, were measured in blood and plasma by a specific and sensitive gas-liquid chromatography assay with an electron capture detector. After an intravenous bolus injection plasma levels of D declined biexponentially in all species examined and the data were analyzed according to the two-compartment open model. The binding of D and desmethyldiazepam has been determined at therapeutic concentrations by equilibrium dialysis in man (96.8 and 96.6%, respectively), dog (96.0 and 95.9%), rabbit (89.9 and 94.7%), guinea pig (91.3 and 78.6%) and rat (86.3 and 90.5%). In man, the elimination half-life, T1/2(beta), increased significantly (P less than .01) with decreasing total plasma clearance (Cl). Plasma binding affected Vd and Cl, but only Cl increased significantly (P less than .05), if more free D was available. This indicates that unbound drug is rate-determining for clearance by the liver, and that D fits into the restrictive elimination class in man. In the four animal species tested, Cl was a direct linear function of the body surface area. T1/2(beta) and the rates of drug clearance were characteristic figures for each species: from 1.1 hours and 81.6 ml/min/kg in the rat to 32.9 hours and 0.35 ml/min/kg in man, whereas T1/2(alpha), the half-life of distribution, varied only approximately 3-fold (0.3-1.0 hours) in the different species. A considerably higher extraction ratio than the unbound fraction of diazepam exists in these animal species, and blood clearance exceeds liver blood flow, giving reason to assume a much higher ability of the liver to metabolize D, and a species-dependent extrahepatic metabolism. The large variations described suggest that pharmacokinetic data or plasma binding results cannot simply be extrapolated to man.
More Related Videos
 10:16
Autoradiography as a Simple and Powerful Method for Visualization and Characterization of Pharmacological Targets
Published on: March 12, 2019
 08:28
Microsurgical Skills of Establishing Permanent Jugular Vein Cannulation in Rats for Serial Blood Sampling of Orally Administered Drug
Published on: December 14, 2021
Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:30
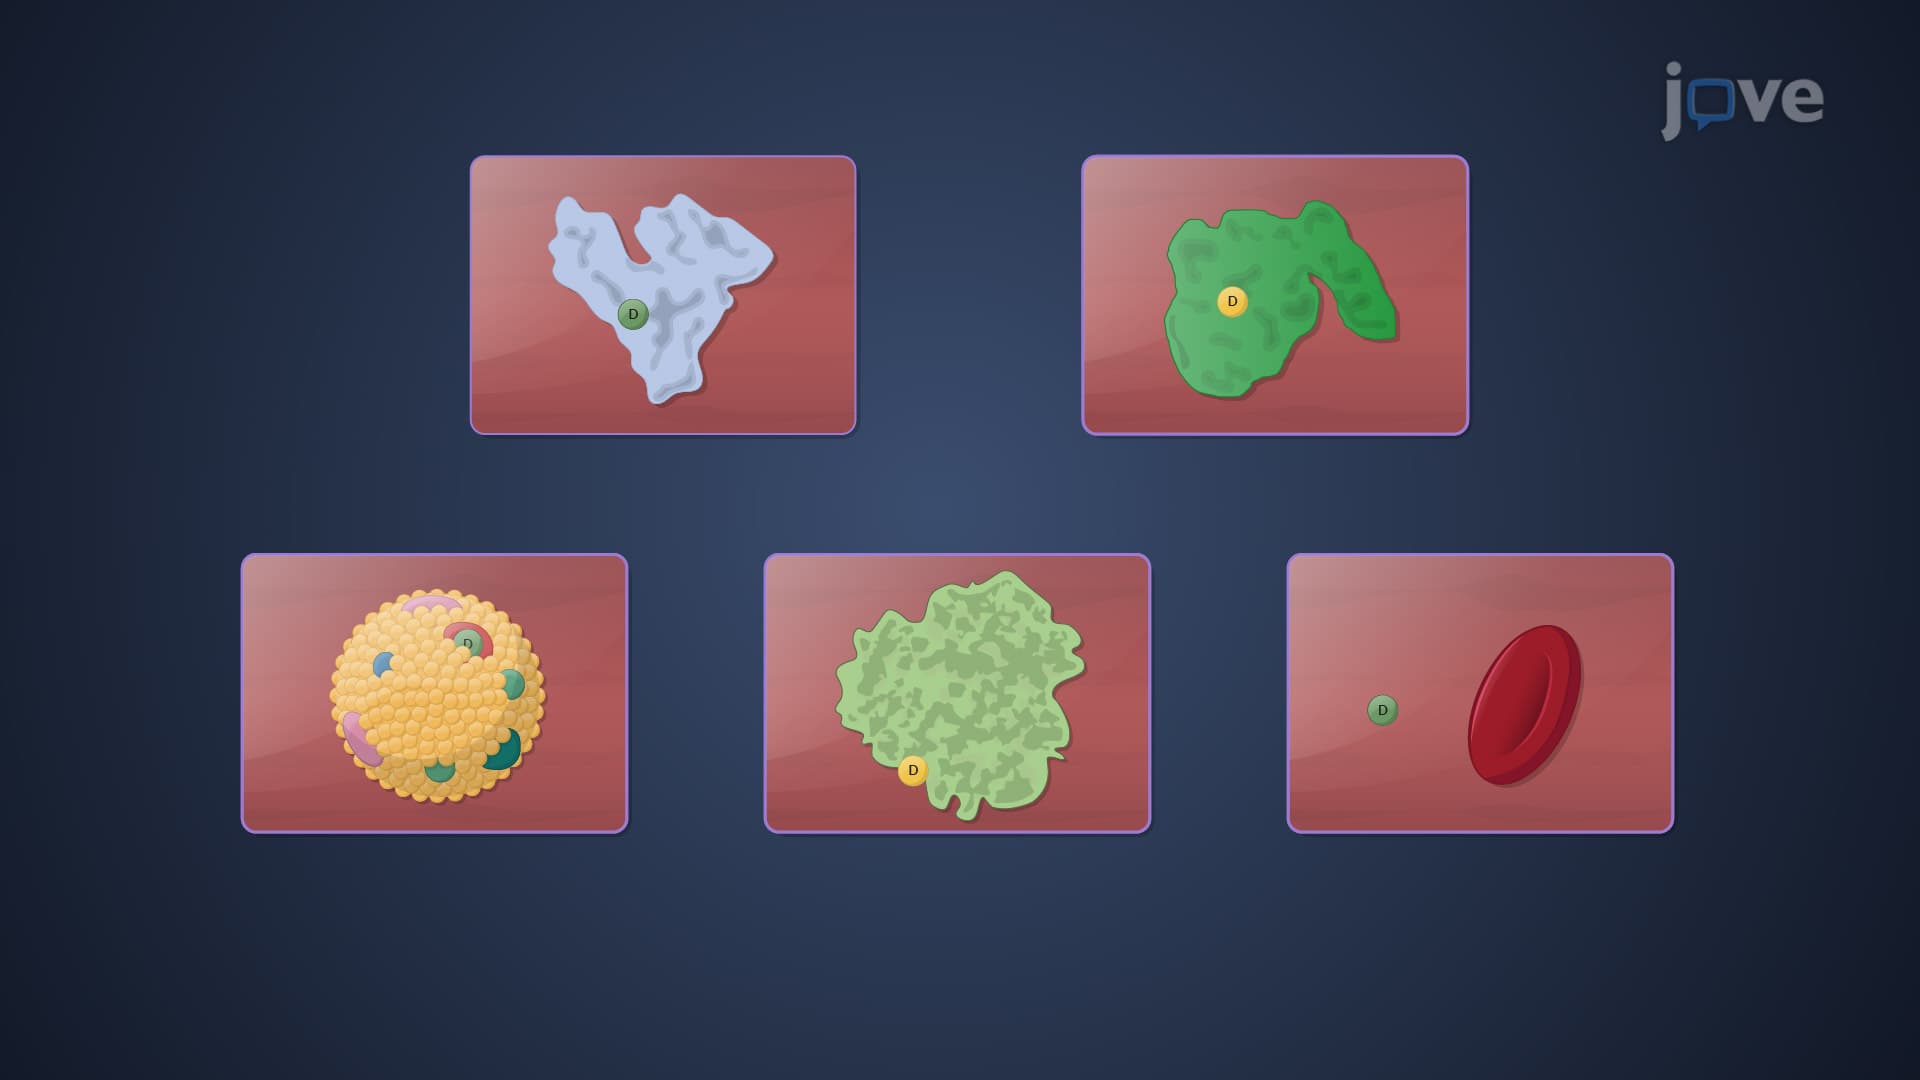
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:09
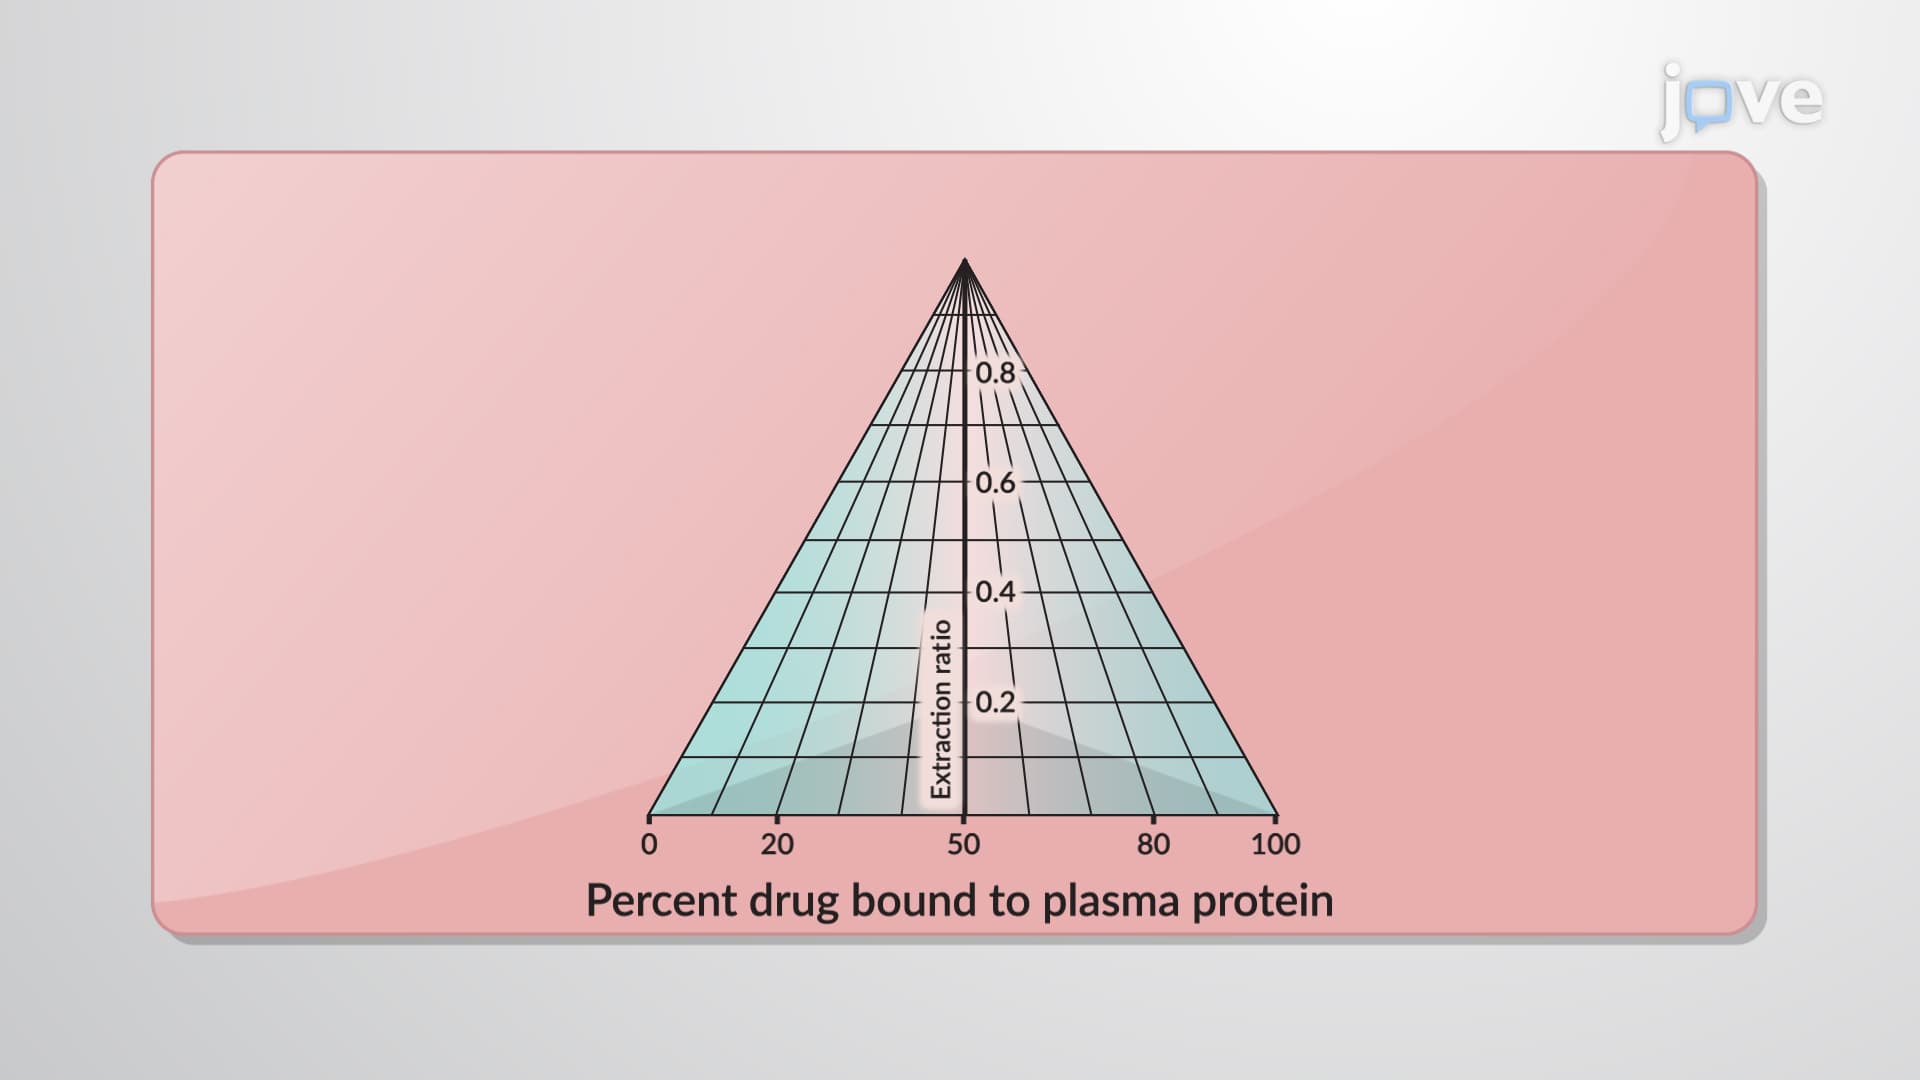
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship
 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems