Related Experiment Video
Updated: Aug 19, 2026
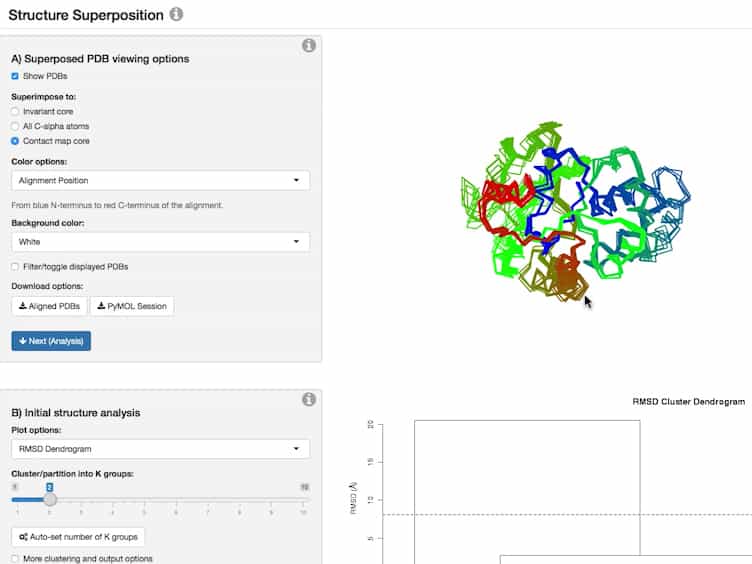

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Backbone dynamics of a highly disordered 131 residue fragment of staphylococcal nuclease
1Department of Biological Chemistry, Johns Hopkins University School of Medicine, Baltimore, MD 21205.

Abstract:
In order to characterize the dynamic properties of the denatured state of staphylococcal nuclease, R1, R2, and NOE relaxation parameters have been measured for the backbone 15N nuclei of a 131 residue fragment that serves as a model of the denatured state under non-denaturing conditions. The relaxation data indicate a wide range of amplitudes for segmental motion and are inconsistent with a random coil conformation. An optimal value of 7.8 ns was obtained for the molecular rotational correlation time tau m based on the analysis of the 79 residues for which R1, R2, and NOE relaxation data could be obtained. This value corresponds roughly to the slowest detectable motion on the nanosecond time scale and is of a magnitude consistent with global tumbling of a large portion of the molecule. For the majority of residues, experimental data could be described most adequately in terms of a modified "model-free" formalism which includes contributions from internal motions on both an intermediate (tau e) and a fast time scale (tau f) in the context of slow overall tumbling (tau m). The generalized order parameters S2, which gives the amplitude of motions on time scales faster than tau m, correlates with sequence hydrophobicity and suggests a relationship between chain flexibility and sequence propensity for hydrophobic collapse. The fractional populations of three alpha-helices in the protein show a stronger correlation with S2 values and hydrophobicities than with intrinsic helix propensities. These observations suggest that secondary structure may be preferentially stabilized in hydrophobic segments of the sequence.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:54
Nucleosome Remodeling
Nucleosome remodeling complex
Eukaryotic cells have specialized enzymes called ATP-dependent nucleosome remodeling enzymes. These enzymes...
 00:55
DNA Helicases
 02:31
Homologous Recombination
 02:37
Restarting Stalled Replication Forks
 01:03
Single-Strand DNA Binding Proteins