Dynamic structure of human lysozyme derived from X-ray crystallography: normal mode refinement
Related Experiment Video
Updated: Aug 14, 2026
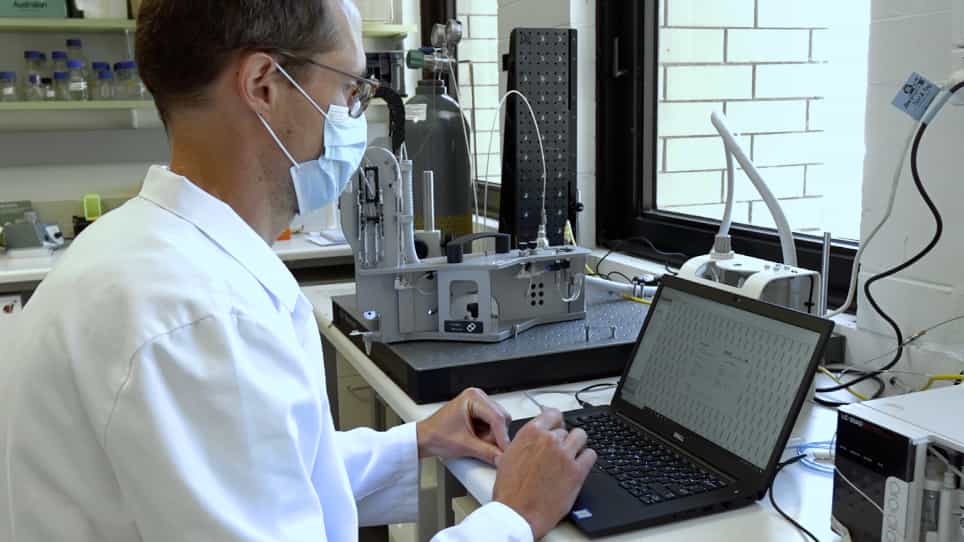
Lipidico Injection Protocol for Serial Crystallography Measurements at the Australian Synchrotron
Published on: September 23, 2020
1Protein Engineering Research Institute, Osaka, Japan.

Abstract:
X-ray crystallography provides a wealth of information about the dynamic as well as static protein structure. A new method of dynamic structure refinement of protein X-ray crystallography, normal mode refinement, is proposed. In this method, the Debye-Waller factor is expanded in terms of the low-frequency internal normal modes and external normal modes, whose amplitudes and couplings are optimized in the process of crystallographic refinement. The internal and external contributions to the atomic fluctuations can be separated. Also, anisotropic atomic fluctuations and their inter-atomic correlations can be determined experimentally even with a relatively small number of adjustable parameters. The method is applied to the analyses of experimental data of human lysozyme and its mutant, C77A/C95A, to reveal its dynamic structure.
More Related Videos

Related Concept Videos
 01:22
Protein Folding
 02:18
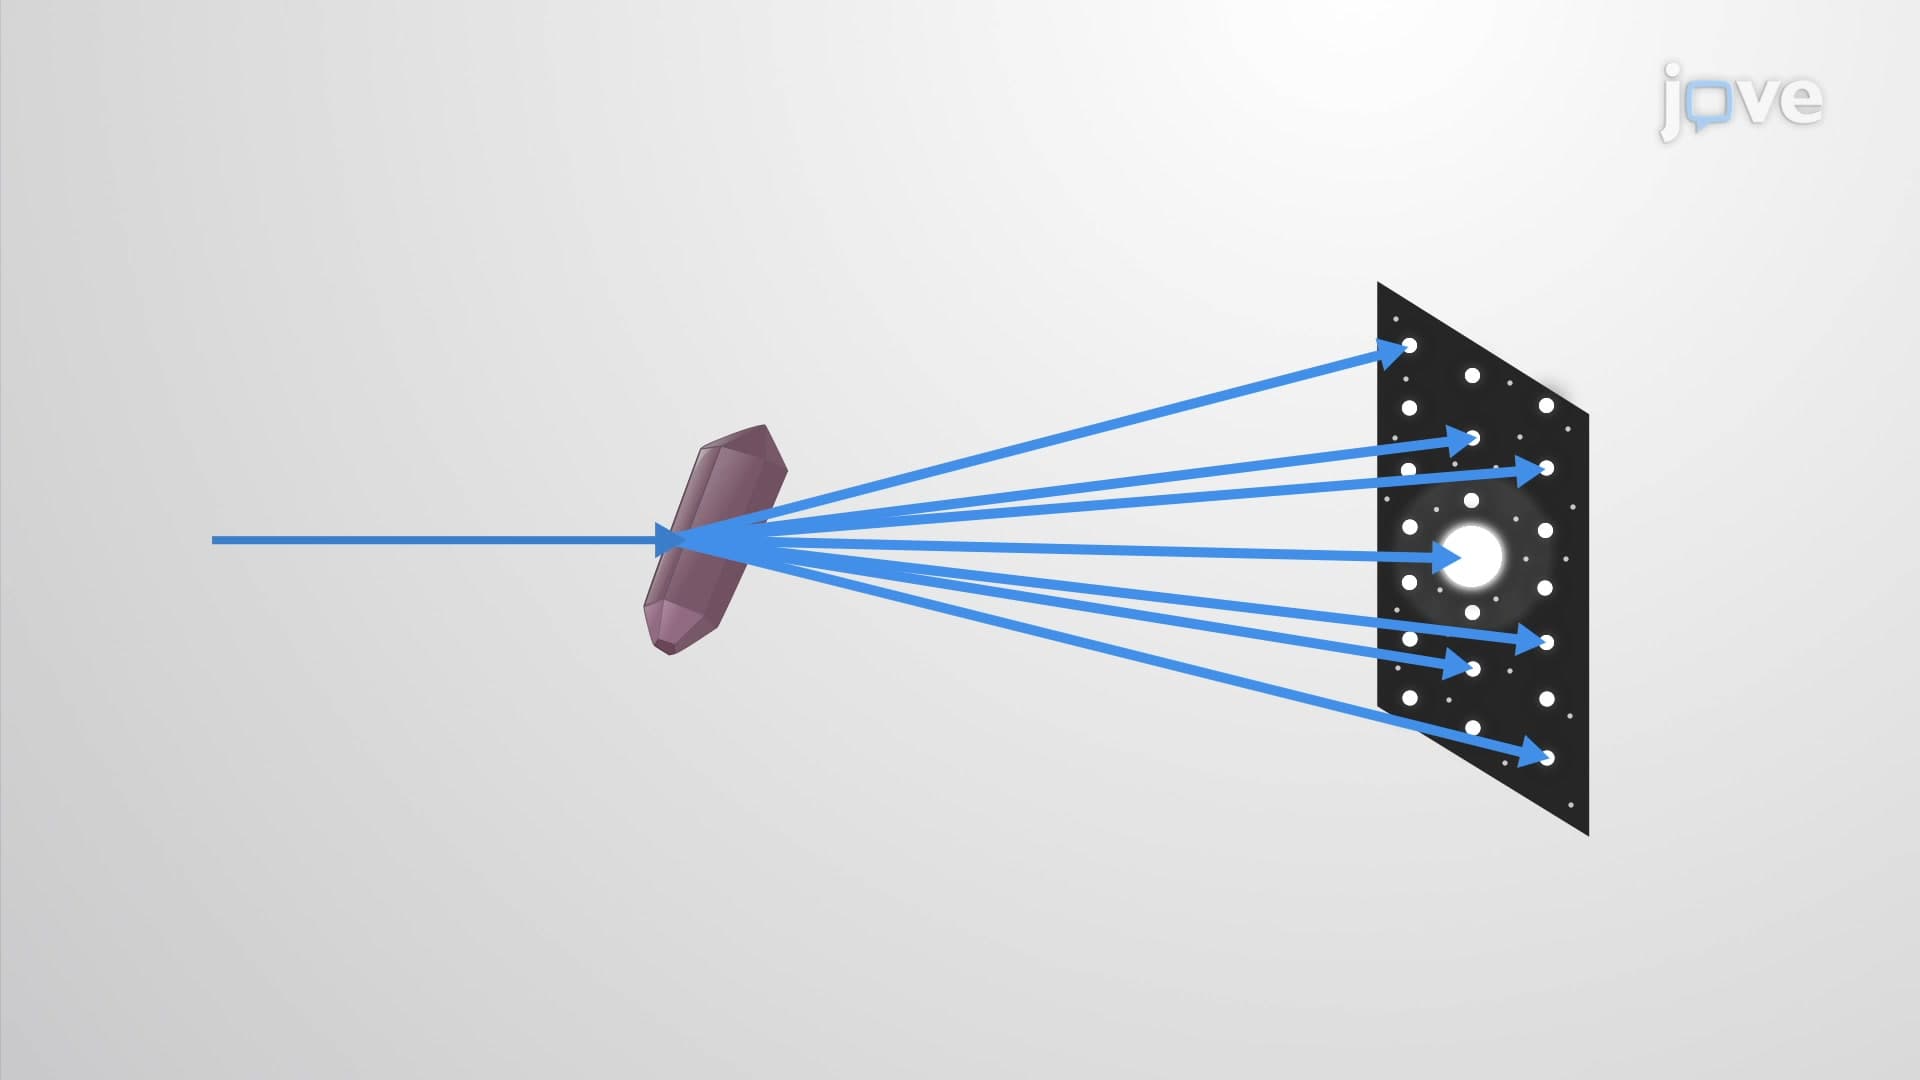
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:10
X-ray Diffraction of Biological Samples
According to Bragg's law, when X-rays strike the sample positioned on a stage, the rays are scattered by the electron clouds around the sample atoms. The X-ray diffraction or scattering is caused by constructive interference of the X-ray waves that reflect off the internal crystal...
 01:29
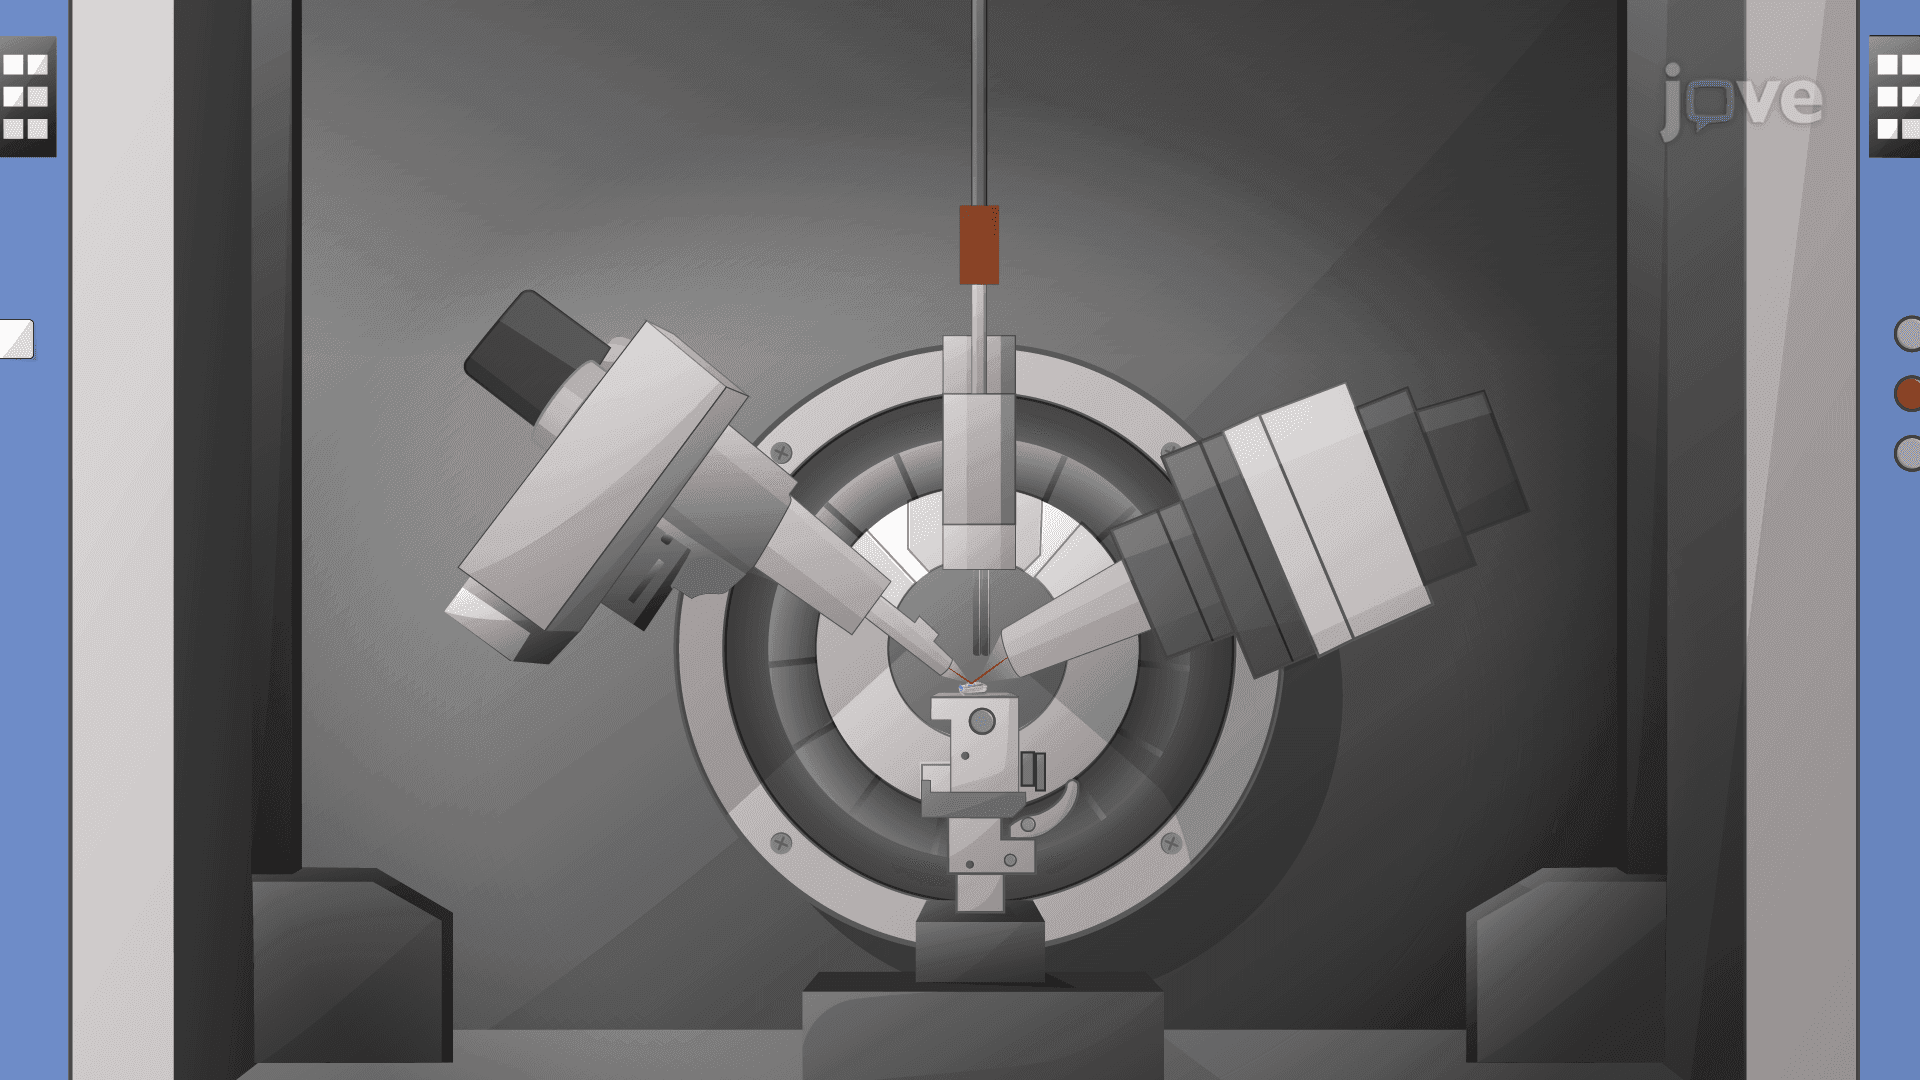
Determination of Crystal Structures