Related Experiment Video
Updated: Aug 5, 2026
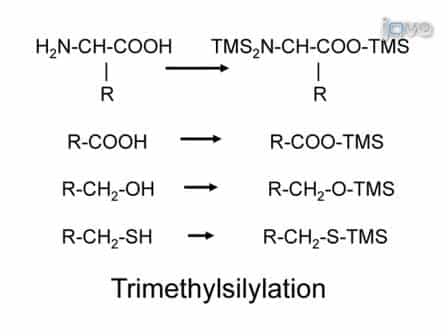
09:28

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Prenatal diagnosis of inherited metabolic diseases
1Department of Obstetrics, Gynecology and Reproductive Sciences, UCSF School of Medicine 94143-0132.
The Western Journal of Medicine
|September 1, 1993

Abstract:
Advances in the prenatal diagnosis of inherited metabolic disease have provided new reproductive options to at-risk couples. These advances have occurred in both sampling techniques and methods of analysis. In this review we present an overview of the currently available prenatal diagnostic approaches for the diagnosis of metabolic disease in a fetus.
Related Concept Videos
 01:35
Pedigree Analysis
Overview
 02:59
Animal Mitochondrial Genetics
Among all the organelles in an animal cell, only mitochondria have their own independent genomes. Animal mitochondrial DNA is a double-stranded, closed-circular molecule with around 20,000 base pairs. Mitochondrial DNA is unique in that one of its two strands, the heavy, or H, -strand is guanine rich, whereas the complementary strand is cytosine rich and called the light, or L, -strand. Compared to nuclear DNA, mitochondrial DNA has a very low percentage of non-coding regions and is marked by...
 02:30
Genomic Imprinting and Inheritance
Diploid organisms inherit genetic material through chromosomes from both parents. Copies of the same gene are known as alleles. In most cases, both alleles are simultaneously expressed and allow various cellular processes to function optimally. If one of the alleles is missing or mutated, the expression of the other allele can compensate; however, this is not true for all genes.
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 01:21
Overview of Protein Metabolism
Proteins are broken down into amino acids during digestion. Unlike fats and carbohydrates, which are stored for later use, proteins are not. Instead, amino acids are either used to produce ATP through oxidation or contribute to the creation of new proteins for the growth and repair of the body. Any surplus amino acids from the diet are converted into glucose or triglycerides rather than excreted.
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...