Molecular genetics of cardiomyopathies
Related Experiment Video
Updated: Nov 7, 2026
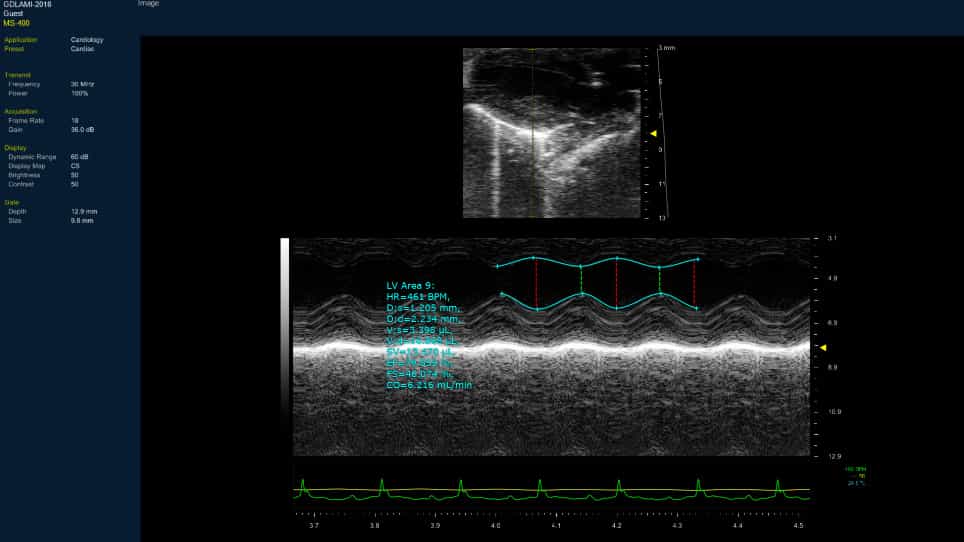
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
1Baylor College of Medicine.

Insights
Familial hypertrophic cardiomyopathy (FHCM) involves genetic mutations, primarily in the beta-myosin heavy chain (beta MHC) gene, influencing disease severity. Ongoing research identifies new genetic loci for FHCM and related cardiomyopathies.
Area of Science:
- Cardiovascular Genetics
- Molecular Cardiology
- Genetic Diseases
Background:
- Familial hypertrophic cardiomyopathy (FHCM) is a genetically heterogeneous disorder with autosomal dominant inheritance and variable penetrance.
- Mutations in the beta-myosin heavy chain (beta MHC) gene are the most common cause, affecting protein structure and function.
- Other genetic loci for FHCM have been identified, indicating complex genetic underpinnings.
Purpose of the Study:
- To review the genetic basis of familial hypertrophic cardiomyopathy (FHCM).
- To discuss the clinical correlations of specific beta MHC gene mutations.
- To highlight the genetic causes of related cardiomyopathies like Duchenne/Becker muscular dystrophy and myotonic dystrophy.
Main Methods:
- Literature review of genetic mutations associated with FHCM.
- Analysis of genotype-phenotype correlations for beta MHC gene mutations.
- Summary of genetic findings in Duchenne/Becker muscular dystrophy and myotonic dystrophy.
Main Results:
- Multiple mutations in the beta MHC gene co-segregate with FHCM, with varying clinical outcomes.
- A missense mutation in exon 13 of the beta MHC gene is linked to severe disease and sudden cardiac death.
- Genetic defects in dystrophin (DMD) and myotonin protein kinase (myotonic dystrophy) cause cardiac complications.
Conclusions:
- The beta MHC gene is a key player in FHCM, with specific mutations dictating disease severity.
- Genetic factors beyond the beta MHC gene contribute to FHCM, with new loci being discovered.
- Understanding the genetic basis of cardiomyopathies is crucial for diagnosis and management.
Abstract:
Familial hypertrophic cardiomyopathy (FHCM) is a heterogeneous disease with an autosomal dominant Mendelian inheritance and variable penetrance. Several mutations in the beta-myosin heavy chain (beta MHC) gene, the first gene identified for this disease, have been described that co-segregate with the inheritance of the disease. All the mutations in the beta MHC gene encode for the globular head of the myosin protein except for the deletion mutation which encodes for the carboxy-terminus (rod) of the protein. The clinical features associated with some of the mutations in the beta MHC gene have been characterized. A missense mutation in exon 13 of the beta MHC gene, is associated with a higher incidence of sudden cardiac death and severe form of the disease, while some others are associated with a more benign form of the disease. Recently, three other loci, on chromosomes 1q3, 11q11 and 15q2, for FHCM have been identified and research is ongoing to identify the candidate genes. Cardiac involvement in Duchenne/Becker muscular dystrophy (DMD), and myotonic dystrophy is common. Heart failure due to dilated cardiomyopathy and sudden cardiac death are the common causes of death in these disorders. The genes responsible for DMD and myotonic dystrophy are dystrophin and myotonin protein kinase genes located on chromosomes X and 19 respectively. The disease in DMD is due to deletion mutations in the dystrophin gene, while myotonic dystrophy is due to expansion of the GCT trinucleotide repeats in the myotonin-protein kinase gene. Familial dilated cardiomyopathy comprises 20% of cases of idiopathic dilated cardiomyopathy.(ABSTRACT TRUNCATED AT 250 WORDS)
Related Concept Videos
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:25
Cardiomyopathy I: Introduction and Classification
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy
 01:29
Cardiomyopathy V: Interprofessional Care
 02:59
Animal Mitochondrial Genetics