Related Experiment Video
Updated: Aug 23, 2026
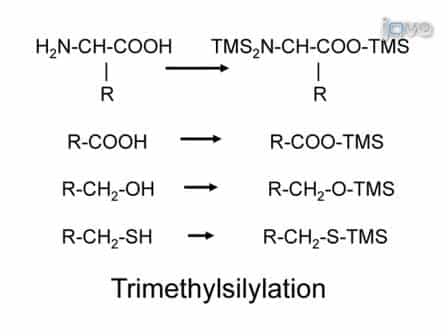
One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
An approach to the diagnosis of overwhelming metabolic disease in early infancy

Insights
Diagnosing rare metabolic diseases in newborns is crucial. This study outlines a systematic approach using clinical clues and laboratory tests for early detection of inborn errors of metabolism.
Area of Science:
- Pediatrics
- Medical Genetics
- Biochemistry
Background:
- Heritable metabolic diseases cause severe illness in infants.
- Many infants with metabolic disorders remain undiagnosed.
- New, unidentified inborn errors of metabolism are suspected in critically ill newborns.
Purpose of the Study:
- To develop a systematic diagnostic approach for critically ill newborns suspected of having metabolic disease.
- To identify early diagnostic clues for metabolic disorders in neonates.
Main Methods:
- Clinical observation for suggestive signs of disordered metabolism.
- Laboratory investigations to confirm clinical suspicions.
- Systematic screening and advanced organic analysis for definitive diagnosis.
Main Results:
- Clinical clues can indicate potential metabolic derangements.
- Laboratory tests are valuable for initial screening and confirmation.
- Diagnostic complexity increases with advanced analytical techniques.
Conclusions:
- A systematic approach is essential for diagnosing metabolic diseases in sick newborns.
- Early diagnosis of inborn errors of metabolism can be achieved through integrated clinical and laboratory methods.
- Further research into novel metabolic pathways is warranted.
Abstract:
Heritable metabolic disease is a significant cause of overwhelming illness in the very young infant. It appears that most patients with well recognized disorders are not being diagnosed, and it is our conviction that there are new, as yet unidentified, inborn errors of metabolism in this population of patients. We have attempted to develop a systematic approach to the seriously ill newborn as a candidate for an early diagnosis of metabolic disease. There are some clinical clues that suggest the presence of disordered metabolism. The laboratory can be useful in confirming initial clinical suspicions and in screening for the presence of abnormality. The complexity of laboratory evaluation increases as one proceeds to definitive diagnosis and modern organic analysis.
Related Concept Videos
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
 01:15
Diabetes: Symptoms, Diagnosis, and Complications
 01:26
Diabetes Mellitus: Introduction
 01:25
Diabetic Ketoacidosis l: Introduction
 01:22
Diabetic Ketoacidosis ll: Pathophysiology