Related Experiment Video
Updated: Sep 6, 2026

Assessment of Immunologically Relevant Dynamic Tertiary Structural Features of the HIV-1 V3 Loop Crown R2 Sequence by ab initio Folding
Published on: September 16, 2010
Determination of the conformation of folding initiation sites in proteins by computer simulation
1Center for Advanced Research in Biotechnology, University of Maryland Biotechnology Institute, Rockville 20850, USA.

Abstract:
Experimental evidence and theoretical models both suggest that protein folding begins by specific short regions of the polypeptide chain intermittently assuming conformations close to their final ones. The independent folding properties and small size of these folding initiation sites make them suitable subjects for computational methods aimed at deriving structure from sequence. We have used a torsion space Monte Carlo procedure together with an all-atom free energy function to investigate the folding of a set of such sites. The free energy function is derived by a potential of mean force analysis of experimental protein structures. The most important contributions to the total free energy are the local main chain electrostatics, main chain hydrogen bonds, and the burial of nonpolar area. Six proposed independent folding units and four control peptides 11-14 residues long have been investigated. Thirty Monte Carlo simulations were performed on each peptide, starting from different random conformations. Five of the six folding units adopted conformations close to the experimental ones in some of the runs. None of the controls did so, as expected. The generated conformations which are close to the experimental ones have among the lowest free energies encountered, although some less native like low free energy conformations were also found. The effectiveness of the method on these peptides, which have a wide variety of experimental conformations, is encouraging in two ways: First, it provides independent evidence that these regions of the sequences are able to adopt native like conformations early in folding, and therefore are most probably key components of the folding pathways. Second, it demonstrates that available simulation methods and free energy functions are able to produce reasonably accurate structures. Extensions of the methods to the folding of larger portions of proteins are suggested.

Related Concept Videos
 01:28
Protein Organization
 01:49
Protein Folding
01:49Protein Folding
 03:00
Molecular Chaperones and Protein Folding
The...
 01:24
Protein Organization
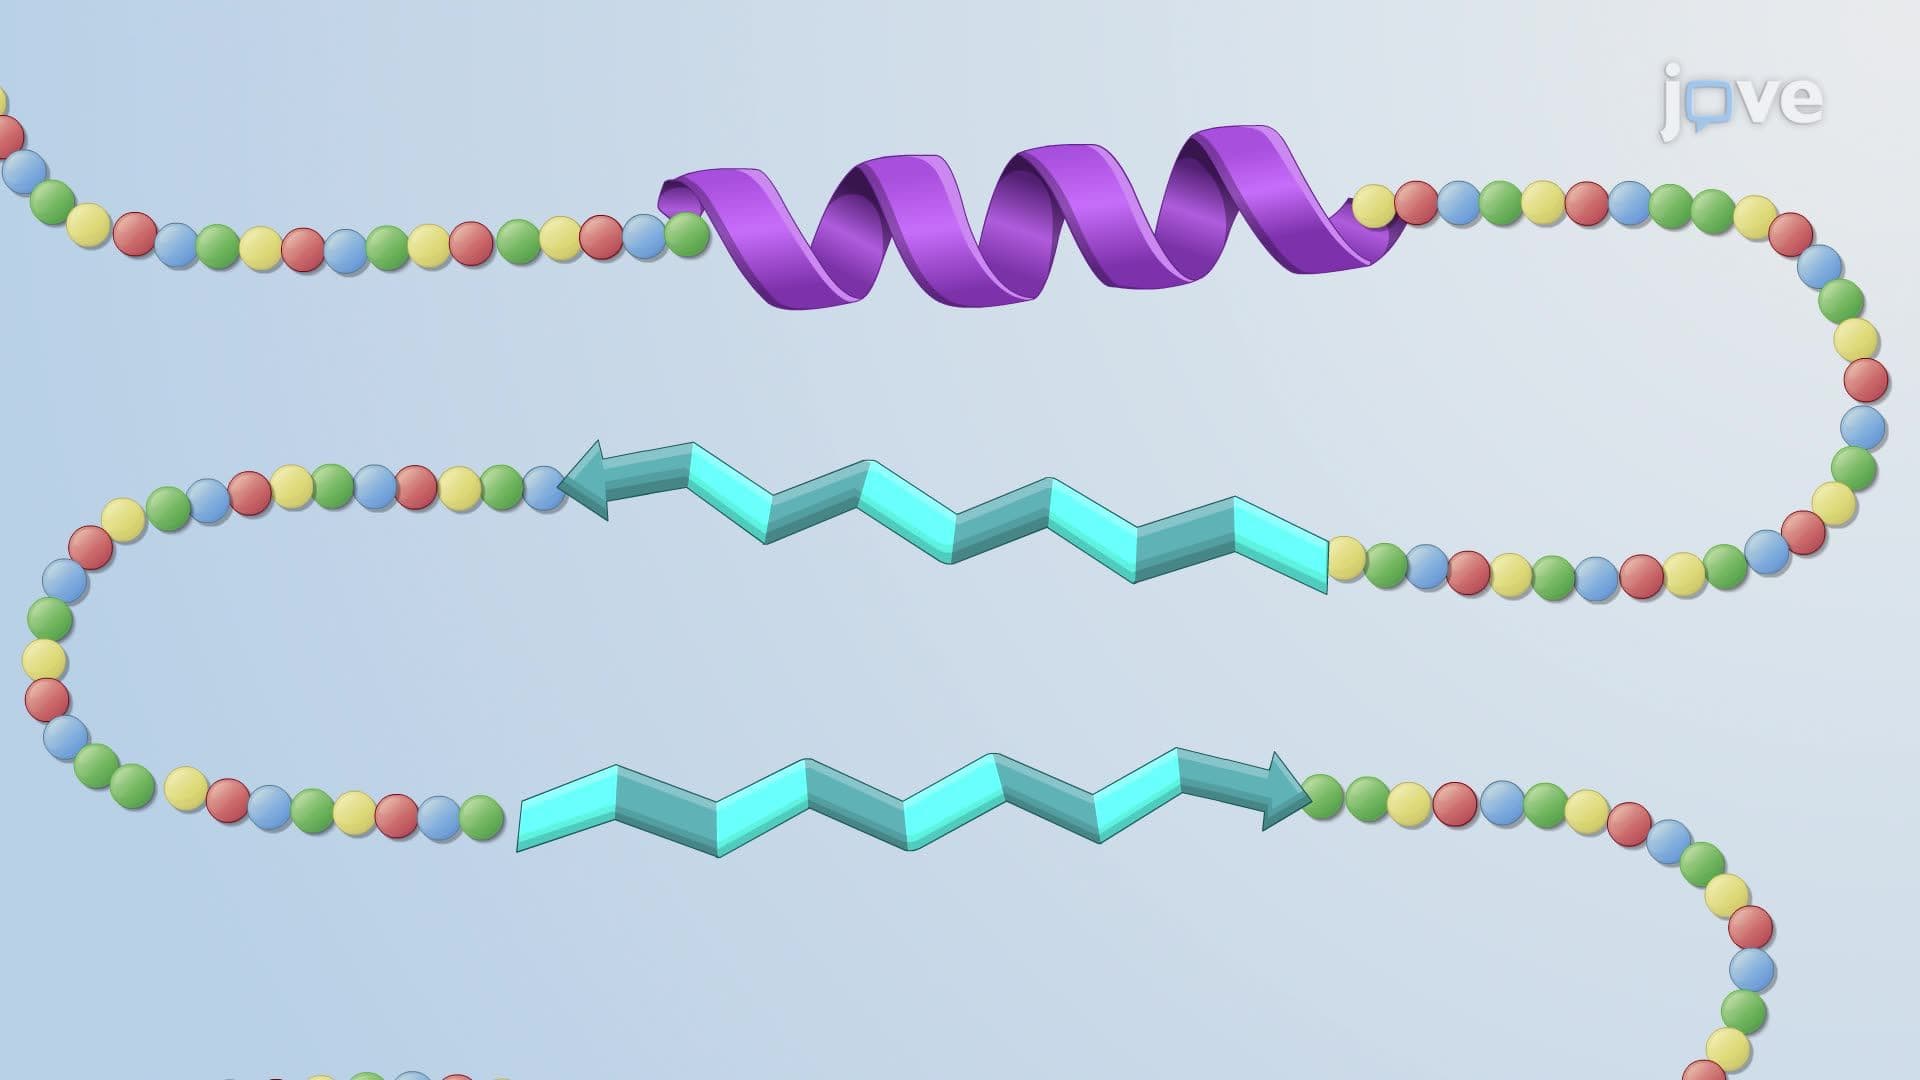
The primary structure of a protein is its amino acid sequence.
 01:25
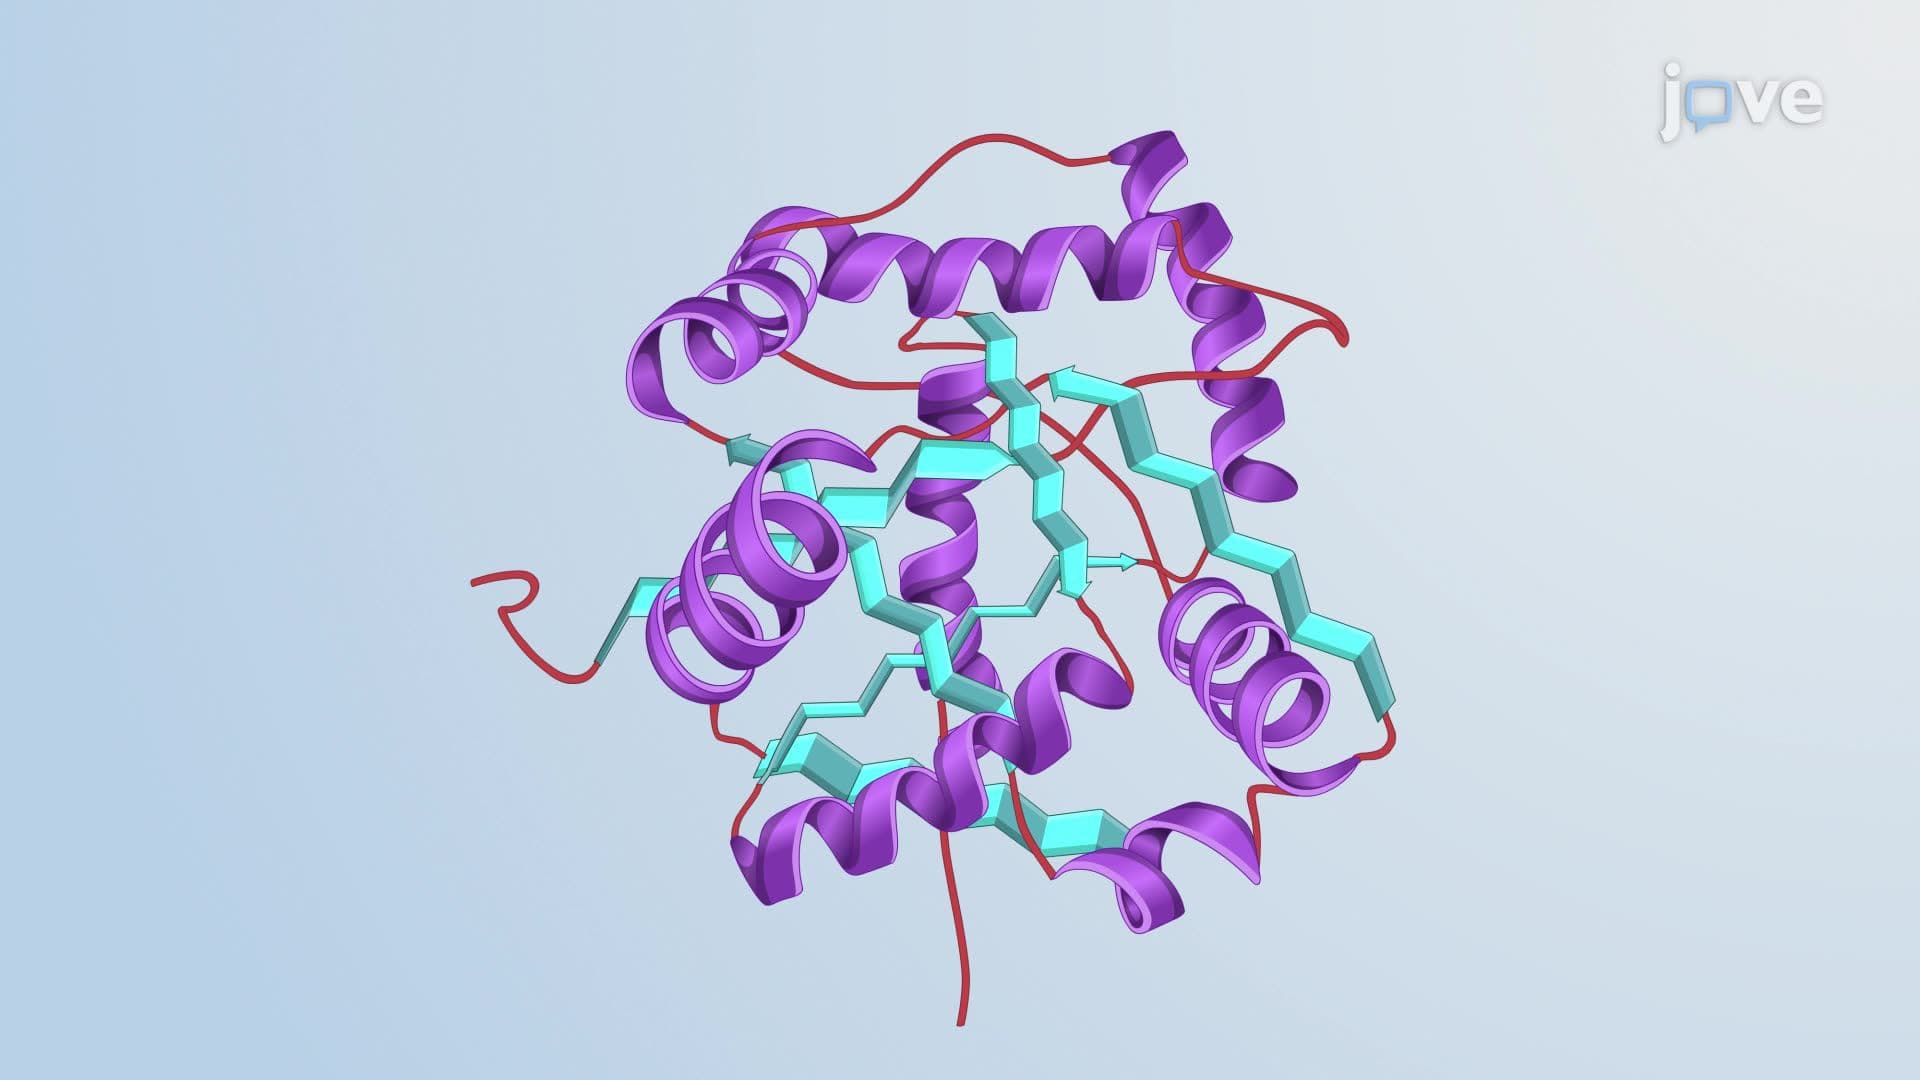
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...