Related Experiment Video
Updated: Aug 12, 2026

Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
A proposed common spatial pharmacophore and the corresponding active conformations of some peptide leukotriene
1Department of Chemical Technology, University of Bombay, Matunga, India.

Abstract:
Molecular modeling studies were carried out by a combined use of conformational analysis and 3D-QSAR methods of identify molecular features common to a series of hydroxyacetophenone (HAP) and non-hydroxyacetophenone (non-HAP) peptide leukotriene (pLT) receptor antagonists. In attempts to develop a ligand-binding model for the pLT receptor, the Apex-3D program was used to identify biophoric structural patterns that are common to 13 diverse sets of compounds showing different levels of biological activity. A systematic conformational analysis was carried out to obtain sterically accessible conformations for these flexible compounds. Apex-3D was then utilized to propose common biophoric regions based on the selection of one of several conformations (MOPAC-minimized AM1) from each compound's data set that best fits the biophoric pattern and the resulting superimposition with all the other data-set compounds. Apex-3D identified three common biophoric features important for activity: one as the hydroxyl, acetyl, carbonyl and carboxyl groups, which mimic the acid-binding region of an agonist, the other as the hydrogen-bond donating site, and the third part is represented by a plane in which lipophilic aromatic groups align. The structure-activity relationships were then assessed by using the 3D-QSAR model. A common biophore model is proposed from the Apex-3D analysis which may be useful in designing new pLT antagonists. Molecular volumes and electrostatic potential similarities were also calculated in order to obtain the important structural requirements for the activity.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:29
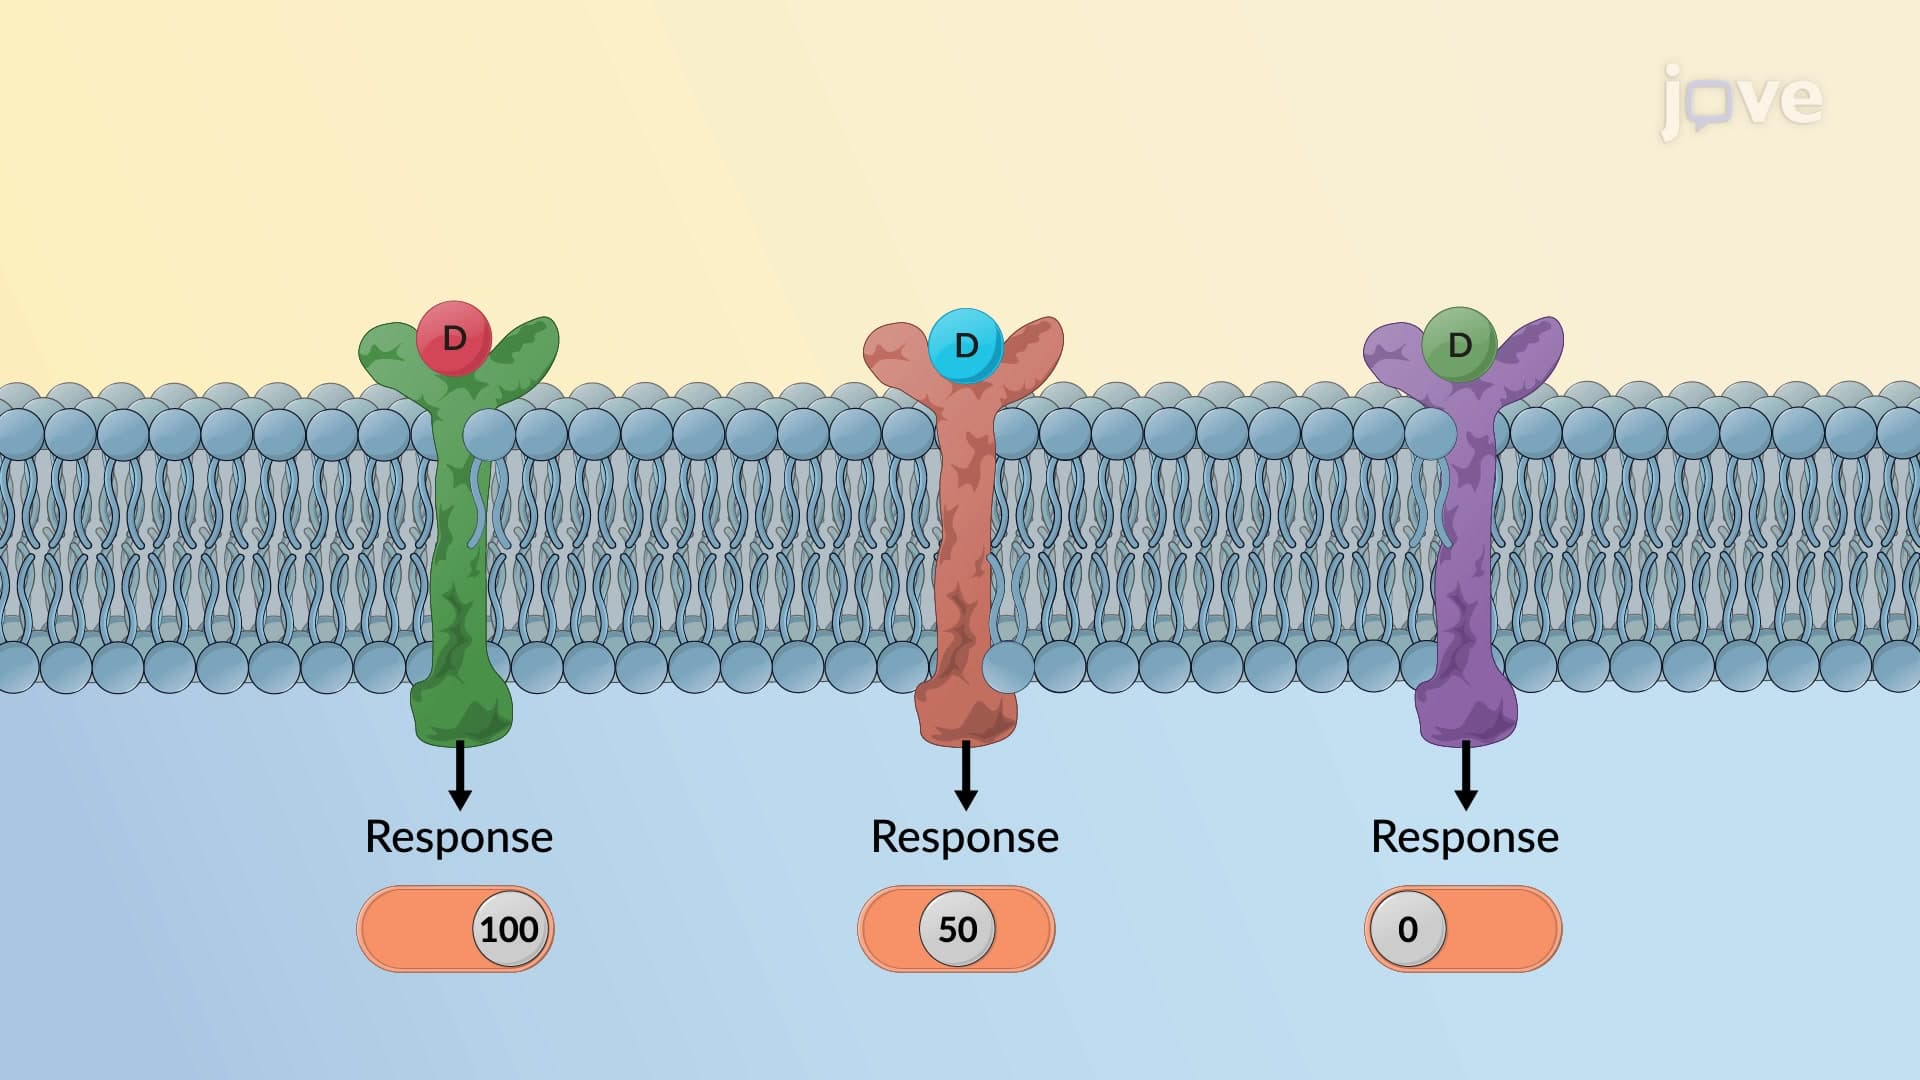
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:25
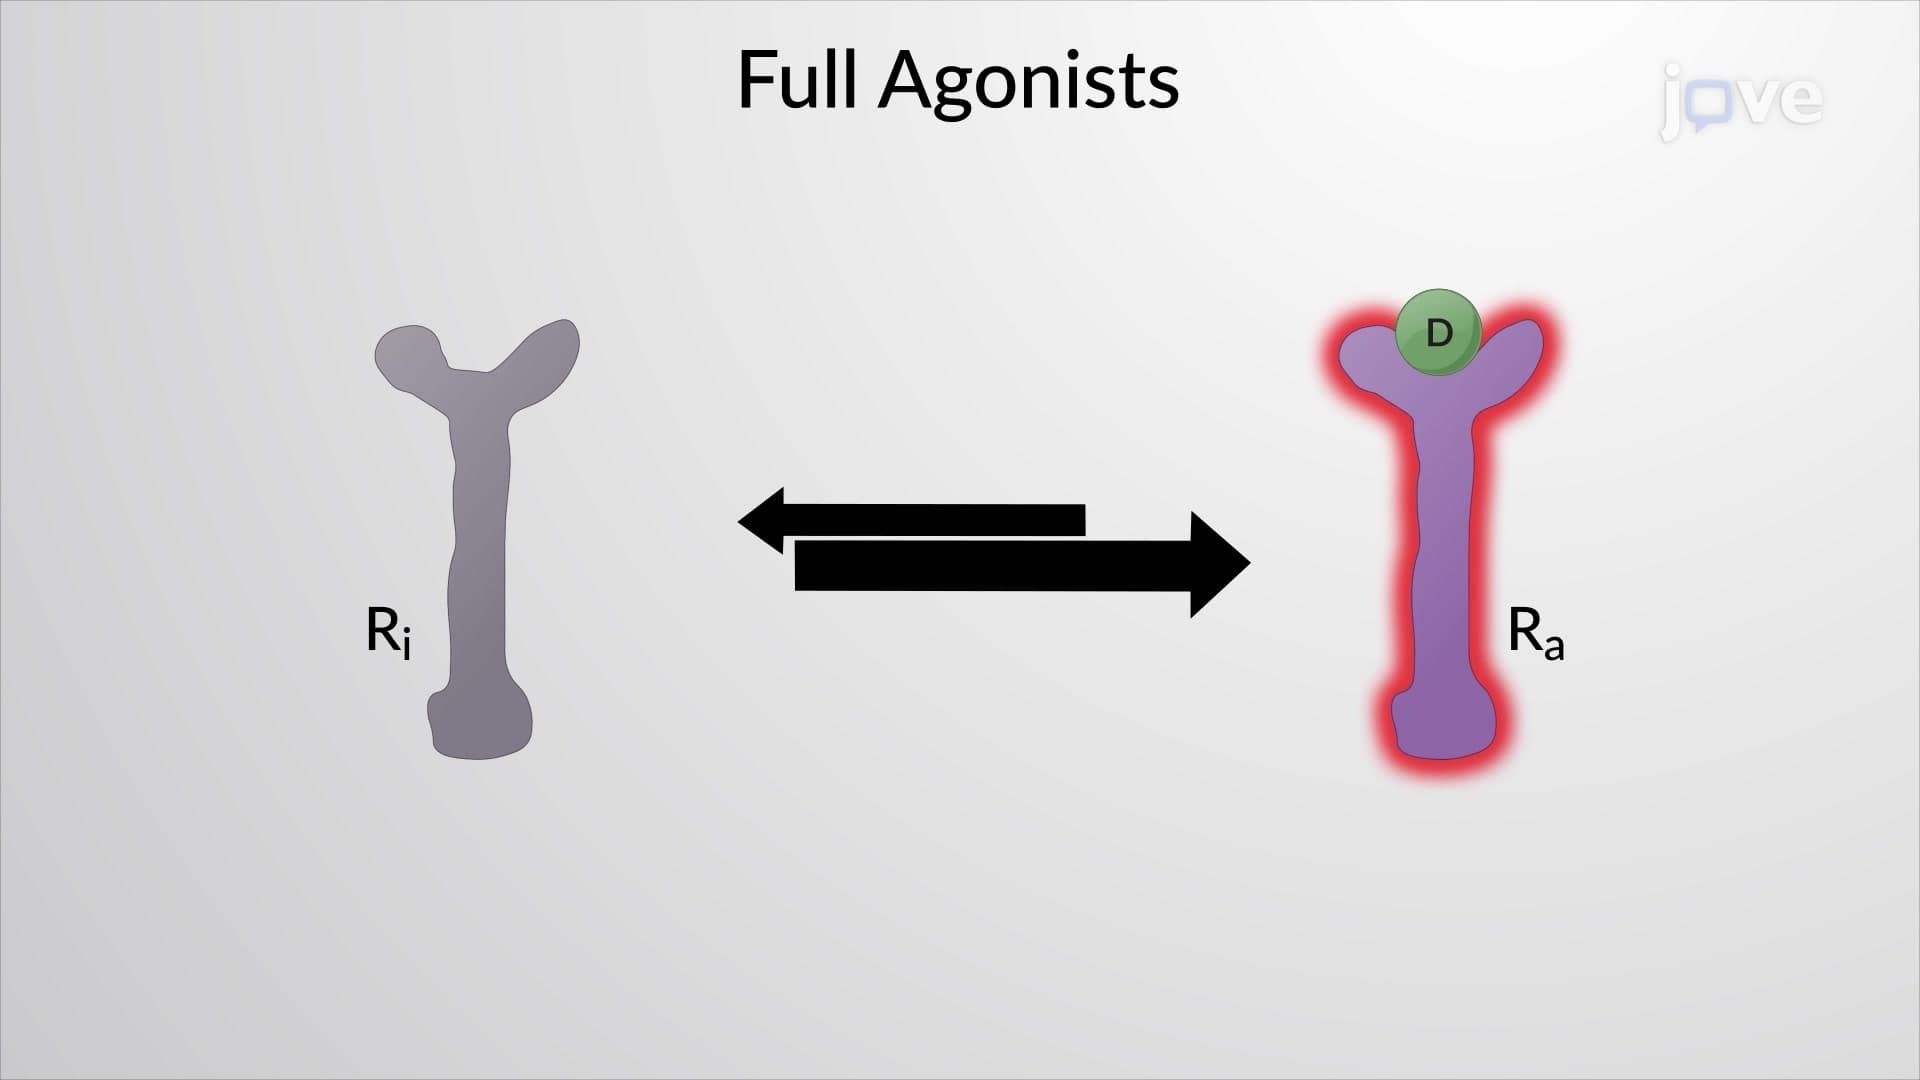
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...