Related Experiment Video
Updated: Aug 16, 2026
Microsurgical Skills of Establishing Permanent Jugular Vein Cannulation in Rats for Serial Blood Sampling of Orally Administered Drug
Published on: December 14, 2021
Enhanced inhibition of microsomal cytochrome P450 3A2 in rat liver during diltiazem biotransformation
1Storr Liver Unit, Department of Medicine, University of Sydney, Westmead Hospital, New South Wales, Australia.

Abstract:
Pharmacokinetic drug interactions involving the calcium channel blocker diltiazem (DTZ) have been attributed to inhibition of microsomal cytochrome P450 (P450)-mediated drug oxidation. Accumulation of certain DTZ metabolites during dosage with the drug, as well as dose-related differences in DTZ pharmacokinetics, suggests that DTZ metabolites may also participate in P450 inhibition. The present study evaluated a series of putative DTZ metabolites as inhibitors of major constitutive P450s in rat liver in vitro, in relation to DTZ biotransformation. The principal finding to emerge was that the N-demethylated metabolite of DTZ was a more potent competitive inhibitor than DTZ of CYP3A2-dependent testosterone 6 beta-hydroxylation. This P450 appeared to be the preferred target for inhibition, because the observed K/K(m) ratio for inhibition of CYP3A2-dependent steroid hydroxylation was approximately 4- and 100-fold lower than those for CYP2C11 and CYP2A1-dependent pathways, respectively. It was also established that N-desmethyl-DTZ was a major metabolite formed during microsomal DTZ biotransformation in rat liver in vitro. The other primary metabolites, desacetyl-DTZ and O-desmethyl-DTZ, were ineffective inhibitors of any pathways of steroid oxidation by P450s, but several other potential metabolites, which were not detected in microsomal incubations, also inhibited P450 activity. Consistent with previous reports, there was no evidence of P450 inactivation or complexation by DTZ, but the drug and its N-desmethyl metabolite generated binding interactions with ferric P450 in rat hepatic microsomes. Considered together, the findings of the present study establish that N-desmethyl-DTZ is a preferential inhibitor of CYP3A2 in rat hepatic microsomes, with greater potency than the parent drug. This is consistent with clinical reports in which this metabolite accumulates during multiple-dose therapy with DTZ. The competitive nature of the inhibitory interaction suggests that the eventual elimination of N-desmethyl-DTZ should restore normal hepatic oxidation capacity.
Related Concept Videos
 01:19
Factors Affecting Drug Biotransformation: Biological
Species differences: Variations in enzyme systems across species can cause disparities in drug metabolism. For instance, humans may metabolize certain drugs faster than rodents, altering therapeutic effects.
Strain differences: Genetic variations within a species can result in differing enzyme activity, impacting drug response and toxicity. For example, some mouse strains may...
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:18
Pharmacokinetics in Geriatric Patients: Effect of Age on Drug Metabolism
 01:26
Effect of Hepatic Disease on Pharmacokinetics: Drug Dosing and Hepatic Blood Flow
 01:14
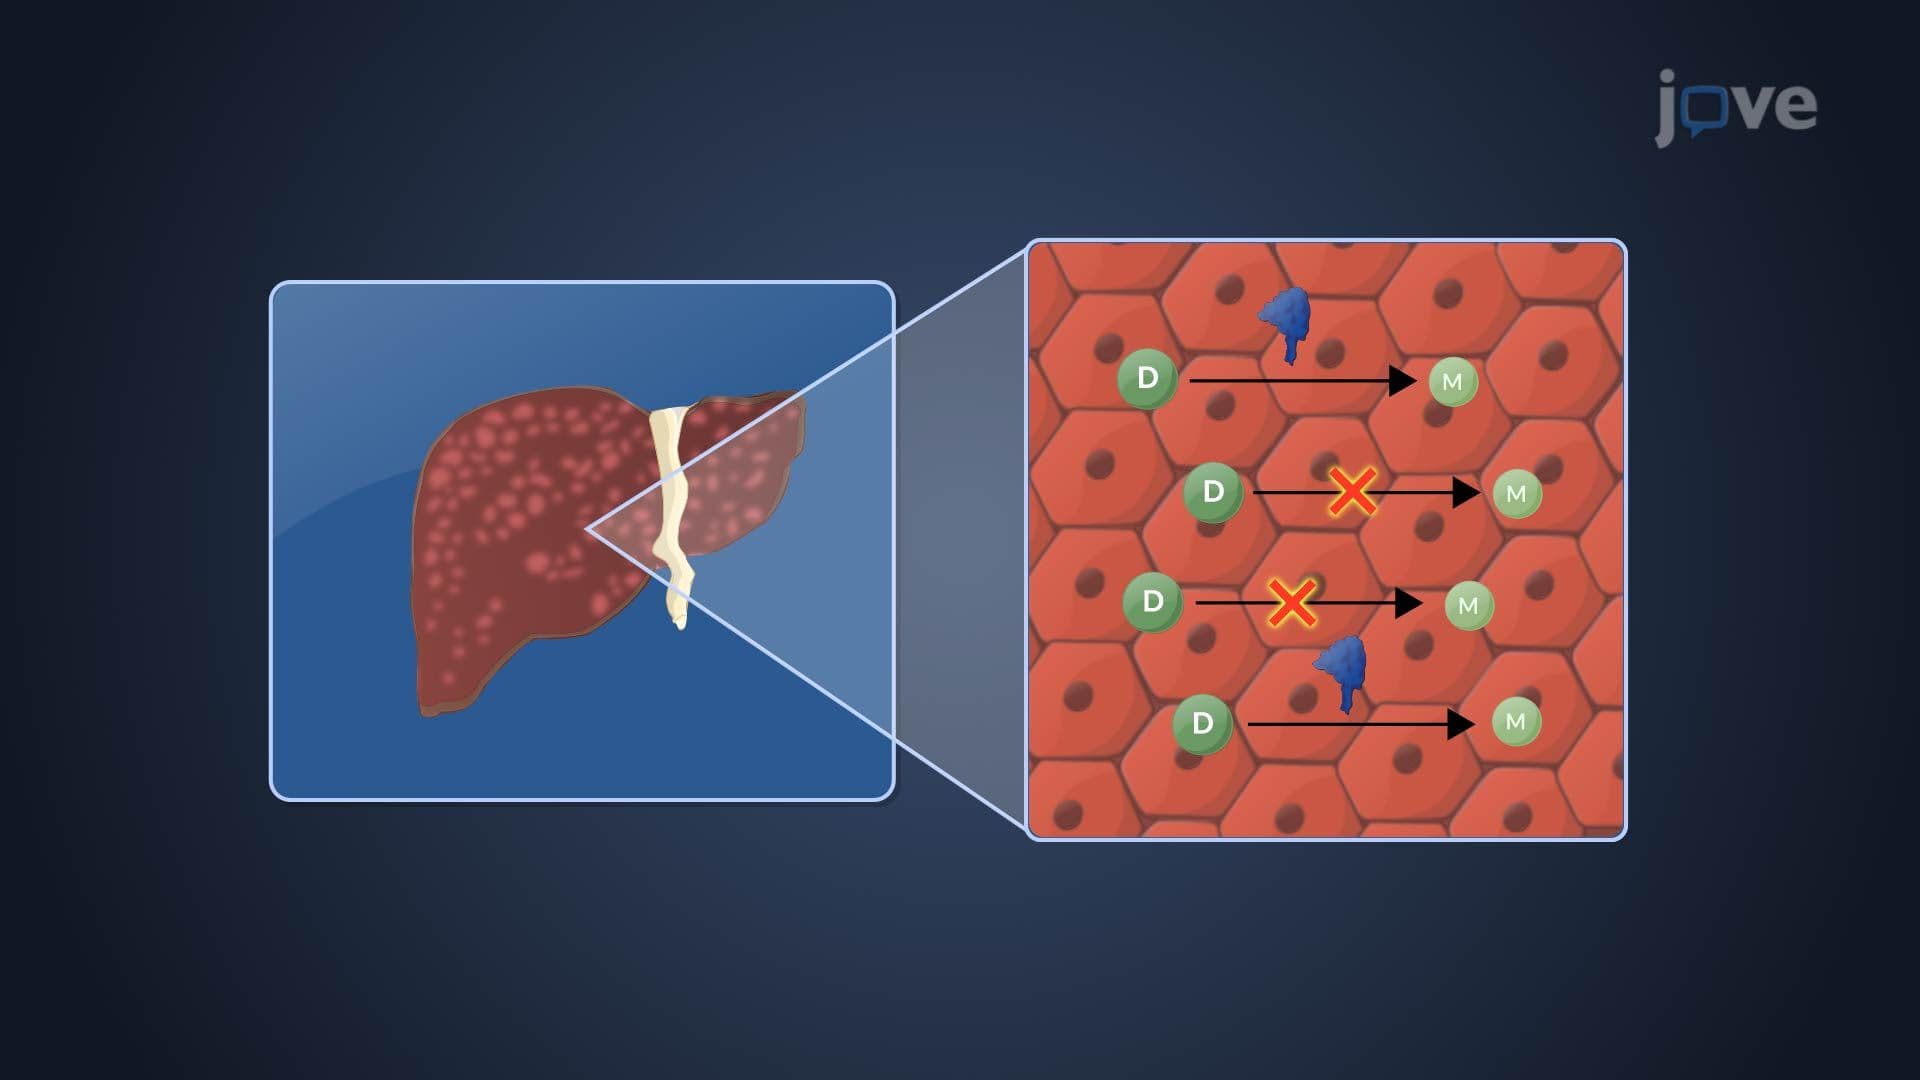
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:08
Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment