Ciprofloxacin decreases plasma phenytoin concentrations in the rat

M S al-Humayyd1

  • 1Department of Medical Pharmacology, College of Medicine, King Saud University, Riyadh, Saudi Arabia.

Related Concept Videos

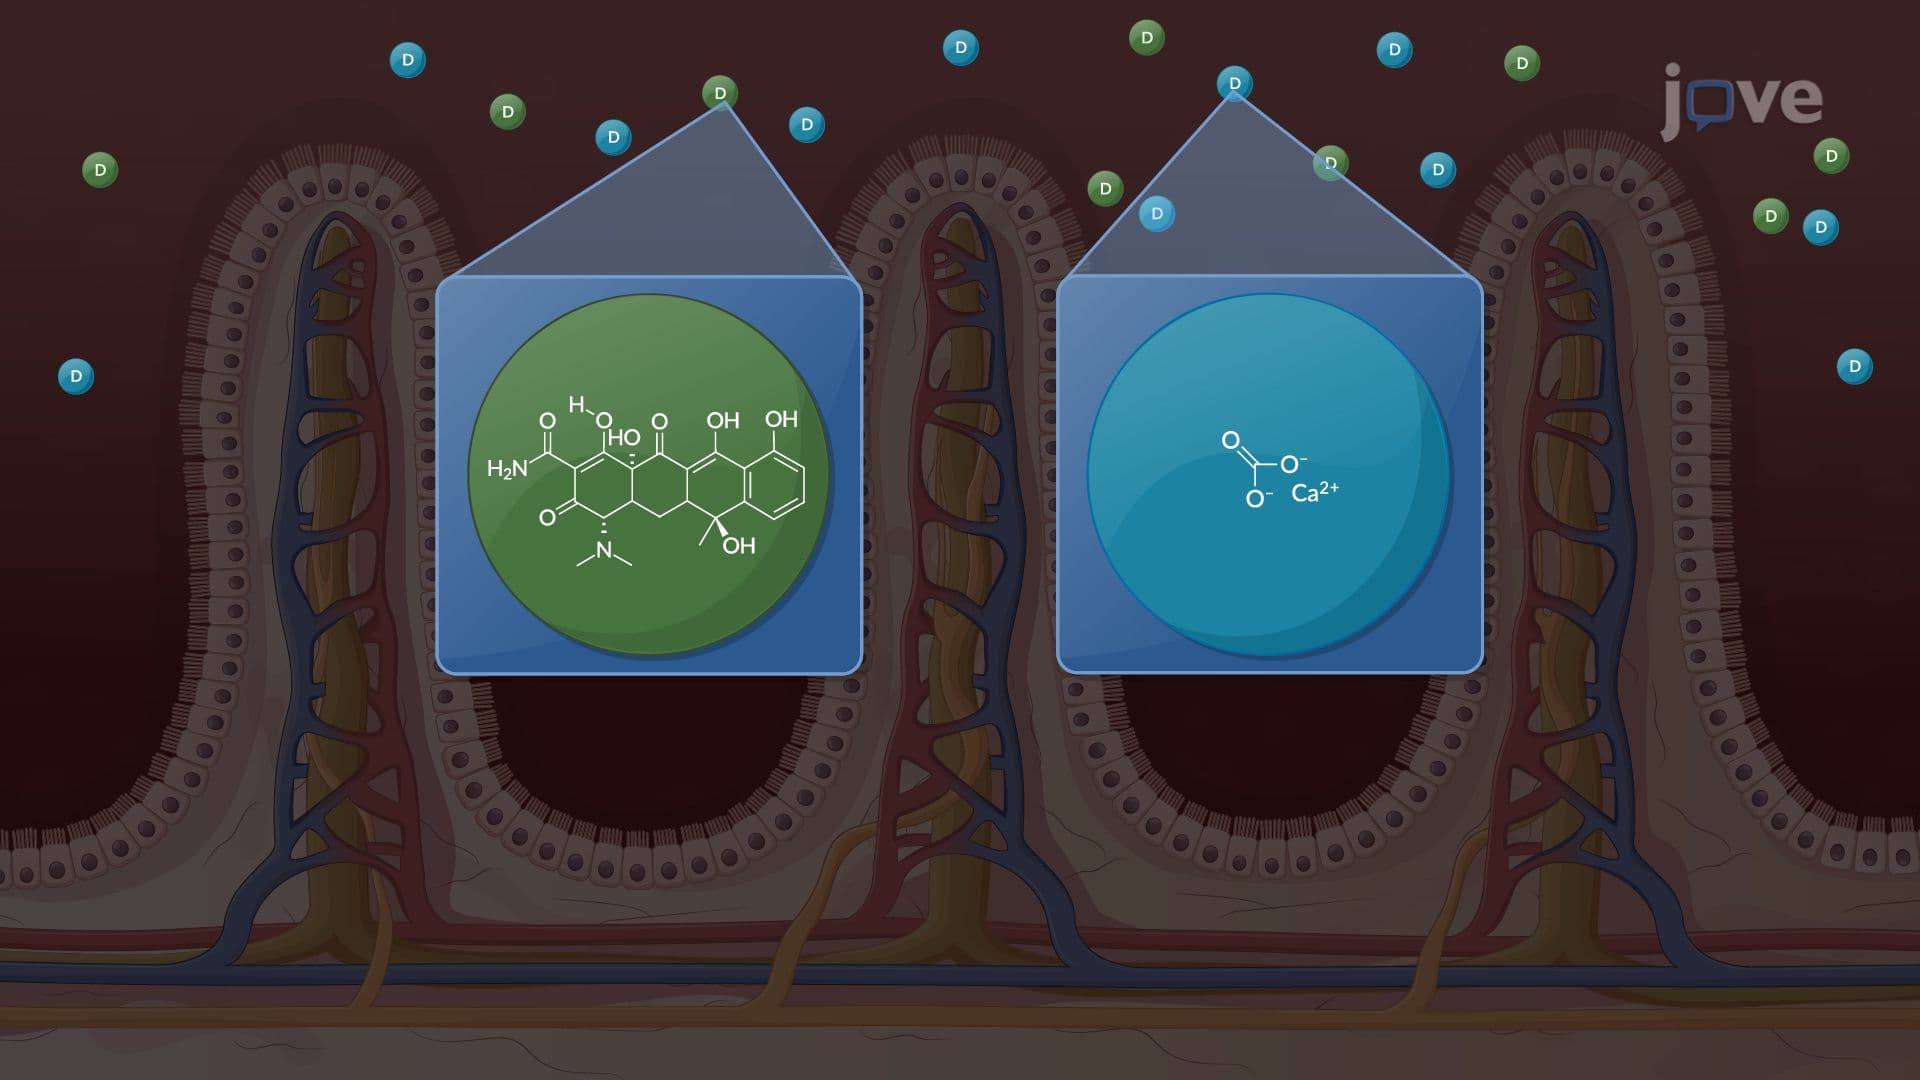
Pharmacokinetics: Drug–Drug Interactions01:25

Pharmacokinetics: Drug–Drug Interactions

Drug interactions occur when the pharmacological effect of one drug is altered by another substance, either enhancing or diminishing its activity. The drug whose activity is altered is known as the object drug, and the substance causing the alteration is called the agent drug or the precipitant. The net effects of these interactions are mostly undesirable, leading to decreased effectiveness or increased adverse effects. In rare cases, interactions can be beneficial, such as the enhanced...
Pharmacokinetics in Pediatric Patients: Drug Distribution01:17

Pharmacokinetics in Pediatric Patients: Drug Distribution
Drug distribution in the pediatric population exhibits unique challenges and considerations due to the physiological differences between children, particularly neonates and infants, and adults. A crucial aspect of pediatric pharmacology is understanding how these differences impact the pharmacokinetics of various drugs, necessitating age-specific dosing strategies to ensure efficacy and safety.Neonates and infants have a higher total body water content, ~75%–90% of their body weight, compared...
Pharmacodynamic Models: Linear Concentration–Effect Model01:15

Pharmacodynamic Models: Linear Concentration–Effect Model

The linear concentration–effect model, underpinned by the principle that pharmacological effect (E) is directly proportional to plasma drug concentration (C), emerges as a pivotal simplification of the Emax model for conditions where C is significantly less than EC50. This model portrays a linear trajectory of the concentration–effect relationship when drug levels are markedly below the EC50 threshold.Despite its inherent assumption of continuous effect augmentation with increasing drug...
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes01:28

Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes

Cytochrome P450 (CYP450) enzymes are a superfamily of heme-containing monooxygenases that play a pivotal role in Phase I drug metabolism by catalyzing oxidation and reduction reactions.These enzymes transform lipophilic xenobiotics into more hydrophilic metabolites, facilitating subsequent Phase II conjugation and eventual excretion. The CYP450 family is classified into families (e.g., CYP1–CYP3) and subfamilies (e.g., CYP2A, CYP2C), based on amino acid sequence homology.CYP450 isoenzymes,...
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance01:23

Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance

The elimination half-life and drug clearance of drugs following nonlinear kinetics can vary with dosage. The Michaelis-Menten parameters and drug concentration influence these factors. As the dose increases, the elimination half-life tends to lengthen, resulting in a reduction in clearance and a disproportionately larger area under the curve. The total clearance can be derived from the Michaelis-Menten equation for drugs following a one-compartment model.
A study on guinea pigs examined the...
Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment01:08

Effect of Hepatic Disease on Pharmacokinetics: Dose Adjustments Due to Hepatic Impairment

Hepatic impairment, characterized by decreased liver function, does not uniformly mandate adjustments in drug dosage. Whether dosage modifications are necessary depends on various factors related to the drug's metabolism and elimination pathways. If a drug is primarily excreted via the kidneys and bypasses significant hepatic processing, if it undergoes minimal metabolic transformation in the liver, or if it is volatile and primarily expelled through the lungs, dose adjustments may not be...