Related Experiment Video
Updated: Aug 7, 2026
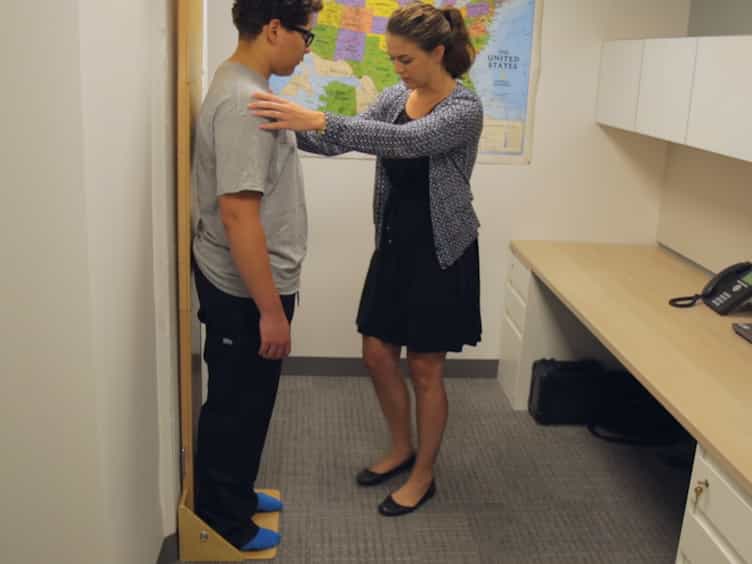

Assessment of Child Anthropometry in a Large Epidemiologic Study
Published on: February 2, 2017
Height and weight in cystic fibrosis: a cross sectional study. UK Cystic Fibrosis Survey Management Committee

Insights
UK patients with cystic fibrosis show improved height and weight in childhood but still experience deficits post-puberty. These findings suggest ongoing challenges in growth maintenance for individuals with cystic fibrosis.
Area of Science:
- Pediatric Endocrinology
- Pulmonology
- Clinical Nutrition
Background:
- Cystic fibrosis (CF) is a genetic disorder affecting multiple organs, notably the lungs and digestive system.
- Growth abnormalities, including reduced height and weight, are common complications in CF patients.
- Previous studies indicated significant growth deficits in CF populations.
Purpose of the Study:
- To present cross-sectional data on anthropometric measurements (height, weight, BMI) in UK patients with cystic fibrosis.
- To assess growth patterns in CF patients across different age groups.
- To identify potential factors influencing growth maintenance in CF.
Main Methods:
- Cross-sectional study design.
- Data collection on height, weight, and body mass index (BMI).
- Analysis of anthropometric data in UK cystic fibrosis patients.
Main Results:
- During the first decade of life, CF patients' height and weight were approximately 0.5 standard deviations below the general population.
- This represents an improvement compared to historical data.
- Substantial deficits in postpubertal stature and weight were observed in the CF cohort.
Conclusions:
- While childhood growth has improved in CF patients, significant challenges remain in achieving optimal growth post-puberty.
- Treatment strategies may need further optimization to address persistent growth deficits.
- Continued monitoring of anthropometric parameters is crucial for managing cystic fibrosis care.
Abstract:
Cross sectional data reporting the height, weight, and body mass index of UK patients with cystic fibrosis are presented. During the first decade of life height and weight in patients with cystic fibrosis are maintained at about 0.5 SD below those of the general population, which reflects an improvement over earlier published observations. Postpubertal stature and weight maintenance in the cystic fibrosis population still show substantial deficits which may be related to treatment.
More Related Videos

Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History