Carbohydrate-deficient glycoprotein syndrome type Ib. Phosphomannose isomerase deficiency and mannose therapy

R Niehues1, M Hasilik, G Alton

  • 1Klinik und Poliklinik für Kinderheilkunde, 48149 Münster, Germany.

Related Concept Videos

Phosphoinositides and PIPs01:42

Phosphoinositides and PIPs

Phosphoinositides are a group of phospholipids containing a glycerol backbone with two fatty acid chains and a phosphate attached to a myoinositol sugar ring. The inositol head group extends into the cytoplasm, where it is modified by adding phosphate groups to form phosphatidylinositol phosphates or PIPs.
Different phosphoinositides are synthesized and recruited on the cytosolic face of the plasma membrane. The localization of specific phosphoinositides concentrated in separate membrane...
7.7K
Oligosaccharide Assembly01:24

Oligosaccharide Assembly

Protein glycosylation starts in the ER lumen and continues in the Golgi apparatus. Glycosyltransferases catalyze the addition of sugar molecules or glycosylation of proteins. Usually, these enzymes add sugars to the hydroxyl groups of selected serine or threonine residues to form O-linked glycans or the amino groups of asparagine residues to form N-linked glycans. Different positions on the same polypeptide chain can contain differently linked glycans.
Multiple sugar molecules that may or may...
2.6K
Proteoglycans01:05

Proteoglycans

Glycans, a class of complex heterogeneous molecules, can be covalently attached to proteins to form glycosylated proteins that regulate various physiological and pathological processes. Glycosylated proteins or glycoproteins comprise N-linked and O-linked oligosaccharides. O-glycosylation is the most common type of protein glycosylation. Here, glycans attach to the oxygen atom of the hydroxyl groups of Serine or Threonine residues. O-linked glycosylation occurs later in protein processing,...
4.1K
Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
1.1K
Type I Diabetes I: Introduction01:12

Type I Diabetes I: Introduction

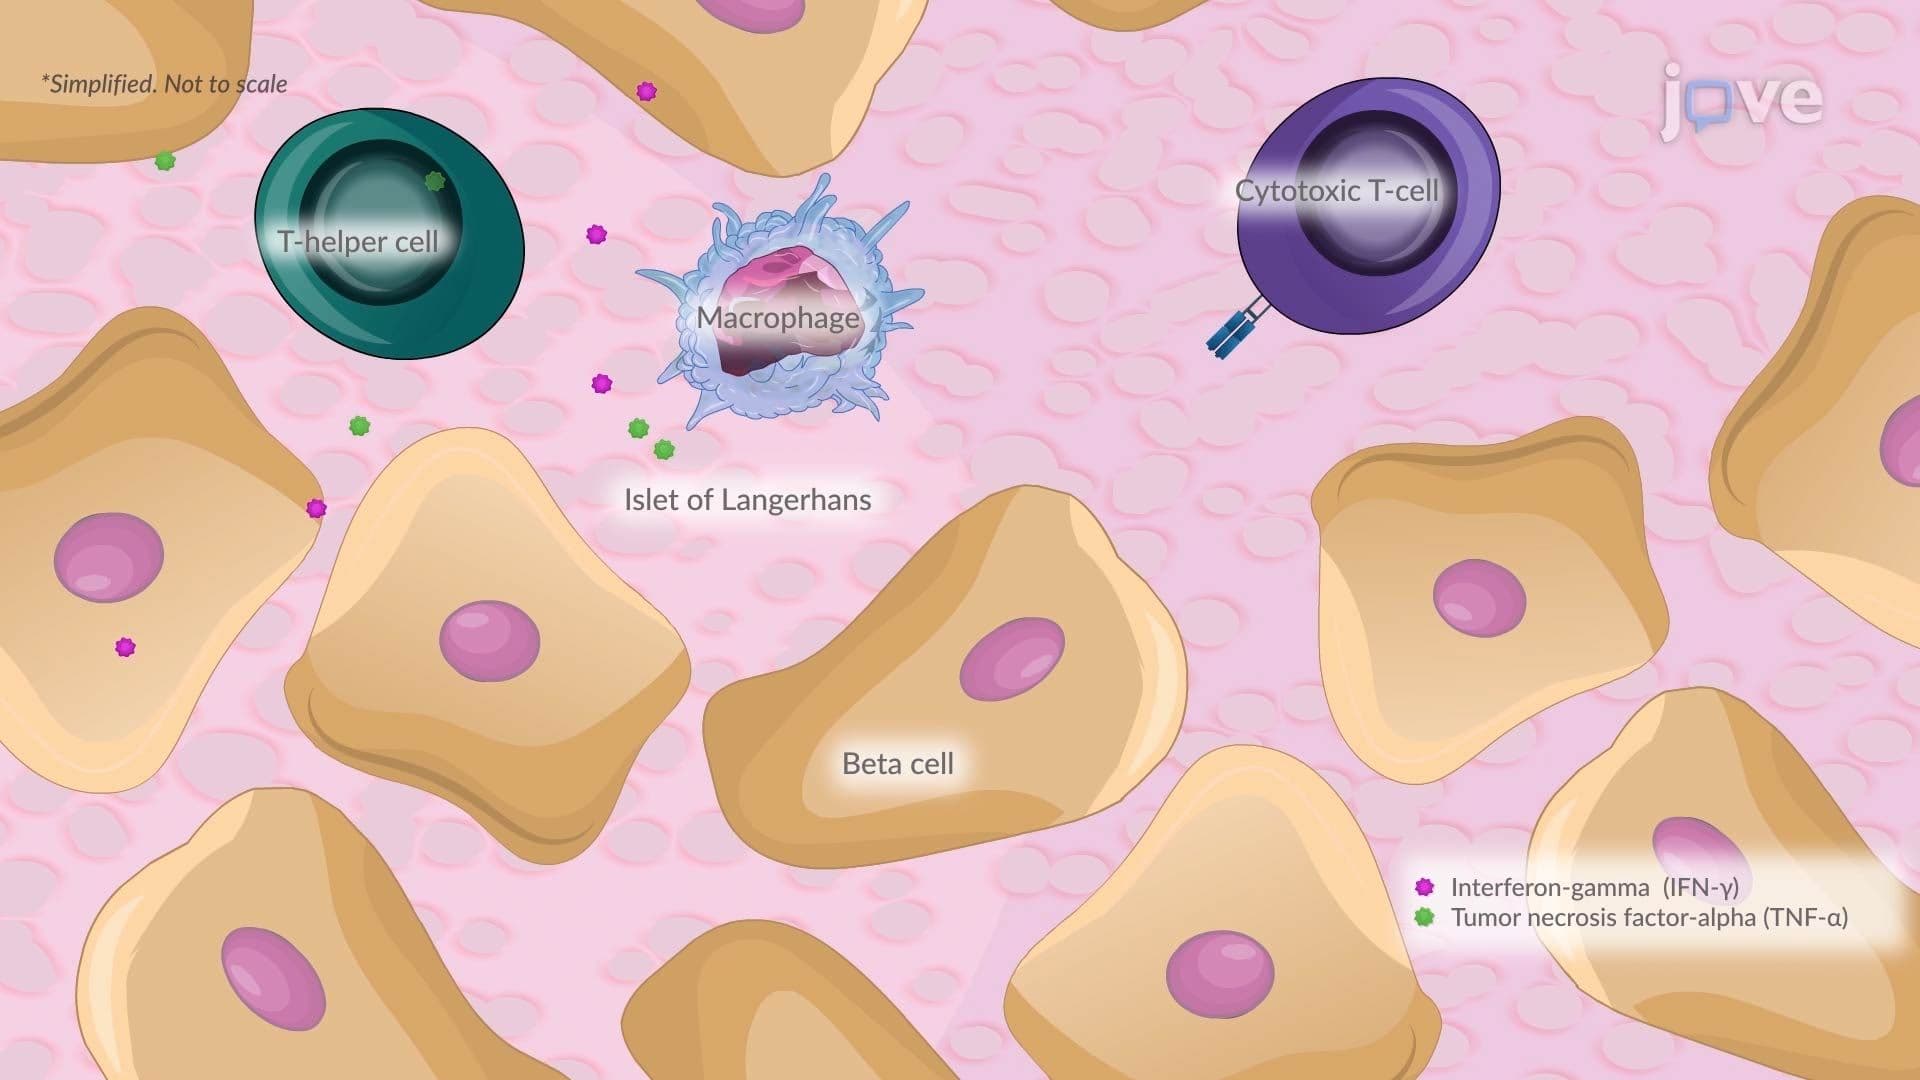
Type 1 diabetes mellitus is a chronic metabolic disorder characterized by an absolute deficiency of insulin resulting from the autoimmune destruction of pancreatic β-cells. Although it can occur at any age, it is most commonly diagnosed in childhood, adolescence, or early adulthood. The loss of insulin production impairs cellular glucose uptake, resulting in persistent hyperglycemia and necessitating lifelong insulin therapy.Autoimmune Destruction of β-CellsThe hallmark of type 1...
45
Type I Diabetes II: Pathophysiology01:26

Type I Diabetes II: Pathophysiology

Type 1 diabetes mellitus arises from an immune-mediated destruction of pancreatic β-cells, resulting in an absolute deficiency of insulin. This process develops in genetically susceptible individuals when autoimmunity, environmental exposures, and immunologic dysregulation converge to trigger a targeted attack on the insulin-producing cells of the pancreas. The β-cells are located within the islets of Langerhans and are essential for regulating blood glucose by facilitating cellular...
75