Related Experiment Video
Updated: Aug 12, 2026
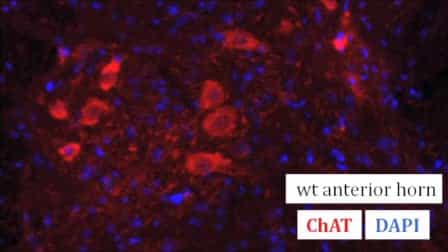

Clinical Testing and Spinal Cord Removal in a Mouse Model for Amyotrophic Lateral Sclerosis (ALS)
Published on: March 17, 2012
Spinal muscular atrophy
1Neuromuscular Disease and Neurorehabilitation, Texas Scottish Rite Hospital for Children, Dallas, USA.

Insights
Spinal muscular atrophy (SMA) is a genetic disorder characterized by progressive muscle weakness. While the survival motor neuron (SMN) gene is implicated in over 98% of cases, effective therapies remain elusive, focusing on complication management.
Area of Science:
- Neurology
- Genetics
- Pediatrics
Background:
- Spinal muscular atrophy (SMA) is a group of inherited neuromuscular disorders.
- Historically described in the 1890s, SMA presents with progressive muscle weakness and floppiness.
- Current classification includes SMA types 1, 2, and 3 based on symptom onset age.
Purpose of the Study:
- To provide a historical overview of Spinal Muscular Atrophy (SMA).
- To outline the current understanding of SMA genetics and classification.
- To describe the current management strategies for SMA.
Main Methods:
- Review of historical medical literature and collaborative research findings.
- Genetic linkage studies identifying chromosome 5q11.2-13.3.
- Analysis of the survival motor neuron (SMN) gene in SMA patients.
Main Results:
- SMA is linked to deletions in the survival motor neuron (SMN) gene in over 98% of patients.
- Mutations in the SMN gene are also found in atypical SMA forms.
- No effective therapy currently exists for SMA.
Conclusions:
- Management of SMA focuses on preventing and treating complications associated with severe muscle weakness.
- Complications include respiratory issues, nutritional deficits, orthopedic deformities, and psychosocial challenges.
- Further research into the SMN gene's function is needed for therapeutic development.
Abstract:
The history of the spinal muscular atrophies (SMA) began in the 1890s with Guido Werdnig and Johann Hoffmann. Together, their papers present a rather complete picture of the clinical and pathologic aspects of infantile SMA: onset during the first year of life, occurrence in siblings with normal parents, progressive floppiness and weakness, hand tremor, and death from pneumonia in early childhood. Based on the work of an international collaboration, the following is current nomenclature: SMA type 1 (or I) for onset of symptoms before age 6 months, SMA type 2 (II) for onset between 6 and 18 months, and SMA type 3 (III) for onset after age 18 months. Linkage of autosomal recessive SMA to chromosome 5q11.2-13.3 was reported by Gilliam et al in 1990. A novel gene, whose function remains unknown, called the survival motor neuron gene (SMN) at 5q13, contains deletions in more than 98% of SMA patients. Some patients with atypical forms of SMA have been shown to have mutations in SMN. Because there is no effective therapy for SMA, management consists of preventing or treating the complications of severe weakness, such as restrictive lung disease, poor nutrition, orthopedic deformities, immobility, and psychosocial problems.
More Related Videos
Related Concept Videos
 01:26
Cross-bridge Cycle
 01:21
Satellite Stem Cells and Muscular Dystrophy
 01:20
Myasthenia Gravis: Overview and Treatment
These antibodies interfere with the function of the nicotinic receptors in three ways: by binding to the receptor and disrupting acetylcholine binding; by causing cross-linking of receptors which leads...
 01:28
Disorders of the Skeletal Muscle
Musculoskeletal disorders
Musculoskeletal disorders involve injuries and conditions affecting the skeletal muscles and associated connective tissues. These disorders can arise from acute biomechanical stresses or chronic overuse and can occur across different age groups. Common injuries include sprains, fractures, and muscular strains, often resulting from...
 01:22
Myasthenia Gravis ll: Pathophysiology
 01:11
Alterations in Muscle Tone lll