PAX8 mutations associated with congenital hypothyroidism caused by thyroid dysgenesis
Related Experiment Video
Updated: Oct 29, 2026
In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
1Stazione Zoologica A. Dohrn, Napoli, Italy.

Abstract:
Permanent congenital hypothyroidism (CH) is a common disease that occurs in 1 of 3,000-4,000 newborns. Except in rare cases due to hypothalamic or pituitary defects, CH is characterized by elevated levels of thyroid-stimulating hormone (TSH) resulting from reduced thyroid function. When thyroid hormone therapy is not initiated within the first two months of life, CH can cause severe neurological, mental and motor damage. In 80-85% of cases, CH is associated with and presumably is a consequence of thyroid dysgenesis (TD). In these cases, the thyroid gland can be absent (agenesis, 35-40%), ectopically located (30-45%) and/or severely reduced in size (hypoplasia, 5%). Familial cases of TD are rare, even though ectopic or absent thyroid has been occasionally observed in siblings. The pathogenesis of TD is still largely unknown. Although a genetic component has been suggested, mutations in the gene encoding the receptor for the thyroid-stimulating hormone (TSHR) have been identified in only two cases of TD with hypoplasia. We report mutations in the coding region of PAX8 in two sporadic patients and one familial case of TD. All three point mutations are located in the paired domain of PAX8 and result in severe reduction of the DNA-binding activity of this transcription factor. These genetic alterations implicate PAX8 in the pathogenesis of TD and in normal thyroid development.
Related Concept Videos
 01:23
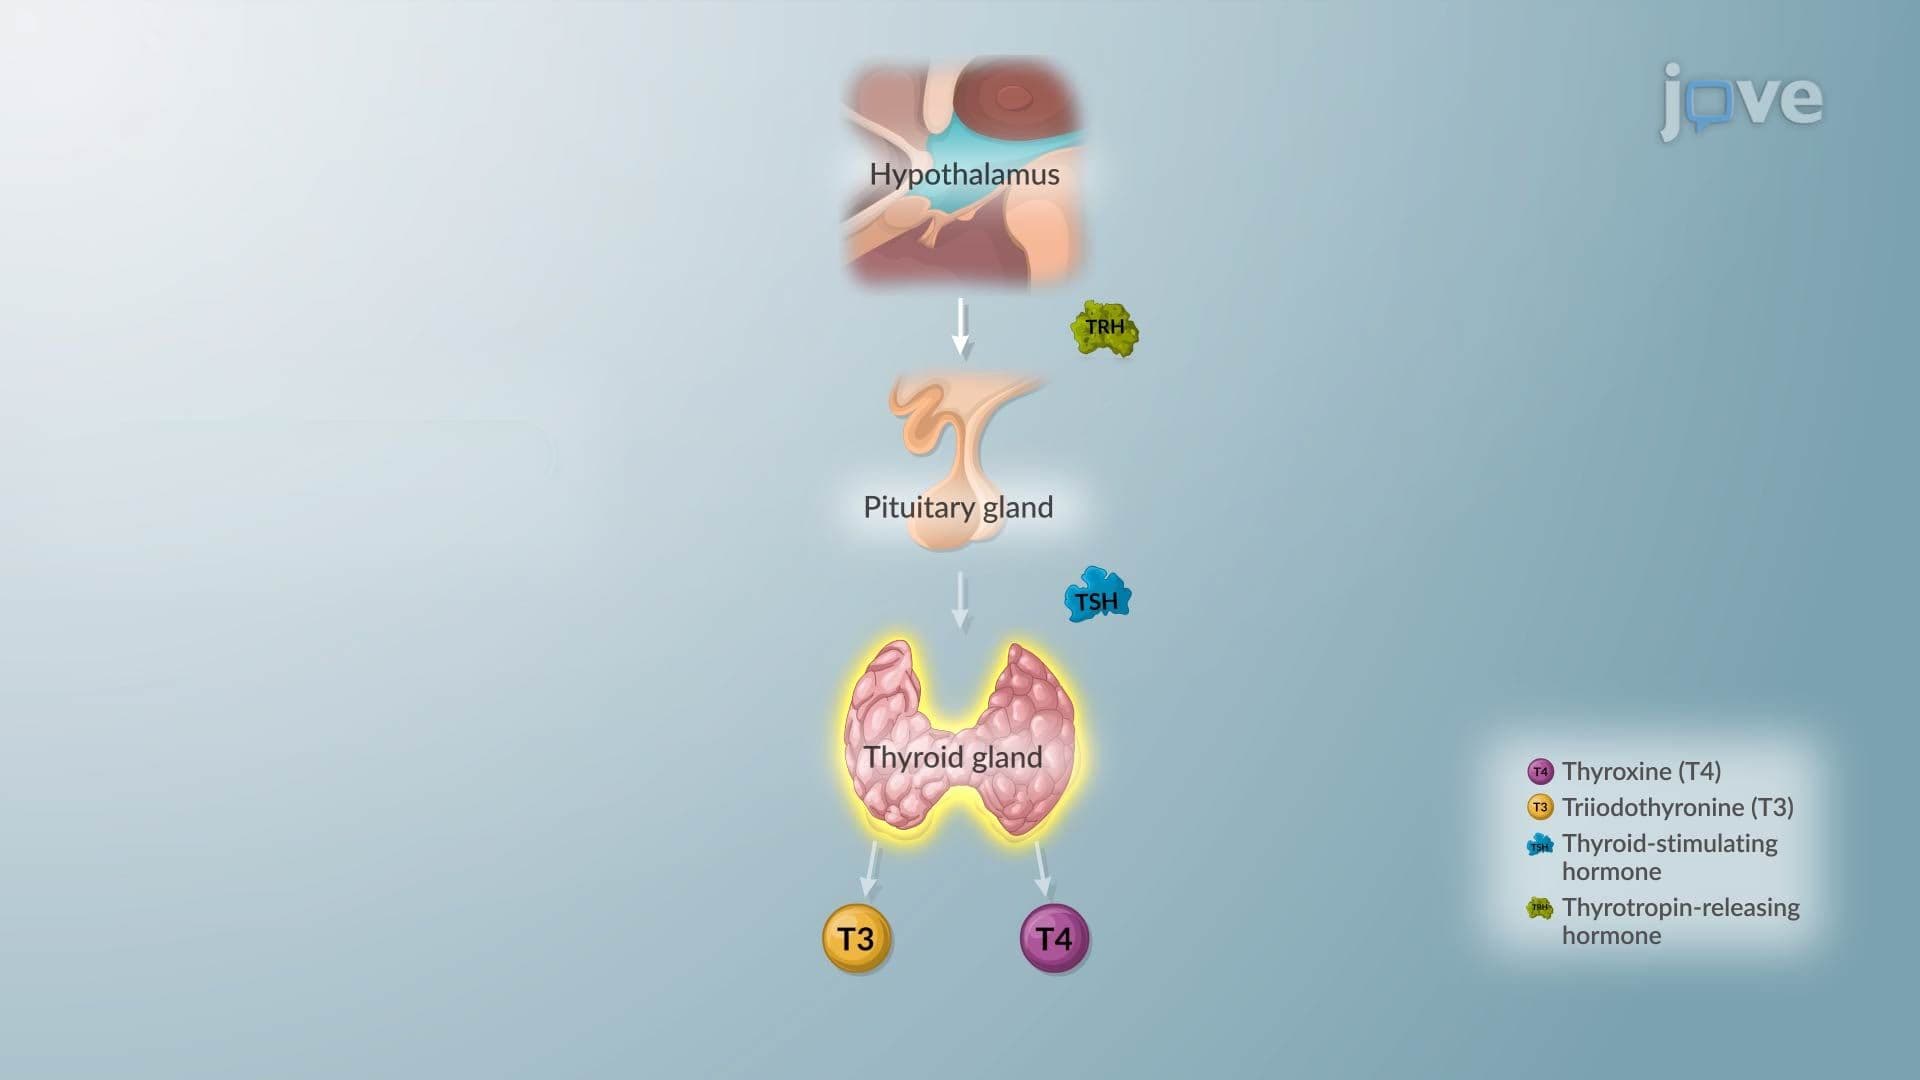
Hypothyroidism II: Pathophysiology
 01:27
Hyperthyroidism II: Pathophysiology
 01:47
Pleiotropy
 01:43
Incomplete Dominance
 01:25
Hyperthyroidism I: Introduction
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...