Extracting contact energies from protein structures: a study using a simplified model
Related Experiment Video
Updated: Aug 12, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
1Department of Biomedical Engineering, Boston University, Massachusetts 02215-2407, USA. czhang@engpub1.bu.edu

Abstract:
In this study, we exploited an elementary 2-dimensional square lattice model of HP polymers to test the premise of extracting contact energies from protein structures. Given a set of prespecified energies for H-H, H-P, and P-P contacts, all possible sequences of various lengths were exhaustively enumerated to find sequences that have unique lowest-energy conformations. The lowest-energy structures (or native structures) of such (native) sequences were used to extract contact energies using the Miyazawa-Jernigan procedure and here-defined reference state. The relative magnitudes of the original energies were restored reasonably well, but the extracted contact energies were independent of the absolute magnitudes of the initial energies. We turned to a more detailed characterization of the energy landscapes of the native sequences in light of a new theoretical framework on protein folding. Foldability of such sequences imposes two limits on the absolute value of the prespecified energies: a lower bound entailed by the minimum requirement for thermodynamic stability and an upper bound associated with the entrapment of the chain to local minima. We found that these two limits confine the prespecified energy values to a rather narrow range which, surprisingly, also contains the extracted energies in all the cases examined. These results indicate that the quasi-chemical approximation can be used to connect quantitatively the occurrence of various residue-residue contacts in an ensemble of native structures with the energies of the contacts. More importantly, they suggest that the extracted contact energies do contain information on structural stability and can be used to estimate actual structural energetics. This study also encourages the use of structure-derived contact energies in threading. The finding that there is a rather narrow range of energies that are optimal for folding a sequence also cautions the use of arbitrary energy Hamiltonion in minimal folding models.
Related Concept Videos
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
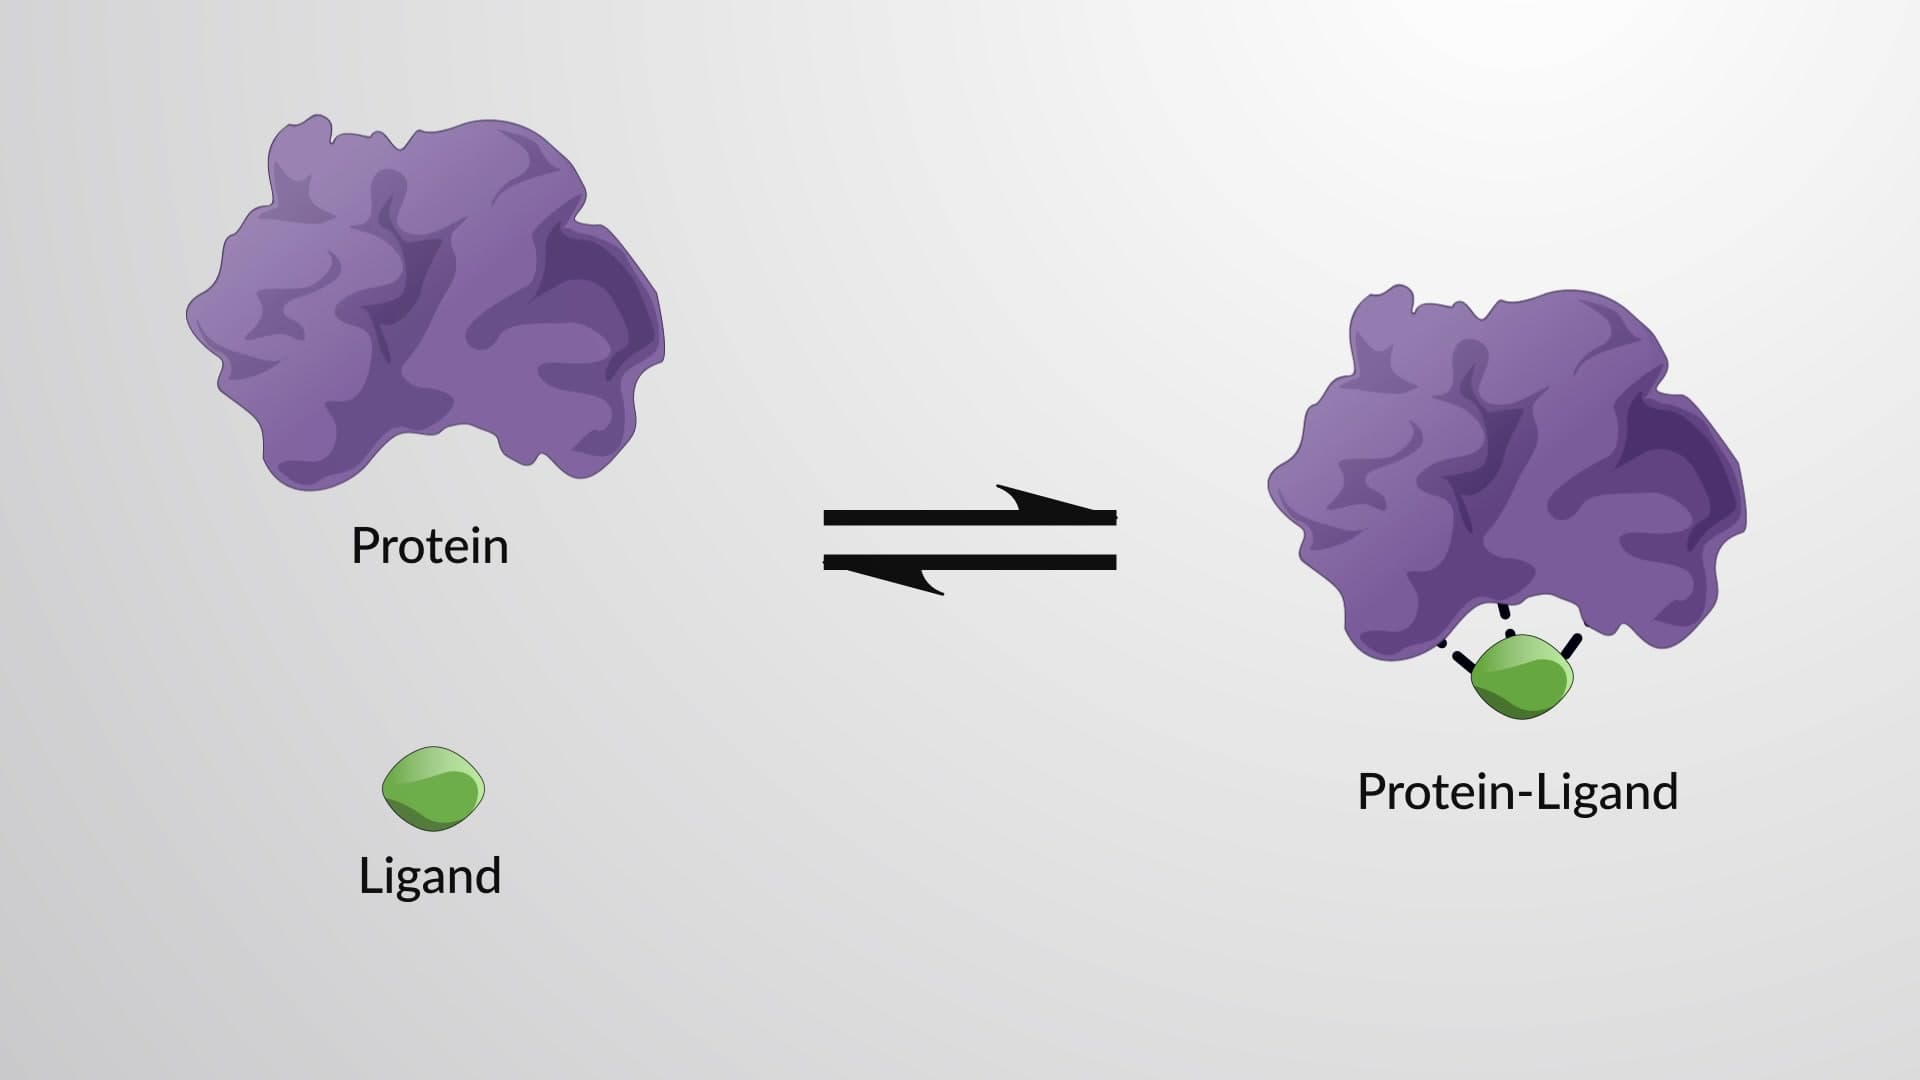
The Equilibrium Binding Constant and Binding Strength
 01:58
Mechanical Protein Functions
 01:24
Protein Organization
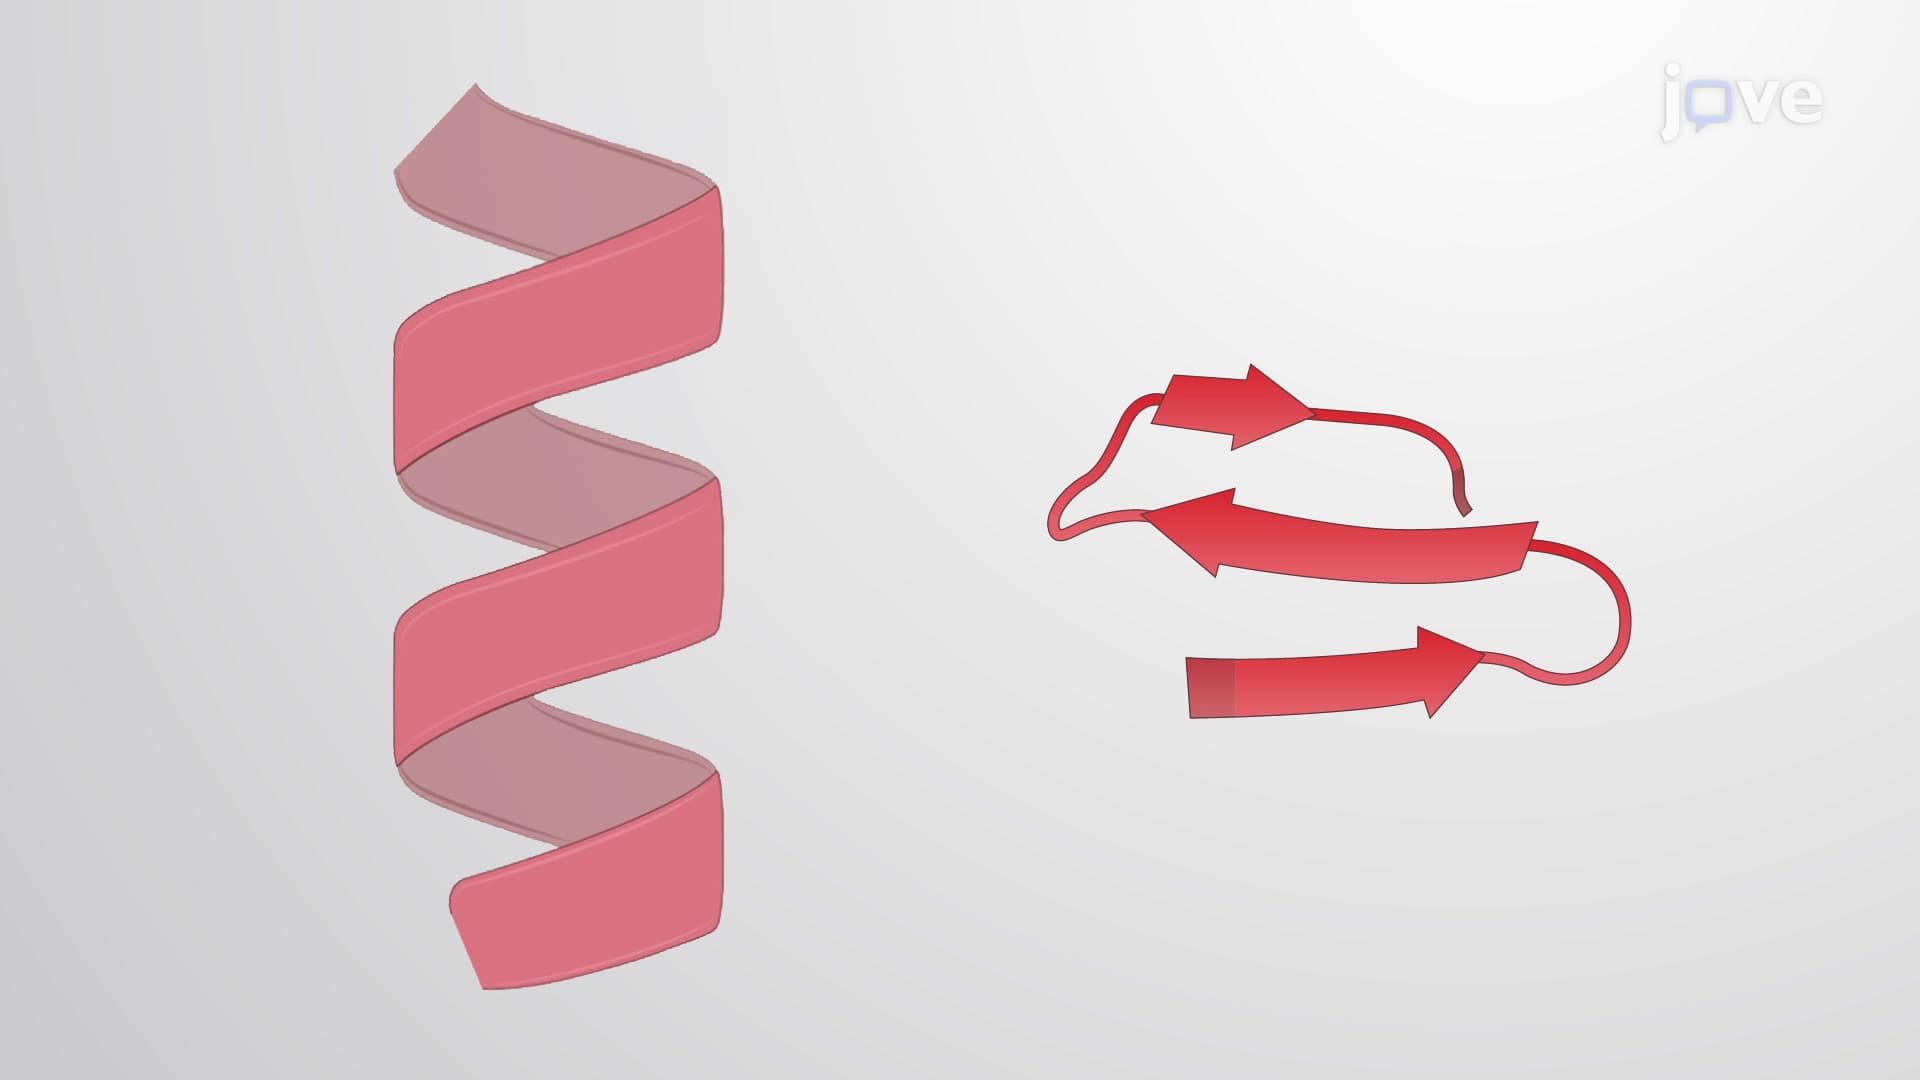
The primary structure of a protein is its amino acid sequence.