Two CPT2 mutations in three Japanese patients with carnitine palmitoyltransferase II deficiency: functional analysis and association with polymorphic haplotypes and two clinical phenotypes
Related Experiment Video
Updated: Oct 21, 2026

Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
K Wataya1, J Akanuma, P Cavadini
1Department of Biochemical Genetics, Tohoku University School of Medicine, Sendai, Japan.

Abstract:
Carnitine palmitoyltransferase II (CPT II) deficiency manifests as two different clinical phenotypes: a muscular form and a hepatic form. We have investigated three nonconsanguineous Japanese patients with CPT II deficiency. Molecular analysis revealed two missense mutations, a glutamate (174)-to-lysine substitution (E174K) and a phenylalanine (383)-to-tyrosine substitution (F383Y) in the CPT II cDNA. Transfection experiments in COS-1 cells demonstrated that the two mutations markedly decreased the catalytic activity of mutant CPT II. Case 1 (hepatic form) was homozygous for the F383Y mutation, whereas case 3 (muscular form) was homozygous for the E174K mutation. Case 2 and her brother, who were compound heterozygotes for E174K and F383Y, exhibited the hepatic phenotype. We also identified a novel polymorphism in the CPT2 gene, a phenylalanine (352)-to-cysteine substitution (F352C), which did not alter CPT II activity in transfected cells. It was present in 21 out of 100 normal alleles in the Japanese population, but absent in Caucasian populations. Genotyping with the F352C polymorphism and the two previously reported polymorphisms, V368I and M647V, allowed normal Japanese alleles to be classified into five haplotypes. In all three families with CPT II deficiency, the E174K mutation resided only on the F1V1M1 allele, whereas the F383Y mutation was observed on the F2V2M1 allele, suggesting a single origin for each mutation.
More Related Videos
 05:58
Digital Polymerase Chain Reaction Assay for the Genetic Variation in a Sporadic Familial Adenomatous Polyposis Patient Using the Chip-in-a-tube Format
Published on: August 20, 2018
 03:45
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
Related Concept Videos
 01:37
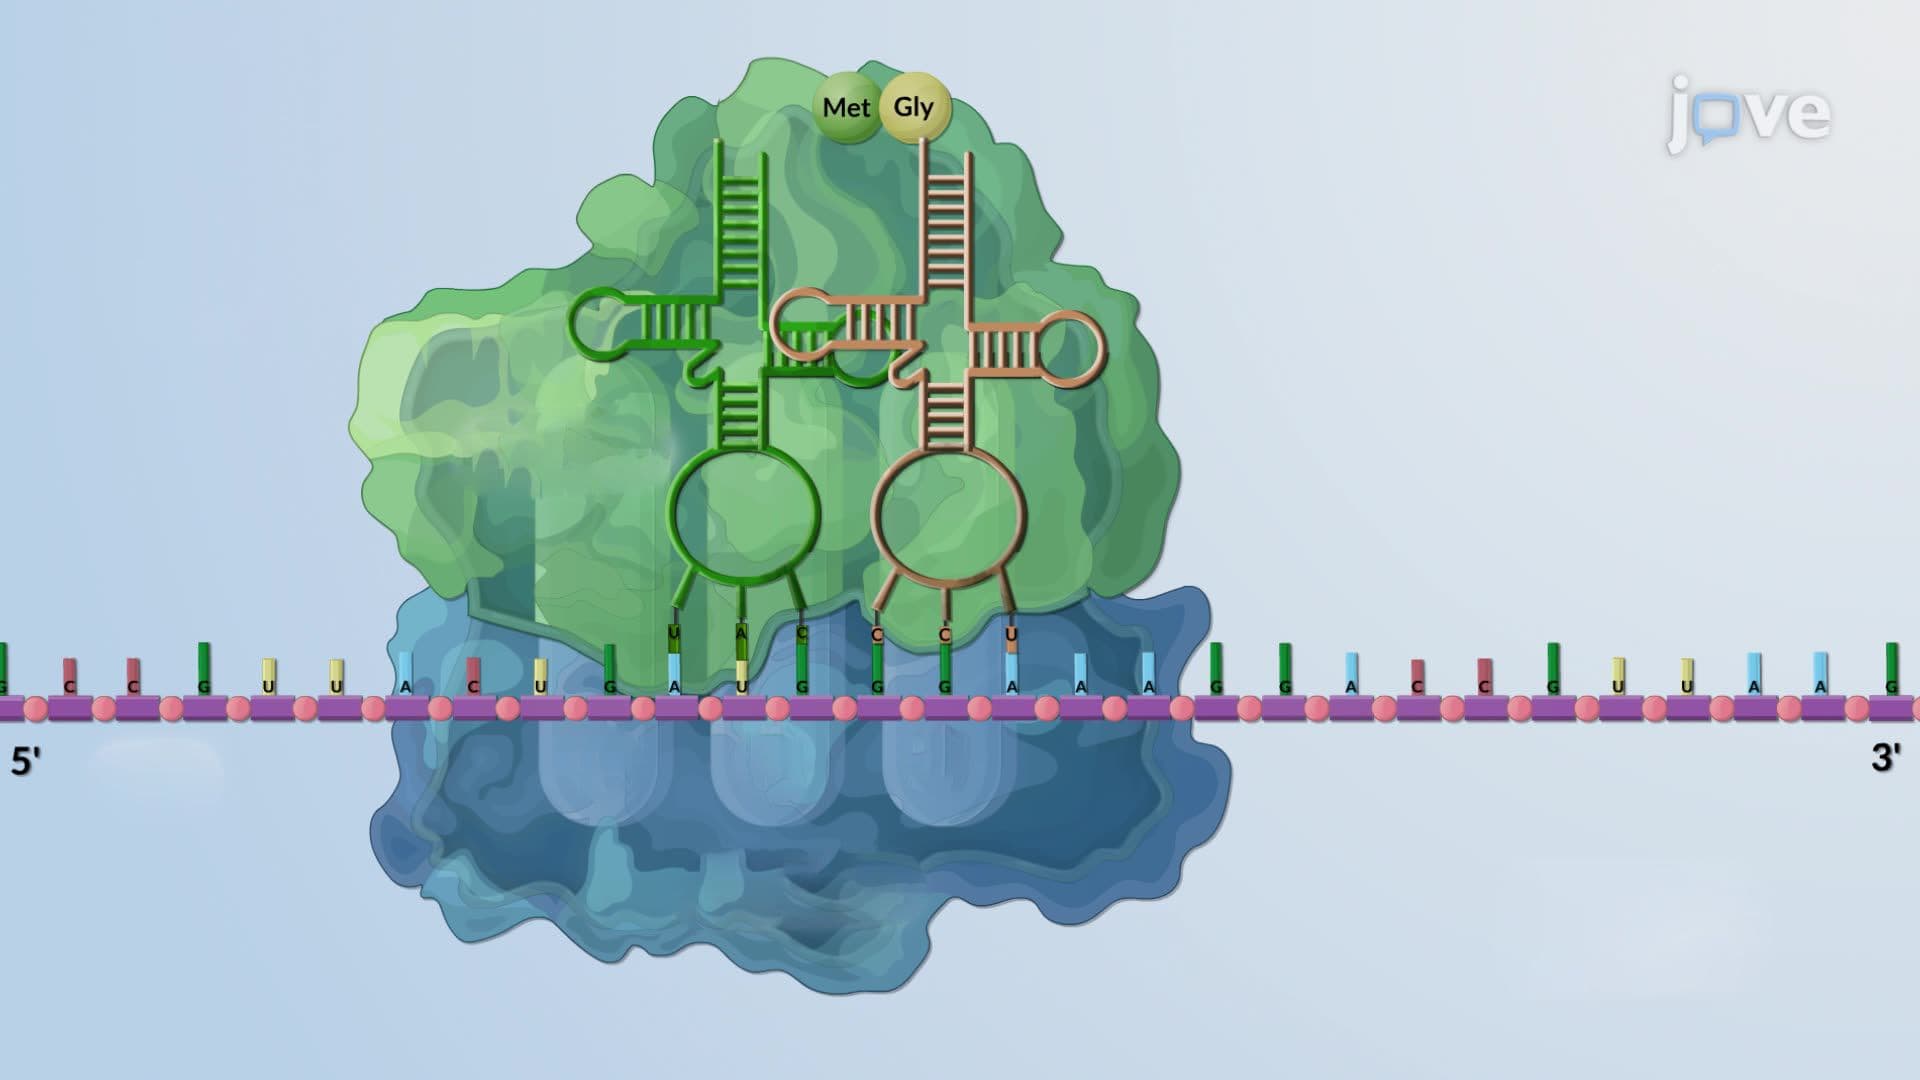
Translation
01:37Translation
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:16
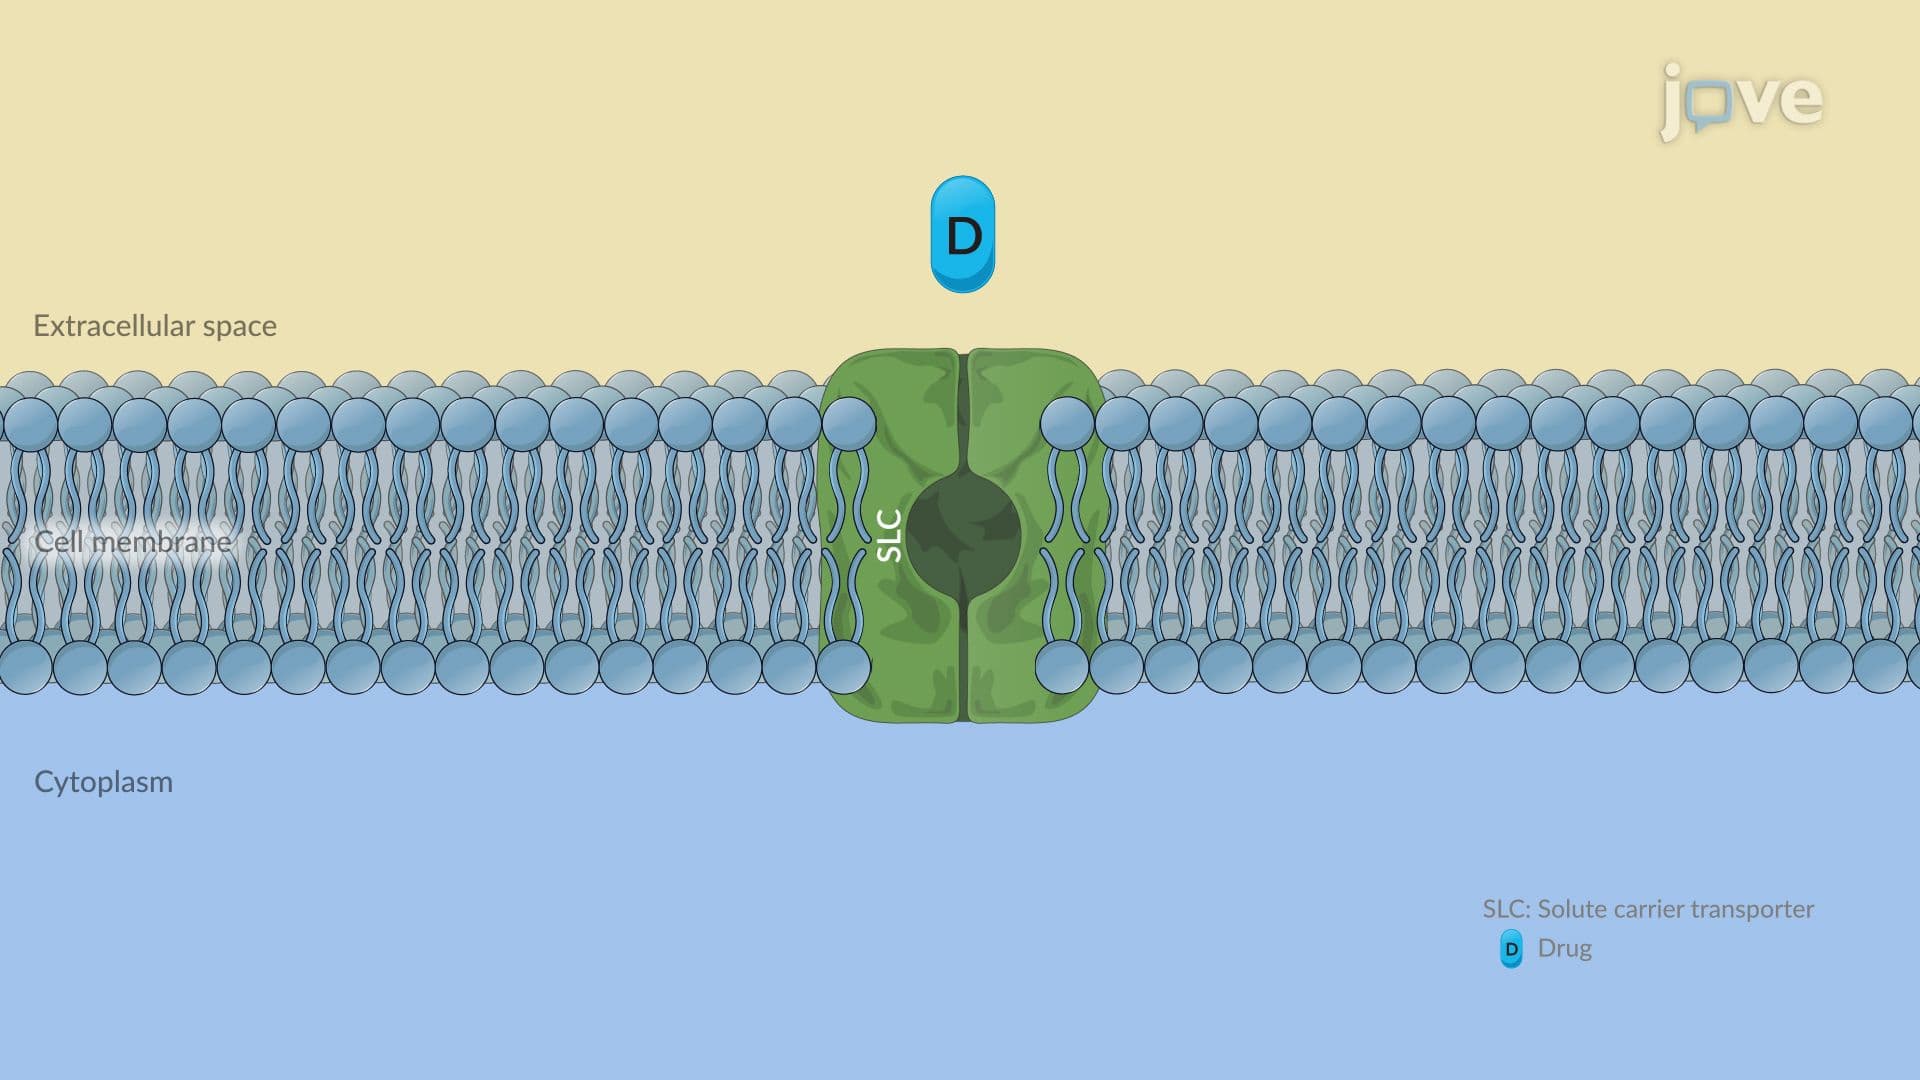
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase