Related Experiment Video
Updated: Aug 18, 2026

In Vitro Aggregation Assays Using Hyperphosphorylated Tau Protein
Published on: January 2, 2015
Mutation in the tau gene in familial multiple system tauopathy with presenile dementia
M G Spillantini1, J R Murrell, M Goedert
1Medical Research Council Centre for Brain Repair and Department of Neurology, University of Cambridge, Robinson Way, Cambridge CB2 2PY, UK. mgsll@cam.ac.uk

Abstract:
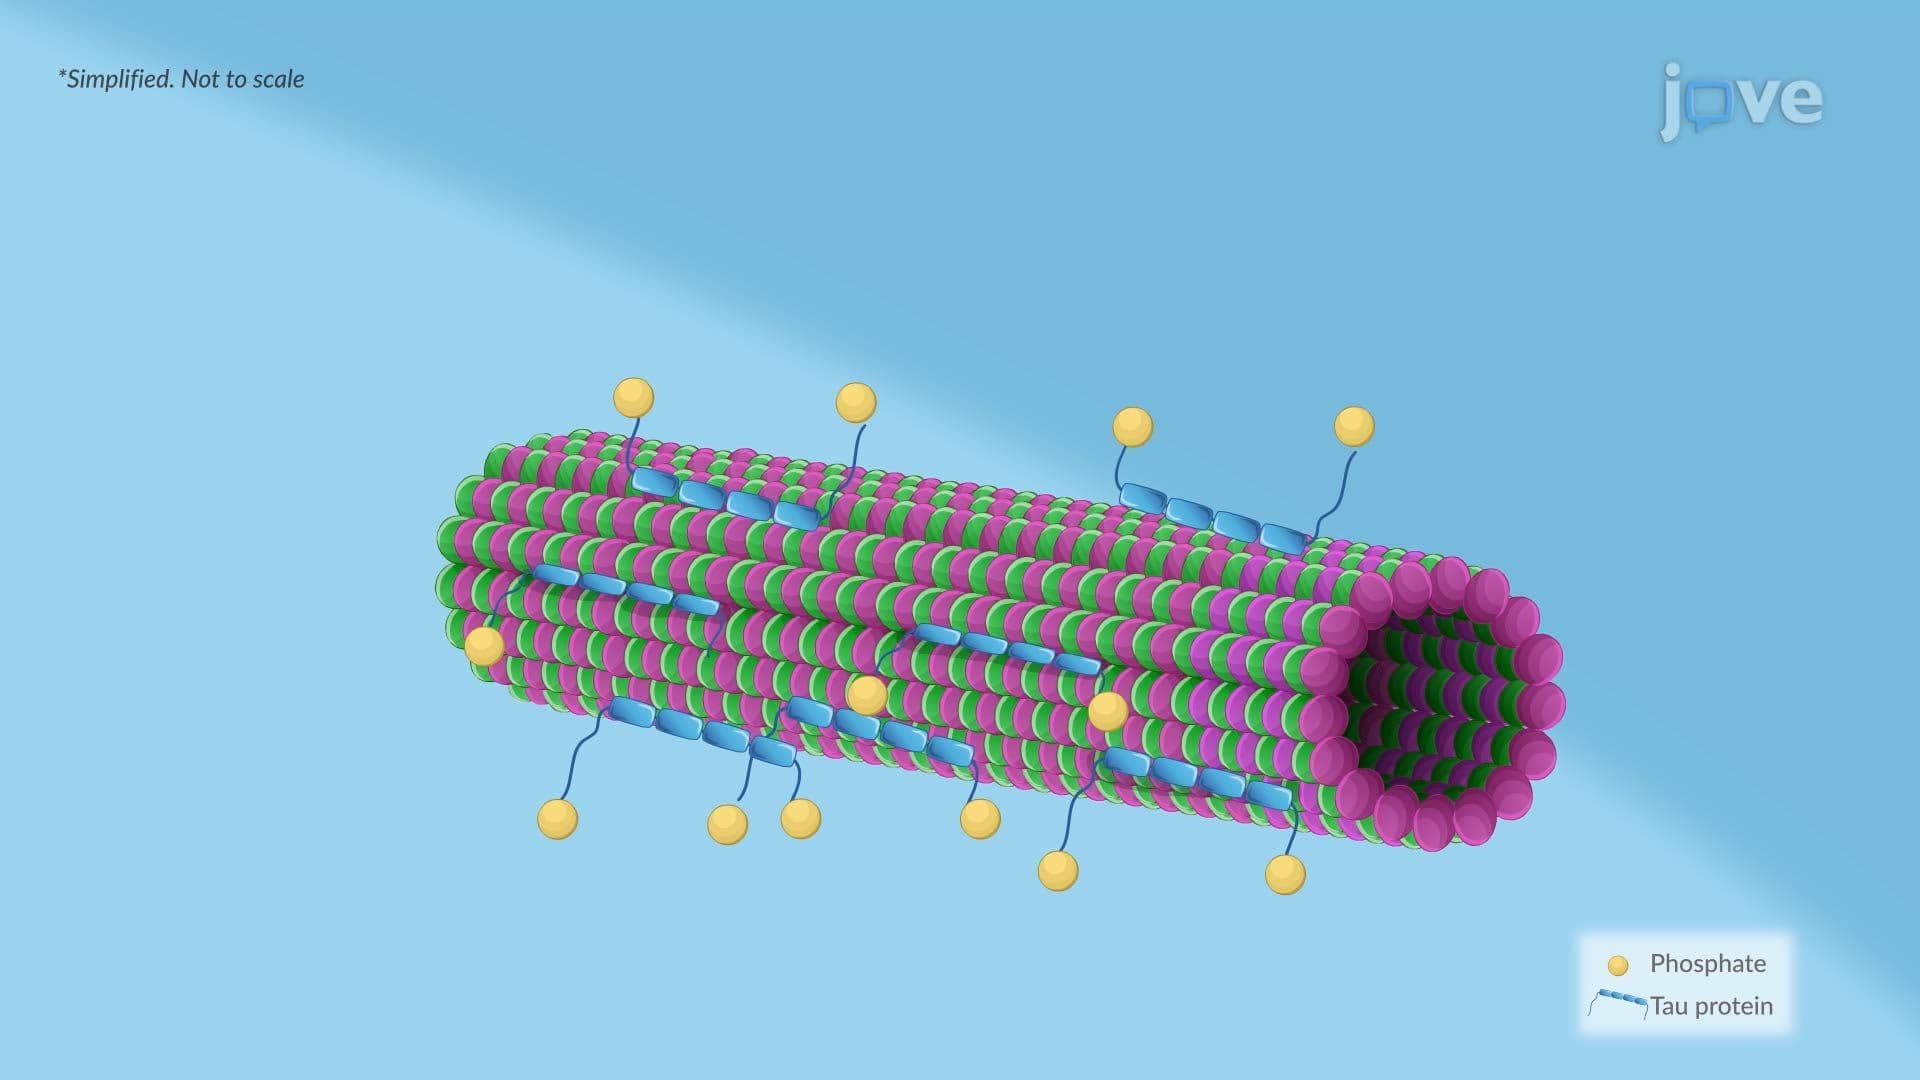
Familial multiple system tauopathy with presenile dementia (MSTD) is a neurodegenerative disease with an abundant filamentous tau protein pathology. It belongs to the group of familial frontotemporal dementias with Parkinsonism linked to chromosome 17 (FTDP-17), a major class of inherited dementing disorders whose genetic basis is unknown. We now report a G to A transition in the intron following exon 10 of the gene for microtubule-associated protein tau in familial MSTD. The mutation is located at the 3' neighboring nucleotide of the GT splice-donor site and disrupts a predicted stem-loop structure. We also report an abnormal preponderance of soluble tau protein isoforms with four microtubule-binding repeats over isoforms with three repeats in familial MSTD. This most likely accounts for our previous finding that sarkosyl-insoluble tau protein extracted from the filamentous deposits in familial MSTD consists only of tau isoforms with four repeats. These findings reveal that a departure from the normal ratio of four-repeat to three-repeat tau isoforms leads to the formation of abnormal tau filaments. The results show that dysregulation of tau protein production can cause neurodegeneration and imply that the FTDP-17 gene is the tau gene. This work has major implications for Alzheimer's disease and other tauopathies.
Insights
A genetic mutation in the tau gene causes familial multiple system tauopathy with presenile dementia (MSTD). This leads to an abnormal ratio of tau protein isoforms, forming toxic tau filaments and causing neurodegeneration.
Area of Science:
- Neuroscience
- Genetics
- Molecular Biology
Background:
- Familial multiple system tauopathy with presenile dementia (MSTD) is a neurodegenerative disease characterized by abundant tau protein pathology.
- MSTD is classified under familial frontotemporal dementias with Parkinsonism linked to chromosome 17 (FTDP-17), a group of inherited dementias with an unknown genetic cause.
Purpose of the Study:
- To identify the genetic basis of familial MSTD.
- To investigate the role of tau protein isoforms in the pathogenesis of MSTD.
Main Methods:
- Genetic analysis to identify mutations in the tau gene.
- Biochemical analysis of tau protein isoforms in affected individuals.
Main Results:
- A G to A transition mutation was identified in the intron following exon 10 of the microtubule-associated protein tau gene in familial MSTD patients.
- This mutation disrupts a predicted stem-loop structure near the splice-donor site.
- An abnormal increase in soluble tau protein isoforms with four microtubule-binding repeats compared to three-repeat isoforms was observed in familial MSTD.
Conclusions:
- The identified mutation in the tau gene is the likely cause of familial MSTD.
- A dysregulated ratio of tau isoforms, specifically a preponderance of four-repeat isoforms, leads to the formation of abnormal tau filaments.
- These findings suggest that tau gene dysregulation can cause neurodegeneration and have implications for Alzheimer's disease and other tauopathies.
More Related Videos
Related Concept Videos
 01:39
Mutations
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
 01:29
Alzheimer Disease l: Introduction
 01:23
Alzheimer Disease ll: Pathophysiology
 01:21
Huntington Disease l: Introduction
 01:22
Dementia l: Introduction