Related Experiment Video
Updated: Aug 2, 2026

Structure and Coordination Determination of Peptide-metal Complexes Using 1D and 2D 1H NMR
Published on: December 16, 2013
Analysis of error propagation from NMR-derived internuclear distances into molecular structure of cyclo-pro-gly
Z Dzakula1, Juranić, M L DeRider
1Department of Biochemistry, University of Wisconsin-Madison, 420 Henry Mall, Madison, Wisconsin, 53706, USA.

Abstract:
Analytical expressions have been derived that translate uncertainties in distance constraints (obtained from NMR investigations) into uncertainties in atom positions in the maximum likelihood (ML) structure consistent with these inputs. As a test of this approach, a comparison was made between test structures reconstructed by the new ML approach, which yields a single structure and a covariance matrix for coordinates, and those reconstructed by metric matrix distance-geometry (MMDG), which yields a family of structures that sample uncertainty space. The test structures used were 560 polyhedra, with edges of arbitrary length containing up to 50 vertices, and one polyhedron, with 100 vertices; randomized distance constraints generated from these structures were used in reconstructing the polyhedra. The uncertainties derived from the two methods showed excellent agreement, and the correlation improved, as expected, with increasingly larger numbers of MMDG structures. This agreement supports the validity of the rapid analytical ML approach, which requires the calculation of only a single structure. As a second test of the ML method, the approach was applied to the determination of uncertainties in the structure of a cyclic dipeptide, cyclo(DL-Pro-Gly) (cPG), derived from NMR cross-relaxation data. The input data were interproton distances calculated from NOEs measured for a solution of the peptide in 2:1 DMSO:H2O at -40 degreesC (so as to yield large negative NOEs). In order to evaluate effects of the quality of the input spectral parameters on the precision of the resulting NMR structure, information from the covalent geometry of cPG was not used in the structure calculations. Results obtained from the analytical ML approach compared favorably with those from the much slower random-walk variant of the Monte Carlo method applied to the same input data. As a third test, the ML approach was used with synthetic structural constraints for a small protein; the results indicate that it will be feasible to use this rapid method to translate uncertainties associated with a given set of distance restraints into uncertainties in atom positions in larger molecules.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Related Concept Videos
 02:29
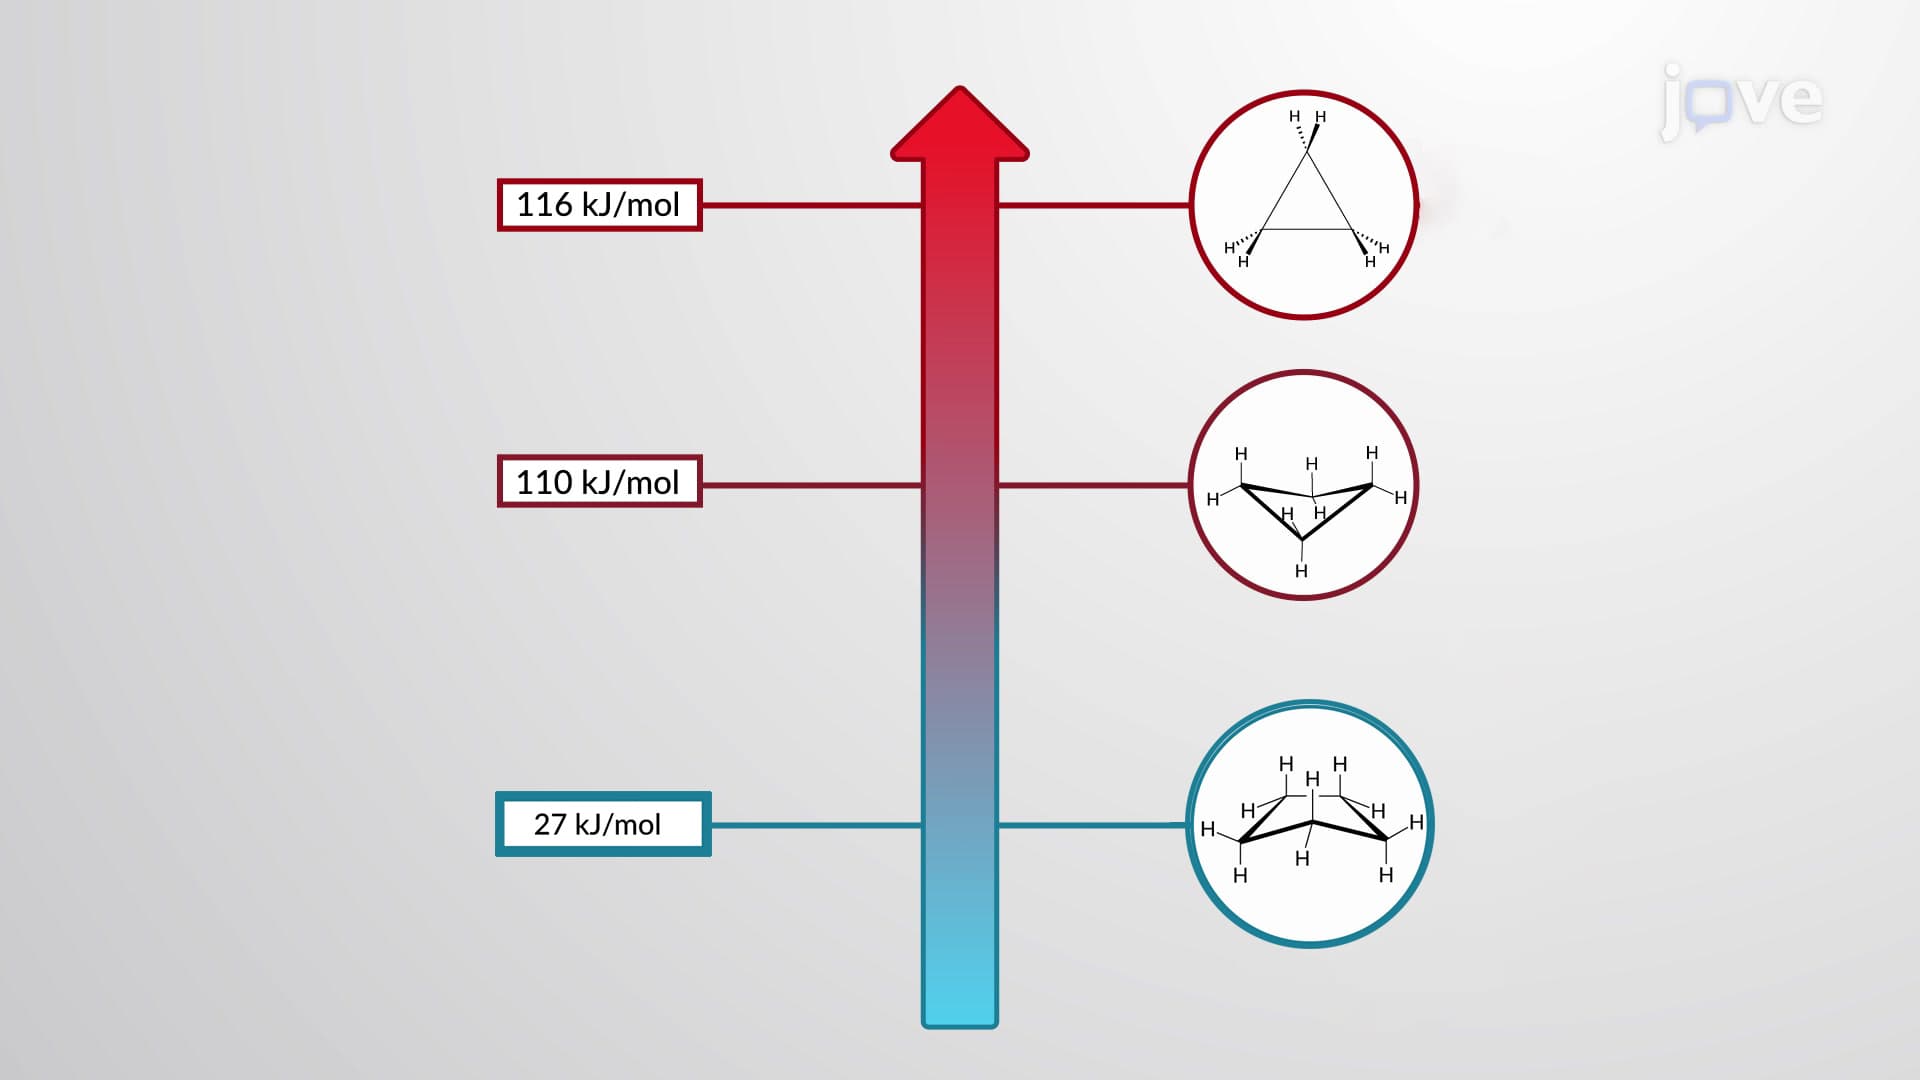
Conformations of Cycloalkanes
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:05
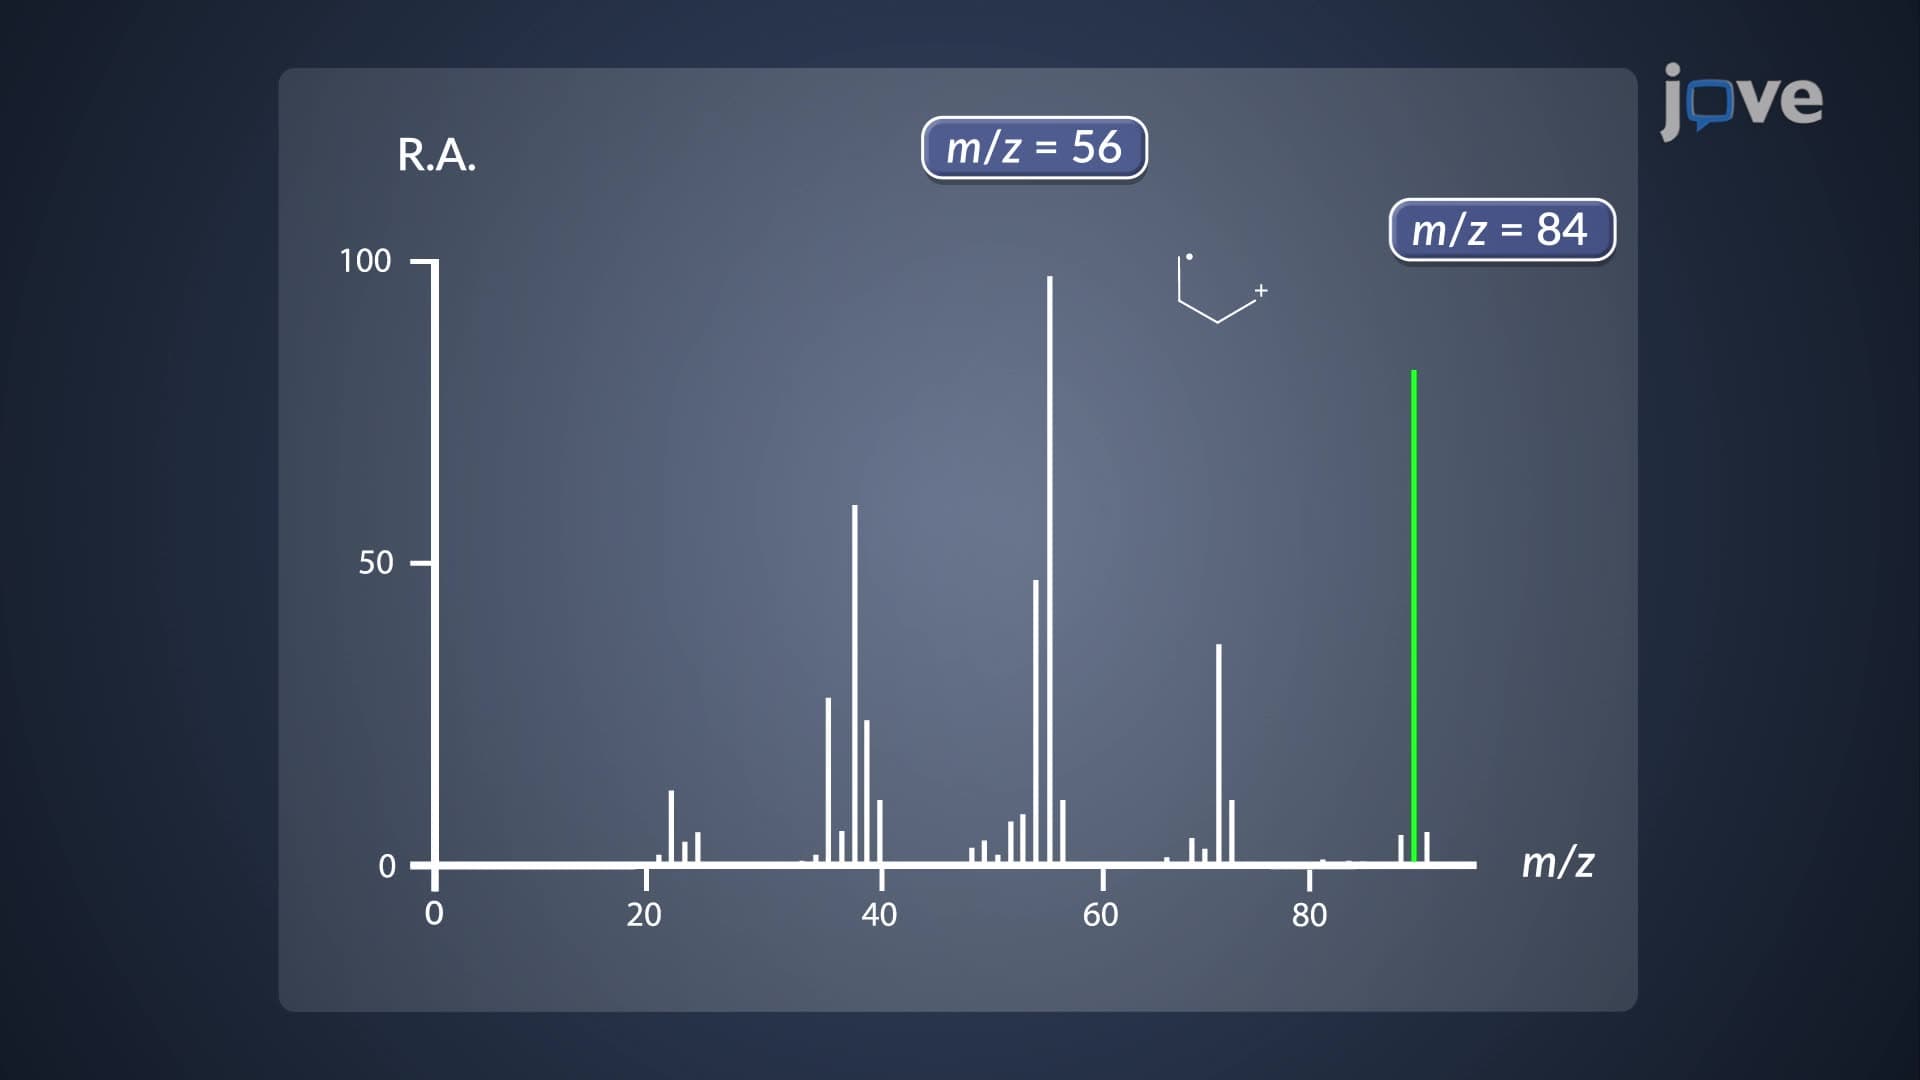
Mass Spectrometry: Cycloalkane Fragmentation
For example, cyclohexane molecular ions have a mass-to-charge ratio (m/z) of 84, which tends to produce a stronger signal than linear alkanes like hexane. This stability comes from...
 00:54
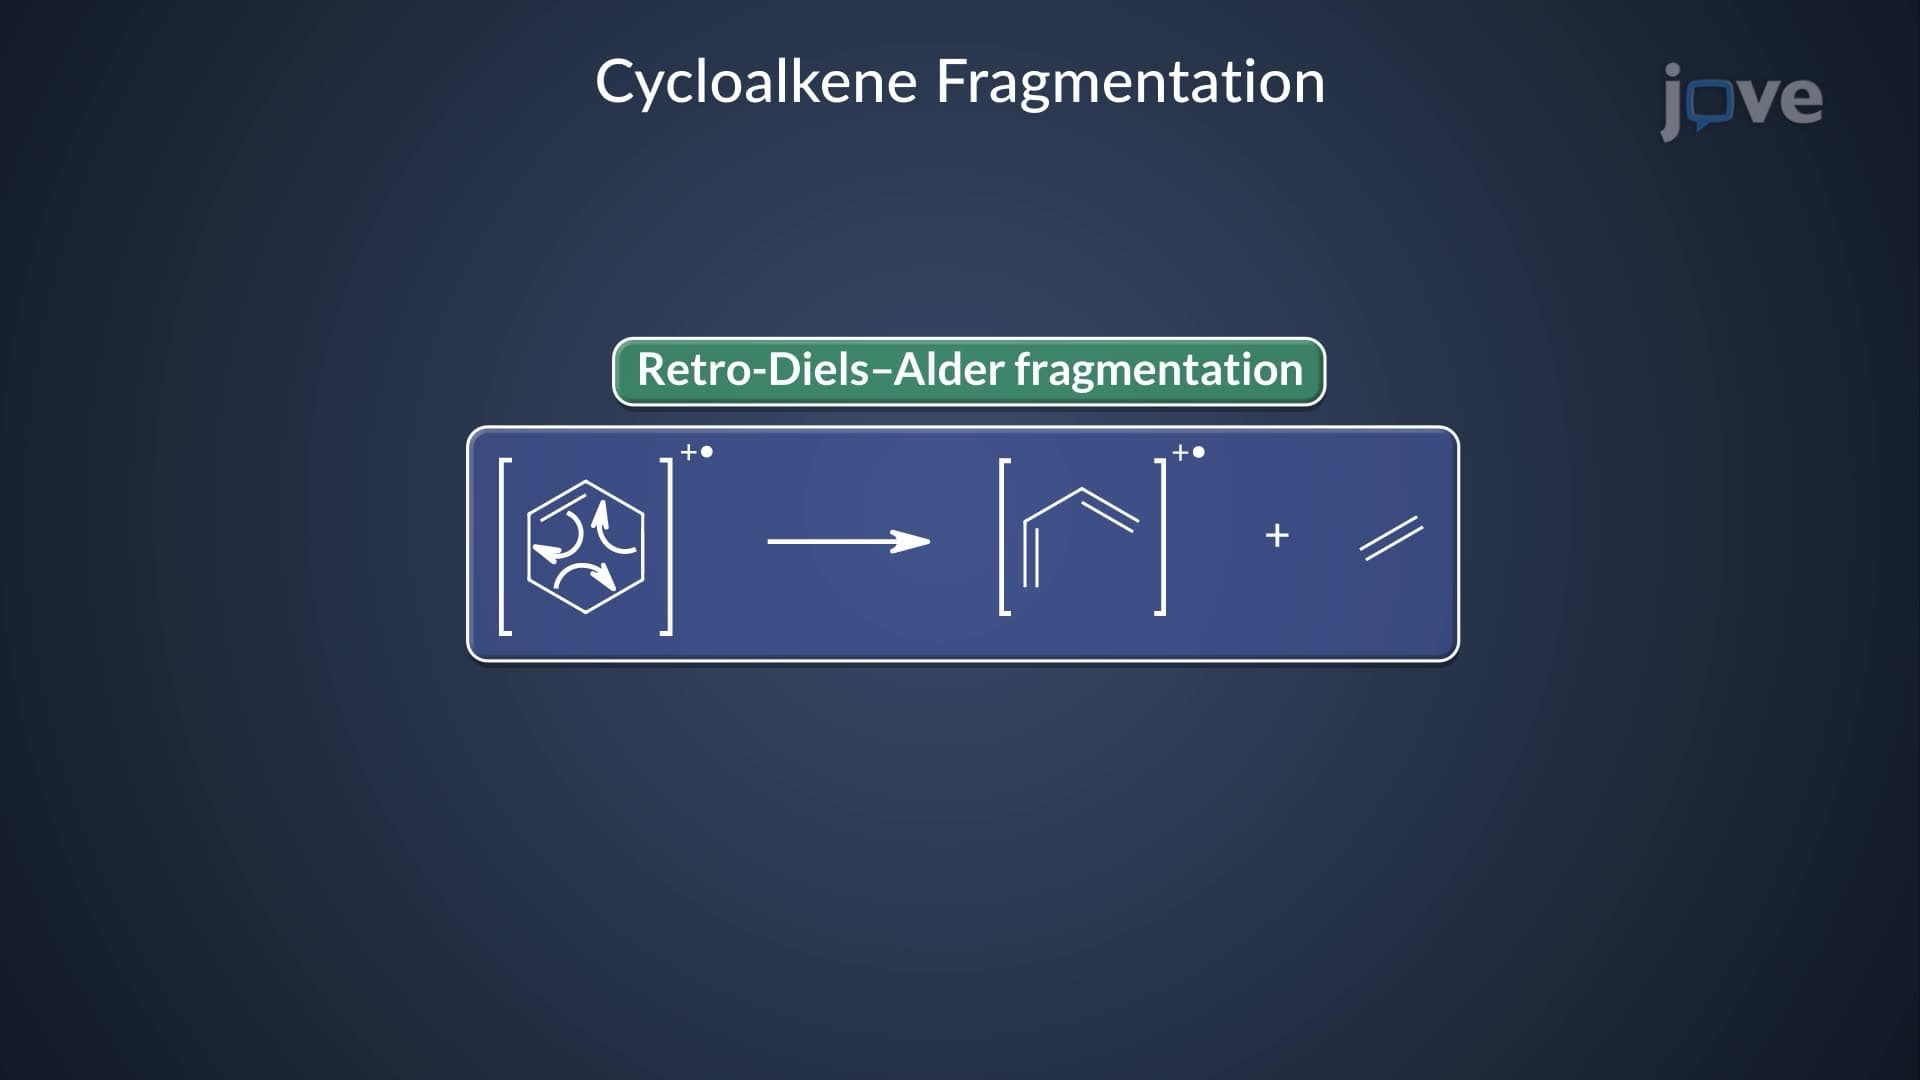
Mass Spectrometry: Cycloalkene Fragmentation