Related Experiment Video
Updated: Aug 27, 2026
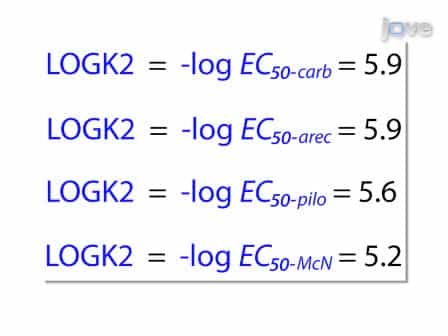
Quantifying Agonist Activity at G Protein-coupled Receptors
Published on: December 26, 2011
Differences in the order of potency for agonists but not antagonists at human and rat adenosine A2A receptors
1Department of Physiology and Pharmacology, Karolinska Institutet, Stockholm, Sweden.

Abstract:
To examine possible species differences in pharmacology, rat adenosine A2A receptors were studied in PC12 (pheochromocytoma) cells, and human receptors in Chinese hamster ovary (CHO) cells transfected with the cloned human A2A receptor cDNA. Using [3H]-5-amino-7(2-phenylethyl)-2-(2-furyl)-pyrazolo[4,3-e]-1,2,4-triazolo [1,5-c]pyrimidine ([3H]-SCH 58261) as radioligand, the estimated Bmax (maximal binding) was 538 and 2085 fmol/mg in CHO and PC12 cells, respectively. The Kd (dissociation constant) values for [3H]-SCH 58261 were 1.05 and 5.6 nM in the two cell types, respectively. The order of potency of antagonists and most agonists was the same in both cell types, but 2-phenylaminoadenosine and 2-chloroadenosine were relatively less potent in PC12 cells than in CHO cells. In the functional assay, using cyclic AMP accumulation, all agonists tested were more potent in CHO than in PC12 cells, but this could not be readily explained by differences in adenylyl cyclase or in the expression of G proteins. As in the case of binding, the relative agonist potencies were similar for most compounds, but 2-phenylaminoadenosine and 2-chloroadenosine were more potent at human A2A receptors in CHO cells than predicted from the data obtained on rat A2A receptors in PC12 cells. Antagonists were approximately equipotent in the two cells. These results show that, despite only small differences in amino acid sequences and no difference in antagonist pharmacology, the relative order of potency of receptor agonists can differ between species homologues of the adenosine A2A receptor.
More Related Videos
Related Concept Videos
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:26
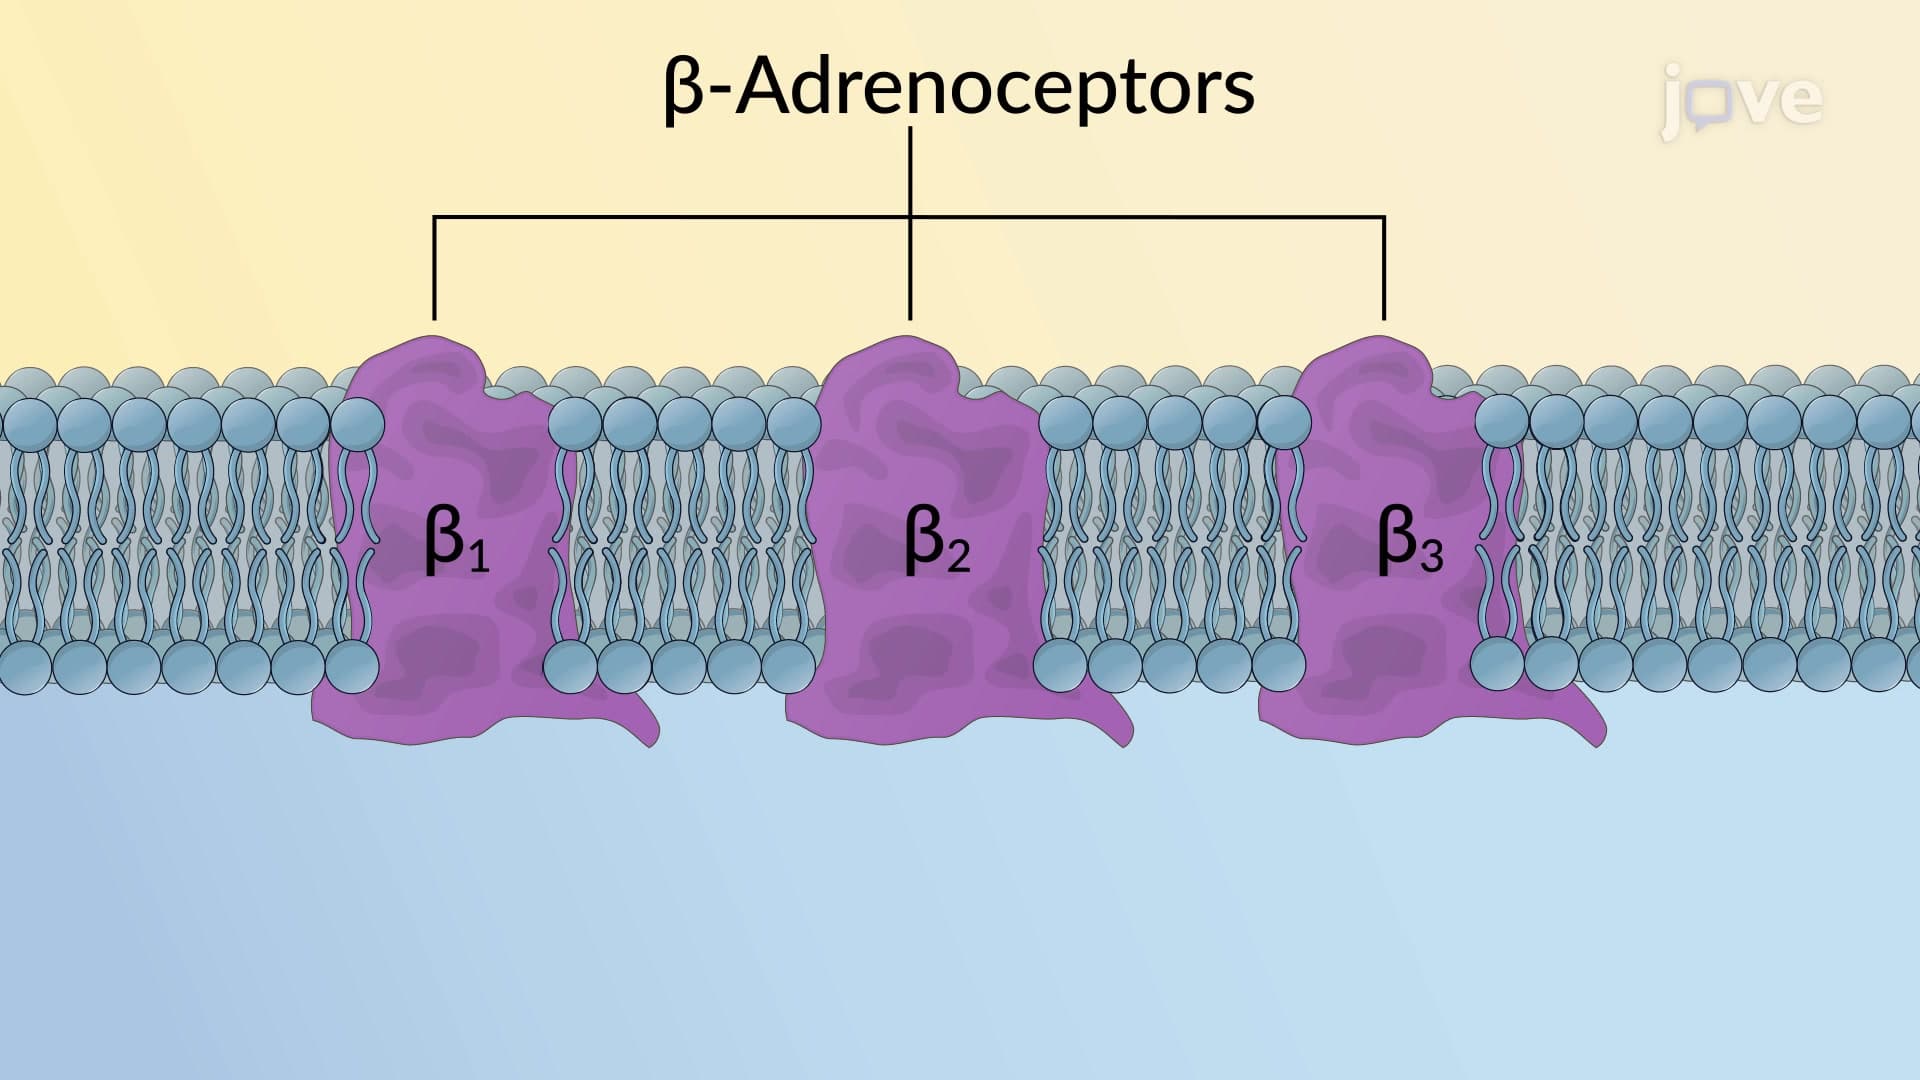
Adrenergic Receptors: β Subtype
Isoprenaline > Adrenaline > Noradrenaline
Neurotransmitter binding to these receptors causes activation of adenylyl cyclase resulting in increased concentrations of cAMP and modulation of calcium ion channels within the cell. They are further classified into β1, β2, and β3 subtypes.
β1-adrenoceptors: β1-adrenoceptors have equal affinities for...
 01:30
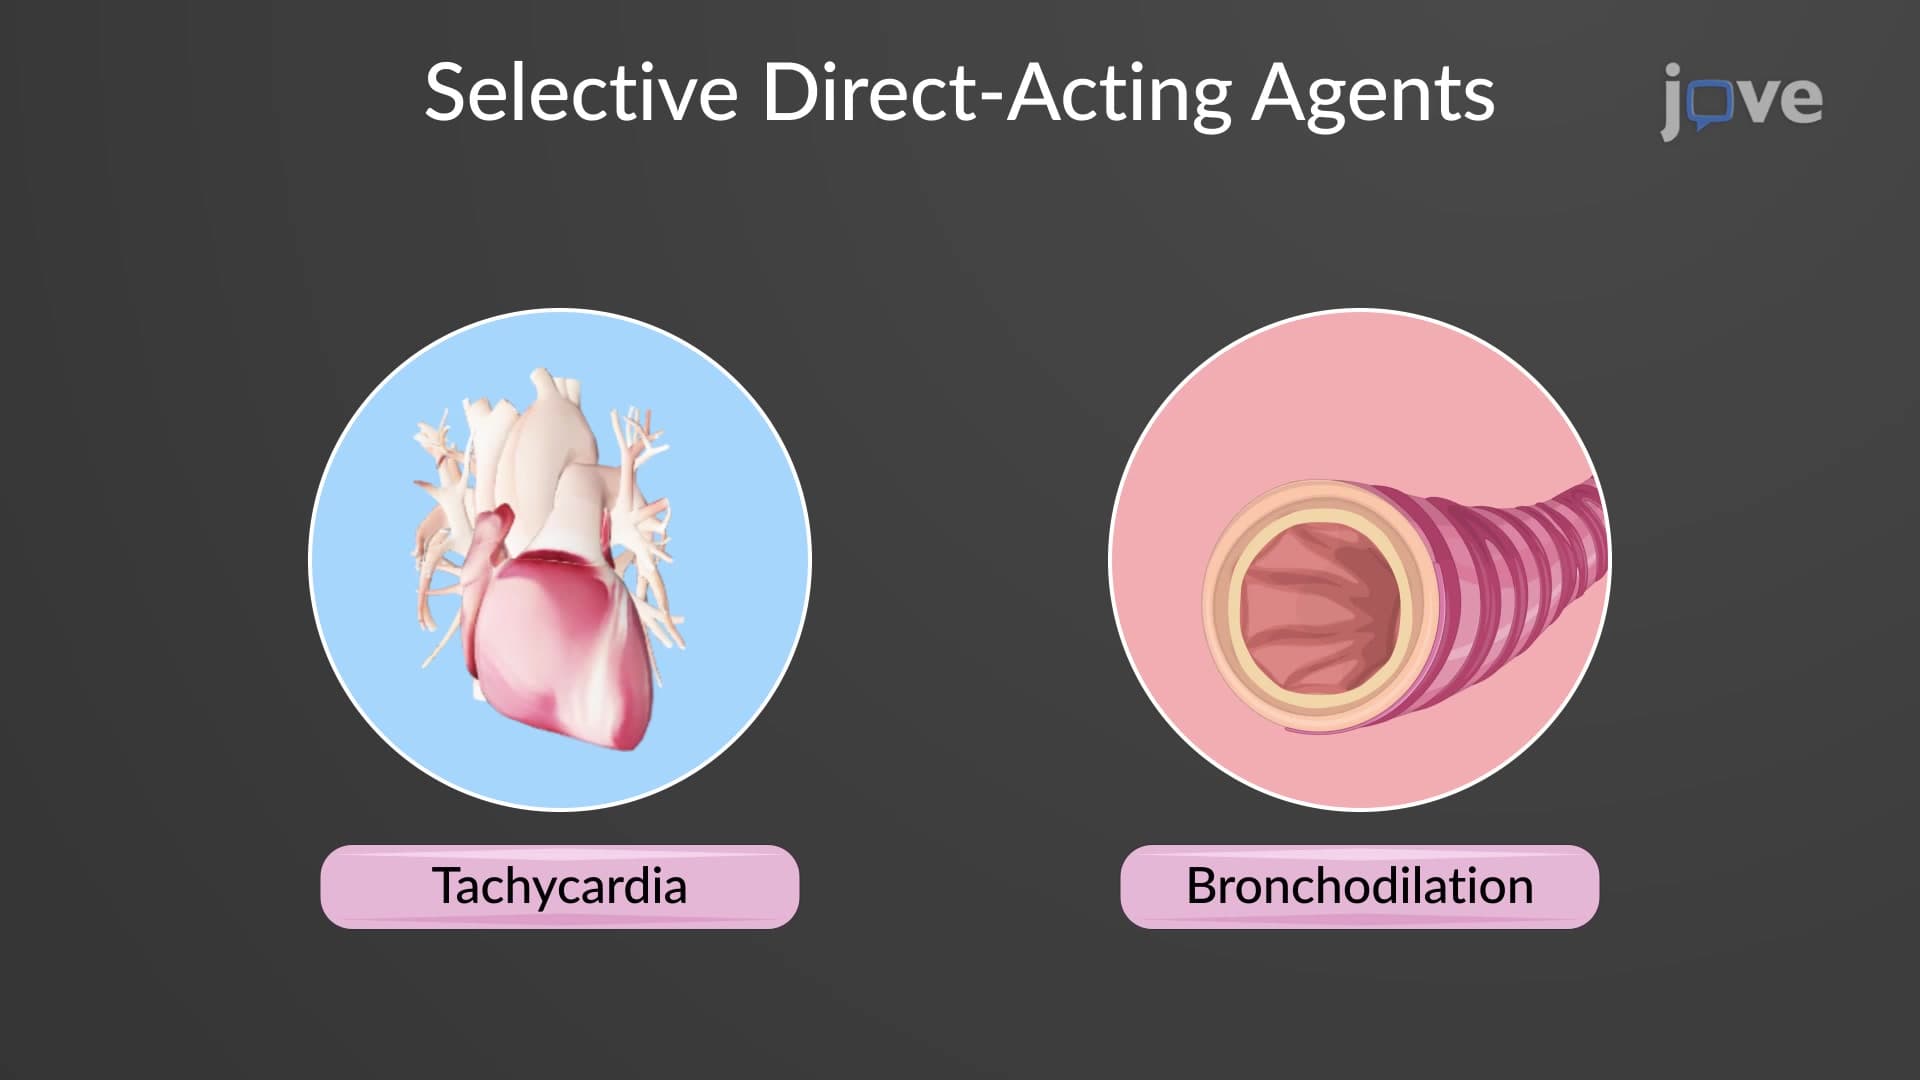
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
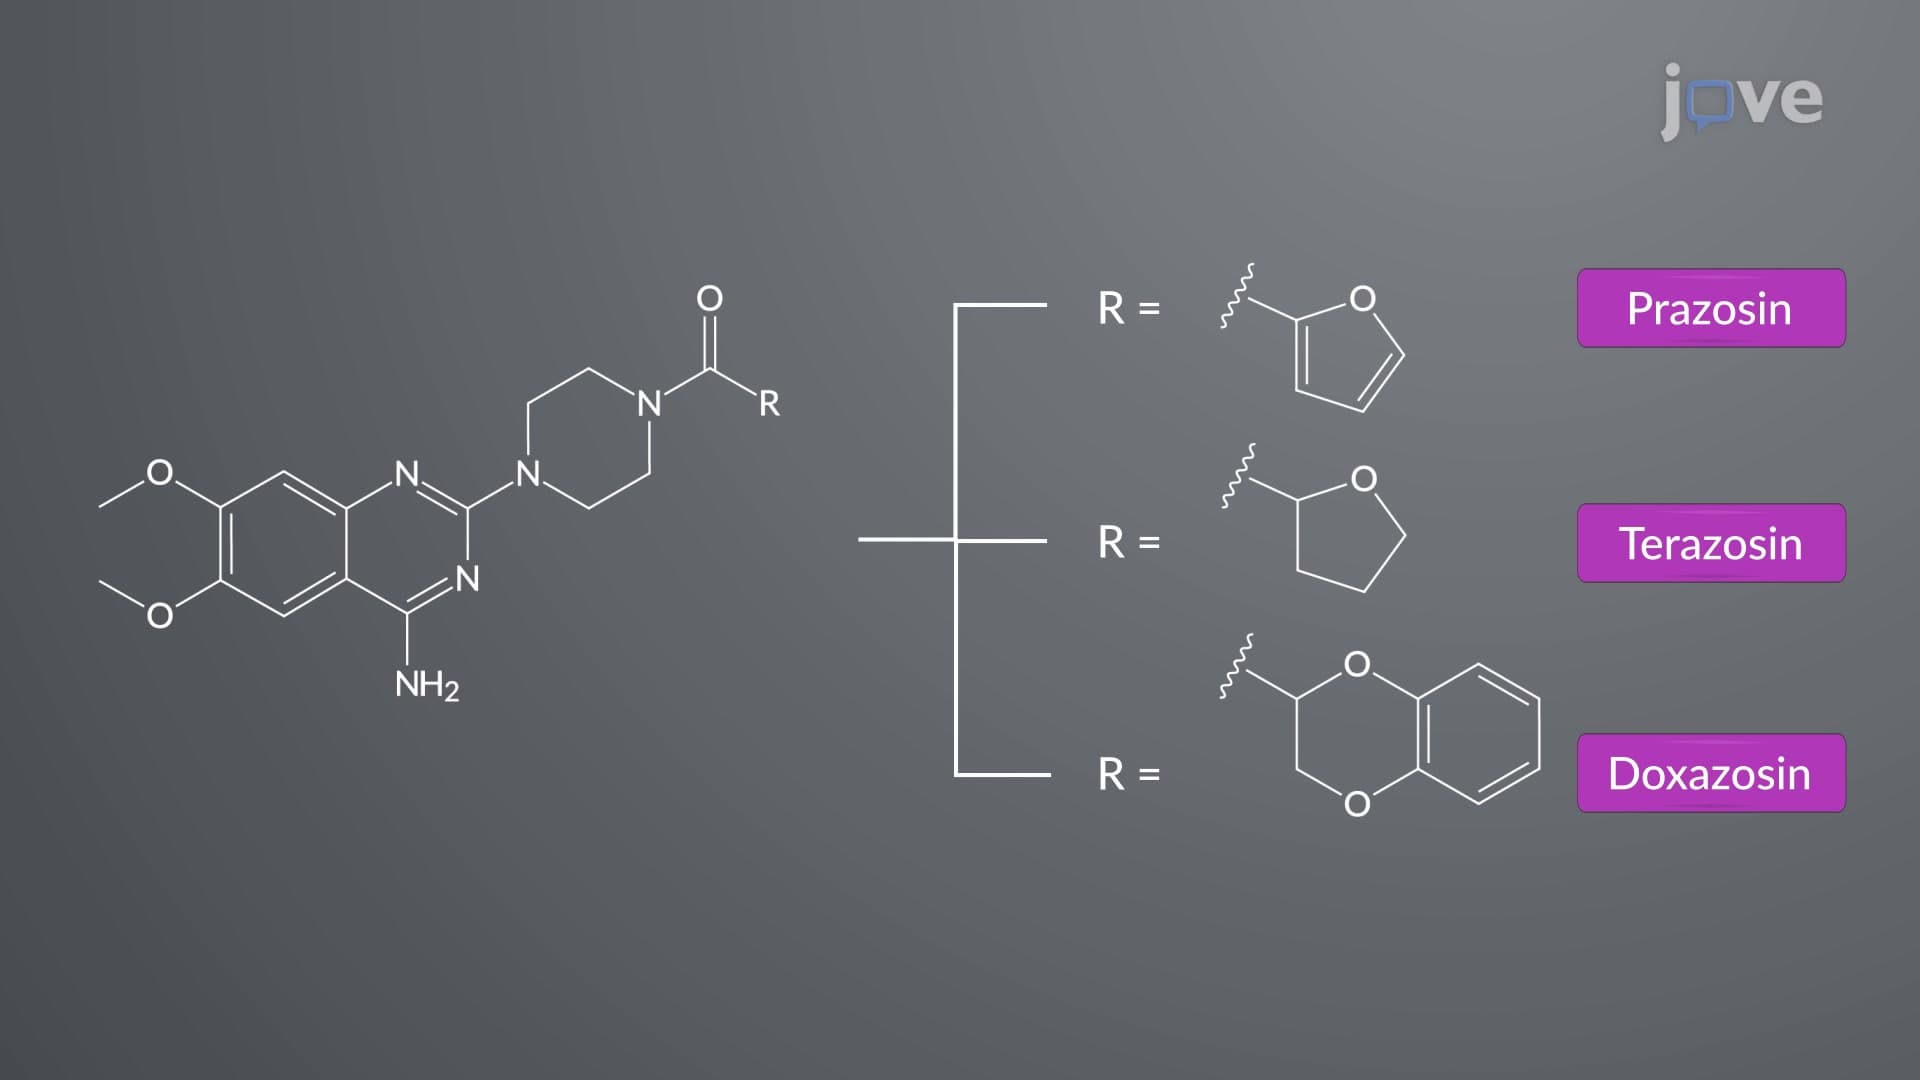
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
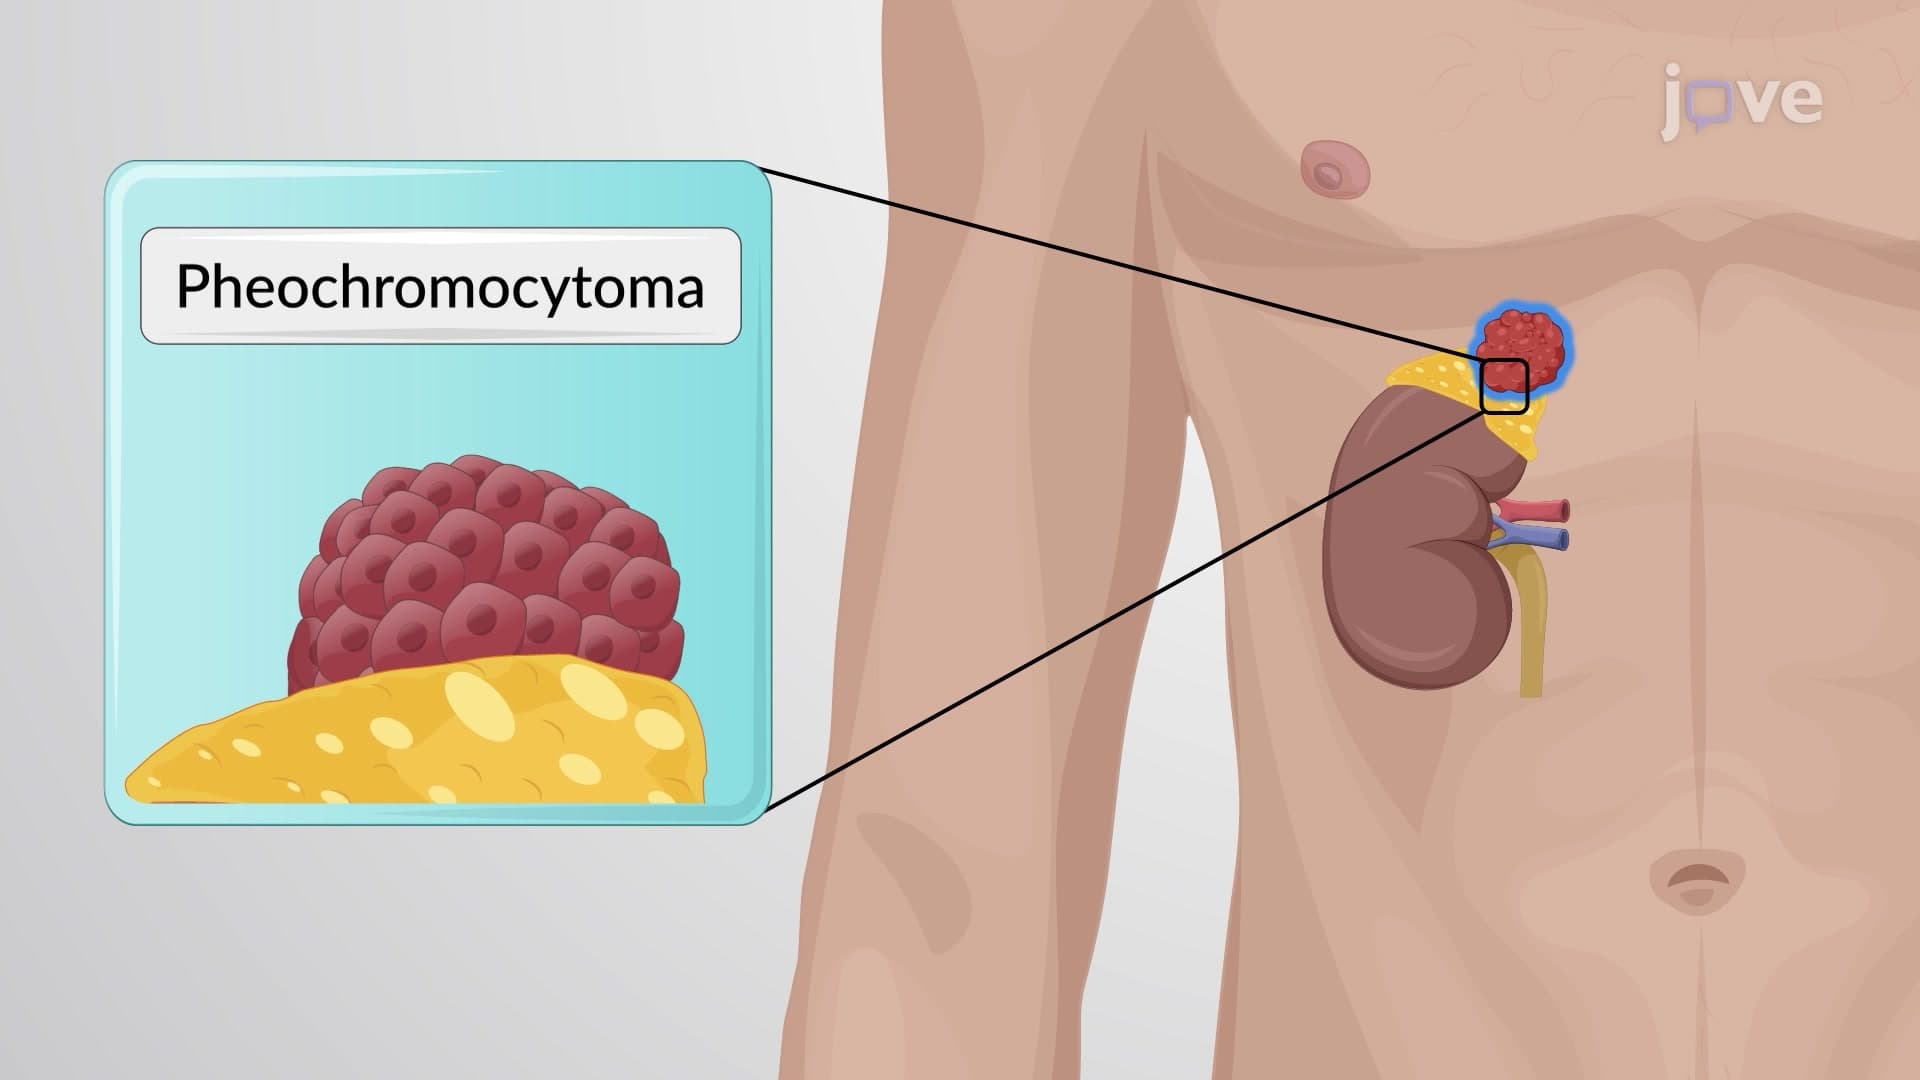
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...