Video Experimental Relacionado
Updated: Feb 25, 2026
09:40

Novel Sequence Discovery by Subtractive Genomics
Published on: January 25, 2019
9.2K
Secuenciación y ensamblaje de novo de 150 genomas de Dinamarca como referencia de población
Lasse Maretty1, Jacob Malte Jensen2,3, Bent Petersen4
1Bioinformatics Centre, Department of Biology, University of Copenhagen, 2200 Copenhagen, Denmark.
Nature
|July 27, 2017

Resumen
La secuenciación de alta cobertura permite excelentes ensamblajes de genoma de novo, revelando extensas variantes estructurales. Esto hace avanzar los estudios de variación genética y las iniciativas de medicina de precisión.
Área de la Ciencia:
- La genómica
- La bioinformática
- La genética humana
Sus antecedentes:
- Los métodos actuales para identificar la variación genética, como el mapeo de lectura corta y el ensamblaje local, son limitados.
- Estos enfoques a menudo pasan por alto las variantes estructurales y las variaciones en regiones genómicas complejas.
- El ensamblaje del genoma de novo a gran escala es crucial para el análisis genético integral.
Objetivo del estudio:
- Demostrar la viabilidad de construir ensamblajes de genoma de novo de alta calidad utilizando bibliotecas de pares de pares.
- Identificar un catálogo completo de variantes estructurales, incluidas las nuevas inserciones.
- Mejorar el poder de los estudios de cartografía de asociación para rasgos y enfermedades complejas.
Principales métodos:
- Utilizó secuenciación de alta cobertura con bibliotecas de pares de mate de hasta 20 kilobases.
- Realizó el ensamblaje de novo en 150 individuos (50 tríos) del proyecto GenomeDenmark.
- Comparación de la calidad de ensamblaje con las tecnologías de secuenciación de larga lectura.
Principales resultados:
- Lograr ensamblajes de novo de calidad comparable a los costosos métodos de lectura larga.
- Identificó un rico conjunto de variantes estructurales, incluyendo numerosas nuevas inserciones.
- Se generaron 100 haplotipos complejos de histocompatibilidad principales completamente resueltos y se resolvieron porciones significativas del cromosoma Y.
Conclusiones:
- La secuenciación de alta cobertura con bibliotecas extendidas de pares de pares es efectiva para el ensamblaje del genoma de novo.
- El catálogo de variantes generado mejora el descifrado de las señales de mapeo de asociaciones.
- El estudio proporciona un genoma de referencia regional valioso, que apoya las iniciativas de medicina de precisión.
Más Videos Relacionados

Videos de Conceptos Relacionados
 03:36
Genome Annotation and Assembly
21.2K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
21.2K
 03:00
Next-generation Sequencing
99.4K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
99.4K
 01:57
Sanger Sequencing
775.9K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
775.9K
 03:21
RNA-seq
12.2K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
12.2K
 02:02
Genomics
41.1K
Genomics is the science of genomes: it is the study of all the genetic material of an organism. In humans, the genome consists of information carried in 23 pairs of chromosomes in the nucleus, as well as mitochondrial DNA. In genomics, both coding and non-coding DNA is sequenced and analyzed. Genomics allows a better understanding of all living things, their evolution, and their diversity. It has a myriad of uses: for example, to build phylogenetic trees, to improve productivity and...
41.1K
 01:05
Maxam-Gilbert Sequencing
13.2K
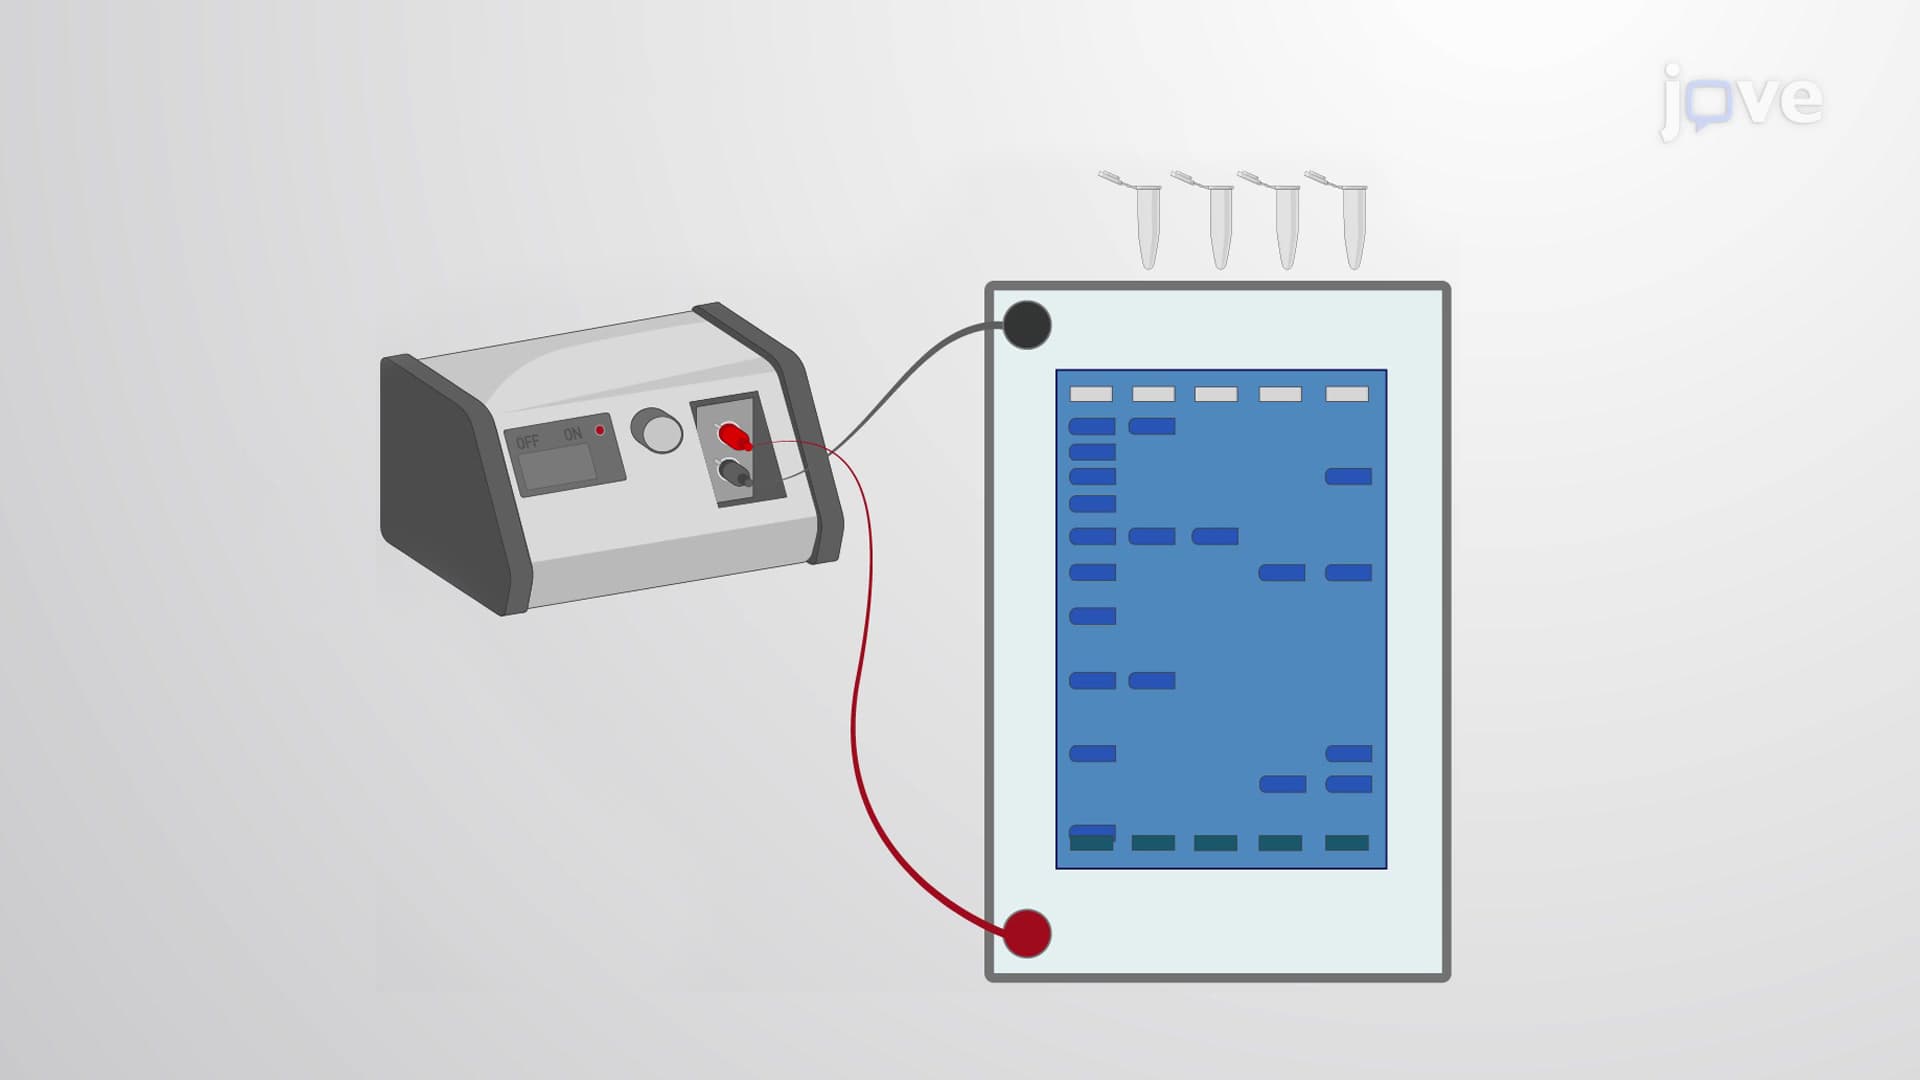
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
13.2K