関連する実験動画
Updated: Jul 15, 2026
10:05

In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
異例のCFTR依存のHCO3-輸送が,システィックフィブロシスに関連した変異で認められる
J Y Choi1, D Muallem, K Kiselyov
1Department of Physiology, The University of Texas Southwestern Medical Center, Dallas 75390, USA.
Nature
|March 10, 2001

まとめ
胞性線維症 (CF) は,胞性線維症のトランスメブラン伝導性調節器 (CFTR) の変異から生じる. CFTRの二酸化炭素 (HCO3-) 輸送の障害は,塩化物 (Cl-) 経路機能だけでなく,CFの病原性には極めて重要です.
科学分野:
- バイオケミストリー バイオケミストリー
- 細胞生物学 細胞生物学
- 遺伝学 遺伝学とは
背景:
- システィック線維症 (CF) は,システィック線維症のトランスメブラン伝導性調節器 (CFTR) 遺伝子の変異によって引き起こされる遺伝疾患です.
- CFTRの主な既知の機能は,循環AMP調節塩化物 (Cl-) チャンネルであるが,CFを引き起こすいくつかの変異は,このチャンネル活動を損なわない.
- CFTRはまた,他のトランスポーター,特に二酸化炭素 (HCO3-) 輸送を調節します,これは,分泌性上皮質における液体のpHと粘度を維持するために重要です.
研究 の 目的:
- CFTR依存バイカーボネート (HCO3-) 輸送がシスティック線維症 (CF) で果たす役割を調査する.
- 異なるCFTR変異,特に臓不全と関連する変異が,HCO3-輸送にどのように影響するかを調べる.
- 分泌性上皮質の機能におけるHCO3-輸送の重要性とそのCF病理生理学への貢献を明らかにする.
主な方法:
- 維持されたまたは正常なCl-チャネル活性を持つCFTR変異体の分析.
- 様々なCFTR変異体におけるCl(-) 結合したHCO3-輸送能力の評価.
- HCO3 - 輸送機能と臨床的なCFのフェノタイプ,特に臓不全および充足の相関.
主要な成果:
- 臓不全に関連したCFTR変異体は,HCO3-輸送の完全な欠如を示した.
- 臓不全に関連したCFTR変異体は,HCO3-輸送が著しく減少したことを示した.
- これらの発見は,保存されたCl-チャネル機能に関係なく,HCO3-輸送の障害がCFの重要な特徴であることを強調しています.
結論:
- CFTR媒介による二酸化炭素 (HCO3-) 輸送は,分泌性上皮質の適切な機能に不可欠である.
- CFTR変異体によるHCO3-輸送の欠陥は,システィック線維症 (CF) の疾患メカニズムに大きく寄与する.
- HCO3 - トランスポートの標的化または理解は,CF患者,特に特定の変異型を有する患者にとって新しい治療の機会を提供することができます.
関連する概念動画
 01:15
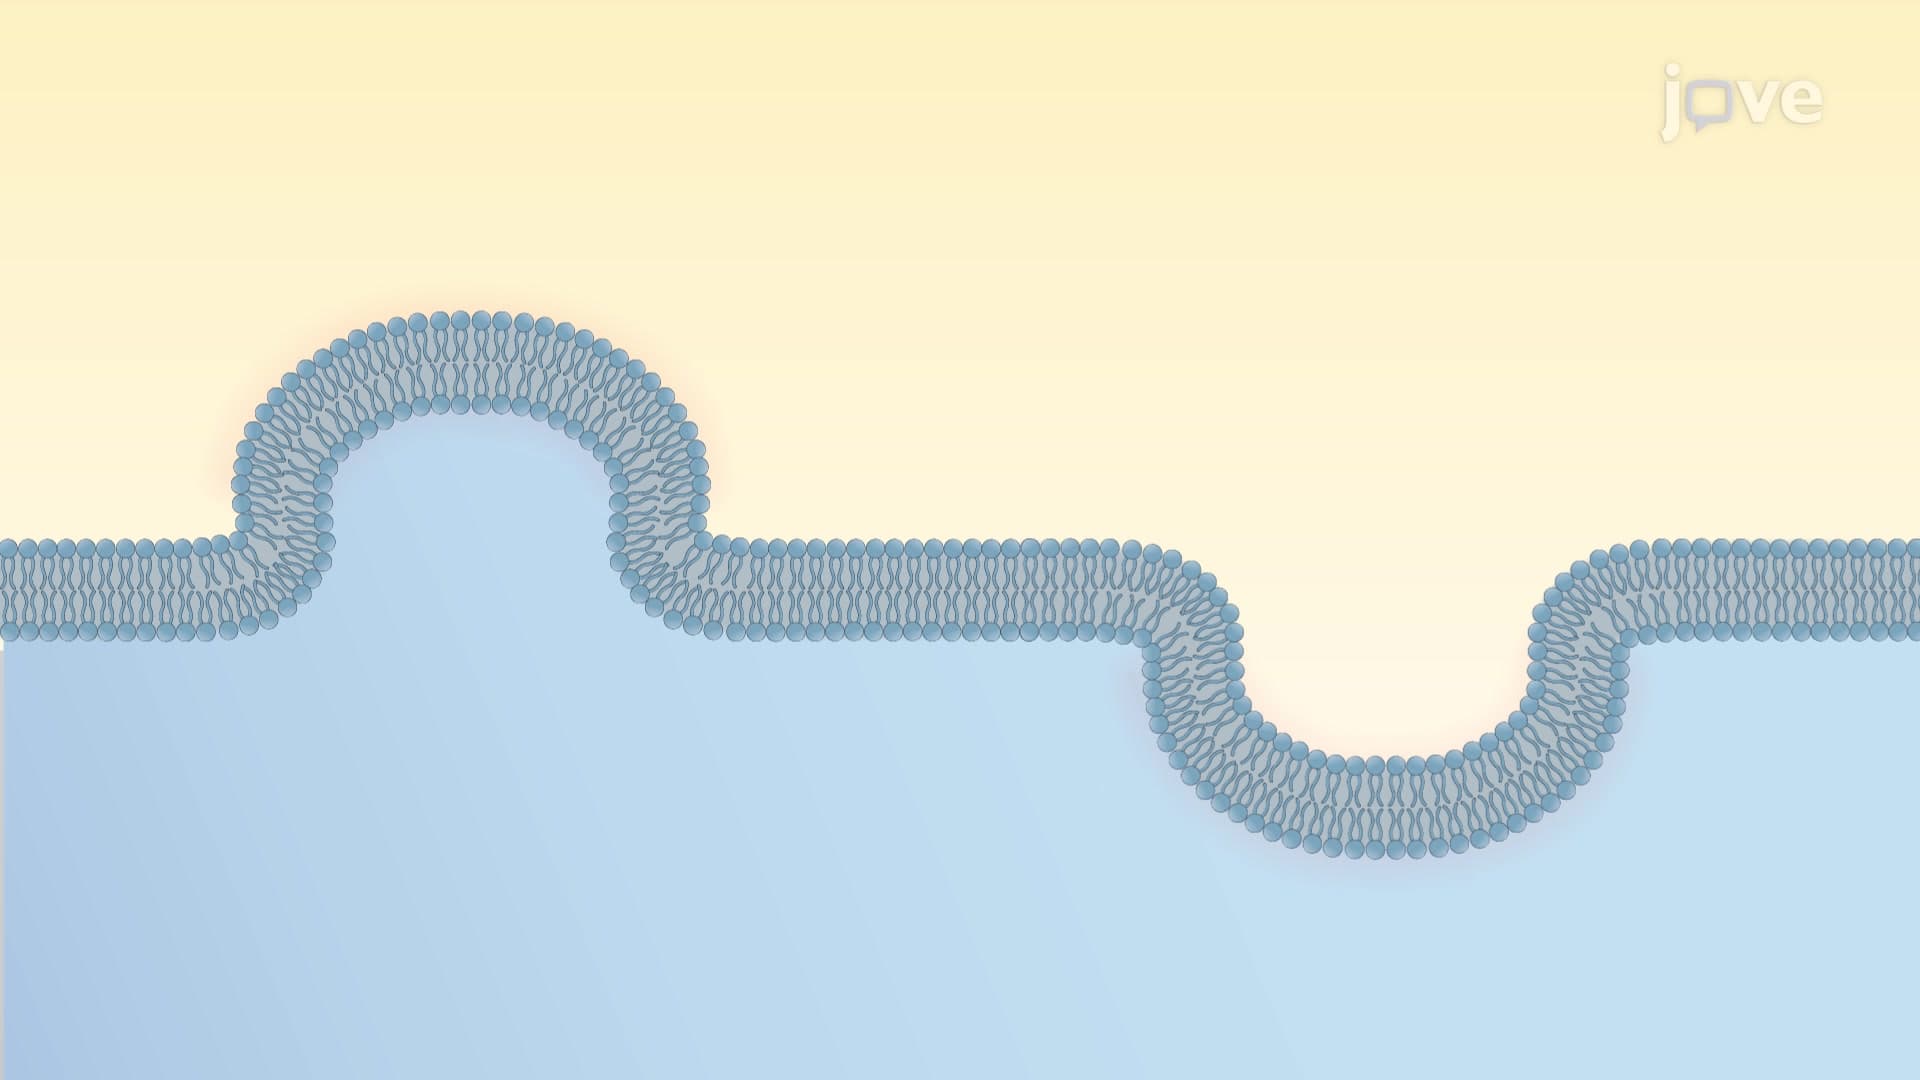
Mechanisms of Membrane-bending
The living membranes are flexible due to their fluid mosaic nature; however, their bending into different shapes is an active process regulated by specific lipids and proteins. The membrane bending can be transient as seen in vesicles or stable for a long time as in microvilli. Cells regulate the size, location, and duration of the membrane curvature.
Membrane bending can happen due to intrinsic changes in lipid composition or extrinsic association with different proteins. The proteins involved...
Membrane bending can happen due to intrinsic changes in lipid composition or extrinsic association with different proteins. The proteins involved...
 01:23
Transcellular Transport of Solutes
Transcellular transport of solutes is the movement of substances like monosaccharides and amino acids through polarized cells. This transport mechanism is primarily seen in epithelial and endothelial cells aided by membrane transport proteins such as channels and transporters. The tight junctions between these cells confine the membrane proteins to the two sides of the cell. The epithelial cells have distinct apical and basolateral domains. In contrast, the endothelial cells show the luminal...
 01:45
Vesicular Tubular Clusters
After budding out from the ER membrane, some COPII vesicles lose their coat and fuse with one another to form larger vesicles and interconnected tubules called vesicular tubular clusters or VTCs. These clusters constitute a compartment at the ER-Golgi interface known as ERGIC (Endoplasmic Reticulum Golgi Intermediate Compartment). The ERGIC is a mobile membrane-bound cargo transport system that sorts proteins secreted from ER and delivers them to the Golgi.
With the help of motor proteins such...
With the help of motor proteins such...
 01:19
Membrane Asymmetry Regulating Transporters
Enzymes like flippase, floppase, and scramblase transfer phospholipids from one layer to another in the membrane, thereby affecting membrane asymmetry.
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...
 01:23
Cystic Fibrosis: Pathogenesis
Cystic fibrosis (CF), an autosomal recessive disorder, significantly affects the function of exocrine glands. This genetically inherited disease is characterized by the production of thick and sticky mucus, which can severely affect various organs and systems in the body.
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Cystic fibrosis (CF) is an autosomal recessive disorder that predominantly affects individuals of Northern European descent, occurring at a rate of 1 in 3500. It is caused by a genetic mutation in a gene on chromosome 7, most commonly the ΔF508 mutation, that codes for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. This results in thicker mucus secretions and obstruction pathologies in multiple organs, including the lungs and sinuses.
Sinus disease and chronic sinusitis...
Sinus disease and chronic sinusitis...