
分子ダイナミクスシミュレーションとドッキング実験によるβ-分泌酵素の活性部位プロトネーション状態の決定:構造ベースの阻害剤設計への影響
1School of Chemistry and Molecular Engineering, and Center for Molecular Catalysis, Seoul National University, Seoul 151-747, South Korea. hwangseo@snu.ac.kr
Journal of the American Chemical Society
|December 25, 2003

まとめ
アルツハイマー病において重要なメマプシン2 (BACE) は,曖昧なアスパルチン酸プロトネーション状態を持っています. シミュレーションにより,Asp32ではなくAsp228が,阻害剤の水素結合受容体として作用し,薬剤設計を導くことが明らかになった.
科学分野:
- バイオケミストリー バイオケミストリー
- 神経科学は神経科学である.
- ドラッグ・ディスカバリー・ドリッグ・ディスカバリー・ドリッグ・ディスカバリー・ドリッグ・ディスカバリー
背景:
- メマプシン2 (BACE) はβ-アミロイドペプチドの生成に関与するβ-セクレターゼであり,アルツハイマー病の病原性における重要な要因である.
- BACEにおける触媒アスパルティック酸残留物 (Asp32およびAsp228) の正確なプロトネーション状態は, BACEの酵素活性には極めて重要であるが,曖昧である.
- これらのプロトネーション状態を理解することは,効果的なBACE阻害剤の開発に不可欠です.
研究 の 目的:
- 分子動力学シミュレーションを用いて,メマプシン2 (BACE) のアスプダイアドの好ましいプロトネーション状態を決定する.
- 強力な阻害剤と結合するAsp32とAsp228の役割を明らかにする.
- 新型BACE阻害剤の構造ベースの設計のための洞察を提供するために.
主な方法:
- BACEの2つの独立した分子動力学 (MD) シミュレーションは,阻害剤OM99-2との複合体である.
- 阻害剤と活性部位残留物の間の水素結合形成と安定性の分析.
- エネルギーと構造の特徴を評価するために,新しい強力な阻害剤とのドッキング実験.
主要な成果:
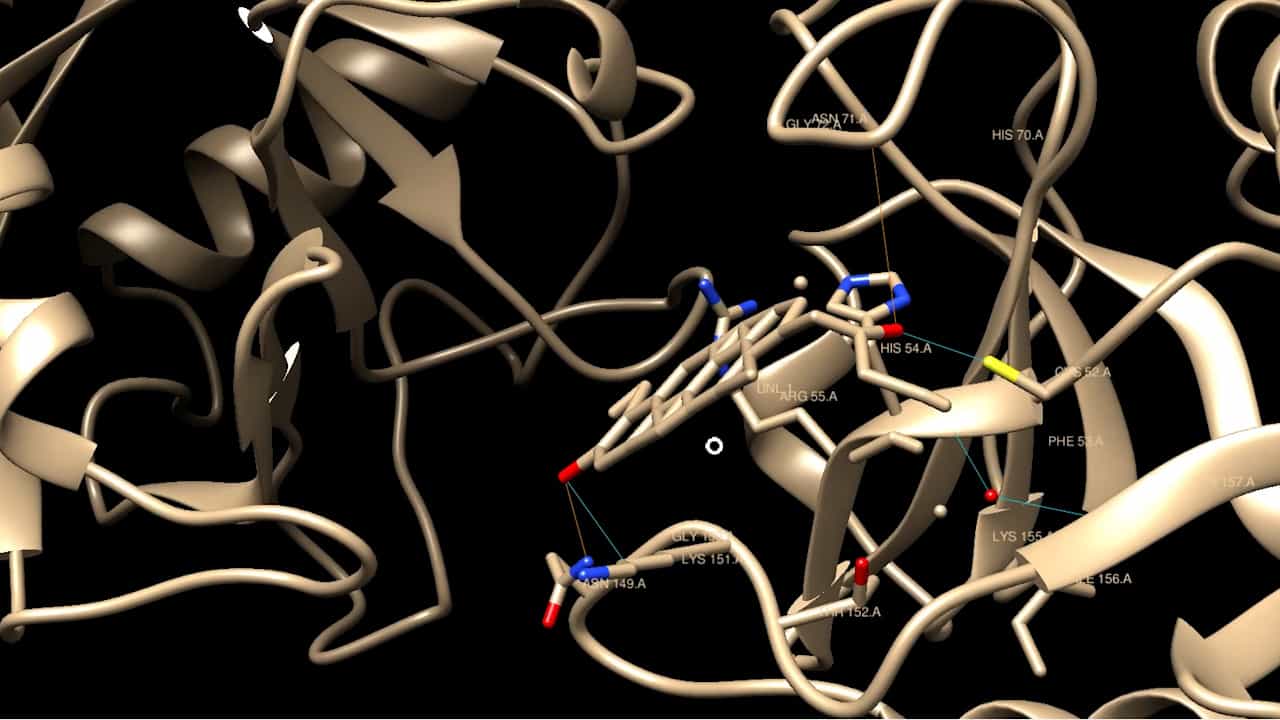
- MDシミュレーションによると,Asp32は中性であり,Asp228はイオン化され,阻害剤のヒドロキシル群との強い水素結合を維持しています.
- このプロトネーション状態は,X線結晶構造と一致して,エネルギー的にも構造的にも好ましい.
- ドッキング研究は,Asp228が主要な水素結合受容体として作用することを示唆し,発見を裏付けています.
結論:
- この研究では,Asp32ではなくAsp228が,BACE阻害剤の主要な水素結合受容体として機能することを示唆しています.
- この発見は,BACEを標的とした新しいアルツハイマー病の治療法の構造ベースの設計と発見に大きな意味を持っています.
- BACE活性部位残留物のプロトネーション状態を明らかにすることは,阻害剤の有効性を最適化するために極めて重要です.
関連する概念動画
 01:08
Allosteric Regulation
Allosteric regulation of enzymes occurs when the binding of an effector molecule to a site that is different from the active site causes a change in the enzymatic activity. This alternate site is called an allosteric site, and an enzyme can contain more than one of these sites. Allosteric regulation can either be positive or negative, resulting in an increase or decrease in enzyme activity. Most enzymes that display allosteric regulation are metabolic enzymes involved in the degradation or...
 01:30
Enzyme Inhibition
Inhibitors are molecules that reduce enzyme activity by binding to the enzyme. In a normally functioning cell, enzymes are regulated by a variety of inhibitors. Drugs and other toxins can also inhibit enzymes. Some inhibitors bind to the enzyme’s active site, while others inhibit enzymatic activity by binding to other sites on the protein structure.
 02:40
Ligand Binding Sites
Proteins are dynamic macromolecules that carry out a wide variety of essential processes; however, the activities of most proteins depend on their interactions with other molecules or ions, known as ligands.
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
The equilibrium binding constant (Kb) quantifies the strength of a protein-ligand interaction. Kb can be calculated as follows when the reaction is at equilibrium:
 01:29
The Two-State Receptor Model
The two-state receptor model explains a drug's interaction with receptors, such as G protein-coupled receptors and ligand-gated ion channels, to induce or inhibit a biological response. When no natural ligands are present, a receptor exists in an equilibrium of inactive (Ri) and active (Ra) conformations. The inactive form does not produce a response, while the active form generates a basal effect known as constitutive activity.
The binding affinity of a drug determines its interaction with one...
The binding affinity of a drug determines its interaction with one...
 01:28
Structure-Activity Relationships and Drug Design
Drug design is a dynamic field that involves discovering and developing new medications based on specific biological targets. This process heavily relies on structure-activity relationships (SAR) and quantitative structure-activity relationships (QSAR) to guide the design and optimization of efficient drugs.
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...