

コンピュータシミュレーションによる分子結晶表面の自由エネルギー: テトラチオフェンへの応用
1Dipartimento di Chimica, Materiali e Ingegneria Chimica Giulio Natta, Politecnico di Milano, Via L. Mancinelli 7, 20131 Milano, Italy.

私たちは,分子結晶の表面自由エネルギーを計算するためのシミュレーションプロトコルを開発しました. この方法は,多形性などの結晶の性質を理解するために不可欠な温度依存のエントロピー効果を明らかにします.
科学分野:
- 材料科学 材料科学とは
- コンピューティング・ケミストリー
- クリスタログラフィーです.
背景:
- 表面自由エネルギーは,ポリモルフィズムと結晶の成長を含む分子結晶の振る舞いを理解するために重要である.
- 既存の静的シミュレーション方法は,温度に依存する効果をしばしば無視しています.
研究 の 目的:
- 分子結晶の表面自由エネルギーを評価するための一般的なシミュレーションプロトコルを提示する.
- 温度に依存するエントロピック貢献が表面エネルギーに及ぼす影響を調査する.
主な方法:
- 一般的なシミュレーションプロトコルを開発し,適用しました.
- 2つのテトラチオフェンポリモルフの選択された表面に対する評価された表面自由エネルギー.
- 温度に依存するエントロピック貢献をシミュレーションに組み込みました.
主要な成果:
- シミュレーションプロトコルは,表面の自由エネルギーを評価することに成功しました.
- 表面自由エネルギーに対する温度に依存する有意なエントロピック貢献が特定されました.
- これらのエントロピー効果は,伝統的な静的シミュレーションではしばしば省略されます.
結論:
- 開発されたシミュレーションプロトコルは,表面自由エネルギーの計算により包括的なアプローチを提供します.
- エントロピーの貢献を考慮することは,分子結晶表面の正確なモデリングに不可欠です.
- この研究は,表面エネルギーによって影響されるポリモルフィズムや結晶の成長などの現象の洞察を提供します.
さらに関連する動画
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
関連する概念動画
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:13
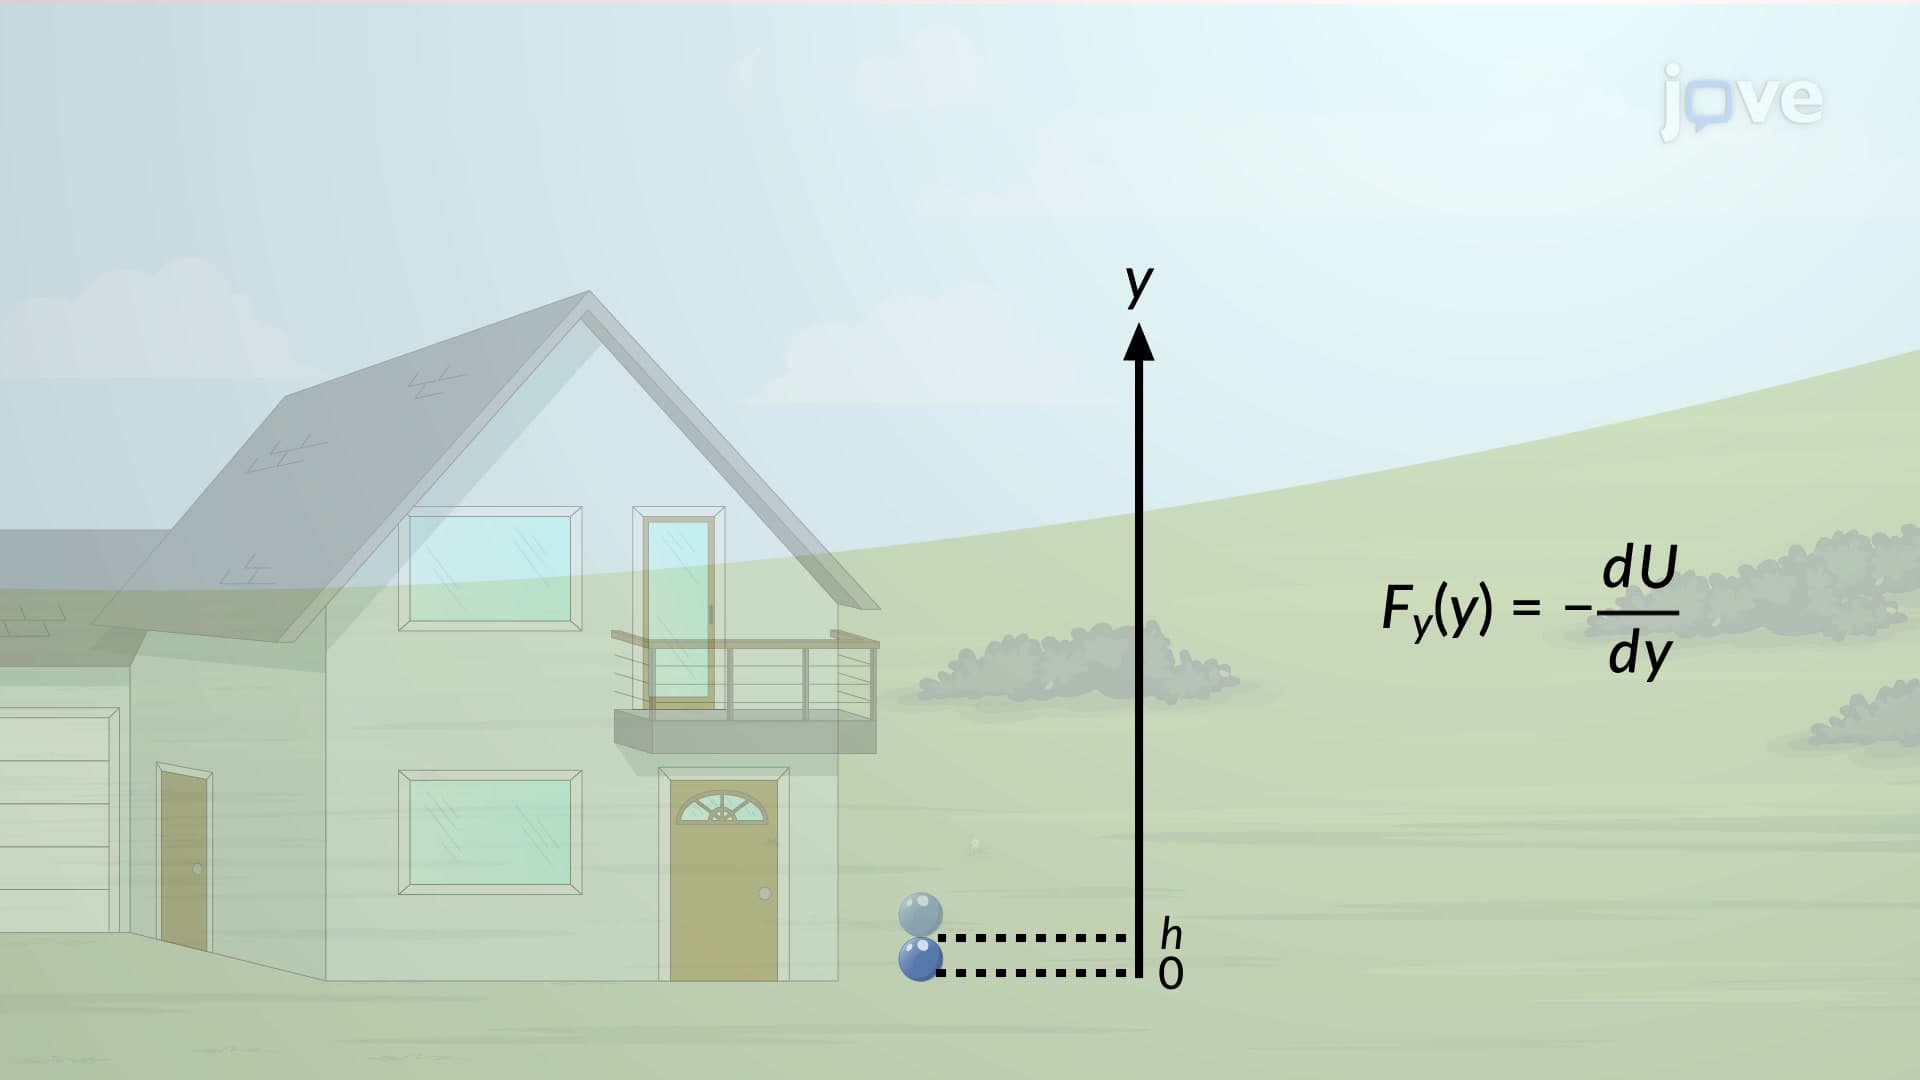
Force and Potential Energy in One Dimension
 01:12
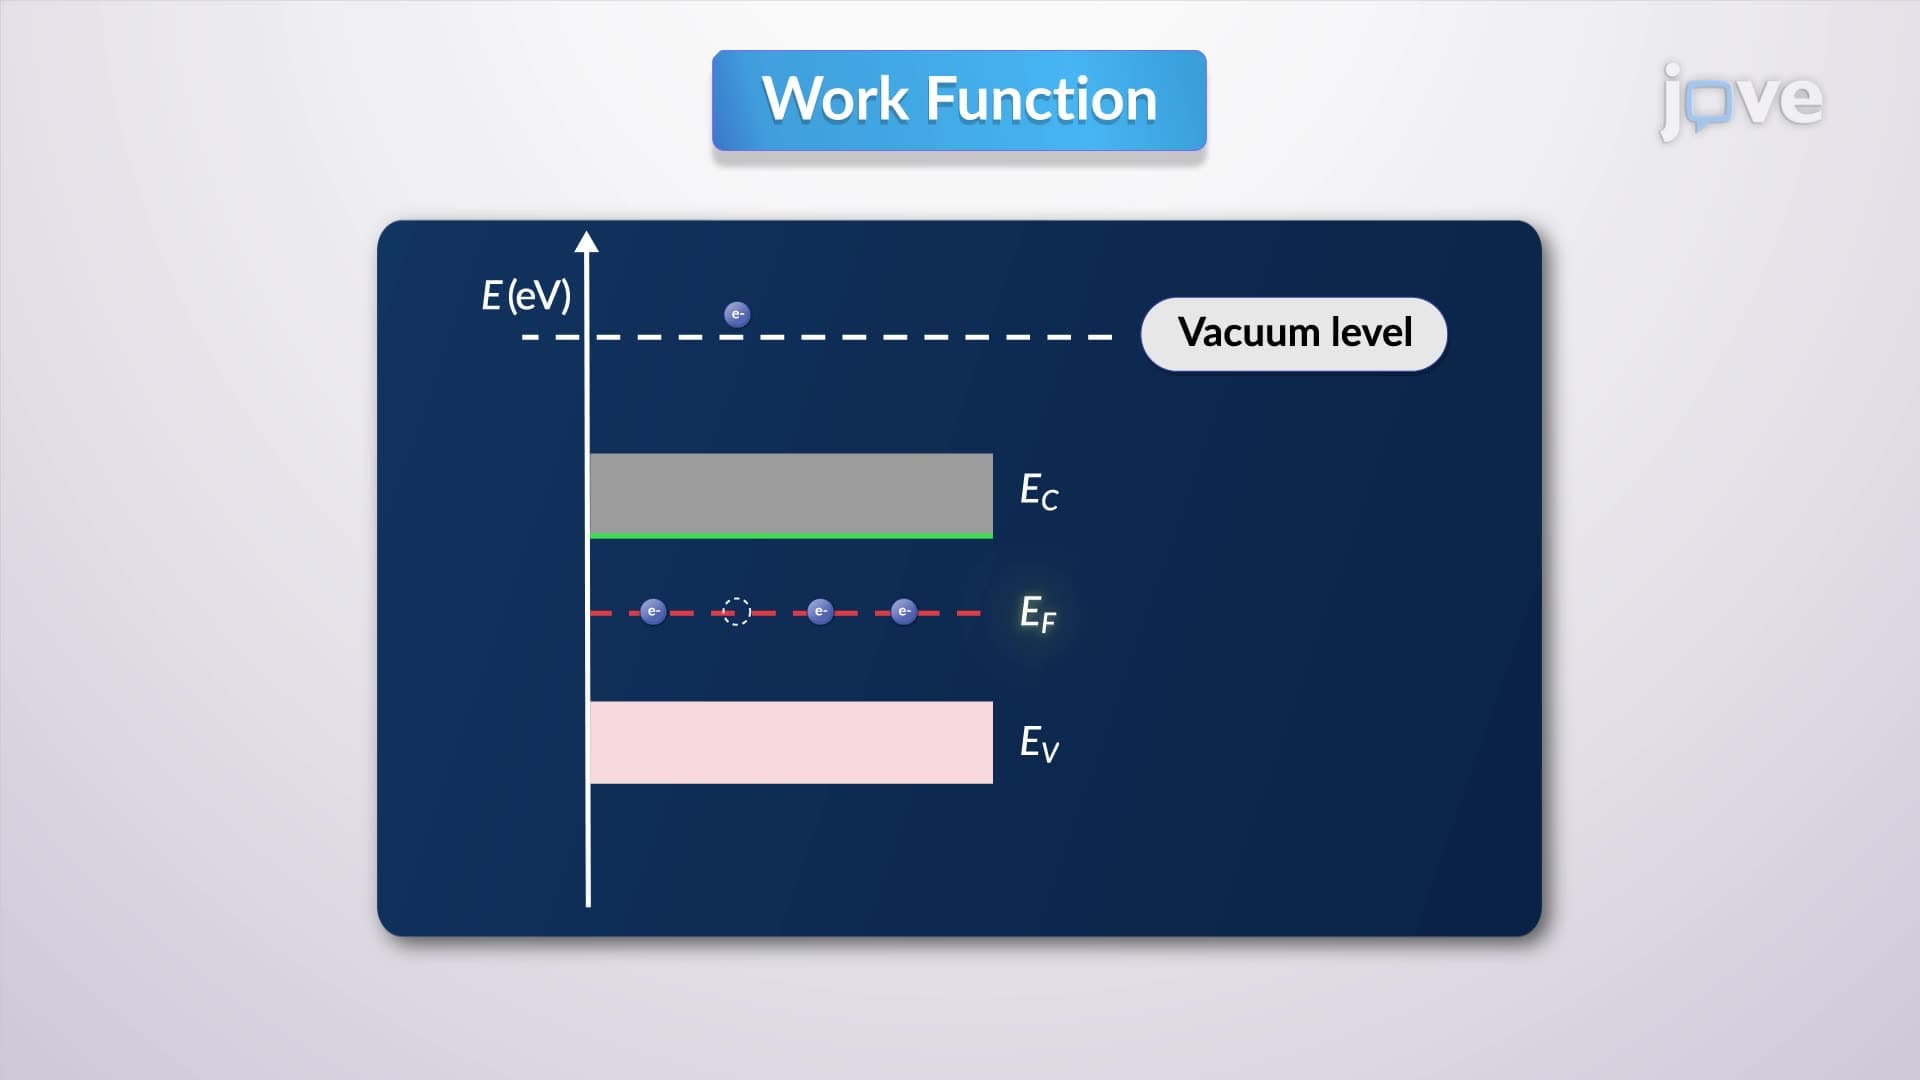
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states that no two...
 01:27
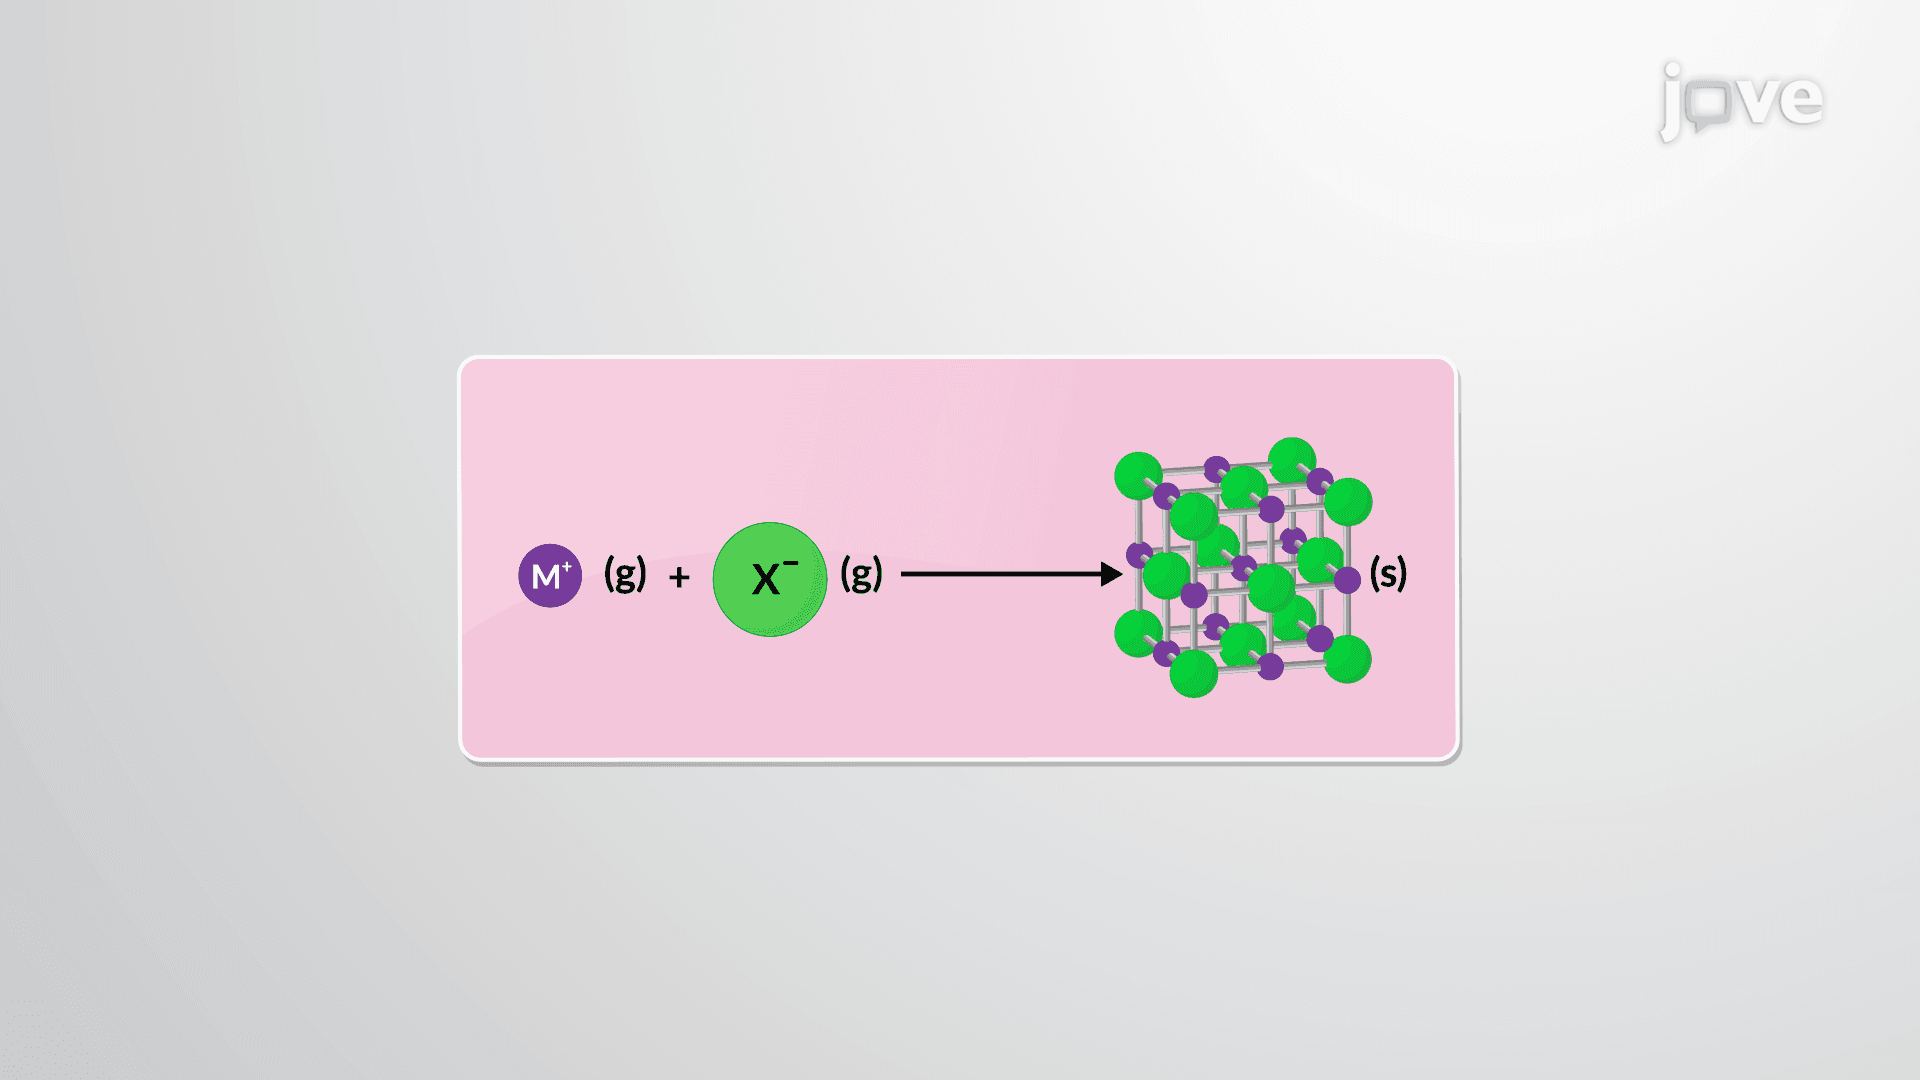
Lattice Energies of Ionic Crystals