
NMRリラクゼーション測定のためのタンパク質におけるアロマティックサイドチェーンの生合成13Cラベリング
Kaare Teilum1, Ulrika Brath, Patrik Lundström
1Department of Biophysical Chemistry, Lund University, Sweden. kaare.tellum@bpc.lu.se
Journal of the American Chemical Society
|February 24, 2006

まとめ
[1-13C]-グルコースを用いたサイト固有の13Cラベリングは,タンパク質における核磁気共鳴 (NMR) のリラクゼーション測定を簡素化します. この方法は,アロマティックアミノ酸からの信号を分離することで,データの解釈を向上させ,タンパク質動態学の研究に役立ちます.
科学分野:
- 生物物理化学 生物物理化学
- 構造生物学 構造生物学とは
- 核磁共振 (NMR) スペクトロスコピー
背景:
- 核磁共振 (NMR) のリラクゼーション実験は,タンパク質のダイナミクスを理解する上で極めて重要です.
- 望ましくないリラクゼーション経路と一貫した磁化移転は,NMRデータ解釈を複雑にします.
- サイト固有の同位体ラベル付けは,信号を隔離することによって,NMR実験を簡素化することができます.
研究 の 目的:
- [1-13C]-グルコースを用いたタンパク質のサイト固有の13Cラベリングの方法を開発および検証.
- アロマティックアミノ酸における13Cリラクゼーション速度の測定と解釈を簡素化するために.
- 化学交換と構造動態の堅実な特徴付けを可能にする.
主な方法:
- [1-13C]-グルコースを唯一の炭素源として使用したEscherichia coliにおけるタンパク質過剰発現.
- アロマティックアミノ酸のサイドチェーンの特定の位置に13Cを組み込む.
- 13C回転フレームのリラクゼーション分散データの取得.
主要な成果:
- アロマティックアミノ酸のサイト固有の13Cラベリングを達成しました.
- 簡素化されたNMRリラクゼーション測定と解釈の実証.
- カルモジュリンの形状交換ダイナミクスを特定し,時間常数71μsでY138.8.の形状交換ダイナミクスを特定した.
結論:
- [1-13C]-グルコースによるサイト固有の13Cラベリングは,NMRリラクゼーション研究を簡素化するための効果的な戦略です.
- この方法は,CPMGまたはスピンロックリラックス技術を使用して,化学交換の特徴を容易にします.
- このアプローチは,タンパク質の構成動態に関する貴重な洞察を提供します.
さらに関連する動画
関連する概念動画
 01:10
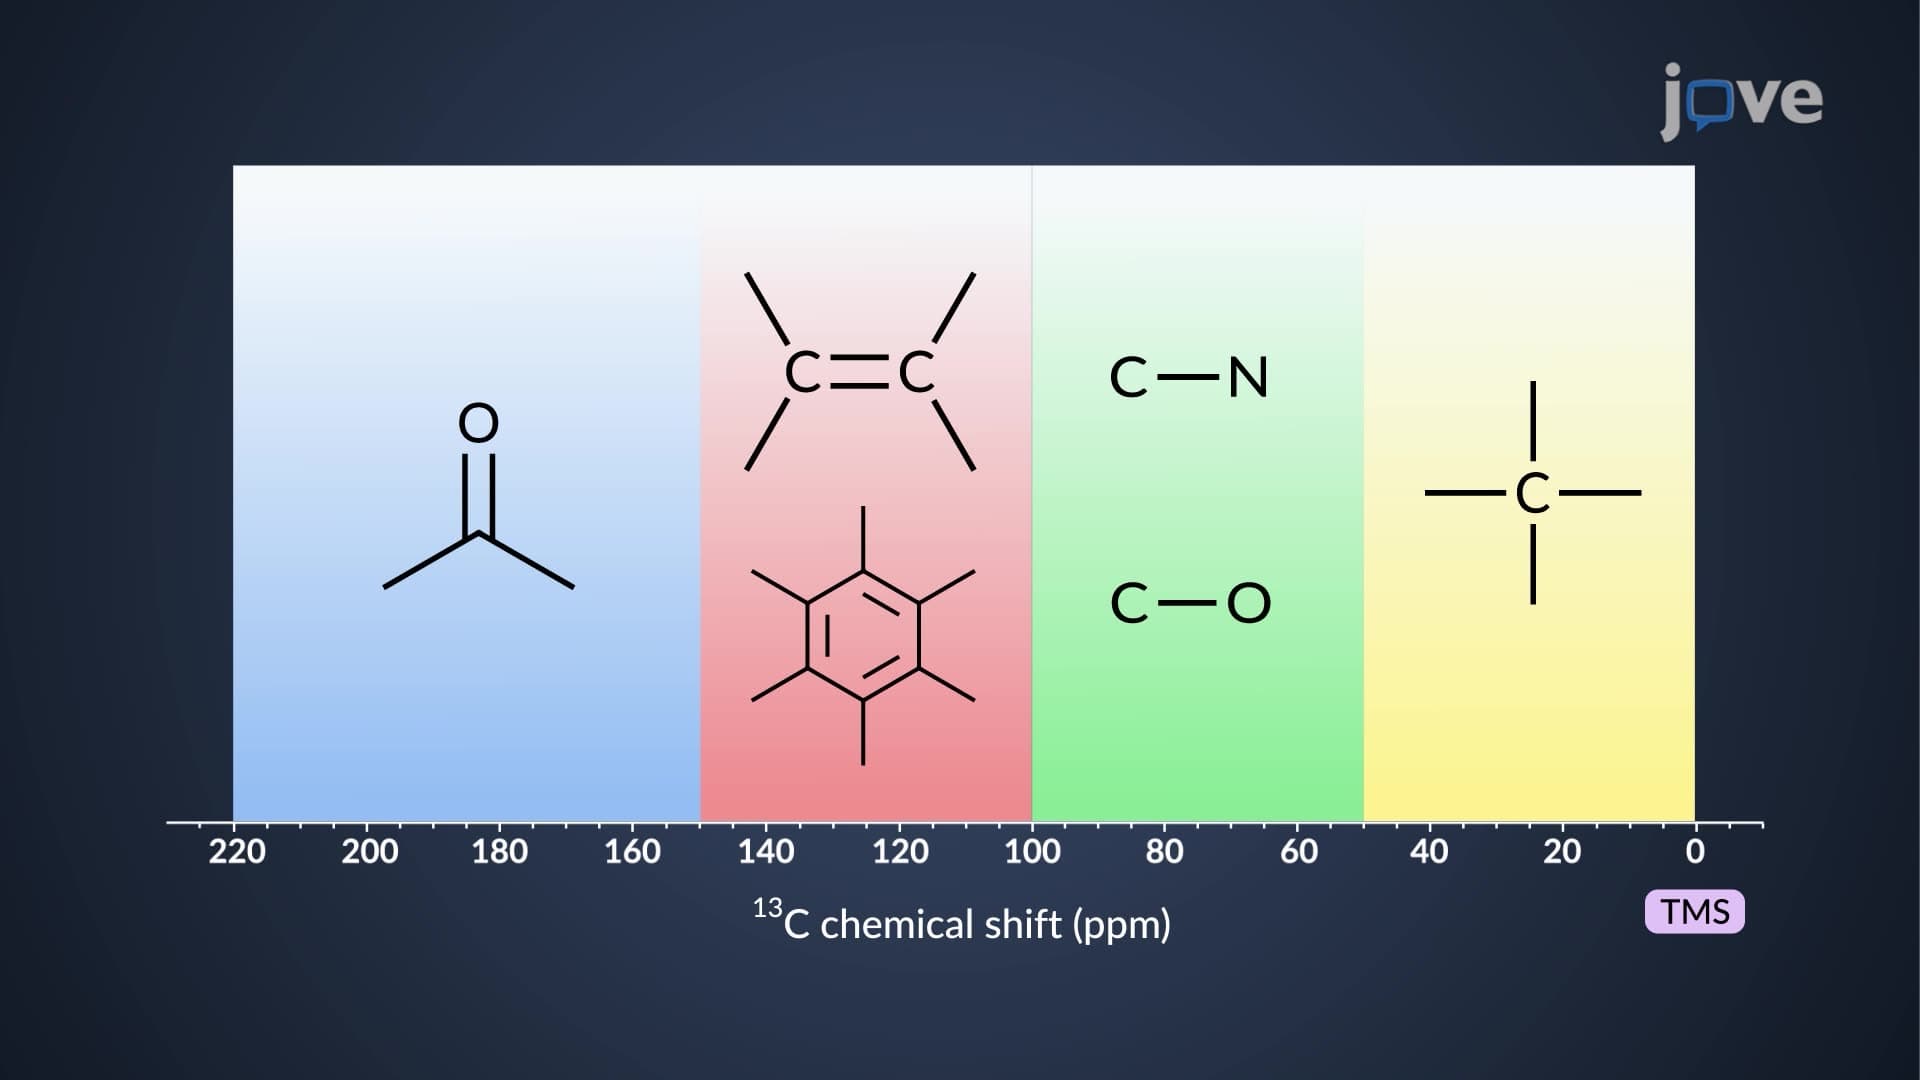
Carbon-13 (¹³C) NMR: Overview
Carbon-13 is a naturally occurring NMR-active isotope of carbon with a low natural abundance of 1.1%. In contrast, carbon-12 is the most abundant isotope of carbon with zero nuclear spin. Therefore, it is NMR inactive. The gyromagnetic ratio of carbon-13 is smaller than that of protons. As a result, carbon-13 resonance is about 6000 times weaker than proton resonance. For a given magnetic field strength, the resonance frequency of carbon-13 is about one-fourth of the resonance frequency for...
 01:20
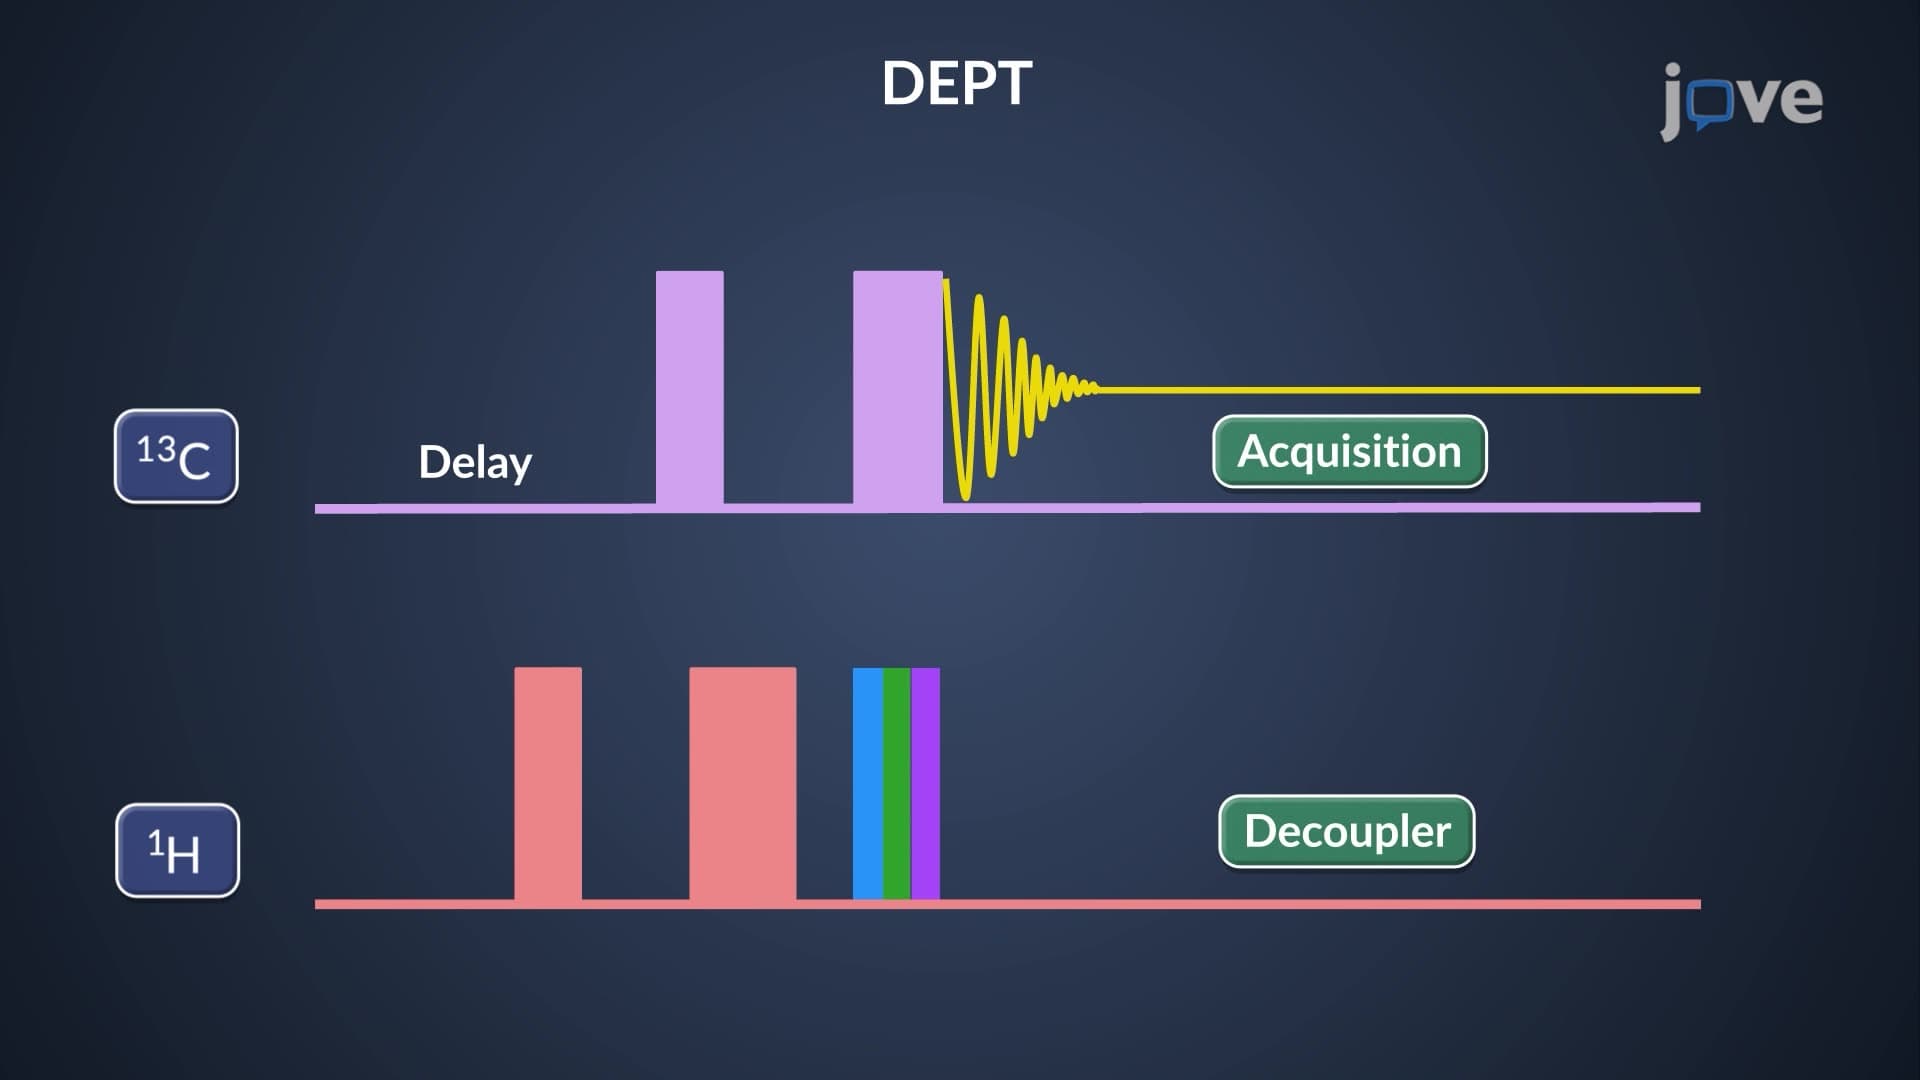
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
When proton-coupled carbon-13 spectra are simplified by a broadband proton decoupling technique, structural information about the coupled protons is lost. Distortionless enhancement by polarization transfer (DEPT) is a technique that provides information on the number of hydrogens attached to each carbon in a molecule. While the DEPT experiment utilizes complex pulse sequences, the pulse delay and flip angle are specifically manipulated. The resulting signals have different phases depending on...
 01:14
NMR Spectroscopy of Aromatic Compounds
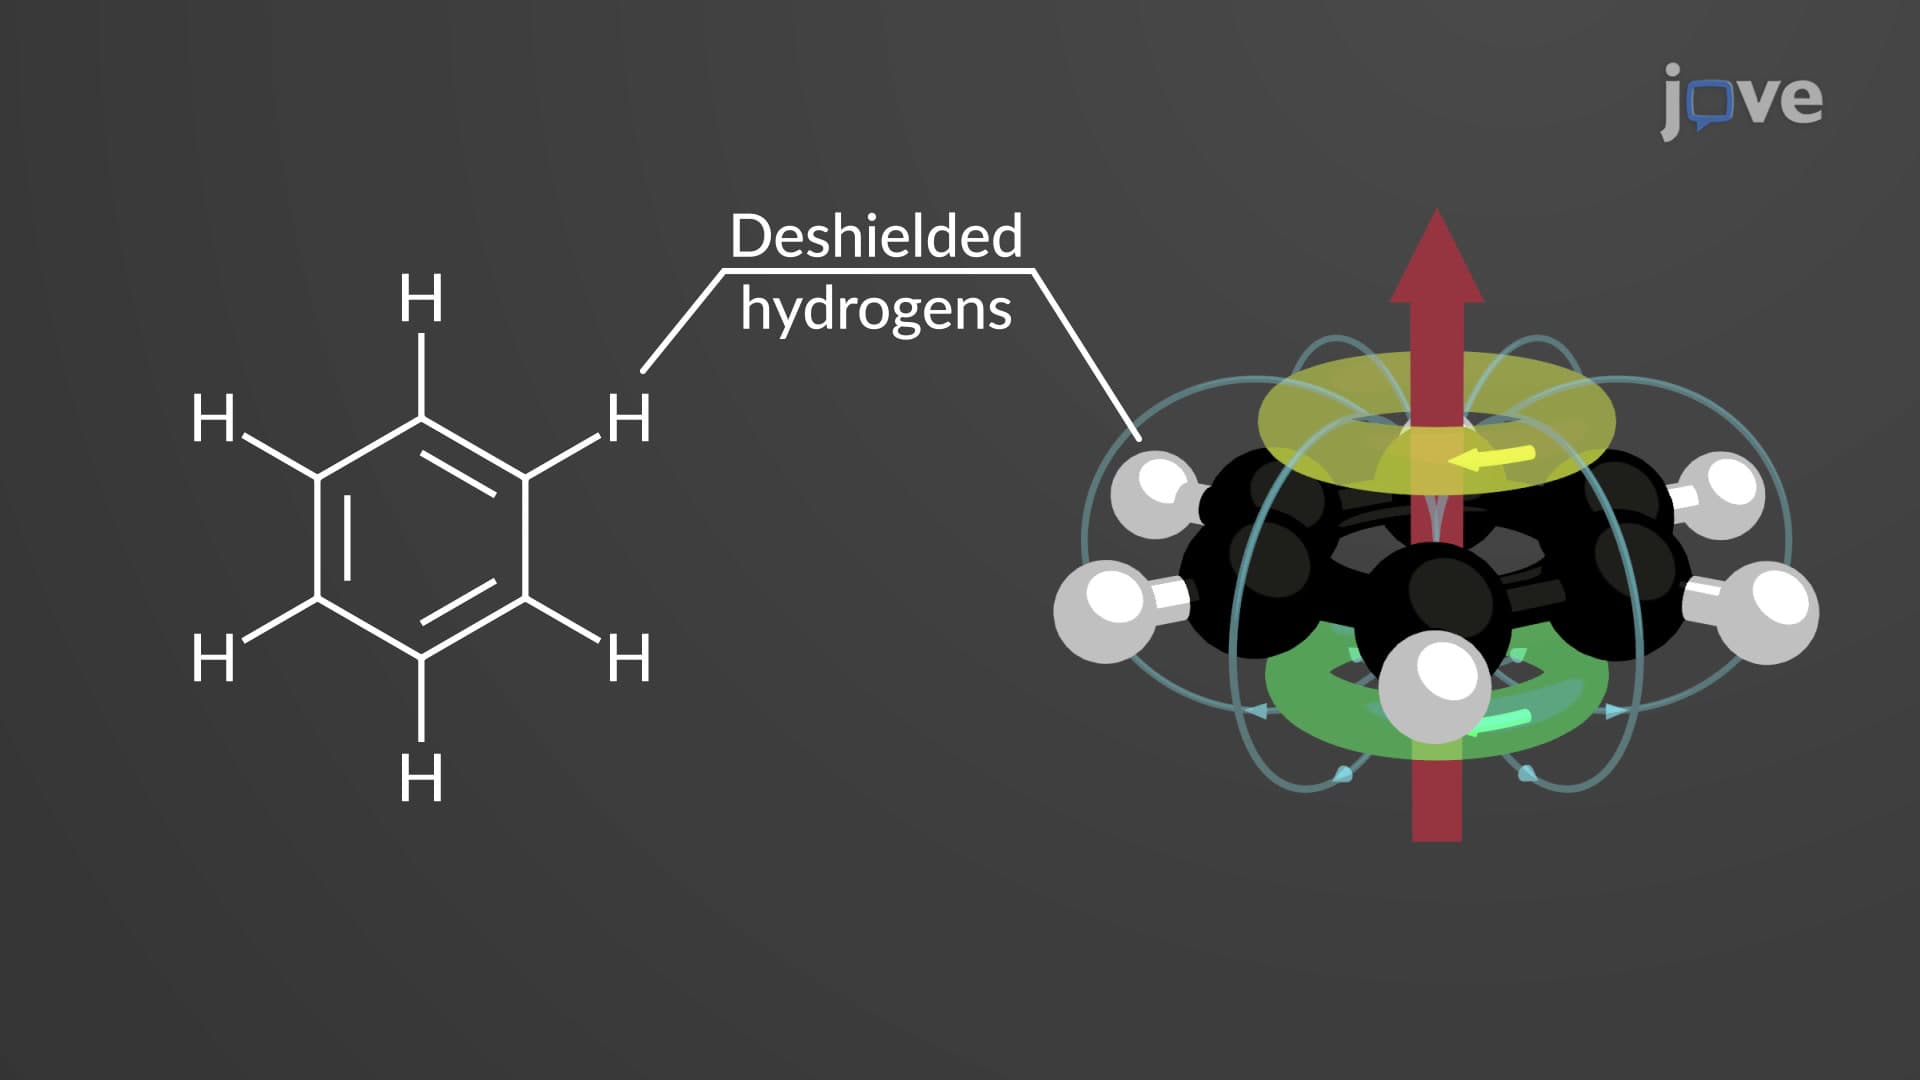
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
 01:37
NMR Spectroscopy of Benzene Derivatives
Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling constants depend...
 01:04
¹³C NMR: ¹H–¹³C Decoupling
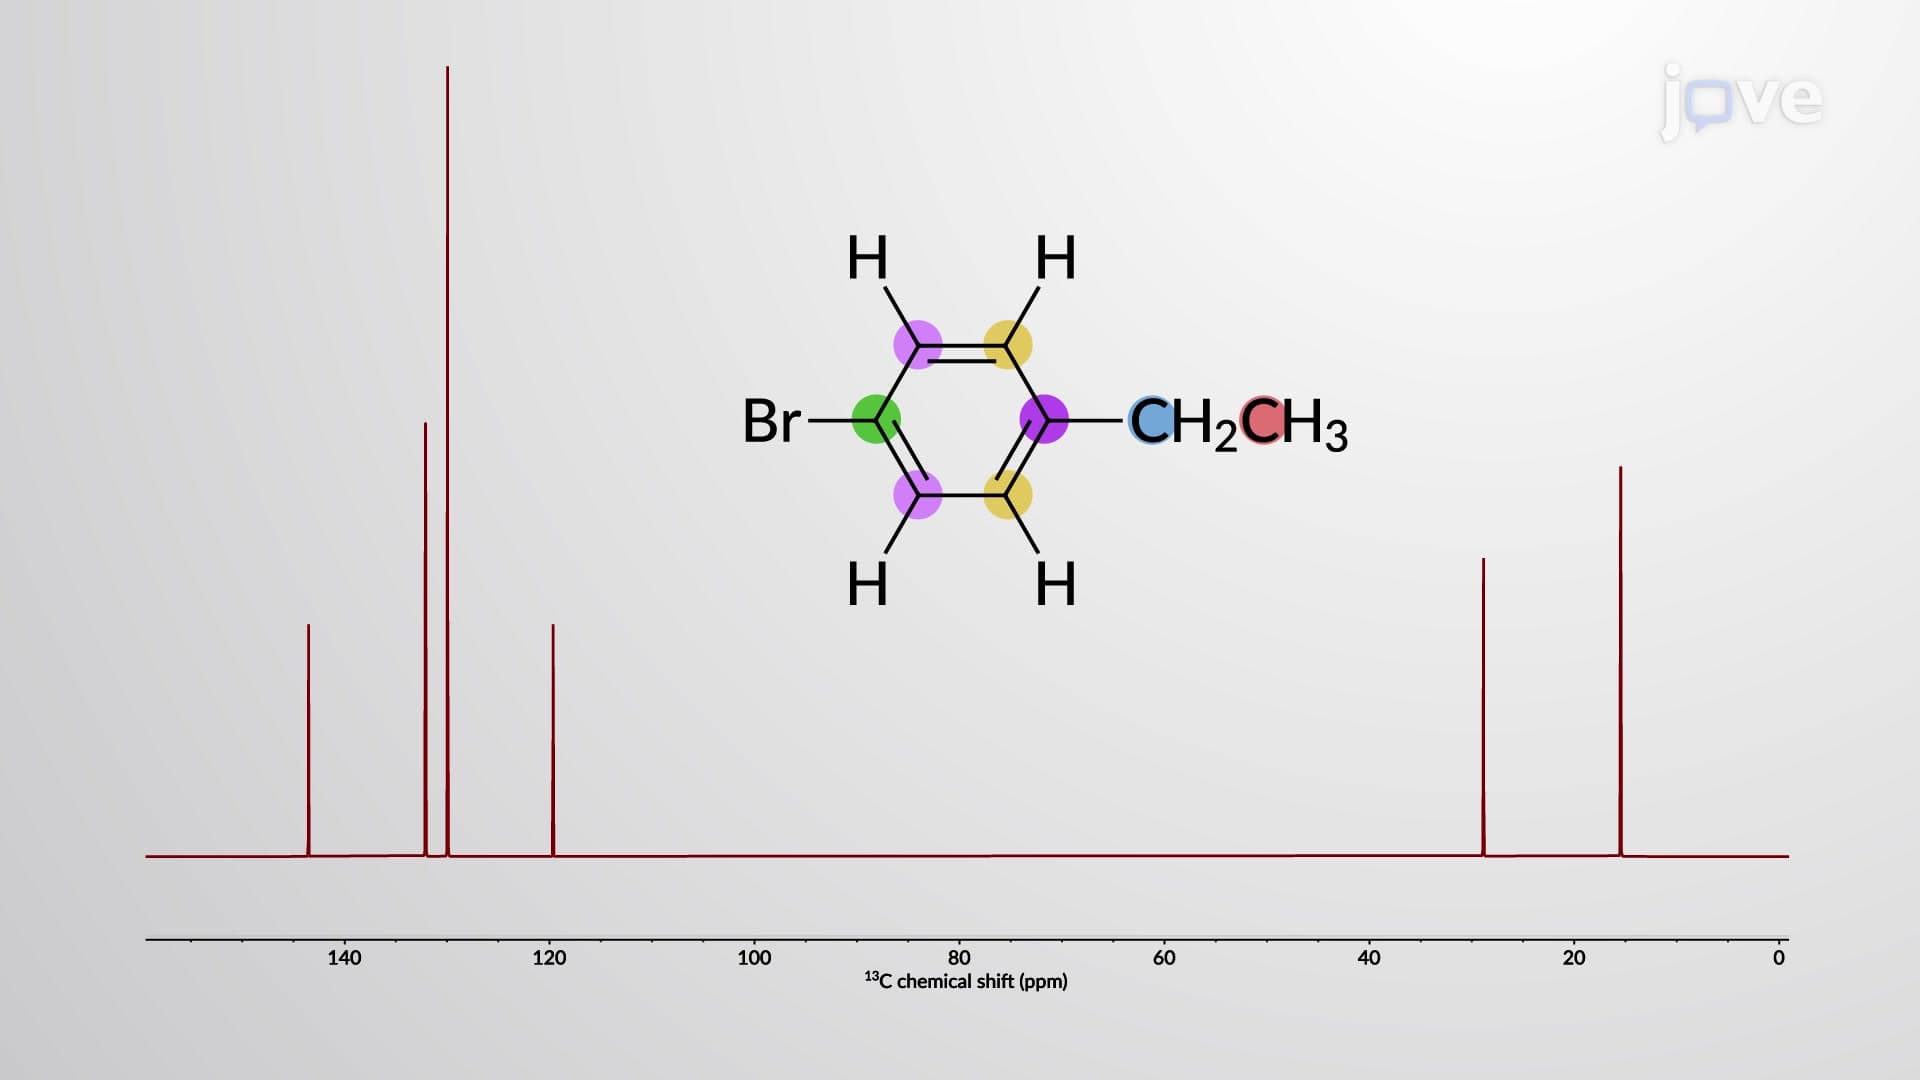
The probability of having two carbon-13 atoms next to each other is negligible because of the low natural abundance of carbon-13. Consequently, peak splitting due to carbon-carbon spin-spin coupling is not observed in spectra. However, protons up to three sigma bonds away split the carbon signal according to the n+1 rule, resulting in complicated spectra.
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...
 01:19
NMR Spectroscopy Of Amines
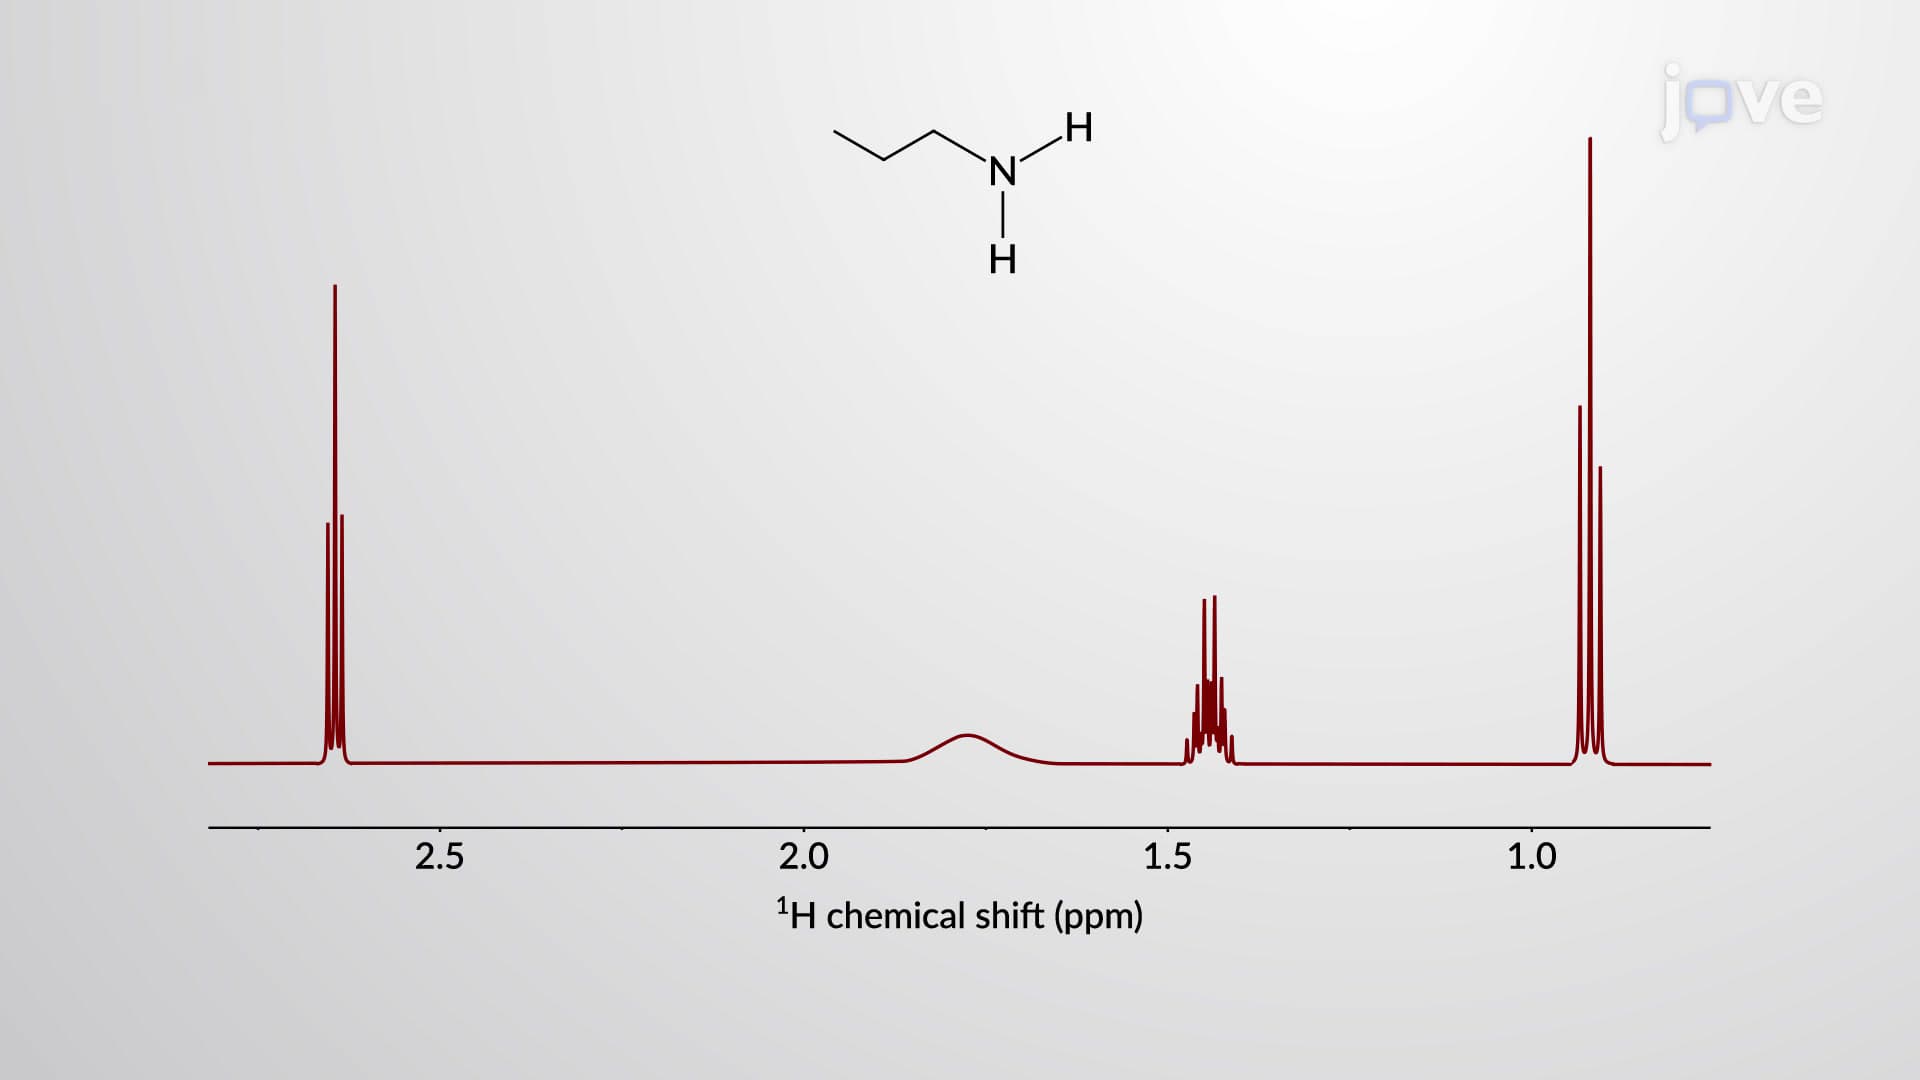
In proton NMR spectroscopy, primary amines and secondary amines showcase their N–H protons as a broad signal in the chemical shift range between δ 0.5 and 5 ppm. The exact position in this range depends on several factors, including sample concentration, hydrogen bonding, and the type of solvent used. Since amine protons undergo fast proton exchange in solution, the protons are labile and therefore do not participate in any splitting with adjacent protons. Thus, the observed peak is broad and...