関連する実験動画
Updated: Jun 10, 2026
08:51
Quantification of Coenzyme A in Cells and Tissues
Published on: September 27, 2019
短鎖アシルコエンザイムAデヒドロゲネーゼ欠乏症における臨床的,生化学的,および遺伝的異質性
Bianca T van Maldegem1, Marinus Duran, Ronald J A Wanders
1Department of Pediatrics, Academic Medical Center, University of Amsterdam, Amsterdam, The Netherlands. b.t.vanmaldegem@amc.uva.nl
JAMA
|August 24, 2006

まとめ
短鎖アシルコエンザイムA (CoA) デヒドロゲナーゼ (SCAD) 欠乏症は,これまで考えられていたよりも一般的であり,特定の遺伝子型に関連していない非特異的な症状があります. 現在の証拠は,新生児のスクリーニングプログラムには適していないことを示唆しています.
科学分野:
- バイオケミストリー バイオケミストリー
- 遺伝学 遺伝学とは
- クリニカル・メディシン 臨床医学
背景:
- 短鎖アシルコエンザイムA (CoA) デヒドロゲナーゼ (SCAD) 欠乏症 (SCADD) は,希少で異質な疾患である.
- SCADDは現在,一部の地域で新生児の拡大スクリーニングに含まれています.
研究 の 目的:
- オランダのSCADD患者や親戚の特徴を明らかにする.
- SCADDにおけるゲノタイプ-フェノタイプ関係を調査する.
主な方法:
- オランダのSCADD患者31人とその親戚8人を対象とした遡及的研究.
- SCADD診断は,生化学マーカー (ブチリルカルニチン,エチルマロン酸) とACADS遺伝子配列解析に基づいています.
- 流行,遺伝子型,生化学レベル,および臨床表現の分析.
主要な成果:
- 計算された出生頻度は少なくとも1:50,000です.
- ほとんどの患者は3歳になる前に非特異的,一時的な症状を発症した.
- ゲノタイプは生化学的レベルと相関しているが,臨床的特徴とは相関しない;親族はほとんど無症状であった.
結論:
- SCADDは,これまで考えられていたよりもはるかに一般的です.
- 臨床症状は非特異的で,一般的に軽度であり,遺伝子型に関連していない.
- SCADDは現在,明確な臨床的意義と差別化課題の欠如のために,新生児のスクリーニングに適していません.
さらに関連する動画
関連する概念動画
 01:35
Pedigree Analysis
Overview
 01:02
Long-patch Base Excision Repair
Since the discovery of the two BER pathways, there has been a debate about how a cell chooses one pathway over the other and the factors determining this selection. Numerous in vitro experiments have pointed out multiple determinants for the sub-pathway selection. These are:
 01:48
ATP Synthase: Mechanism
In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased ATP...
 01:18
ATP Synthase: Structure
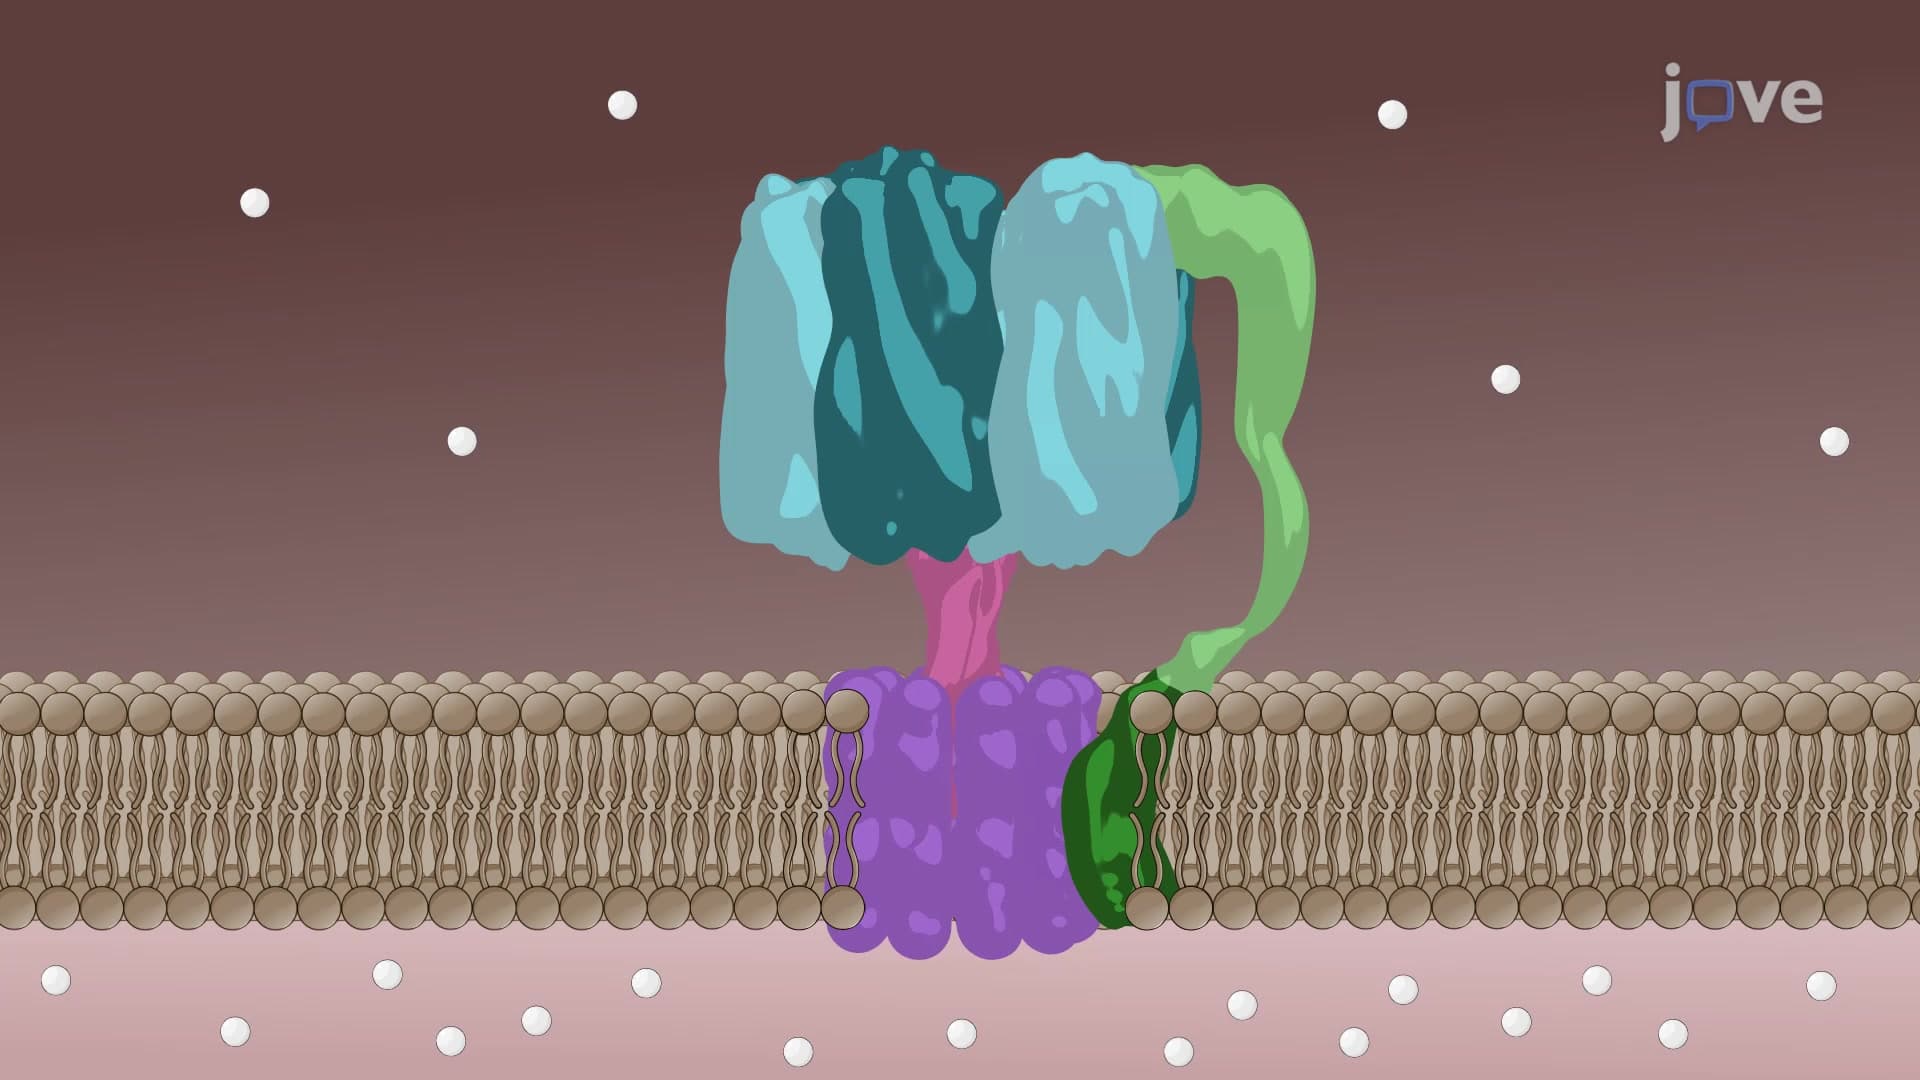
ATP synthase or ATPase is among the most conserved proteins found in bacteria, mammals, and plants. This enzyme can catalyze a forward reaction in response to the electrochemical gradient, producing ATP from ADP and inorganic phosphate. ATP synthase can also work in a reverse direction by hydrolyzing ATP and generating an electrochemical gradient. Different forms of ATP synthases have evolved special features to meet the specific demands of the cell. Based on their specific feature, ATP...
 01:22
Lysosomal Hydrolases
Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...
 01:20
Inborn Errors of Metabolism
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...