
トップダウン質量スペクトロメトリを200キロダルトン以上の質量を持つタンパク質に拡張する
Xuemei Han1, Mi Jin, Kathrin Breuker
1Department of Chemistry and Chemical Biology, Baker Laboratory, Cornell University, Ithaca, NY 14853, USA.
Science (New York, N.Y.)
|October 7, 2006

まとめ
トップダウン質量スペクトロメトリーは,以前のサイズ制限を克服して,より大きなタンパク質を分析します. この高度な方法は,タンパク質の詳細な特徴化のために,高度に特異的な分子および断片イオンデータを提供します.
科学分野:
- プロテオミクスはプロテオミクスを用います.
- マススペクトロメトリーによる質量スペクトロメトリーです.
- バイオケミストリー バイオケミストリー
背景:
- 伝統的なタンパク質分析はペプチド質量データに依存しており,これは分子および断片イオンデータよりも特異性が低い.
- トップダウン質量スペクトロメトリーは,より高い特異性を提供しますが,より小さなタンパク質 (約. 50 kDa) である.
研究 の 目的:
- トップダウンの質量スペクトロメトリの適用範囲を,より大きなタンパク質に拡大する.
- タンパク質の配列と翻訳後の改変の特徴づけを強化する.
主な方法:
- 電気スプレーの添加物,加熱した蒸発,および分離された非共振性および共振性結合解離技術を使用しました.
- トップダウン質量スペクトロメトリーの約500残留解離制限を克服するために,これらの方法を適用しました.
主要な成果:
- 1314残基 (144-kD) のタンパク質で287の残基間結合を成功裏に割った.
- 1714残留 (200-kD) のタンパク質内の27のシステインのうち8の未知の二硫化結合を特定しました.
- 2つの大きなタンパク質のシーケンスの予測を修正し,そのうち1つは2153個の残基 (229 kD) を有する.
結論:
- 強化されたトップダウン質量スペクトロメトリのアプローチは,分析可能なタンパク質のサイズ範囲を大幅に拡張します.
- この方法論は,配列および二硫化結合分析を含む,タンパク質の特徴づけのためのより具体的で包括的な方法を提供します.
関連する概念動画
 01:21
Tandem Mass Spectrometry
Tandem mass spectrometry is a technique that uses multiple mass analyzers in series to obtain a higher selectivity and reduce chemical noise during analyte detection. Instruments with multiple analyzers separated by an interaction cell enable secondary fragmentation and selected study of the fragment ions.Secondary fragmentations occur in the interaction cell and can be induced by various factors. Fragmentation induced by collision with inert gases, such as N2, Ar, He, etc., is called...
 01:33
Peptide Identification Using Tandem Mass Spectrometry
Tandem mass spectrometry, also known as MS/MS or MS2, is an analytical technique that employs two mass analyzers. Essentially it is a series of mass spectrometers that helps isolate a particular biomolecule and then helps study its chemical properties.
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:19
MALDI-TOF Mass Spectrometry
Mass spectrometry is a powerful characterization technique that can identify and separate a wide variety of compounds ranging from chemical to biological entities, based on their mass-to-charge ratio (m/z). The instruments that allow this detection, known as mass spectrometers, have three components: an ion source, a mass analyzer, and a detector. These spectrometers differ based on the nature of their ion source and analyzers.Matrix-assisted laser desorption ionization (MALDI) is a commonly...
 01:21
Mass Spectrometry: Complex Analysis
Mass spectrometry is an important technique for the identification of pure compounds. However, it has some limitations for the analysis of complex mixtures, often due to excessive fragmentation making the spectrum too complicated to decipher. Mass spectrometry can be combined with suitable separation methods in sequence, forming hyphenated methods, which are useful in the analysis of complex mixtures.
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:19
Mass Spectrometry: Overview
Mass spectrometry is an analytical technique used to determine the molecular mass and molecular formula of a compound. The basic principle of mass spectrometry is to generate ions from the analyte molecule and measure these ion abundances against their molecular mass. One common type of ionization, known as electron ionization or EI, bombards the analyte molecules in the gas phase with high-energy electron beams. The electron beams displace an electron from the molecule and leave behind a...
 01:15
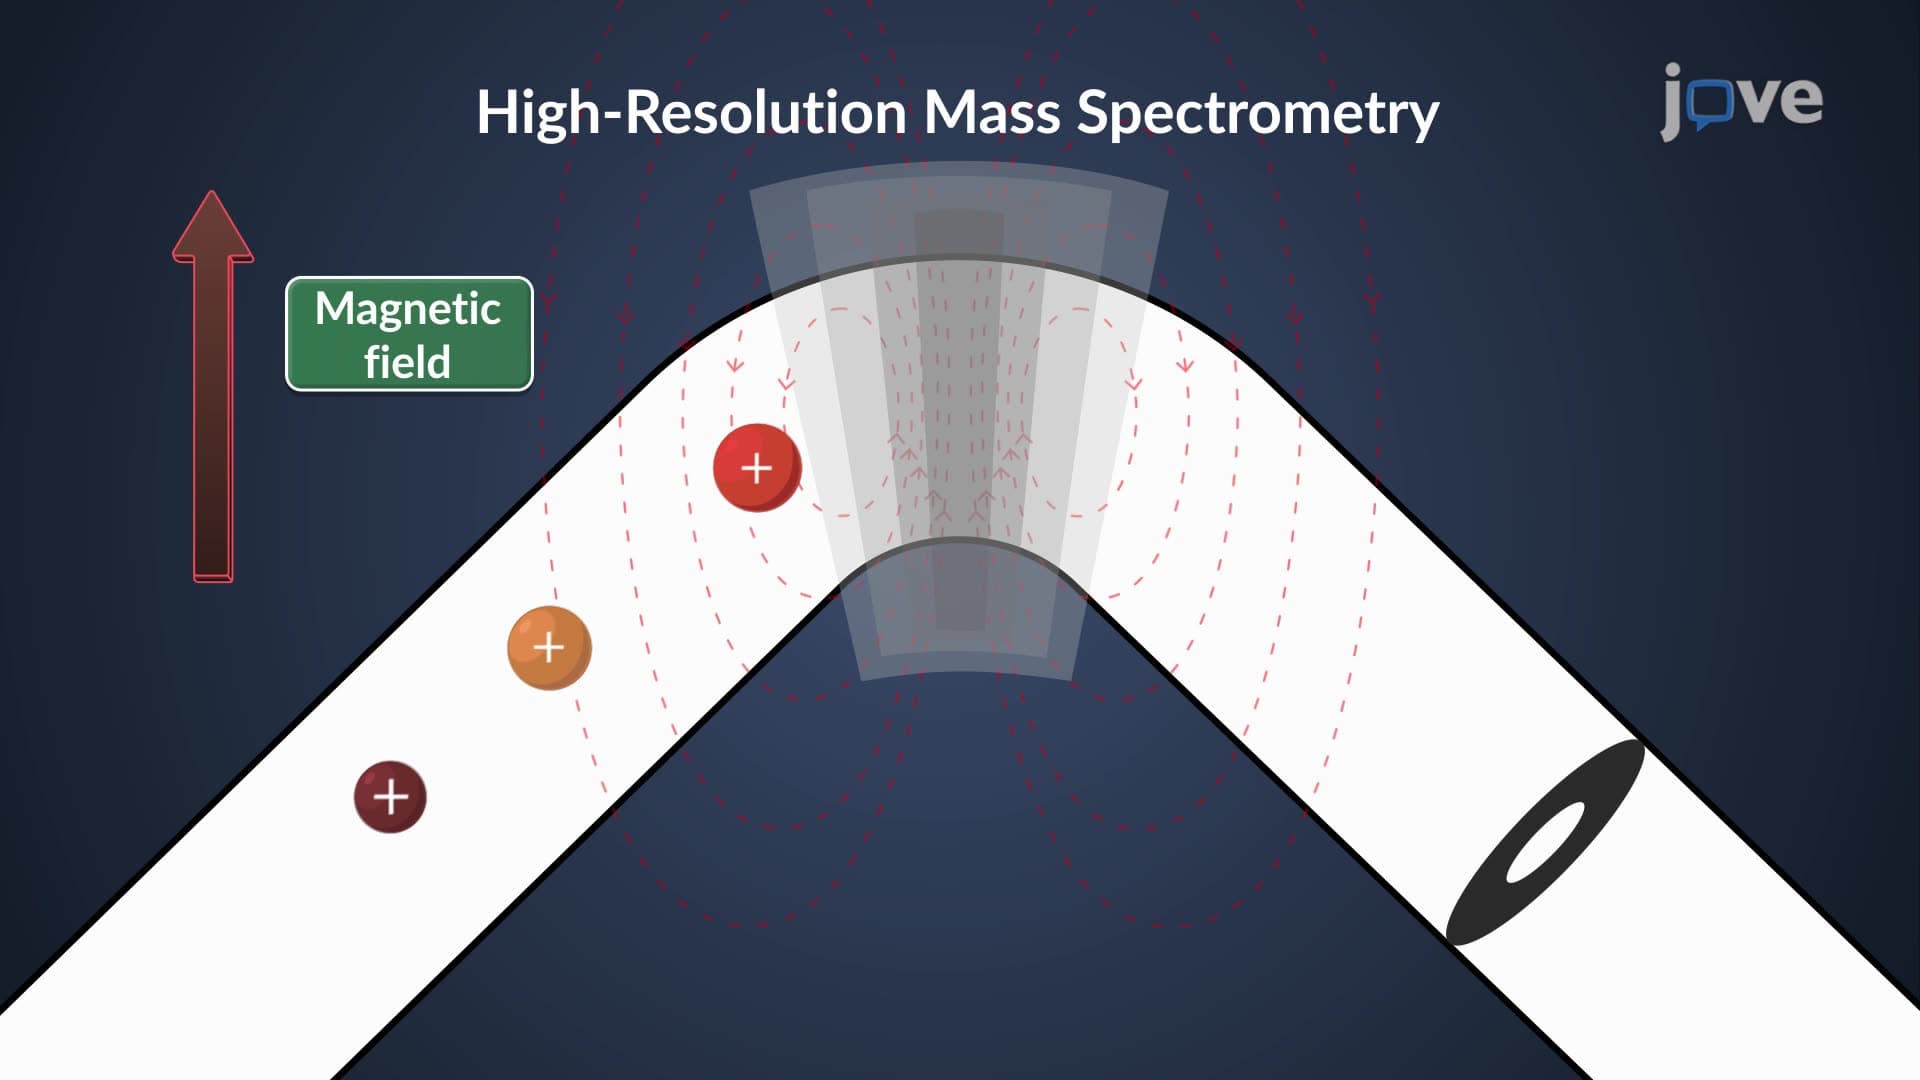
High-Resolution Mass Spectrometry (HRMS)
The resolution of a mass spectrometer depends on the efficiency of separating ions with different ion masses. The mass of an atom is approximated to the sum of the masses of protons and neutrons inside, considering the masses of protons and neutrons as equal. However, the masses of the proton (1.6726 × 10⁻²⁴ g) and neutron (1.6749 × 10⁻²⁴ g) are not truly equal. There is a minor error in the expression of atomic masses relative to the simplest atom of hydrogen. For example, the mass of helium...