
ヴァンコマイシンの触媒部位選択型チオカルボニル化および脱酸素化は,ヒドロキシル依存型構成効果を明らかにする
Brandon S Fowler1, Kai M Laemmerhold, Scott J Miller
1Department of Chemistry, Yale University, P.O. Box 208107, New Haven, Connecticut 06520-8107, USA.
Journal of the American Chemical Society
|May 25, 2012

まとめ
ペプチド触媒は,バンコマイシンのサイト選択的チオカルボニル化に使用され,2つの新しいデオキシバンコマイシン誘導体が得られました. これらの化合物は,驚くべき形状の変化と生物学的活動の変化を明らかにし,ヒドロキシル群の構造的役割を強調しました.
科学分野:
- 薬用化学 薬用化学について
- オーガニック・シンセシス オーガニック・シンセシス
- カタリシス カタリシス カタリシス
背景:
- バンコマイシンは,深刻な細菌感染症の治療に不可欠な抗生物質です.
- バンコマイシンを改変することで,耐性を克服し,有効性を向上させることができます.
- サイト選択的機能化は,新しいバンコマイシンアナログを生成するための鍵です.
研究 の 目的:
- バンコマイシンのサイト選択性チオカルボニレーションのためのペプチドベースの触媒を開発する.
- 新型デオキシバンコマイシン誘導体を合成する.
- 脱酸素化の構造的および生物学的影響を調査する.
主な方法:
- ペプチド触媒のスクリーニングと合理的な設計.
- 保護されたバンコマイシンのサイト選択的チオカルボニル化.
- 主要な中間物質のスケールアップ合成 (0.50gスケール).
- 新種の同類物を生成するための脱酸素化反応.
- 適合分析と生物学的活動測定法.
主要な成果:
- バンコマイシンチオカルボニレーションの選択性を強化または変更するペプチド触媒を特定した.
- 2つの新しいデオキシバンコマイシン誘導体を成功裏に合成しました.
- デオキシバンコマイシンアナログで予期せぬ形状変化が観察されました.
- これらの形状の変化が,変化した生物学的活動と相関することを示した.
結論:
- ペプチド触媒は,選択的バンコマイシン改変のための実行可能な戦略を提供します.
- バンコマイシンの脱酸素化は,重要な形状および機能的な変化につながる可能性があります.
- 特定のヒドロキシルグループは,ネイティブバンコマイシンの活性において重要な構造的役割を果たします.
関連する概念動画
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
The rate of acid-catalyzed hydration of alkenes depends on the alkene's structure, as the presence of alkyl substituents at the double bond can significantly influence the rate.
 02:36
Regioselectivity and Stereochemistry of Hydroboration
A significant aspect of hydroboration–oxidation is the regio- and stereochemical outcome of the reaction.
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:14
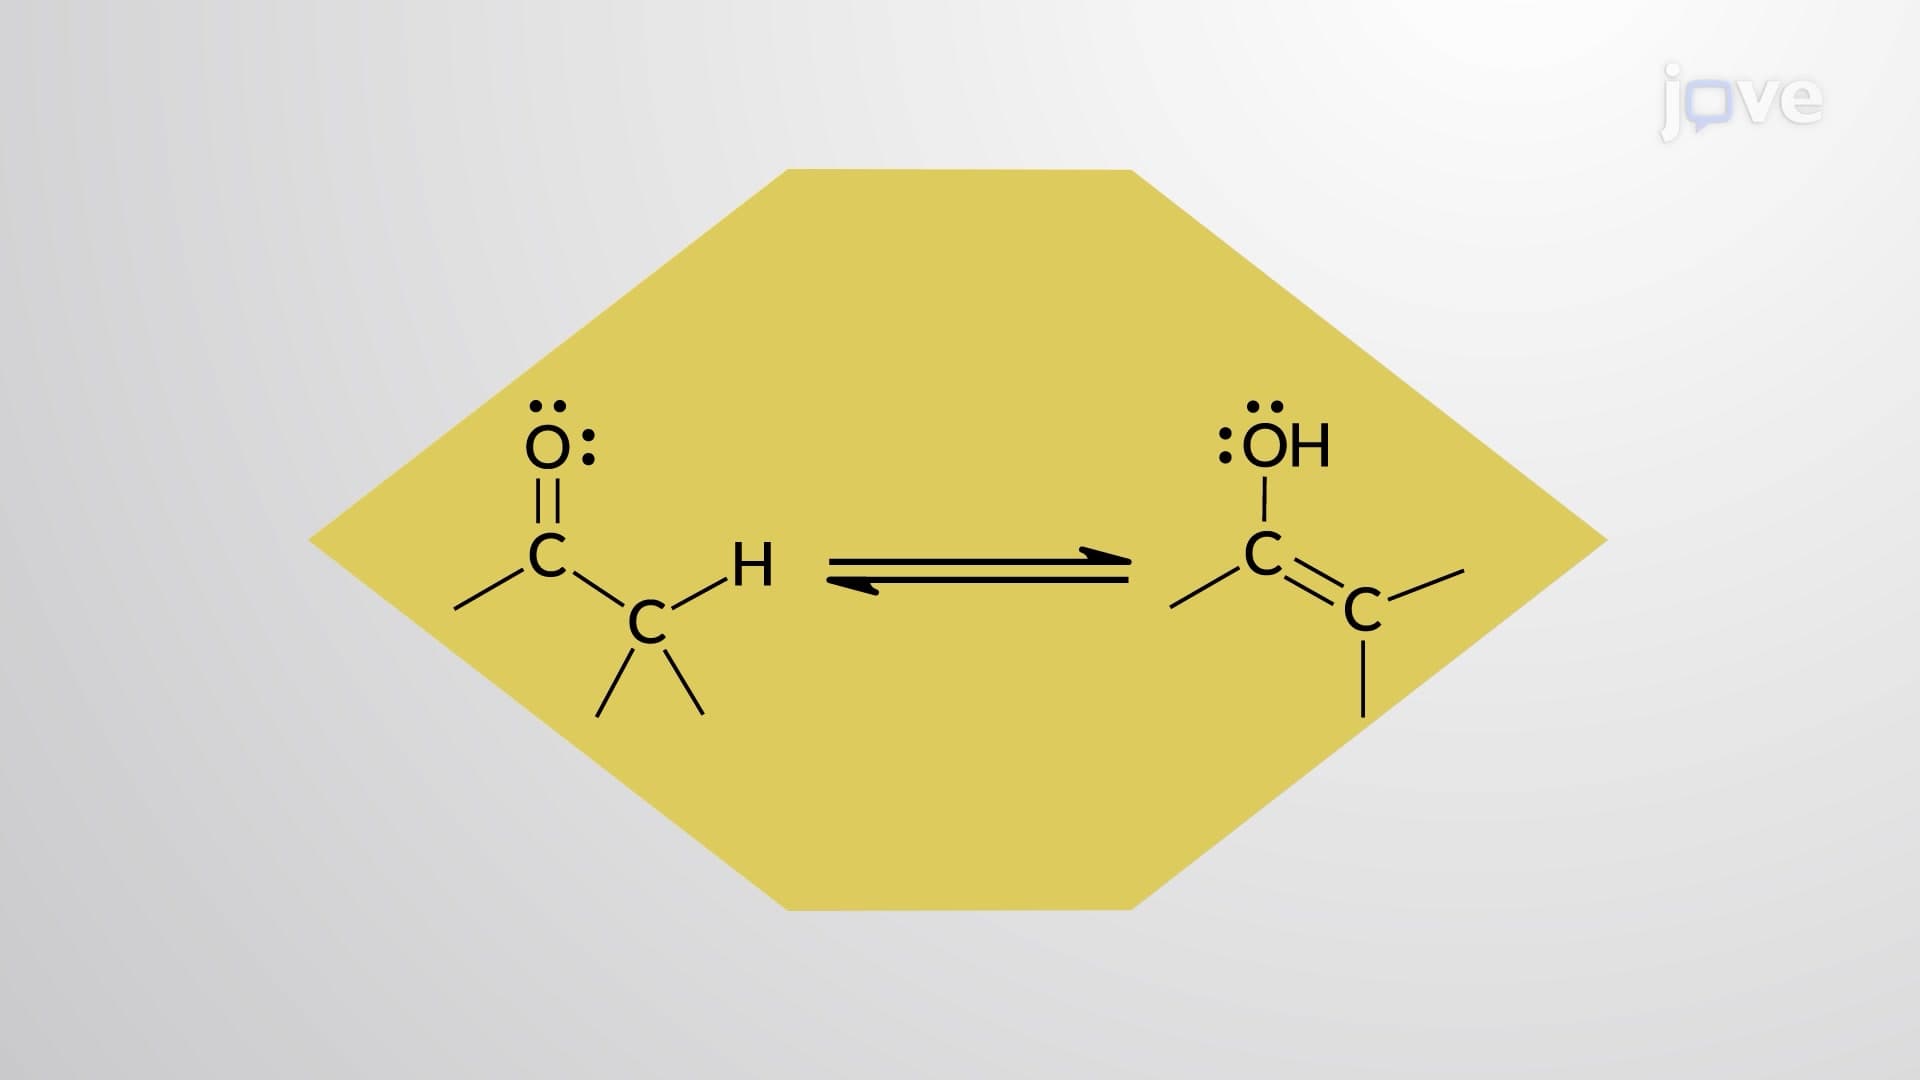
Keto–Enol Tautomerism: Mechanism
The keto and enol forms are known as tautomers and they constantly interconvert (or tautomerize) between the two forms under acid or base catalyzed conditions. Both the reactions involve the same steps—protonation and deprotonation— although in the reverse order.
 03:08
Hydroboration-Oxidation of Alkenes
In addition to the oxymercuration–demercuration method, which converts the alkenes to alcohols with Markovnikov orientation, a complementary hydroboration-oxidation method yields the anti-Markovnikov product. The hydroboration reaction, discovered in 1959 by H.C. Brown, involves the addition of a B–H bond of borane to an alkene giving an organoborane intermediate. The oxidation of this intermediate with basic hydrogen peroxide forms an alcohol.
 01:35
Loss of Carboxy Group as CO2: Decarboxylation of Malonic Acid Derivatives
Just like β-keto acids—which upon thermal decarboxylation form ketones—β-dicarboxylic acids undergo decarboxylation to generate monocarboxylic acids with the liberation of carbon dioxide.
 01:22
Reactions of Aldehydes and Ketones: Baeyer–Villiger Oxidation
Baeyer–Villiger oxidation converts aldehydes to carboxylic acids and ketones to esters. The reaction uses peroxy acids or peracids and is often catalyzed by acid. The reaction is named after its pioneers, Adolf von Baeyer and Victor Villiger. The reaction is achieved by a wide range of peracids such as m-chloroperoxybenzoic acid (mCPBA), perbenzoic acid (C6H5COOOH), peracetic acid (CH3COOOH), hydrogen peroxide (H2O2), and tert-butyl hydroperoxide (t-BuOOH).
The carbonyl center is activated by...
The carbonyl center is activated by...