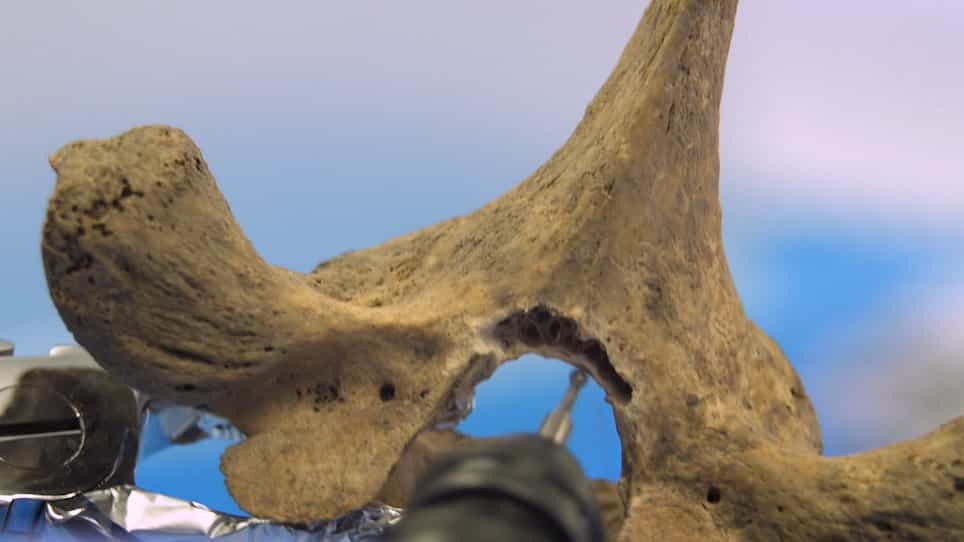

中期プレイストセンのシマ・デ・ロス・ウエソスのヒトの核DNA配列
Matthias Meyer1, Juan-Luis Arsuaga2,3, Cesare de Filippo1
1Department of Evolutionary Genetics, Max Planck Institute for Evolutionary Anthropology, Deutscher Platz 6, 04103 Leipzig, Germany.
Nature
|March 16, 2016

まとめ
スペインのプレイストセンのホミニンは ネアンデルタール人と 関連していました デニソワ人とは違います ネアンデルタール人とデニソワ人の 人口が 43万年以上前に 分かったことを 核DNA分析で明らかにしました
科学分野:
- 古人類学
- 古代のDNA分析
- 人類の進化
背景:
- スペインのシマ・デ・ロス・ウエソス遺跡には 約43万年前の 28匹のヒト類が発見されています
- ネアンデルタール人やデニソヴァ人のような 後期の人類との関係を 理解することが重要です
研究 の 目的:
- シマ・デ・ロス・ホメソスのヒトと ネアンデルタール人と デニソヴァン人の進化的関係を調べる
- 古代のDNAを使って 人類の多様性を調査する
主な方法:
- ホミニンの放射性年代測定
- ミトコンドリアDNA (mtDNA) のシーケンシング
- 複数の個体からの核DNA (nDNA) 配列解析
主要な成果:
- 一人のヒトのミトコンドリアDNAは デニソヴァンと密接な関係を示した
- 2人のDNAの配列は ネアンデルタール人との 明確な関係を明らかにしました
- ネアンデルタール人とデニソワ人の 人口の相違は 43万年前に遡ります
結論:
- シマ・デ・ロス・ウエソスのヒト類は ネアンデルタール人の初期の人類か 密接に関連しています
- ミトコンドリアDNAの系統は必ずしも完全な種関係を反映するものではありません.
- ネアンデルタール人のmtDNAは 進化の過程で後になって 置き換えられたのかもしれません
関連する概念動画
 00:58
Genomic DNA in Eukaryotes
50.4K
Eukaryotes have large genomes compared to prokaryotes. To fit their genomes into a cell, eukaryotic DNA is packaged extraordinarily tightly inside the nucleus. To achieve this, DNA is tightly wound around proteins called histones, which are packaged into nucleosomes that are joined by linker DNA and coil into chromatin fibers. Additional fibrous proteins further compact the chromatin, which is recognizable as chromosomes during certain phases of cell division.
50.4K
 02:51
Multi-species Conserved Sequences
4.4K
Next-generation sequencing technologies have created large genomic databases of a variety of animals and plants. Ever since the human genome project was completed, scientists studied the genome of primates, mammals, and other phylogenetically distant living beings. Such large-scale studies have provided new insights into the evolutionary relationship between organisms.
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved...
4.4K
 01:57
Sanger Sequencing
763.9K
DNA sequencing is a fundamental technique that is routinely used in the biological sciences. This method can be applied to a range of questions at different scales - from the sequencing of a cloned DNA fragment or the study of a mutation in a gene up to whole-genome sequencing. However, despite the widespread use of sequencing today, it was not until 1977 that Fredrick Sanger and his collaborators developed the chain-termination method to decode DNA sequences. It relies on the separation of a...
763.9K
 02:54
Evolutionary Relationships through Genome Comparisons
6.5K
Genome comparison is one of the excellent ways to interpret the evolutionary relationships between organisms. The basic principle of genome comparison is that if two species share a common feature, it is likely encoded by the DNA sequence conserved between both species. The advent of genome sequencing technologies in the late 20th century enabled scientists to understand the concept of conservation of domains between species and helped them to deduce evolutionary relationships across diverse...
6.5K
 03:00
Next-generation Sequencing
95.2K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
95.2K
 01:05
Maxam-Gilbert Sequencing
11.8K
In the same year as the discovery of the Sanger sequencing method, another group of scientists, Allan Maxam and Walter Gilbert, demonstrated their chemical-cleavage method for DNA sequencing. The Maxam-Gilbert method relies on using different chemicals that can cleave the DNA sequence at specific sites, the separation of resulting DNA fragments of variable size using electrophoresis, and deciphering the DNA sequence from the resulting gel bands.
Challenges of the Maxam-Gilbert Method
The...
Challenges of the Maxam-Gilbert Method
The...
11.8K