
触媒的交叉メタテシスによるZアルケニルハリドの直接合成
Ming Joo Koh1, Thach T Nguyen1, Hanmo Zhang1
1Department of Chemistry, Merkert Chemistry Center, Boston College, Chestnut Hill, Massachusetts 02467, USA.
Nature
|March 25, 2016

まとめ
研究者は,Z-アルケニルハライドを効率的に生成するオレフィン転化のための新しいモリブデン基触媒を開発しました. この突破は複雑な有機分子を合成し 選択的な化を可能にするために 汎用的な方法を提供します
科学分野:
- 有機化学
- カタリシス
- 有機金属化学
背景:
- オレフィン変異は強力な合成ツールですが,アサイクリックアルケニルハライドを生成することは依然として困難です.
- この変換のための既存のルテニウムベースの触媒は,しばしば不安定,低活性,および貧弱なステレオ選択性に苦しんでいます.
研究 の 目的:
- アサイクリックZ-アルケニルハライドを合成するための効率的で選択的な方法を開発する.
- オレフィンメタテシスにおける新しいホイロ置換モリブデンアルキリデン種の反応性を調査する.
主な方法:
- 新型ハロー置換モリブデンアルキリデン触媒のインサイト生成
- 軽度な条件下 (環境温度,4時間) で,商用 1,2-ジハロエテンの反応剤で反応する.
- 単純で使いやすいため,未浄化液体反応剤を使用します.
主要な成果:
- モリブデン触媒の 極めて高い反応性を示した.
- アサイクリック1,2-非置換アルケニルハリド (塩化物,ブロミド,およびフッ素) に対して,高い収量 (最大91%) と完全なZ選択性を達成した.
- 生物学的活性化合物の成功合成と複雑な分子のサイト/ステレオ選択的化.
結論:
- ハロで置換されたモリブデンアルキリデン種は,オレフィン転移のための非常に効果的な触媒クラスです.
- この方法は,価値あるアルケニルハリドの構成要素に堅牢で効率的でステレオ選択的な経路を提供します.
- 開発された方法論は,複雑な有機分子と先進的なフッ素化合物の合成に重要な意味を持つ.
関連する概念動画
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
8.9K
Introduction
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
8.9K
 02:35
Electrophilic Addition to Alkynes: Hydrohalogenation
11.3K
Electrophilic addition of hydrogen halides, HX (X = Cl, Br or I) to alkenes forms alkyl halides as per Markovnikov's rule, where the hydrogen gets added to the less substituted carbon of the double bond. Hydrohalogenation of alkynes takes place in a similar manner, with the first addition of HX forming a vinyl halide and the second giving a geminal dihalide.
11.3K
 02:34
Preparation of Alkynes: Dehydrohalogenation
17.9K
Introduction
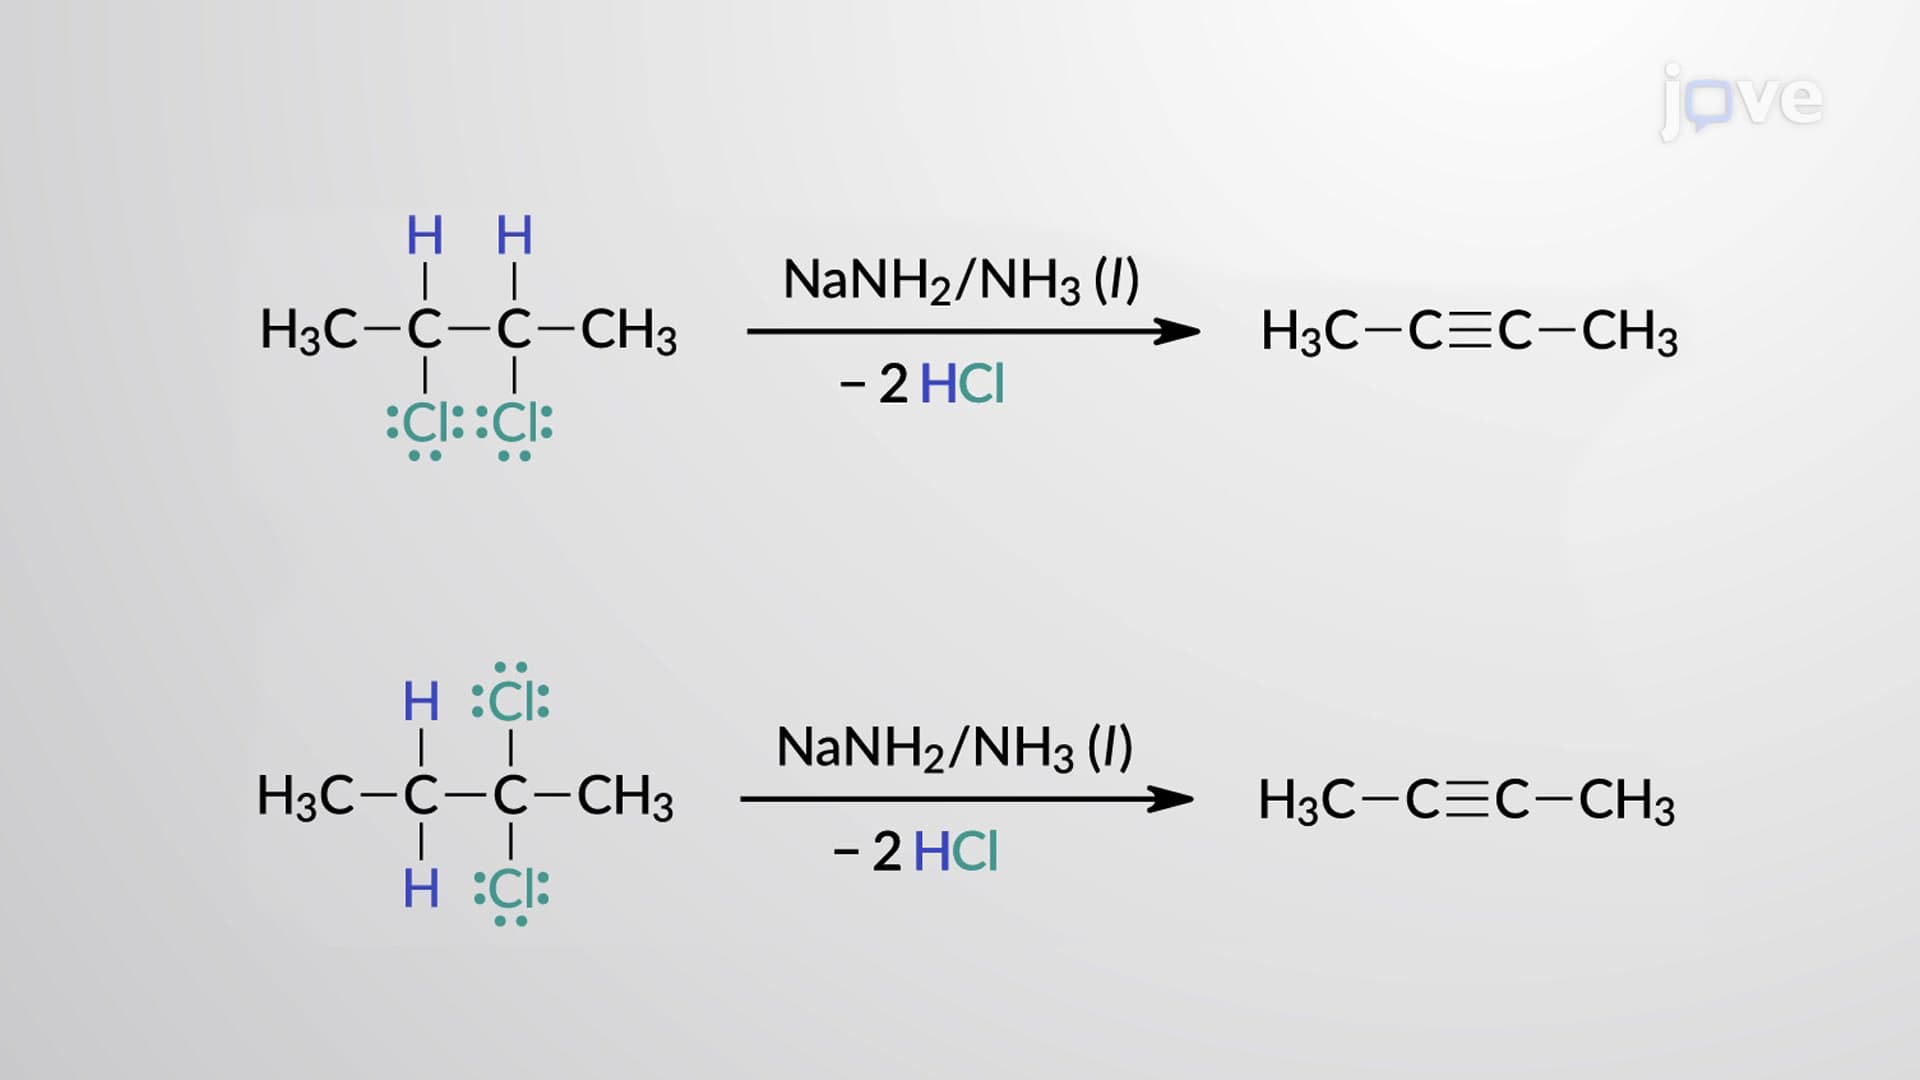
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.
17.9K
 02:27
Preparation of Alkynes: Alkylation Reaction
11.9K
Introduction
Alkylation of terminal alkynes with primary alkyl halides in the presence of a strong base like sodium amide is one of the common methods for the synthesis of longer carbon-chain alkynes. For example, treatment of 1-propyne with sodium amide followed by reaction with ethyl bromide yields 2-pentyne.
Alkylation of terminal alkynes with primary alkyl halides in the presence of a strong base like sodium amide is one of the common methods for the synthesis of longer carbon-chain alkynes. For example, treatment of 1-propyne with sodium amide followed by reaction with ethyl bromide yields 2-pentyne.
11.9K
 02:46
Halogenation of Alkenes
18.3K
Halogenation is the addition of chlorine or bromine across the double bond in an alkene to yield a vicinal dihalide. The reaction occurs in the presence of inert and non-nucleophilic solvents, such as methylene chloride, chloroform, or carbon tetrachloride.
Consider the bromination of cyclopentene. Molecular bromine is polarized in the proximity of the π electrons of cyclopentene. An electrophilic bromine atom adds across the double bond, forming a cyclic bromonium ion intermediate.
Consider the bromination of cyclopentene. Molecular bromine is polarized in the proximity of the π electrons of cyclopentene. An electrophilic bromine atom adds across the double bond, forming a cyclic bromonium ion intermediate.
18.3K
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
4.7K
By replacing an α-hydrogen with a halogen, acid-catalyzed α-halogenation of aldehydes or ketones yields a monohalogenated product
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
4.7K