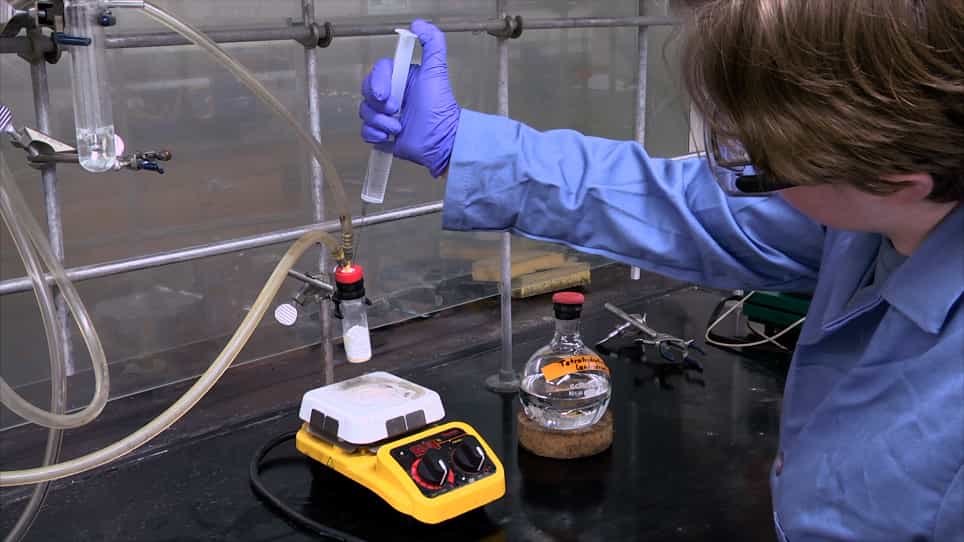
ピリジンのC-α-モノボリレーションによる化学分岐性オルガノランタニド触媒
Jacob O Rothbaum1, Alessandro Motta2, Yosi Kratish1
1Department of Chemistry, Northwestern University, Evanston, Illinois 60208-3113, United States.
Journal of the American Chemical Society
|September 8, 2022

まとめ
オルゴランタニド触媒は,ピリジンの選択的アルファ・モノボリレーションを可能にします. この化学分岐アプローチは 価値ある有機分子と 機能的な材料を合成するための 新しい経路を提供します
科学分野:
- 有機金属化学
- カタリシス
- 合成有機化学
背景:
- ピリディノイドアジンのC-H活性化と機能化は,天然製品,医薬品,および材料の合成に不可欠である.
- アジンのアルファ位置の機能化は,触媒の調整とピリジンの電子欠乏を含むステリックおよび電子的要因のために困難である.
研究 の 目的:
- アジンの選択的アルファ単一機能化のための地球豊富な触媒を開発する.
- 有機製品に構造的多様性を導入するための合成化学的方法論を実現する.
主な方法:
- 18種類のピリジンのアルファ・モノボリレーションのために,Cp*2LuCH(TMS) 2を用いたオルゴランタニド触媒を用いた.
- 反応経路を明らかにするために,実験的および理論的メカニズム研究を使用した.
- ランタニド同一性と基質の電子特性による触媒選択性の影響を調査した.
主要な成果:
- オルゴランタニド触媒を用いた多様なピリジンの選択的アルファ・モノボリレーションを達成した.
- ランタニドイオンと基質の電子特性によって前例のない触媒化学分岐を証明した.
- C-Hで活性化されたeta(2) -ランタニド-アジン複合体の中間物質を特定し,シグマ結合転移を含むメカニズムを提案した.
結論:
- ピリジンの選択的アルファ・モノボリレーションのための新しいオルガノランタニド触媒法を開発した.
- 選択的なC-H機能化またはB-N結合形成の脱オロマ化を可能にするランタニド系触媒化学分岐を展示した.
- 有機合成におけるさらなる機能化のための貴重な前駆物質を提供した.
関連する概念動画
 02:36
Regioselectivity and Stereochemistry of Hydroboration
8.4K
A significant aspect of hydroboration–oxidation is the regio- and stereochemical outcome of the reaction.
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
8.4K
 03:08
Hydroboration-Oxidation of Alkenes
8.7K
In addition to the oxymercuration–demercuration method, which converts the alkenes to alcohols with Markovnikov orientation, a complementary hydroboration-oxidation method yields the anti-Markovnikov product. The hydroboration reaction, discovered in 1959 by H.C. Brown, involves the addition of a B–H bond of borane to an alkene giving an organoborane intermediate. The oxidation of this intermediate with basic hydrogen peroxide forms an alcohol.
8.7K
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
18.7K
Introduction
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
18.7K
 00:51
Base-Promoted α-Halogenation of Aldehydes and Ketones
3.6K
α-Halogenation of aldehydes and ketones is a reaction involving the substitution of α hydrogens with halogens in the presence of a base. The reaction begins with the abstraction of α hydrogen by the base to produce a nucleophilic enolate ion. This intermediate undergoes a subsequent nucleophilic substitution with the halogen to produce a monohalogenated carbonyl compound. If the starting substrate has more than one α hydrogen, it is difficult to stop the reaction...
3.6K
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
6.2K
All ortho–para directors, excluding halogens, are activating groups. These groups donate electrons to the ring, making the ring carbons electron-rich. Consequently, the reactivity of the aromatic ring towards electrophilic substitution increases. For instance, the nitration of anisole is about 10,000 times faster than the nitration of benzene. The electron-donating effect of the methoxy group in anisole activates the ortho and para positions on the ring and stabilizes the corresponding...
6.2K
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
4.0K
By replacing an α-hydrogen with a halogen, acid-catalyzed α-halogenation of aldehydes or ketones yields a monohalogenated product
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...
4.0K