関連する実験動画
Updated: Apr 17, 2026
22:27

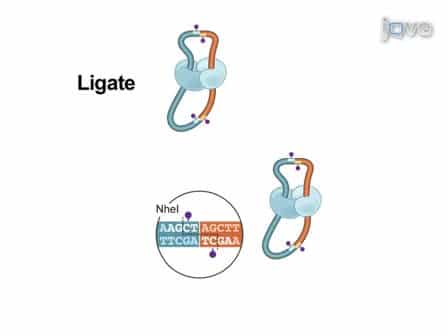
Hi-C: A Method to Study the Three-dimensional Architecture of Genomes.
Published on: May 6, 2010
409.6K
ロングリースシーケンシングとHi-C技術の組み合わせで,がんにおけるクロモアゲネシス現象を特定する
Marius-Konstantin Klever1,2, Julius Jungnitsch3,4, Lars Bullinger5,6
1Division of Hematology, Oncology, and Cancer Immunology, Medical Department, Charité-Universitätsmedizin Berlin, Corporate Member of Freie Universität Berlin, Humboldt-Universität zu Berlin, and Berlin Institute of Health, Berlin, Germany. marius-konstantin.klever.01@regionh.dk.
Methods in molecular biology (Clifton, N.J.)
|August 30, 2025

まとめ
この研究は,がん遺伝子の複雑な構造変異を正確に検出するために,ロングリードとHi-C配列を組み合わせた新しいワークフローを導入し,がん研究に高解像度を提供します.
科学分野:
- ゲノミクス
- 癌 の 遺伝子
- バイオ情報学
背景:
- 構造変異 (SV) はがん遺伝学において極めて重要です.
- 配列CGH,カリオタイプ,全ゲノム配列などの既存の検出方法は,特にクロモアゲネシスでは,複雑なSVの解像度と精度が欠けている.
- 複雑なSVを理解することは,がんの病理メカニズムを解明するために不可欠です.
研究 の 目的:
- 新しい構造変異検出のワークフローを提示します.
- 非常に高い解像度で高信頼性SVを呼び出すために.
- 癌におけるクロモアゲネシスの複雑さについて 新たな洞察を得るために
主な方法:
- ゲノムDNA (gDNA) の長読み配列の統合
- Hi-C 配列の統合
- 新しいSV検出ワークフローの開発
主要な成果:
- このワークフローは,非常に高い解像度で高信頼性の構造変数呼び出しを可能にします.
- 複雑なカリオタイプを持つ急性骨髄性白血病 (AML) に適用すると,クロモアゲネシスの複雑性に関する新しい洞察が明らかになりました.
- この方法は,その後の機能研究の可能性を高めます.
結論:
- この新しいワークフローは複雑な構造変異の検出を大幅に改善します
- この進歩により,AMLのような癌におけるクロモアゲネシスの理解が深まります.
- この発見は将来のがん病理学的研究を導くことができます.
関連する概念動画
 03:00
Next-generation Sequencing
102.1K
The first human genome sequencing project cost $2.7 billion and was declared complete in 2003, after 15 years of international cooperation and collaboration between several research teams and funding agencies. Today, with the advent of next-generation sequencing technologies, the cost and time of sequencing a human genome have dropped over 100 fold.
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
102.1K
 03:21
RNA-seq
12.7K
RNA sequencing, or RNA-Seq, is a high-throughput sequencing technology used to study the transcriptome of a cell. Transcriptomics helps to interpret the functional elements of a genome and identify the molecular constituents of an organism. Additionally, it also helps in understanding the development of an organism and the occurrence of diseases.
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
12.7K
 03:36
Genome Annotation and Assembly
22.2K
The genome refers to all of the genetic material in an organism. It can range from a few million base pairs in microbial cells to several billion base pairs in many eukaryotic organisms. Genome assembly refers to the process of taking the DNA sequencing data and putting it all back together in a correct order to create a close representation of the original genome. This is followed by the identification of functional elements on the newly assembled genome, a process called genome annotation.
22.2K