
適度なCYP3A阻害剤との薬物相互作用の評価のためのヒトの血内の内生性バイオマーカーとしての1β-Hydroxydeoxycholic acid
Yizhe Chen1, Runlan Huo1, Brian Melo1
1Translational Medicine and Clinical Pharmacology, Bristol-Myers Squibb, Princeton, New Jersey.
Drug metabolism and disposition: the biological fate of chemicals
|September 6, 2025

まとめ
血中の総1β- 酸化塩酸 (1β- OH DCA) は,薬物相互作用 (DDI) のバイオマーカーとしての可能性を示している. このバイオマーカーは,他の方法とは異なり,中程度のCYP3A阻害と強いCYP3A阻害の両方を予測することができます.
科学分野:
- 薬理学と薬物の代謝
- バイオマーカーの発見
- 薬物相互作用 (DDI) の評価
背景:
- 1β-Hydroxydeoxycholic acid (1β-OH DCA) は,CYP3A誘導体および抑制剤を含む薬物相互作用 (DDI) を評価するための潜在的なバイオマーカーです.
- 4β-ヒドロキシコレステロールのような既存のバイオマーカーは,抑制剤ではなくCYP3A誘発剤を特定することに限定されています.
- 1β-OH DCAの合計は,CYP3A調節に対する反応として,4β-ヒドロキシコレステロールよりも有意な変化を示した.
研究 の 目的:
- 適度なCYP3A阻害剤によるDDIのバイオマーカーとしての合計1β-OHDCAの有用性を評価する.
- 1β-OH DCAおよびその結合体に対する定量測定法を改善する.
- DDI中の合計1β-OH DCA暴露とCYP3A基質 (フェドラチニブ) 曝露の相関を評価する.
主な方法:
- 安定した内定基準を用いて,総1β-OH DCAの強化された定量分析を開発した.
- 抽出とクロマトグラフィの条件を最適化して,サンプル浄化と分離を改善します.
- フェドラチニブ (CYP3A基質) をフルコナゾール (中程度のCYP3A阻害剤) と併用して投与し,バイオマーカーの反応を評価した.
主要な成果:
- フルコナゾルの抑制により,合計1β-OH DCAのCmax (39%) とAUC (0~24hでは39%,0~216hでは18%) が著しく低下した.
- 合計1β-OH DCA被曝の幾何学平均比率は,フェドラチニブ被曝比率と良好に相関していた.
- これらの発見は,総1β-OH DCAが中程度の阻害剤によるCYP3A阻害を反映していることを示唆している.
結論:
- 血における総1β-OH DCAは,CYP3Aの活動範囲におけるDDIを予測する多用途のバイオマーカーとして有望である.
- このバイオマーカーは,特に中程度のCYP3A阻害に関して,既存のDDI評価戦略を補完します.
- 予備的なデータによると,総1β-OH DCAは,CYP3Aの中等および強い阻害の両方の予測を可能にし,これは他のバイオマーカーにはない能力である.
さらに関連する動画
関連する概念動画
 01:16
Hepatic Drug Excretion: Influencing Factors
214
The biliary system of the liver, crucial for bile secretion and drug excretion, comprises intrahepatic bile ducts that merge to form the common hepatic duct. This duct, carrying hepatic bile, combines with the cystic duct, draining the gallbladder and forming the common bile duct, which empties into the duodenum. Bile, produced by hepatic cells lining the bile canaliculi, is composed primarily of water, bile salts, pigments, electrolytes, and lesser amounts of cholesterol and fatty acids. Bile...
214
 01:14
Time Course of Drug Effect
2.2K
The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50...
2.2K
 01:17
Hepatic Drug Excretion: Enterohepatic Cycling
1.7K
Enterohepatic cycling involves the active secretion of drugs and their metabolites into the bile via transporters in the canalicular membrane of hepatocytes. This secretion is an integral part of the digestive process, releasing these substances into the gastrointestinal (GI) tract.
Post-release drugs and metabolites can be reabsorbed into the body from the intestine. For conjugated metabolites like glucuronides, reabsorption requires enzymatic hydrolysis by intestinal microflora. This...
Post-release drugs and metabolites can be reabsorbed into the body from the intestine. For conjugated metabolites like glucuronides, reabsorption requires enzymatic hydrolysis by intestinal microflora. This...
1.7K
 01:09
Hepatic Drug Clearance: Effect of Protein Binding
284
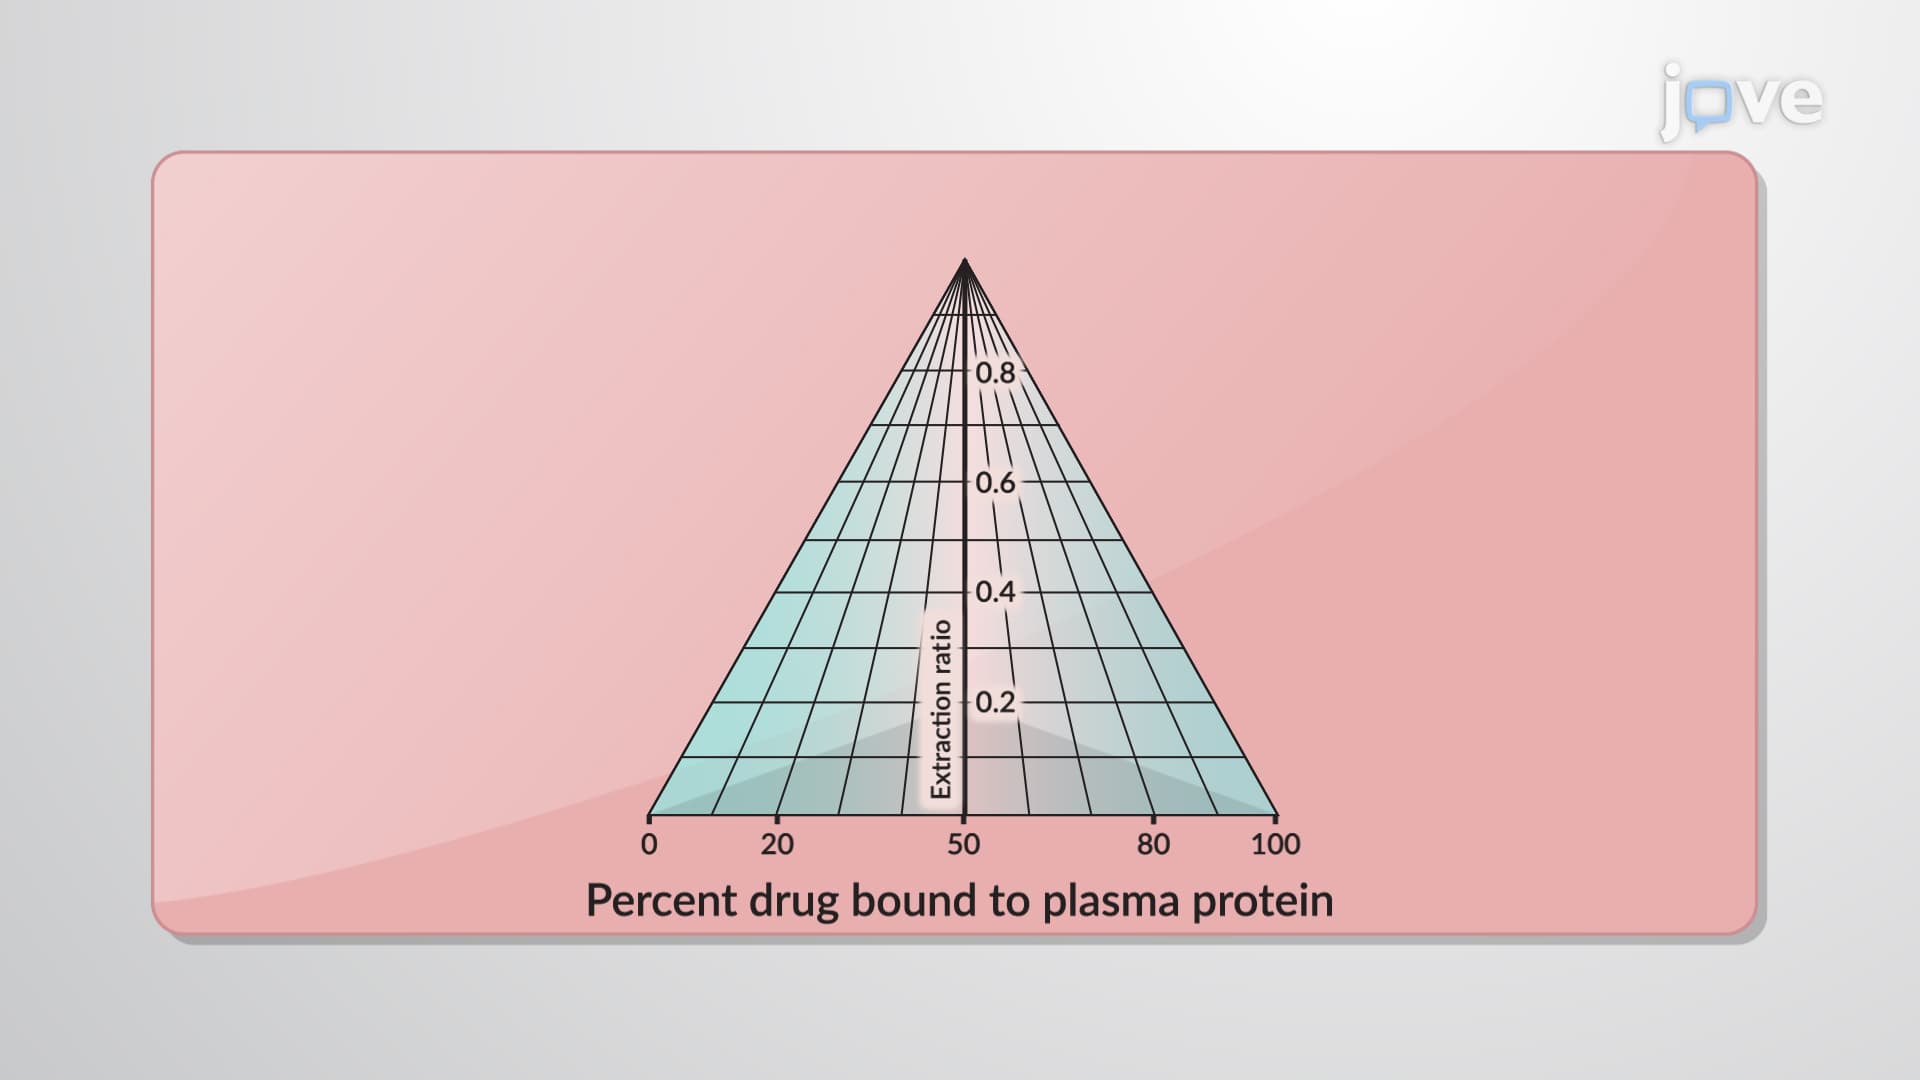
Hepatic clearance is influenced by protein binding based on the drug's extraction ratio. Drugs with high extraction ratios are considered flow-limited and remain unaffected by protein binding during hepatic clearance. On the other hand, drugs with low extraction ratios may be impacted by plasma protein binding, although the extent of this influence depends on the fraction of the drug bound.
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
284
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
78
Drug transporters are critical in drug absorption, distribution, and excretion processes. They should be included in physiological-based pharmacokinetic (PBPK) models, which help predict human drug disposition. However, predicting this is challenging during drug development, especially when liver transport is involved. However, with a realistic representation of body transport processes, an accurate model may be possible.
A recent model describes pravastatin's hepatobiliary excretion,...
A recent model describes pravastatin's hepatobiliary excretion,...
78
 01:14
Hepatic Drug Clearance: Role of Transporters
100
In the liver and bile canaliculi, influx and efflux transporters modification can influence intrinsic clearance. Transporters play a significant role in moving drugs within liver cells. Elaborate models, such as the Biopharmaceutical Classification System (BCS), are essential to relate transporters to drug disposition. This system categorizes drugs into four classes based on solubility and permeability, providing insights into elimination routes and the effects of transporters following oral...
100