

複雑な系の第一原理分子動力学熱伝導率計算の加速
Sandro Wieser1, Yu-Jie Cen1, Georg K H Madsen1
1Institute of Materials Chemistry, TU Wien, A-1060 Vienna, Austria.

Abstract:
Atomistic simulations of heat transport in complex materials are costly and hard to converge. This has led to the development of several noise-reduction techniques applicable to equilibrium molecular-dynamics (MD) simulations. We analyze the performance of those strategies, taking InAs nanowires as our benchmark due to the diverse structures and complex phonon spectra of these quasi-1D systems. We demonstrate how, for low-thermal-conductivity systems, cepstral analysis can reduce computational demands while still delivering accurate results that do not require discarding arbitrary parts of the data set. However, issues with this approach are revealed when treating high-thermal-conductivity systems, where the thermal conductivity is significantly underestimated. We discuss alternative methods to be used in that situation, relying on uncertainty propagation from independent simulations. We show that the contributions of the covariance matrix have to be included for a quantitative assessment of the error. The combination of these strategies with machine-learning interatomic potentials (MLIPs) provides an accelerated, robust workflow applicable to a diverse set of systems, as our examples using a highly transferable MACE potential illustrate.
さらに関連する動画
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
関連する概念動画
 01:17
Theory of Metallic Conduction
In this theory, Newton's second law of motion is used to determine the acceleration of an electron in the presence of an applied electric field. Then, its velocity is expressed via this acceleration.
An electron moves through the crystal, containing positive ions,...
 01:24
Thermodynamics: Activity Coefficient
The activity coefficient is a measure of the deviation from ideal behavior. When the ionic strength of the solution is minimal, the activity coefficient of an ionic species is close to unity, making...
 01:26
Thermodynamic Potentials
 01:16
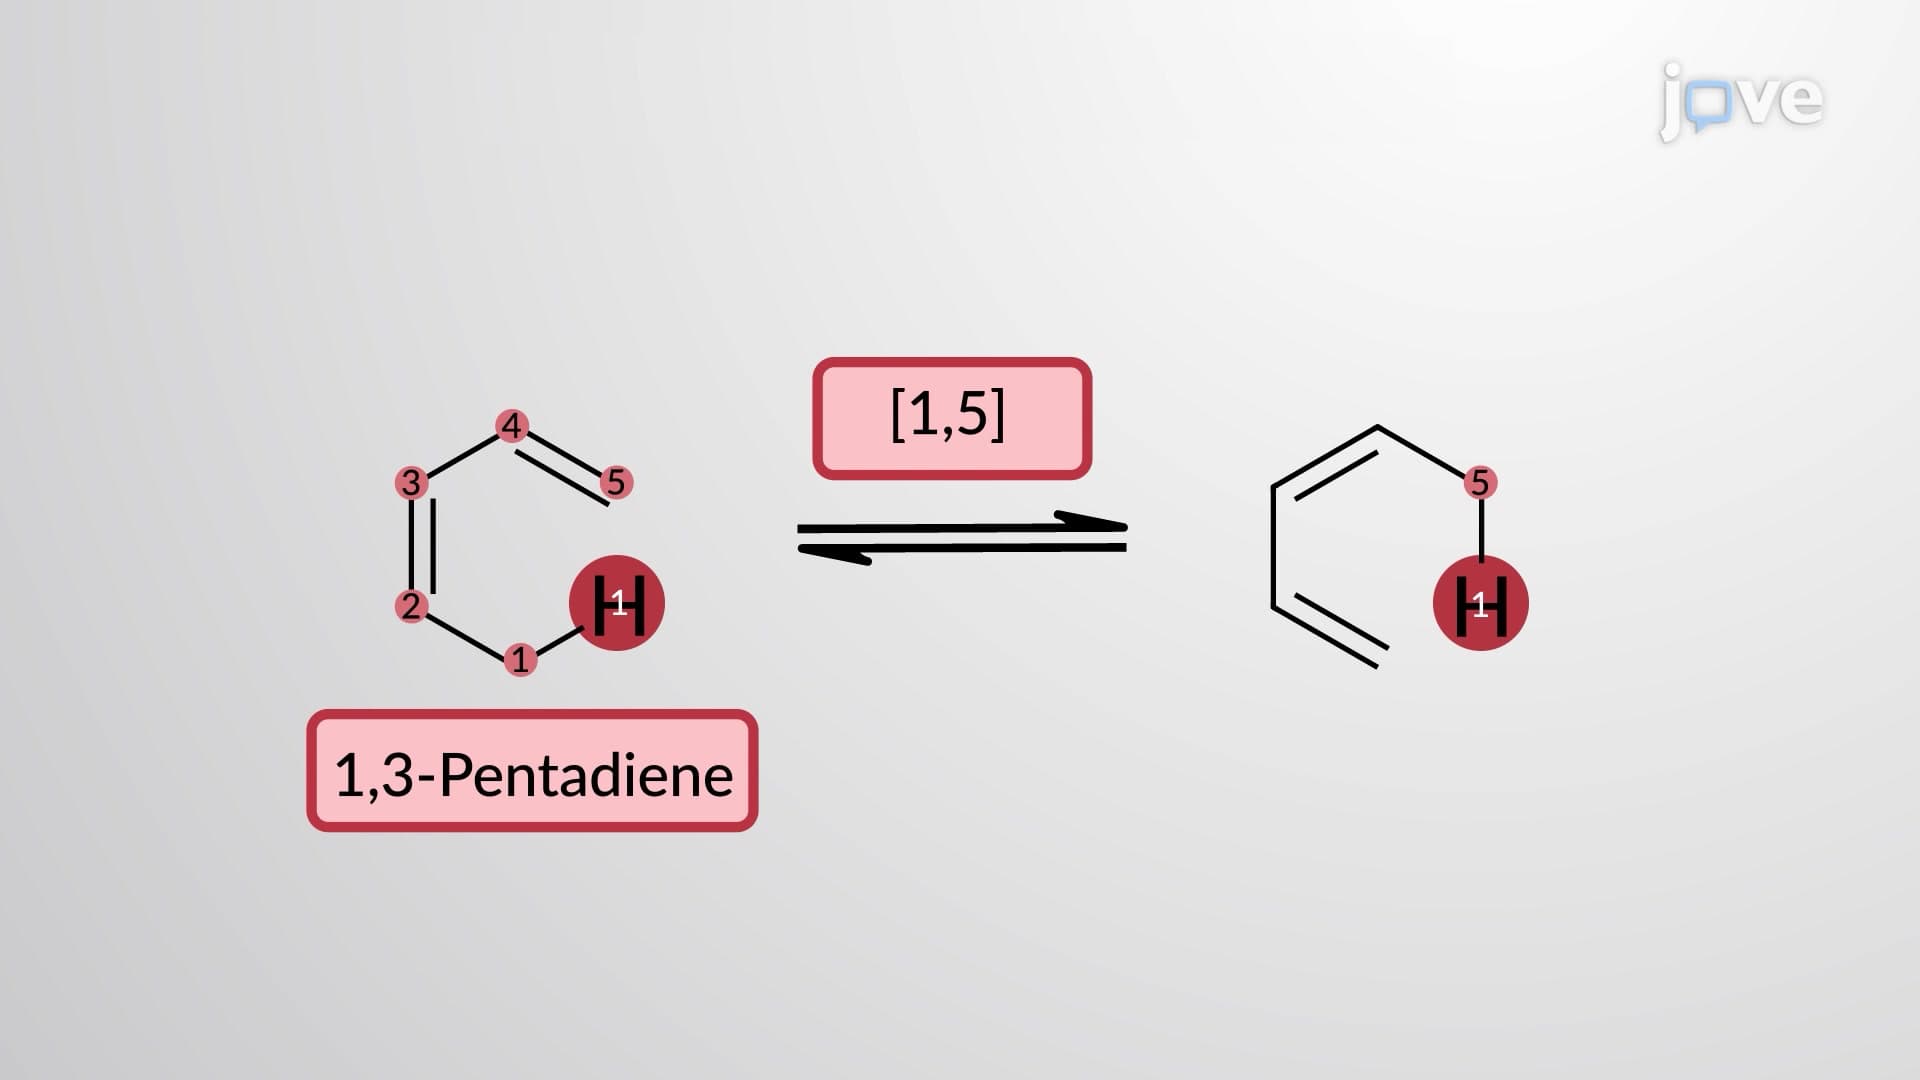
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 01:20
Mechanisms of Heat Transfer II
 01:14
Mechanisms of Heat Transfer I