AI駆動のBRAF阻害剤のde novo設計:結合親和性と薬剤様性の向上
Zuokun Lu1,2, Aili Zhang1
1Food and Pharmacy College, Xuchang University, Xuchang, Henan, China.

Background:
Traditional drug discovery methods, such as high-throughput screening (HTS), are often inefficient and costly, especially in complex areas like oncology. The BRAF V600E mutation is a validated therapeutic target in cancers such as melanoma, thyroid carcinoma, and colorectal cancer. However, existing BRAF inhibitors face challenges like acquired resistance and off-target toxicity. Artificial intelligence (AI) has emerged as a transformative tool for designing novel inhibitors more efficiently.
Methods:
This study employed REINVENT 4, an advanced machine learning (ML) framework using recurrent neural networks and transformer architectures, for targeted generation and property optimization of BRAF V600E inhibitors, integrating reinforcement learning (RL) for drug-likeness optimization and transfer learning (TL) for mutation-specific design. Molecular docking and dynamics simulations were used to evaluate binding affinity and stability.
Results:
The AI-driven approach generated 41,721 novel BRAF V600E inhibitor candidates with enhanced drug-likeness (mean Quantitative Estimate of Drug-likeness (QED) score: 0.61 ± 0.17 vs. the training set 0.40 ± 0.13) and predicted inhibitory activity (83.8% with predicted pIC50 > 6). The generated compounds showed a 32% reduction in mean molecular weight (326.8 ± 45.6 g/mol vs. 480.8 ± 84.2 g/mol in the training set) while maintaining inhibitory potency. Pharmacokinetic analysis revealed that 99.7% of generated compounds satisfied Lipinski's Rule of Five criteria, suggesting favorable absorption and distribution profiles. Molecular docking analysis of selected compounds revealed strong binding affinities, with an average free energy of -8.03 ± 1.12 kcal/mol and top-performing compounds reaching -11.5 kcal/mol. Molecular dynamics simulations conducted over 200 ns confirmed complex stability, demonstrating protein backbone RMSD values of 0.35-0.55 nm and ligand RMSD values of 0.086-0.161 nm. Structural novelty assessment using Tanimoto similarity coefficients showed values below 0.45 when compared with FDA-approved BRAF inhibitors (including Sorafenib and Vemurafenib).
Discussion:
This work highlights a reproducible, integrated AI-driven workflow demonstration for targeted inhibitor generation. The generated inhibitors exhibit favorable drug-like properties and inhibitory activity, offering a scalable solution for designing safer cancer therapies. Experimental validation is needed to address potential discrepancies between computational predictions and biological behavior.
さらに関連する動画
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 05:54
Generation of High-Throughput Three-Dimensional Tumor Spheroids for Drug Screening
Published on: September 5, 2018
関連する概念動画
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against...
 02:50
Combination Therapies and Personalized Medicine
The combination of the drug acetazolamide and sulforaphane is a good example of combination therapy to treat cancer. The cells in the interior of a large tumor often die due to the hypoxic and...
 01:26
Drug Discovery: Overview
 01:16
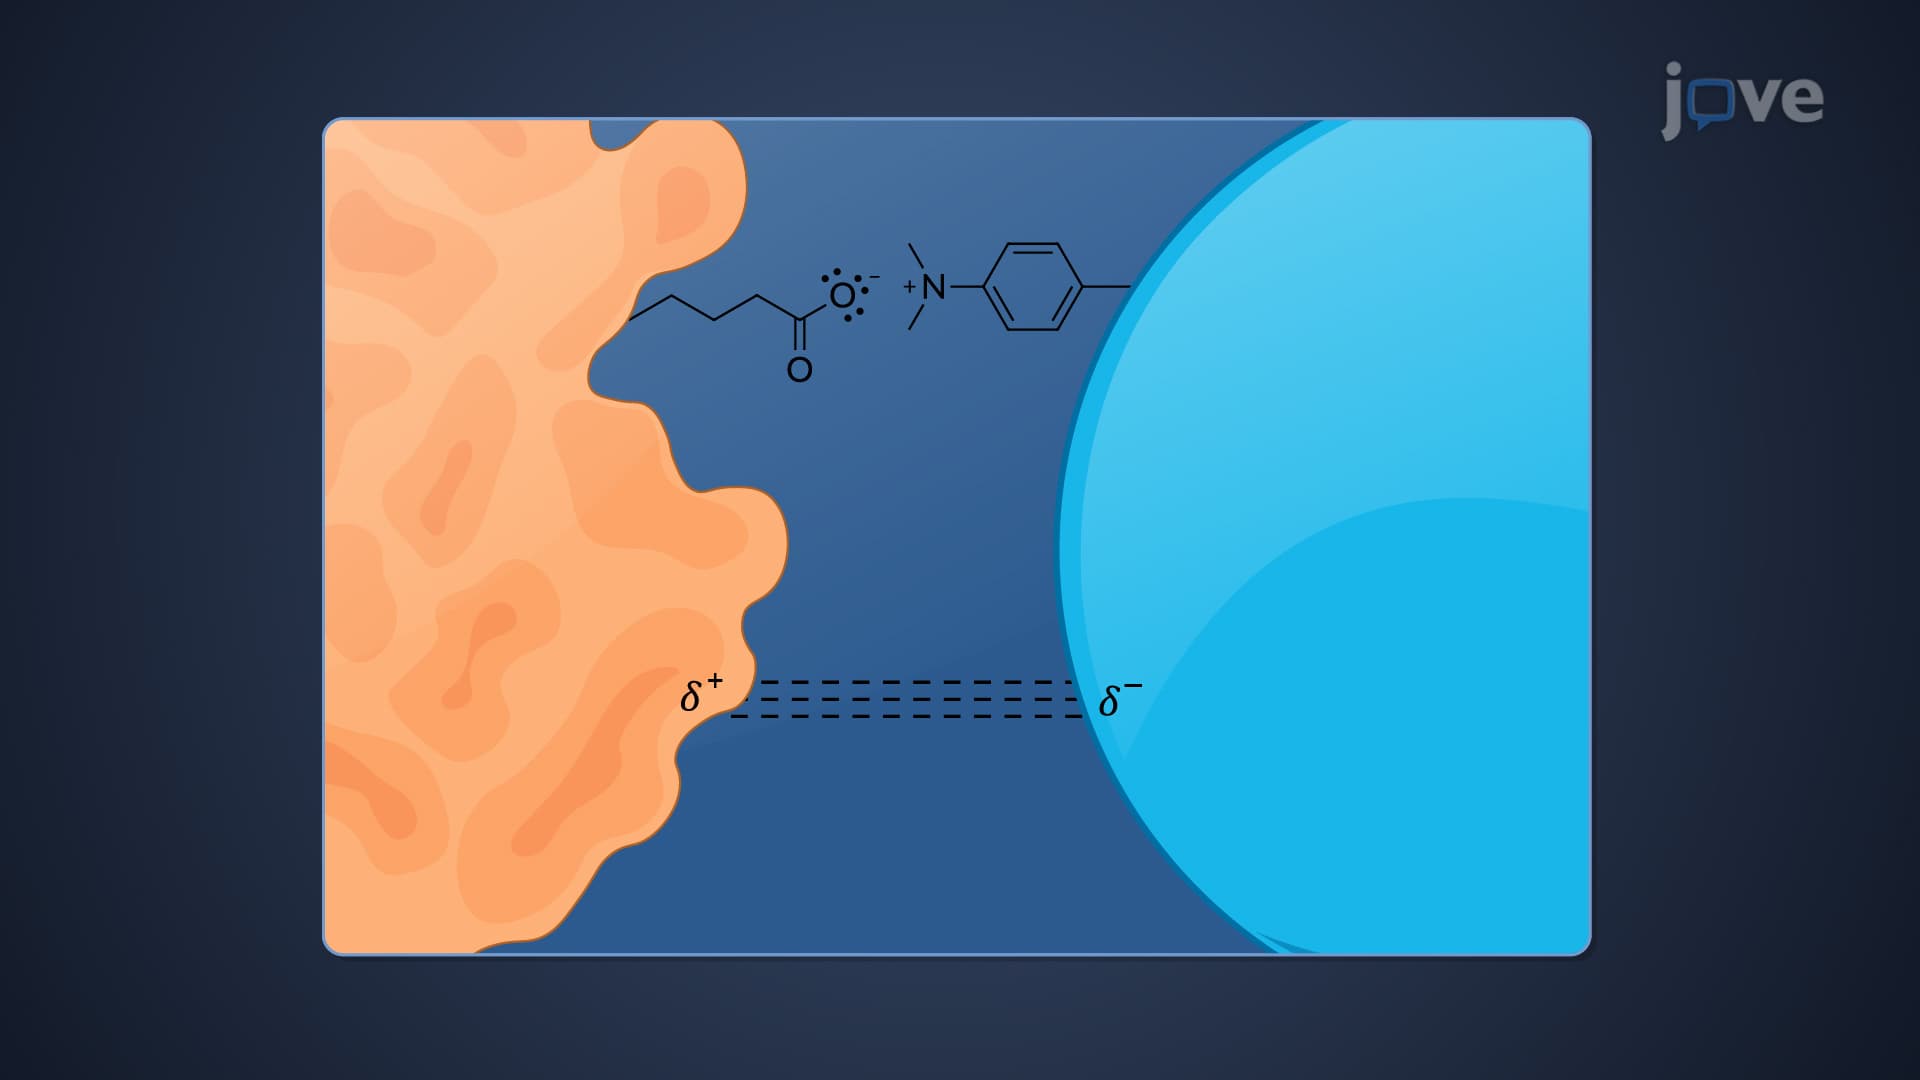
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:27
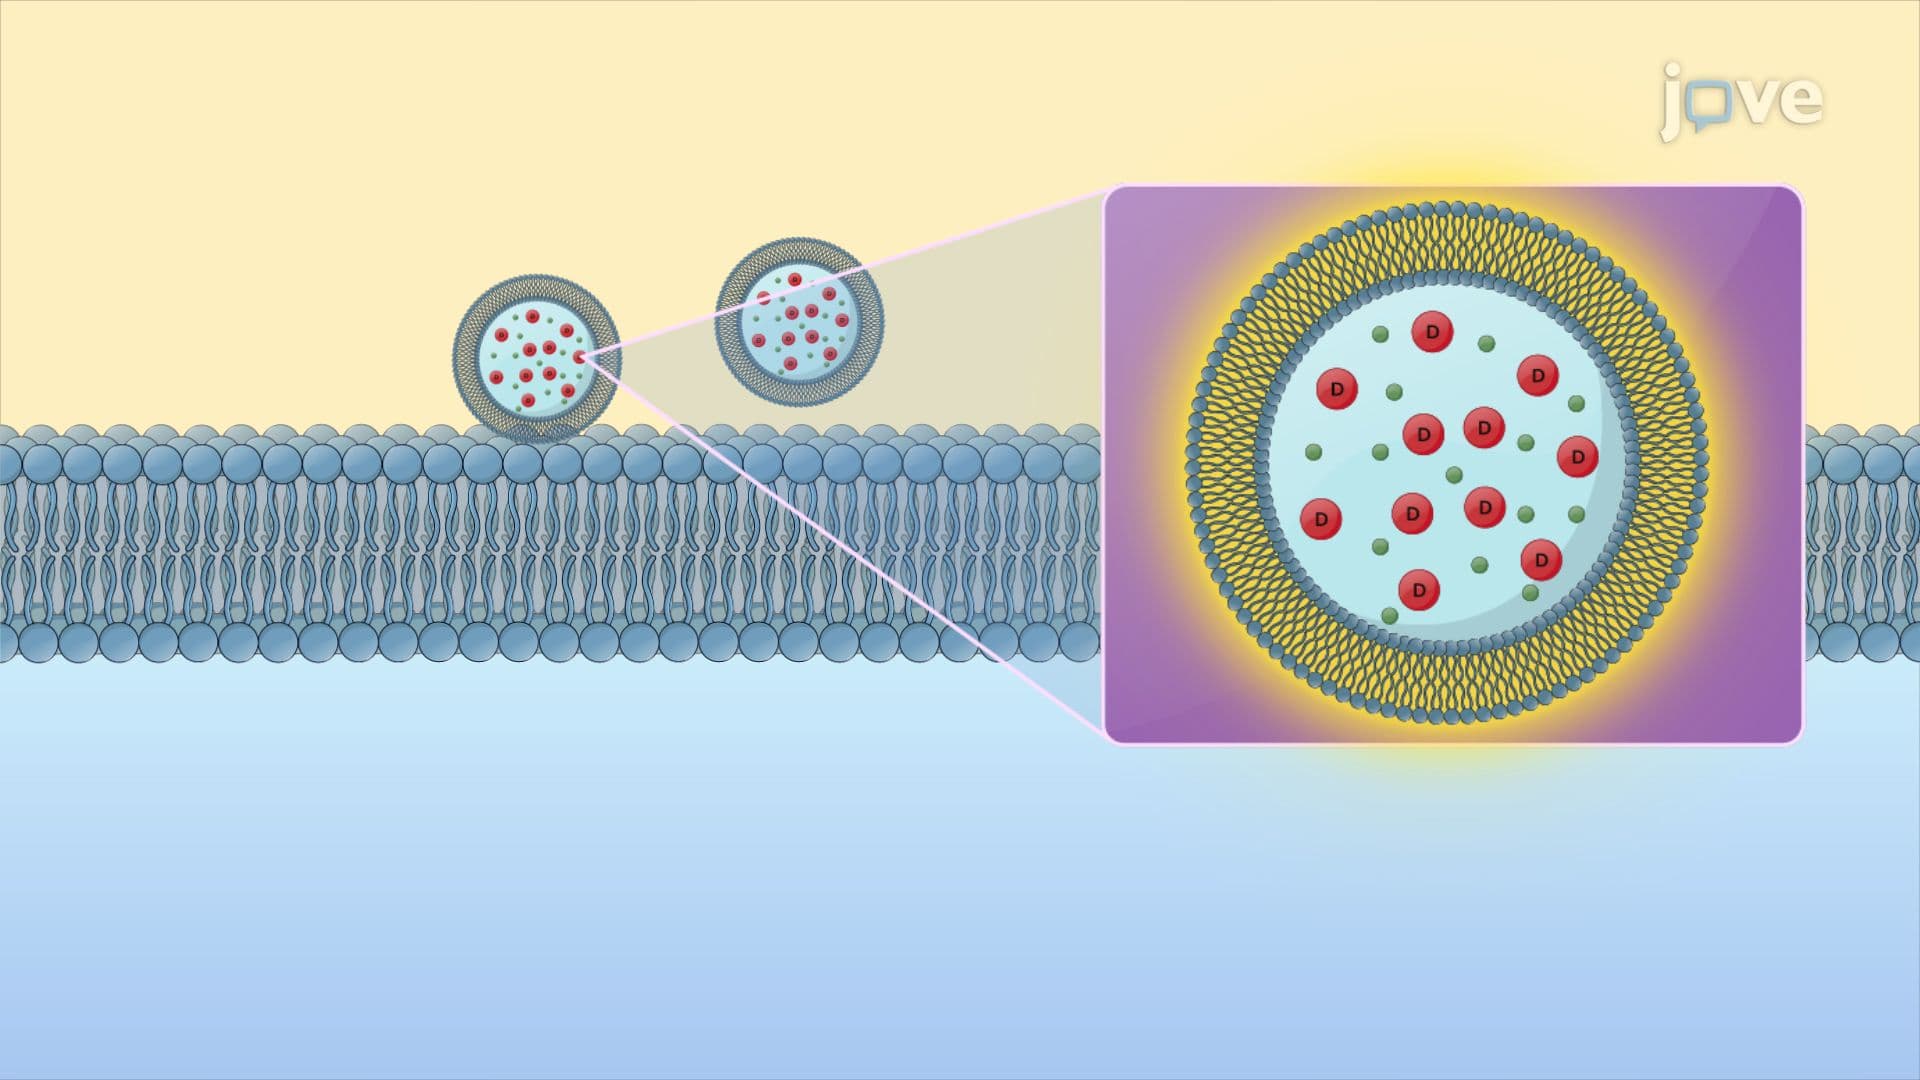
Bioavailability Enhancement: Drug Permeability Enhancement