効率的なタンパク質-リガンド結合自由エネルギー推定のための粗視化ファネルメタダイナミクス

Andrea Grazzi1, Chelsea M Brown2, Maurizio Sironi1
1Department of Chemistry, University of Milan, Via C. Golgi 19, 20133 Milan, Italy.

Abstract:
Despite considerable advances in computational chemistry, bridging the gap between the accuracy of all-atom molecular dynamics (AA-MD) and the high-throughput capabilities of docking remains an unsolved problem in protein-ligand binding free energy predictions. In this work, we propose to address this challenge through coarse-grained funnel metadynamics (CG-FMD) with the Martini 3 force field. This approach combines the reduced computational cost of a CG representation with state-of-the-art enhanced sampling techniques and the interpretability of a physics-based force field. Specifically, the binding of colchicine to two different protein targets was modeled at both AA and CG resolutions, and the corresponding ΔGbind predictions were compared with experimental references. Additionally, the robustness of CG-FMD-based ΔGbind predictions was evaluated with respect to various aspects of the simulation setup by collecting more than 7 ms of CG-FMD simulations. The optimal simulation protocol has been further validated against a limited set of compounds chemically different from colchicine. The results demonstrate that CG-FMD yields ΔGbind estimates comparable to experimental values while requiring only a fraction of the computational cost of AA-MD simulations. Moreover, the extensive sampling achievable with CG-FMD reduces statistical uncertainty in the final predictions, effectively compensating for the simplified system representation. Future work should build upon these methodological insights to broaden the scope of ligands and targets explored.
関連する概念動画
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
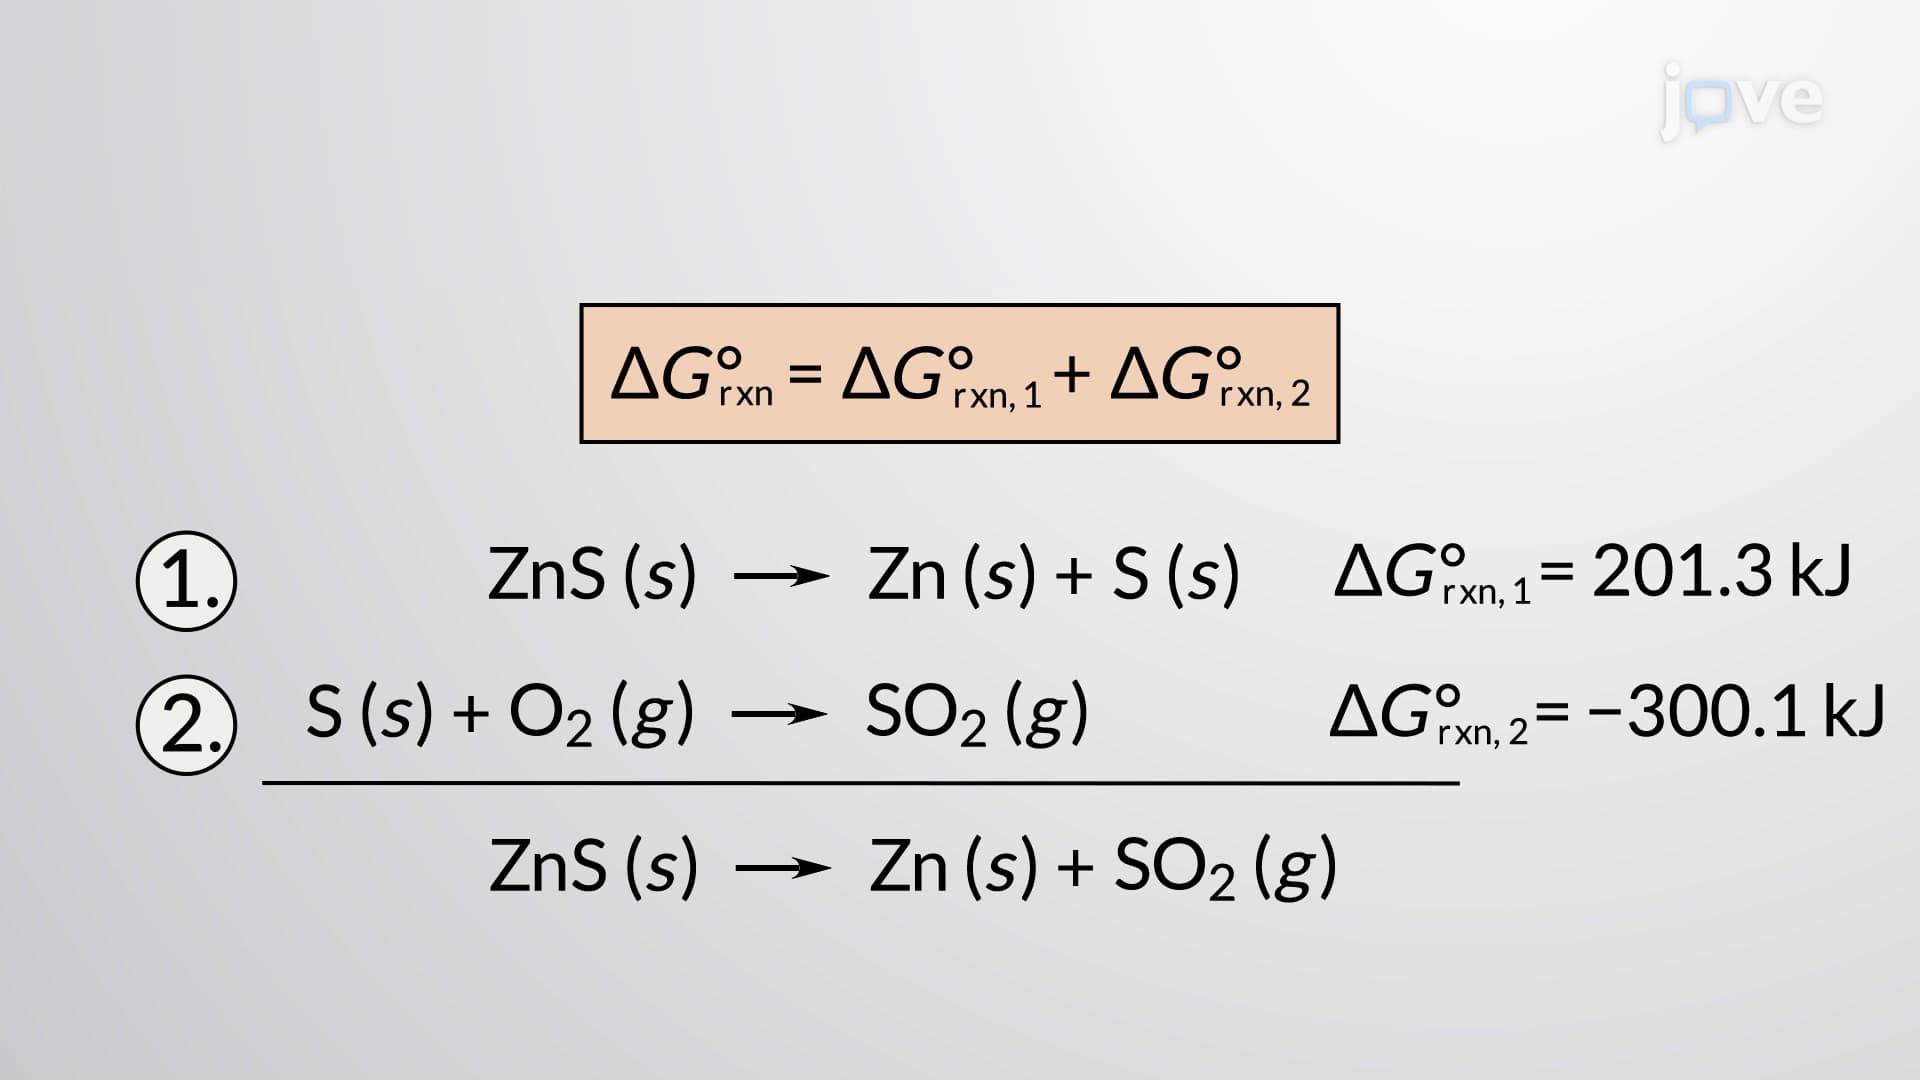 02:49
Calculating Standard Free Energy Changes