
プロピオン酸血症のアミッシュ患者における心臓および心肝移植
Evan H Whitehead1, Jeffrey Bennett2, Pavan Bhat1
1Department of Cardiovascular Medicine, Heart and Vascular Institute, Cleveland Clinic, Cleveland, OH, USA.

Background And Aim:
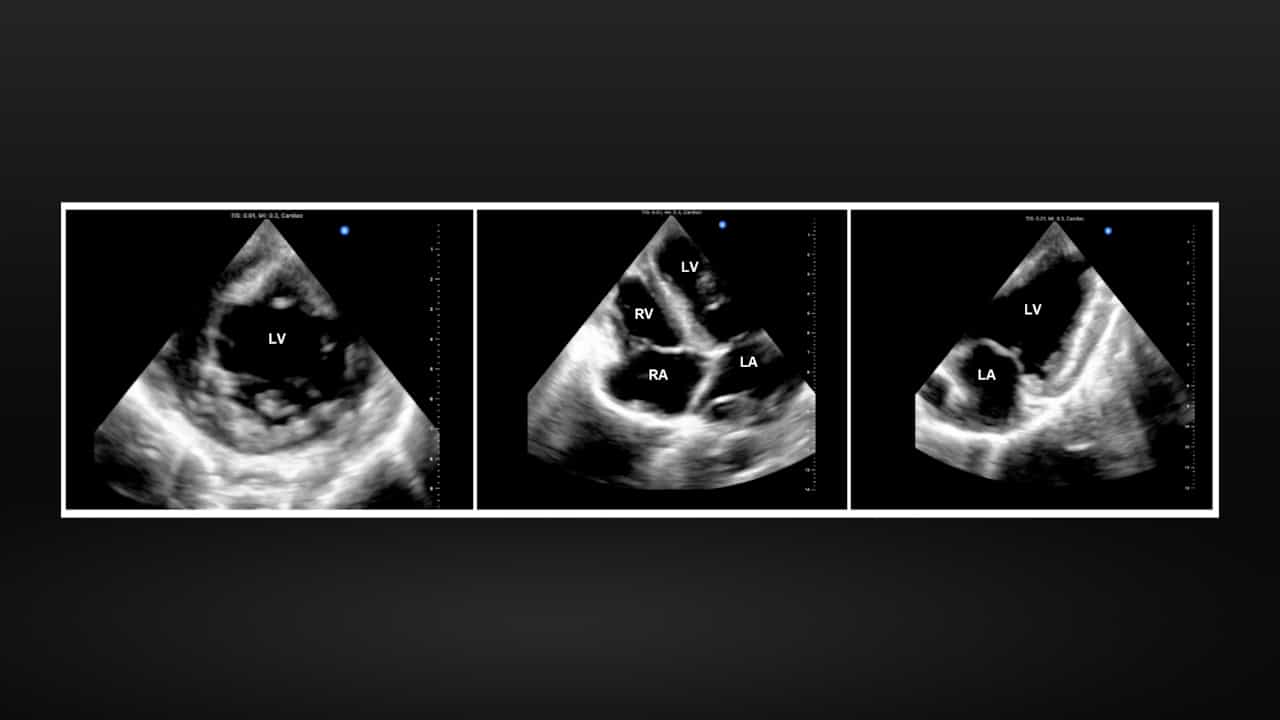
Propionic acidemia (PA) is a genetic metabolic disorder caused by deficient activity of the enzyme propionyl-CoA carboxylase, resulting in accumulation of toxic metabolites during catabolism of odd-chain fatty acids and branched-chain amino acids. Most PA occurs in compound heterozygotes, typically presenting with metabolic acidosis and seizures in infancy. A milder phenotype of PA is prevalent in the Amish population due to a founder missense variant in PCCB (c.1606 A > G; p.Asn536Asp) and is frequently present as an isolated dilated cardiomyopathy in adolescence.
Methods And Results:
Here we report our experience with three Amish patients with genetically confirmed PA and end-stage heart failure. While one patient underwent successful heart transplantation with no complications, another developed recurrent cardiogenic shock after transplant due to metabolic decompensation. Based on this experience, a subsequent patient was treated with combined heart/liver transplantation.
Conclusions:
These cases highlight unique challenges in managing patients with metabolic cardiomyopathies and emphasize the importance of a multidisciplinary approach to achieve the best possible outcomes.