
光学特性の計算のための密度機能的緊密結合分子力学アプローチの適用性に関する研究
Tengxiang Li1, Ruicheng Li1, Keisuke Kameda1
1School of Materials and Chemical Technology, Institute of Science Tokyo, Tokyo, Japan.
Journal of computational chemistry
|February 17, 2026

まとめ
分子力学 (DFTB-MM) による密度関数的緊密結合は,大規模光学特性計算に適しています. 非結合パラメータを省略すると,スペクトルの精度が最小限に影響され,DFTBアプリケーションが拡張されます.
科学分野:
- 計算化学はコンピュータ化学である.
- マテリアルサイエンス 材料科学
- 量子力学は,量子力学という
背景:
- 密度関数緊固結合 (DFTB) と時間依存型 (TD) -DFTBは,Kohn-Sham密度関数理論 (DFT) よりも大きなスケールでの電子および光学特性研究を可能にします.
- 分子力学によるDFTB (DFTB-MM) は,特定の原子組の分子力学を組み込むことにより,正確なスレーター-コスター (SK) パラメータが欠けているシステムにDFTBの適用性を拡張します.
研究 の 目的:
- 光学特性の計算のための線形応答TD-DFTBと組み合わせたDFTB-MMの有効性を評価する.
- 非結合原子ペアのSKパラメータがスペクトル精度に与える影響を評価する.
- 分子-TiO2ナノ粒子システムのためのDFT計算に対してTD-DFTB-MMを検証する.
主な方法:
- 分子力学による線形応答時間依存型DFTB (TD-DFTB-MM).
- インタフェースの電荷移転を示す分子-TiO2ナノ粒子システムのモデリング.
- TD-DFTB-MMの結果と,さまざまな関数を使用したDFT計算の比較.
主要な成果:
- TD-DFTB-MMは,インターフェイスの電荷伝送帯やハイブリッド化効果を含む光学特性を正確に捉えます.
- 結合しない原子ペアのSKパラメータを省略すると,スペクトル品質にはほとんど影響はなかった.
- TD-DFTB-MMの性能は,研究されたシステムのDFTに匹敵しました.
結論:
- TD-DFTB-MMは,大規模の電子構造と光学特性計算のための計算効率的で正確な方法です.
- この方法は,複雑な分子・ナノ粒子系を研究するための有望な方法を示しています.
- DFTB-MMでのパラメータ化の簡素化は,スペクトルの精度を大幅に損なわない.


関連する概念動画
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
3.1K
A covalently bonded heteronuclear diatomic molecule can be modeled as two vibrating masses connected by a spring. The vibrational frequency of the bond can be expressed using an equation derived from Hooke's law, which describes how the force applied to stretch or compress a spring is proportional to the displacement of the spring. In this case, the atoms behave like masses, and the bond acts like a spring.
According to Hooke's law, the vibrational frequency is directly proportional to...
According to Hooke's law, the vibrational frequency is directly proportional to...
3.1K
 03:51
Molecular Orbital Theory II
27.7K
Molecular Orbital Energy Diagrams
27.7K
 02:40
MO Theory and Covalent Bonding
14.2K
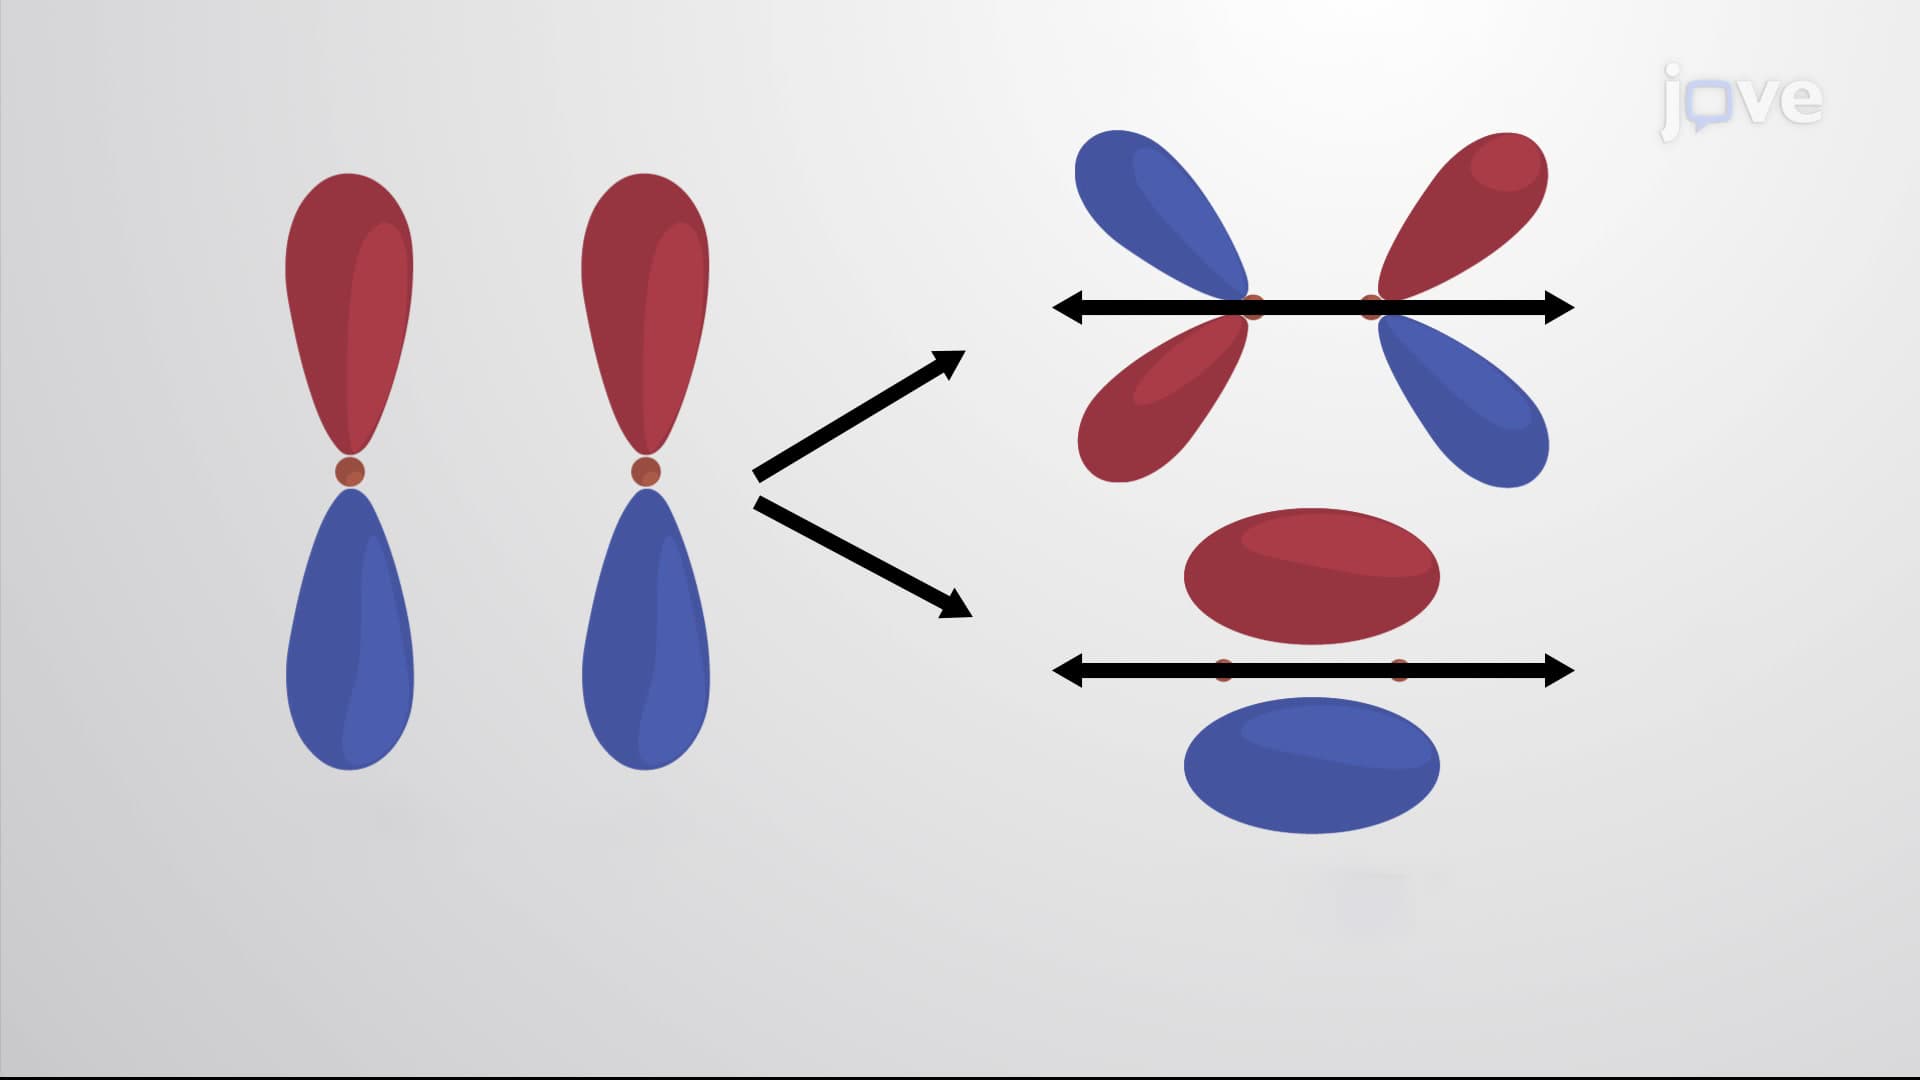
The molecular orbital theory describes the distribution of electrons in molecules in a manner similar to the distribution of electrons in atomic orbitals. The region of space in which a valence electron in a molecule is likely to be found is called a molecular orbital. Mathematically, the linear combination of atomic orbitals (LCAO) generates molecular orbitals. Combinations of in-phase atomic orbital wave functions result in regions with a high probability of electron density, while...
14.2K
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
28.7K
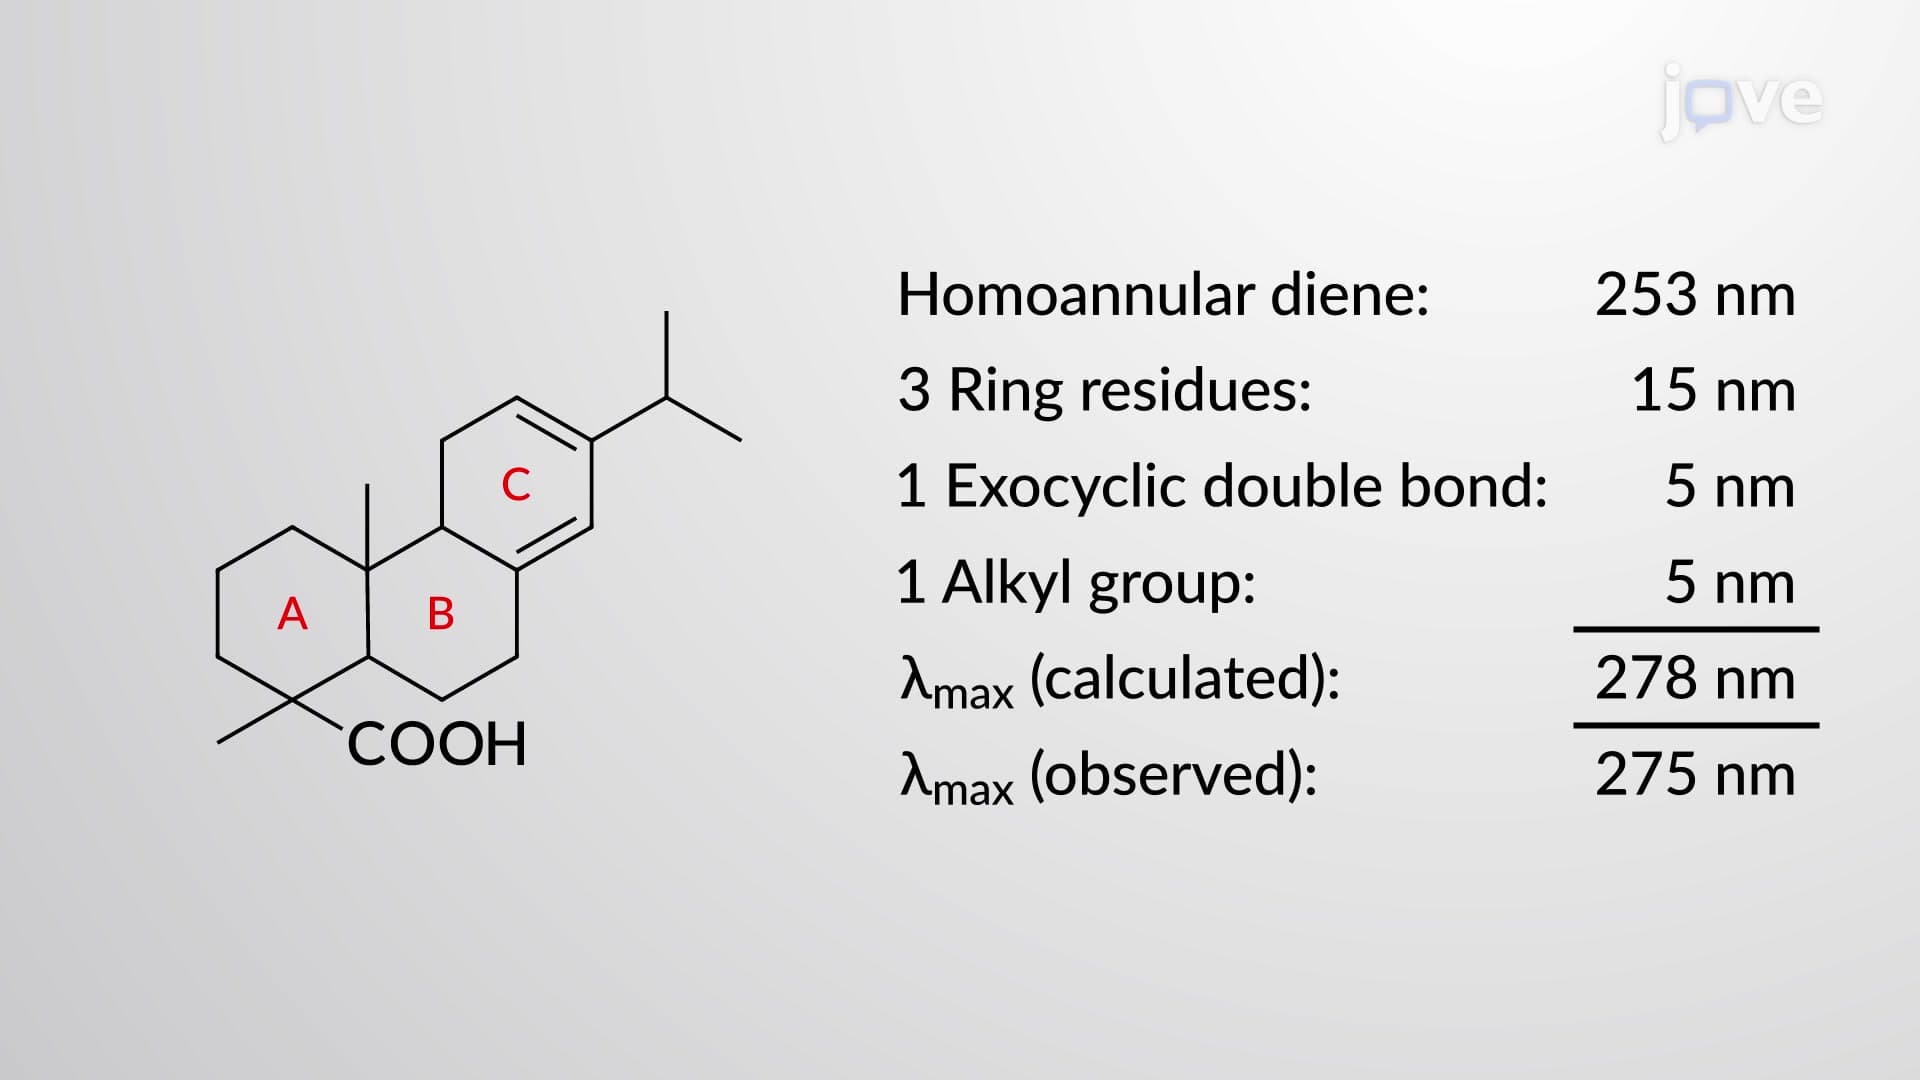
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the...
28.7K
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
39.4K
Thus far, the ideal gas law, PV = nRT, has been applied to a variety of different types of problems, ranging from reaction stoichiometry and empirical and molecular formula problems to determining the density and molar mass of a gas. However, the behavior of a gas is often non-ideal, meaning that the observed relationships between its pressure, volume, and temperature are not accurately described by the gas laws.
39.4K
 02:36
Molecular Geometry and Dipole Moments
19.3K
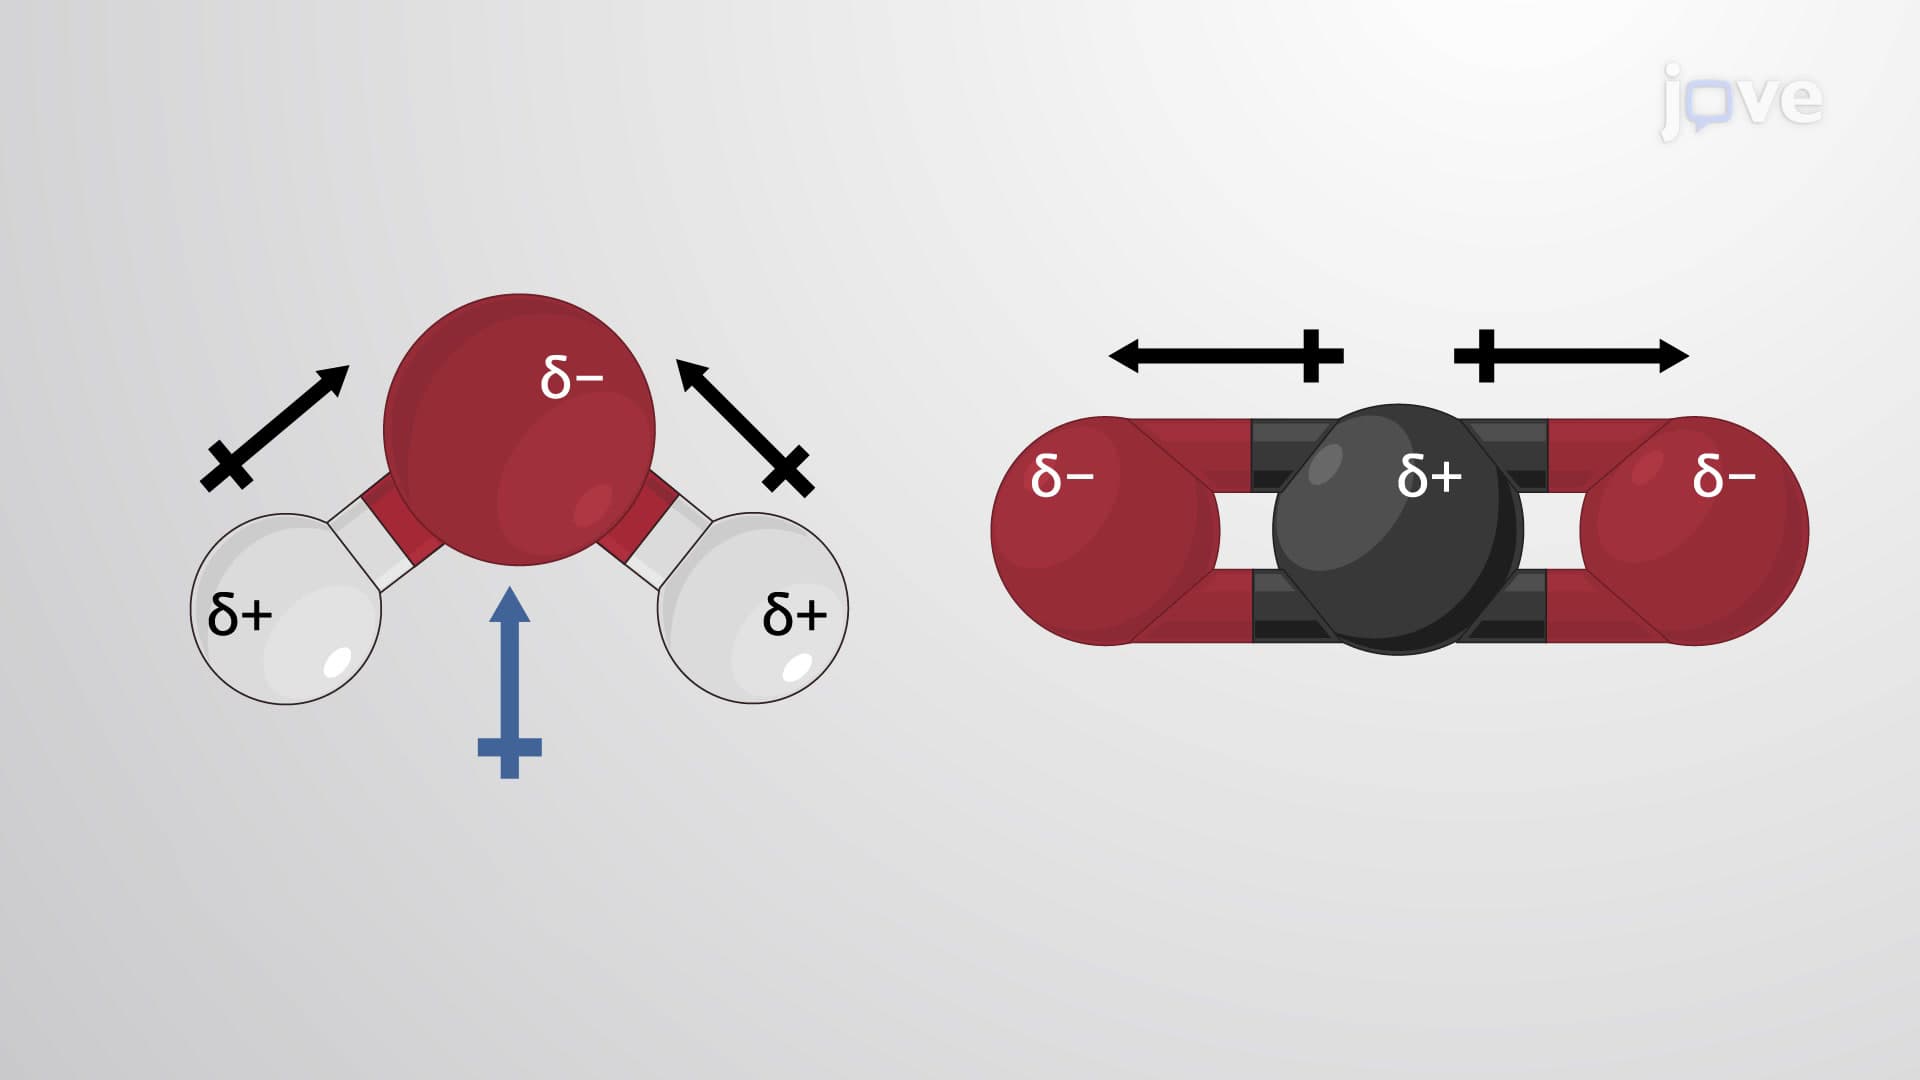
The VSEPR theory can be used to determine the electron pair geometries and molecular structures as follows:
19.3K